Related Experiment Video
Updated: Feb 12, 2026

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
The relationship between complement C3 expression and the MUC5B genotype in pulmonary fibrosis
Tsukasa Okamoto1, Susan K Mathai1, Corinne E Hennessy1
1Department of Medicine, University of Colorado , Aurora, Colorado.

Abstract:
The common gain-of-function MUC5B promoter variant ( rs35705950 ) is the strongest risk factor for the development of idiopathic pulmonary fibrosis (IPF). While the role of complement in IPF is controversial, both MUC5B and the complement system play a role in lung host defense. The aim of this study was to evaluate the relationship between complement component 3 (C3) and MUC5B in patients with IPF and in bleomycin-induced lung injury in mice. To do this, we evaluated C3 gene expression in whole lung tissue from 300 subjects with IPF and 175 healthy controls. Expression of C3 was higher in IPF than healthy controls {1.40-fold increase [95% confidence interval (CI) 1.31-1.50]; P < 0.0001} and even greater among IPF subjects with the highest-risk IPF MUC5B promoter genotype [TT vs. GG = 1.59-fold (95% CI 1.15-2.20); P < 0.05; TT vs. GT = 1.66-fold (95% CI 1.20-2.30); P < 0.05]. Among subjects with IPF, C3 expression was significantly higher in the lung tissue without microscopic honeycombing than in the lung tissue with microscopic honeycombing [1.40-fold increase (95% CI 1.23- 1.59); P < 0.01]. In mice, while bleomycin exposure increased Muc5b protein expression, C3-deficient mice were protected from bleomycin-induced lung injury. In aggregate, our findings indicate that the MUC5B promoter variant is associated with higher C3 expression and suggest that the complement system may contribute to the pathogenesis of IPF.
Insights
The MUC5B promoter variant, a key risk factor for idiopathic pulmonary fibrosis (IPF), is linked to increased complement component 3 (C3) expression. C3-deficient mice showed protection against lung injury, suggesting complement
Area of Science:
- Pulmonology and Immunology
- Genetic Epidemiology
- Host Defense Mechanisms
Background:
- The MUC5B promoter variant (rs35705950) is the leading genetic risk factor for idiopathic pulmonary fibrosis (IPF).
- The roles of MUC5B and the complement system in lung host defense are established, but their interplay in IPF remains unclear.
- Previous research suggests a controversial role for the complement system in IPF pathogenesis.
Purpose of the Study:
- To investigate the association between complement component 3 (C3) and MUC5B in patients with IPF.
- To evaluate the relationship between C3 and MUC5B in a mouse model of bleomycin-induced lung injury.
- To explore the potential contribution of the complement system to IPF development.
Main Methods:
- Quantified C3 gene expression in whole lung tissue from 300 IPF patients and 175 healthy controls.
- Analyzed C3 expression in relation to IPF MUC5B promoter genotypes (TT, GT, GG) and microscopic honeycombing.
- Assessed lung injury and Muc5b expression in C3-deficient and wild-type mice following bleomycin exposure.
Main Results:
- C3 expression was significantly elevated in IPF lungs compared to controls (1.40-fold increase, P < 0.0001).
- IPF patients with the highest-risk MUC5B genotype (TT) exhibited greater C3 expression compared to other genotypes (1.59-1.66-fold increase, P < 0.05).
- C3 expression was higher in IPF lungs without honeycombing than in those with honeycombing (1.40-fold increase, P < 0.01).
- Bleomycin increased Muc5b in mice, but C3-deficient mice displayed reduced bleomycin-induced lung injury.
Conclusions:
- The common MUC5B promoter variant is associated with increased C3 expression in IPF.
- The complement system, specifically C3, may play a role in the pathogenesis of IPF.
- Targeting the complement system could be a potential therapeutic strategy for IPF.
Related Concept Videos
 01:27
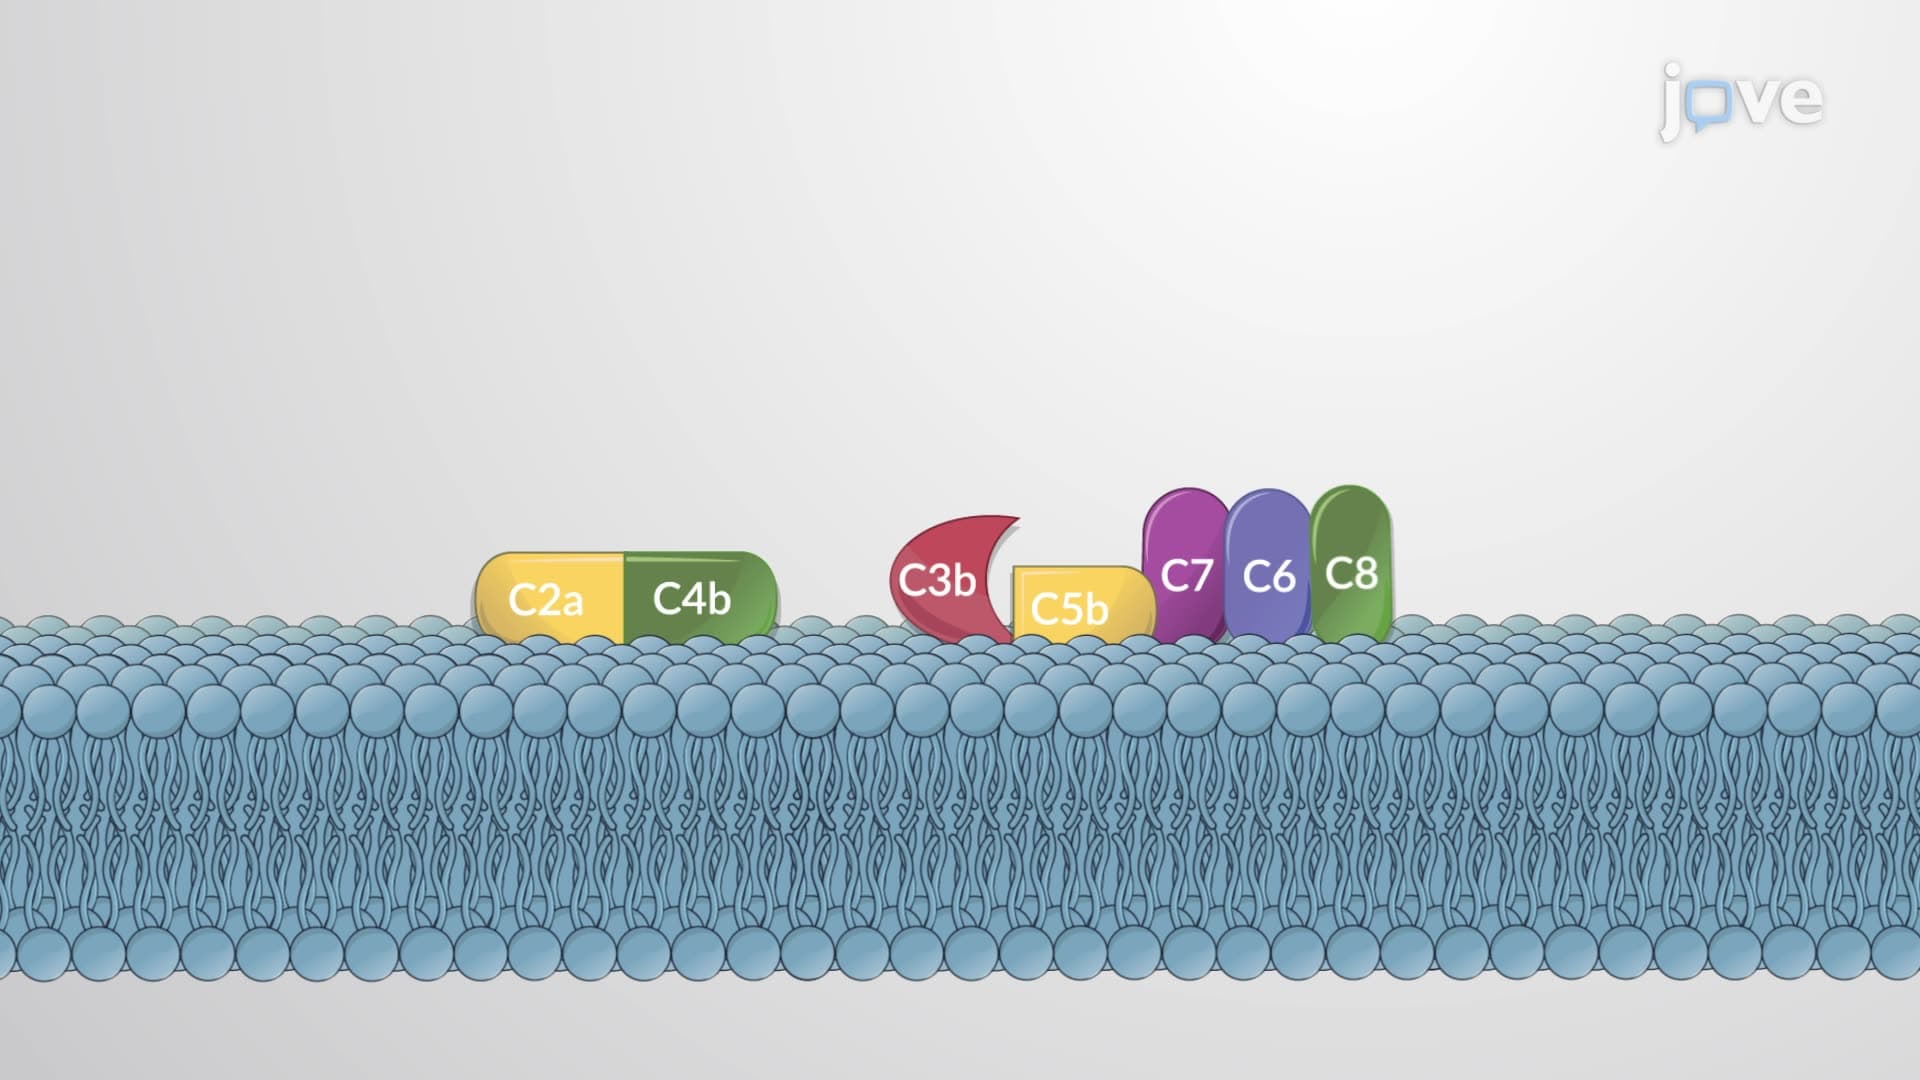
Complement System
 02:12
Relationship Formation
 00:49
Complementation Tests
Organisms heterozygous for different mutations are crossed pairwise in all combinations. If present on different genes, the mutations can complement each other by providing the missing...
 01:28
Ending Relationships
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:14
Pharmacokinetic–Pharmacodynamic Relationship: Duration of Dose-Effect Relationship