Related Experiment Video
Updated: Feb 12, 2026

Accessing Valuable Ligand Supports for Transition Metals: A Modified, Intermediate Scale Preparation of 1,2,3,4,5-Pentamethylcyclopentadiene
Published on: March 20, 2017
Calculation of Ligand Dissociation Energies in Large Transition-Metal Complexes
Tamara Husch1, Leon Freitag1, Markus Reiher1
1Laboratorium für Physikalische Chemie , ETH Zürich , Vladimir-Prelog-Weg 2 , 8093 Zürich , Switzerland.

Abstract:
The accurate calculation of ligand dissociation (or equivalently, ligand binding) energies is crucial for computational coordination chemistry. Despite its importance, obtaining accurate ab initio reference data is difficult, and density-functional methods of uncertain reliability are chosen for feasibility reasons. Here, we consider advanced coupled-cluster and multiconfigurational approaches to reinvestigate our WCCR10 set of 10 gas-phase ligand dissociation energies [ J. Chem. Theory Comput. 2014, 10, 3092]. We assess the potential multiconfigurational character of all molecules involved in these reactions with a multireference diagnostic [ Mol. Phys. 2017, 115, 2110] in order to determine where single-reference coupled-cluster approaches can be applied. For some reactions of the WCCR10 set, large deviations of density-functional results including semiclassical dispersion corrections from experimental reference data had been observed. This puzzling observation deserves special attention here, and we tackle the issue (i) by comparing to ab initio data that comprise dispersion effects on a rigorous first-principles footing and (ii) by a comparison of density-functional approaches that model dispersion interactions in various ways. For two reactions, species exhibiting nonnegligible static electron correlation were identified. These two reactions represent hard problems for electronic structure methods and also for multireference perturbation theories. However, most of the ligand dissociation reactions in WCCR10 do not exhibit static electron correlation effects, and hence, we may choose standard single-reference coupled-cluster approaches to compare with density-functional methods. For WCCR10, the Minnesota M06-L functional yielded the smallest mean absolute deviation of 13.2 kJ mol-1 out of all density functionals considered (PBE, BP86, BLYP, TPSS, M06-L, PBE0, B3LYP, TPSSh, and M06-2X) without additional dispersion corrections in comparison to the coupled-cluster results, and the PBE0-D3 functional produced the overall smallest mean absolute deviation of 4.3 kJ mol-1. The agreement of density-functional results with coupled-cluster data increases significantly upon inclusion of any type of dispersion correction. It is important to emphasize that different density-functional schemes available for this purpose perform equally well. The coupled-cluster dissociation energies, however, deviate from experimental results on average by 30.3 kJ mol-1. Possible reasons for these deviations are discussed.
Related Concept Videos
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 02:58
Properties of Transition Metals
 02:49
Calculating Standard Free Energy Changes
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
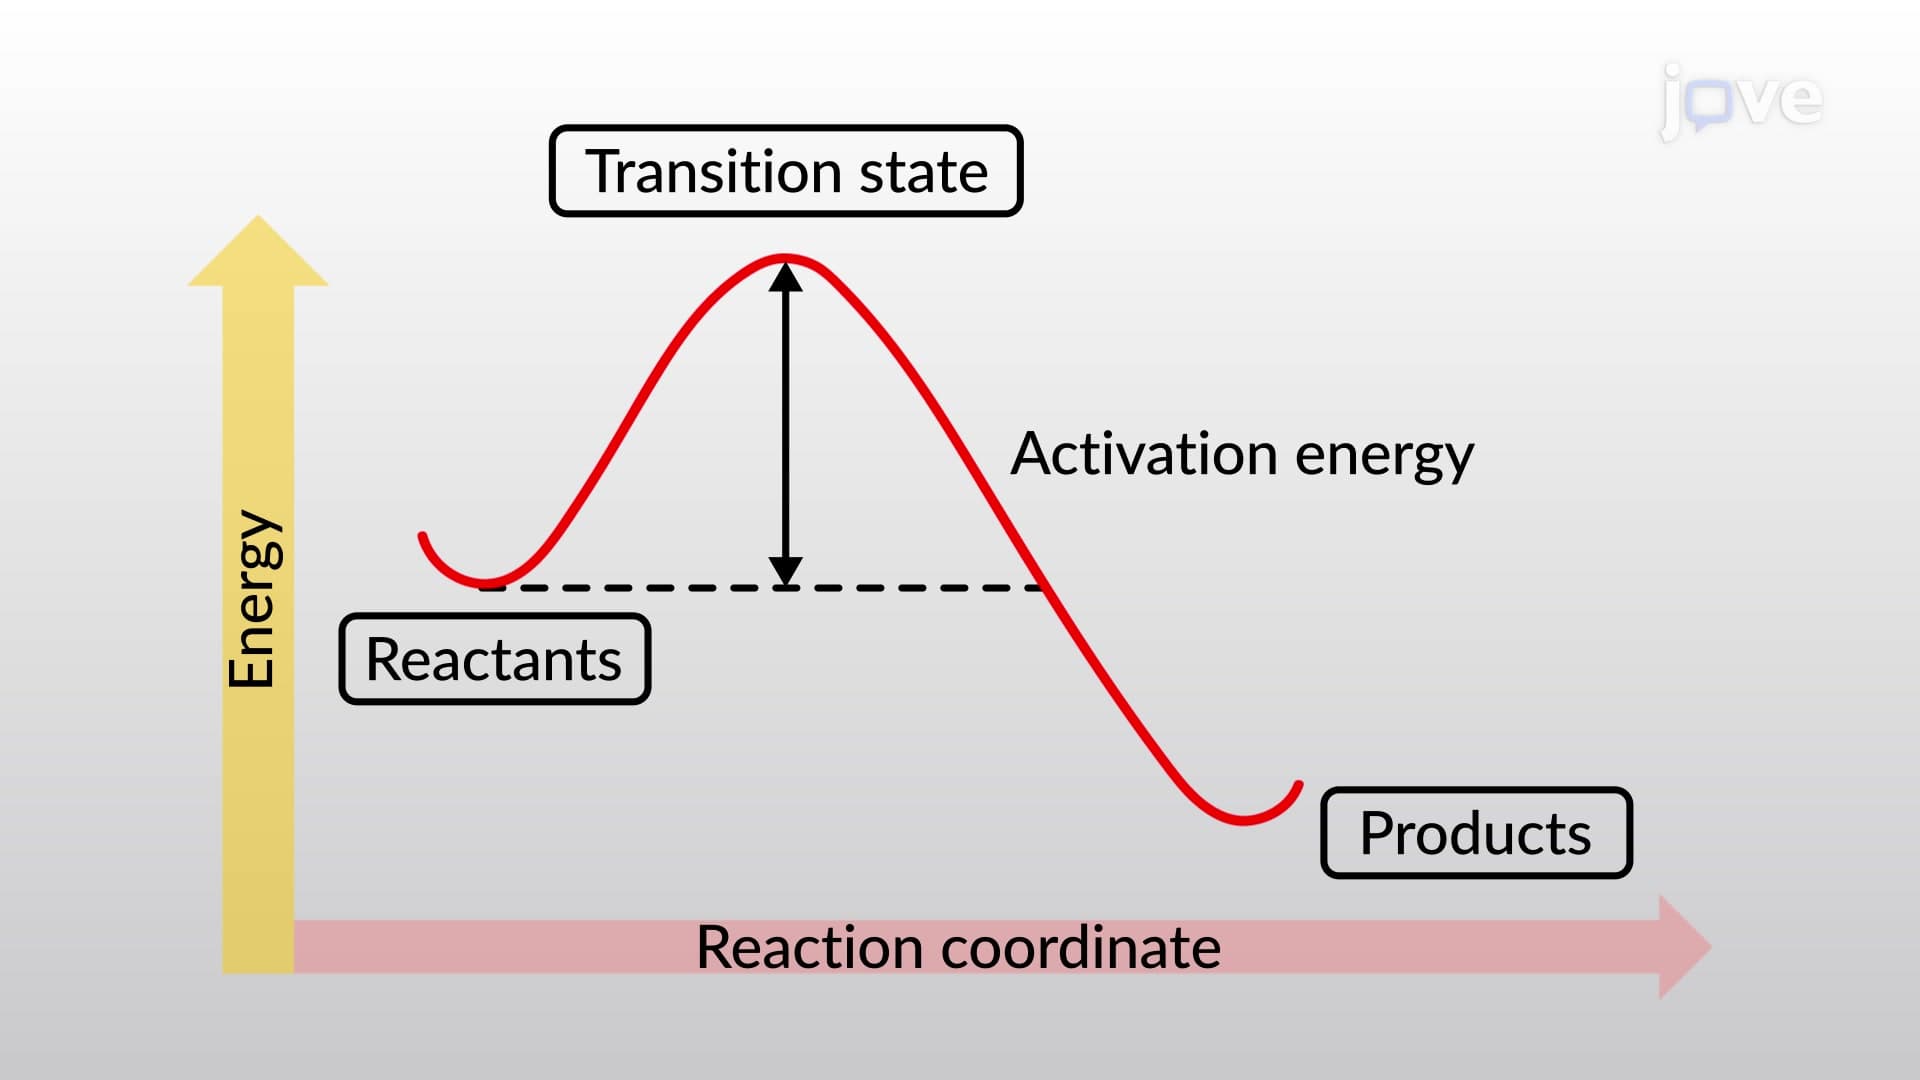 02:13
Bond Dissociation Energy and Activation Energy
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...