Related Experiment Video
Updated: Aug 30, 2026
The Use of Primary Human Fibroblasts for Monitoring Mitochondrial Phenotypes in the Field of Parkinson's Disease
Published on: October 3, 2012
Genotype-Phenotype Relations for the Parkinson's Disease Genes Parkin, PINK1, DJ1: MDSGene Systematic Review
Meike Kasten1,2, Corinna Hartmann1, Jennie Hampf1
1Institute of Neurogenetics, University of Lübeck, Lübeck, Germany.

Abstract:
This first comprehensive MDSGene review is devoted to the 3 autosomal recessive Parkinson's disease forms: PARK-Parkin, PARK-PINK1, and PARK-DJ1. It followed MDSGene's standardized data extraction protocol and screened a total of 3652 citations and is based on fully curated phenotypic and genotypic data on >1100 patients with recessively inherited PD because of 221 different disease-causing mutations in Parkin, PINK1, or DJ1. All these data are also available in an easily searchable online database (www.mdsgene.org), which also provides descriptive summary statistics on phenotypic and genetic data. Despite the high degree of missingness of phenotypic features and unsystematic reporting of genotype data in the original literature, the present review recapitulates many of the previously described findings including early onset (median age at onset of ∼30 years for carriers of at least 2 mutations in any of the 3 genes) of an overall clinically typical form of PD with excellent treatment response, dystonia and dyskinesia being relatively common and cognitive decline relatively uncommon. However, when comparing actual data with common expert knowledge in previously published reviews, we detected several discrepancies. We conclude that systematic reporting of phenotypes is a pressing need in light of increasingly available molecular genetic testing and the emergence of first gene-specific therapies entering clinical trials. © 2018 International Parkinson and Movement Disorder Society.
Insights
This review details 3 autosomal recessive Parkinson's disease (PD) forms caused by mutations in Parkin, PINK1, and DJ1. Findings reveal typical PD symptoms with early onset, but highlight discrepancies with existing knowledge.
Area of Science:
- Genetics
- Neurology
- Molecular Biology
Background:
- Parkinson's disease (PD) has genetic forms, including autosomal recessive types.
- Mutations in Parkin, PINK1, and DJ1 genes are known causes of recessive PD.
- Comprehensive data on these genetic forms are crucial for understanding disease mechanisms.
Purpose of the Study:
- To conduct a comprehensive review of autosomal recessive Parkinson's disease linked to Parkin, PINK1, and DJ1.
- To curate and analyze phenotypic and genotypic data from over 1100 patients.
- To compare findings with existing literature and identify discrepancies.
Main Methods:
- Systematic data extraction from 3652 citations following the MDSGene protocol.
- Analysis of phenotypic and genotypic data from >1100 patients with mutations in Parkin, PINK1, or DJ1.
- Creation of a searchable online database (www.mdsgene.org) for patient data.
Main Results:
- Identified 221 distinct disease-causing mutations across the three genes.
- Observed a median age at onset of ~30 years for carriers of at least two mutations.
- Found clinically typical PD with good treatment response, common dystonia/dyskinesia, and uncommon cognitive decline.
Conclusions:
- Recapitulated known findings but identified discrepancies with common expert knowledge.
- Emphasized the need for systematic phenotype reporting due to increasing genetic testing.
- Highlighted the importance of data for emerging gene-specific therapies in clinical trials.
Related Concept Videos
 01:43
Incomplete Dominance
 01:15
Parkinson's Disease: Overview
 01:24
Parkinson's Disease: Treatment
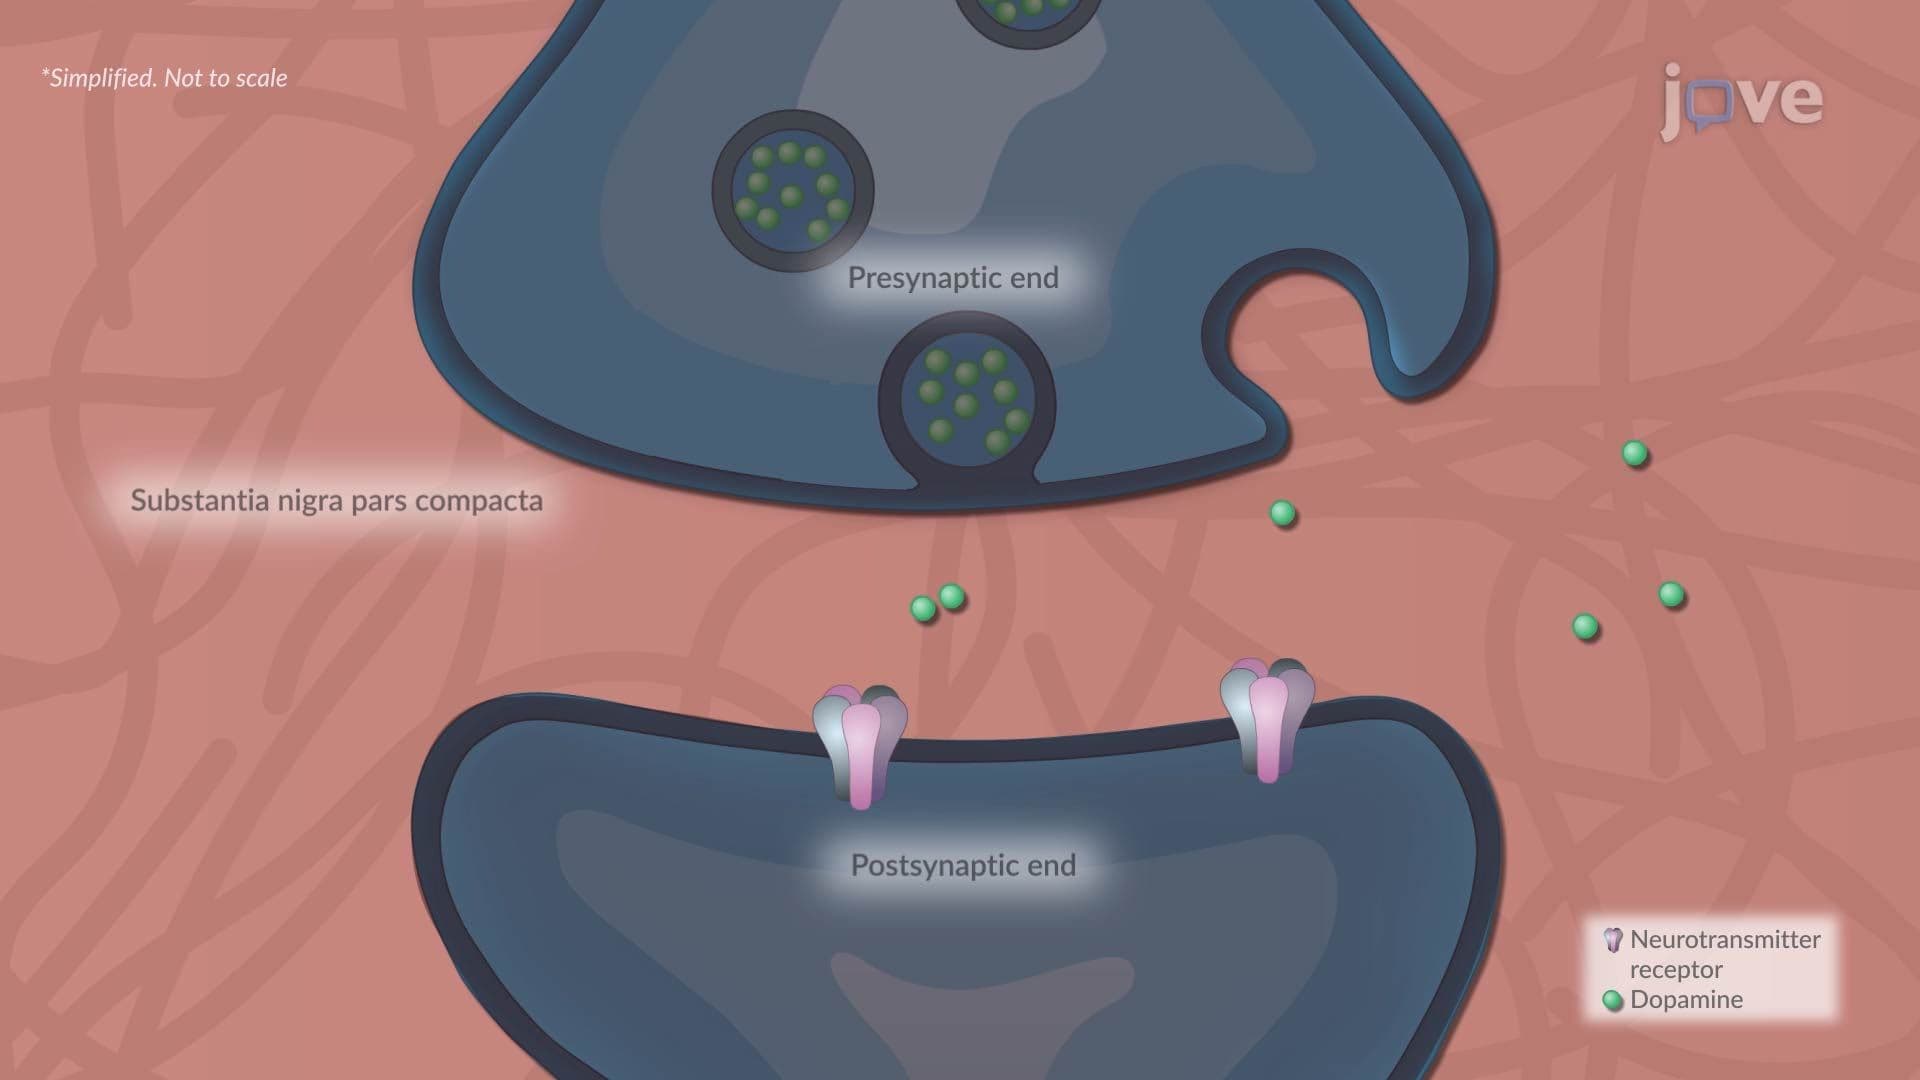
Parkinson's Disease is primarily a result of the loss of dopaminergic neurons in the substantia nigra pars compacta. The cornerstone of its...
 01:28
Human Genetics
The complex relationship between genetics and psychology is observable through common biological components such...
 01:24
Parkinson Disease l: Introduction
 01:24
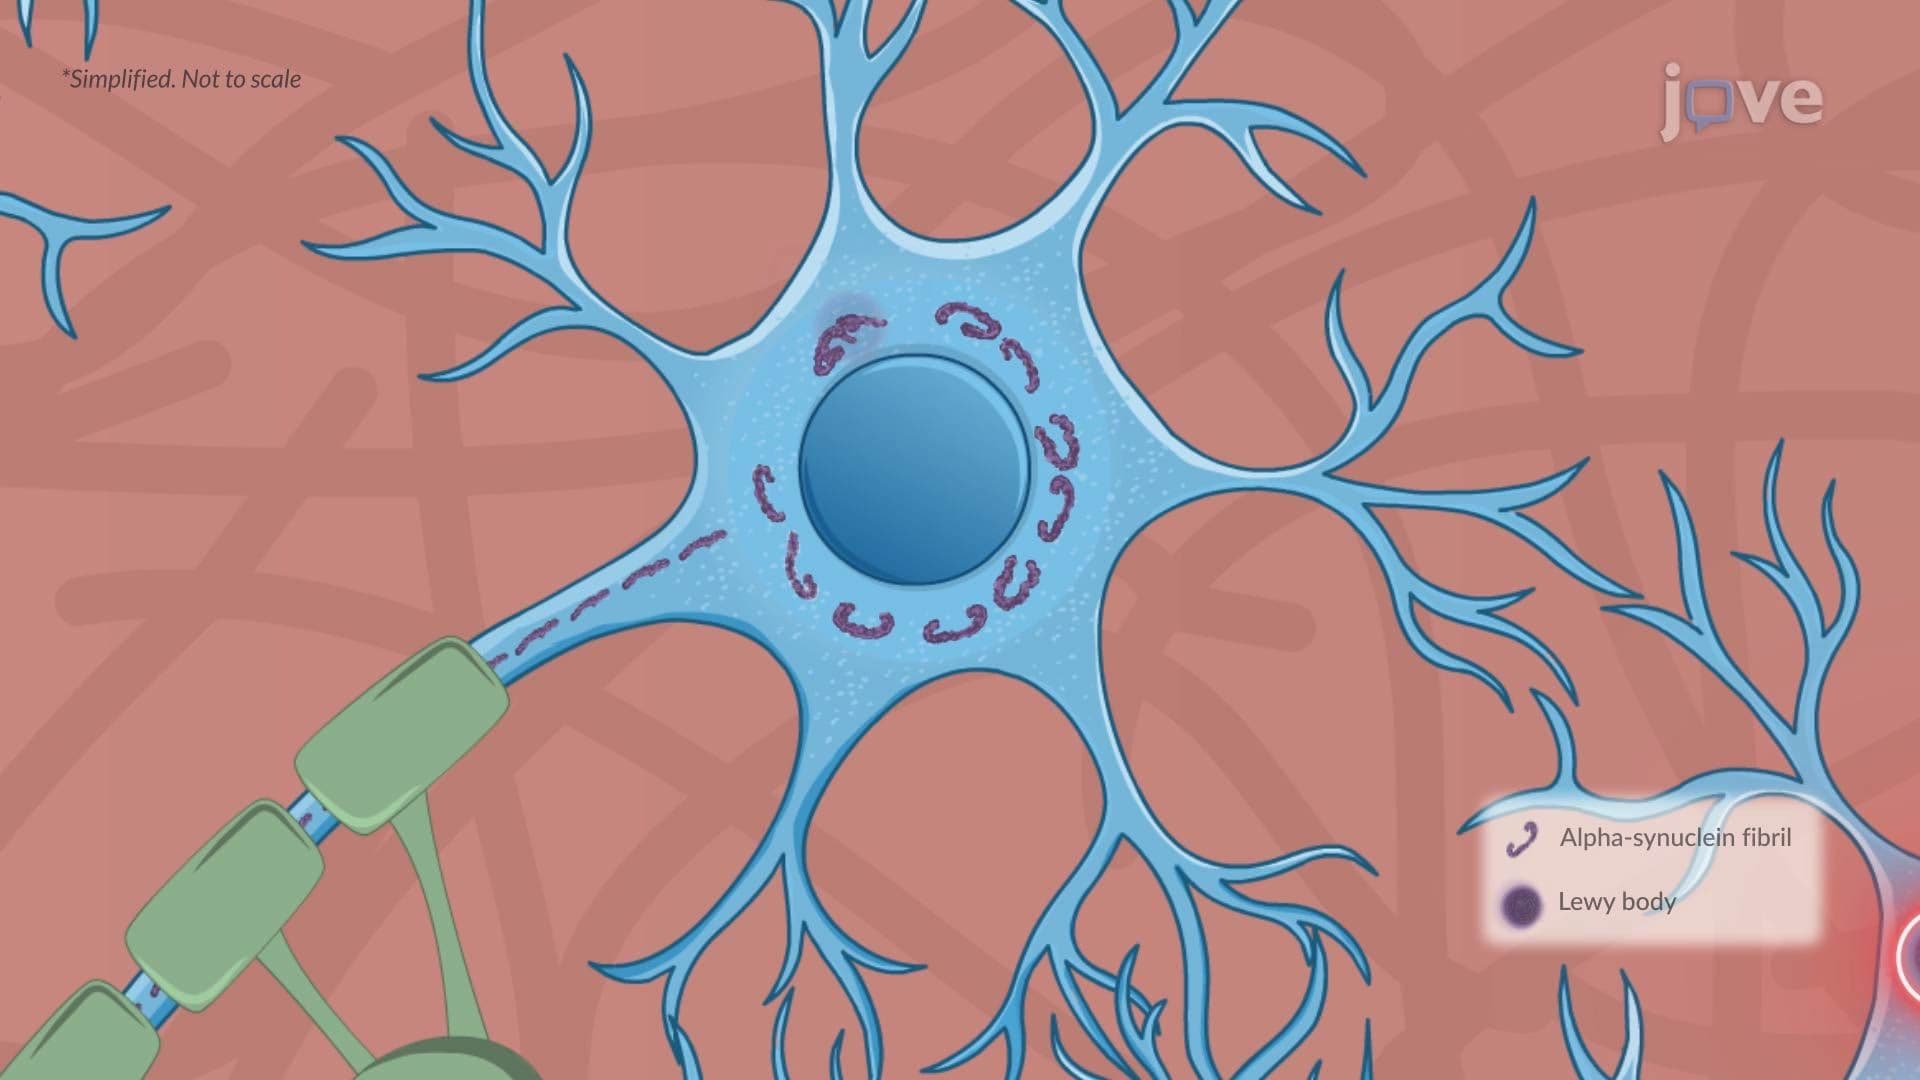
Parkinson Disease ll: Pathophysiology