Physiologically based pharmacokinetic modeling of nanoceria systemic distribution in rats suggests dose- and
Ulrika Carlander1, Tshepo Paulsen Moto2, Anteneh Assefa Desalegn1
1Unit of Work Environment Toxicology, Institute of Environmental Medicine, Karolinska Institutet, Solna, Sweden.

Background:
Cerium dioxide nanoparticles (nanoceria) are increasingly being used in a variety of products as catalysts, coatings, and polishing agents. Furthermore, their antioxidant properties make nanoceria potential candidates for biomedical applications. To predict and avoid toxicity, information about their biokinetics is essential. A useful tool to explore such associations between exposure and internal target dose is physiologically based pharmacokinetic (PBPK) modeling. The aim of this study was to test the appropriateness of our previously published PBPK model developed for intravenous (IV) administration when applied to various sizes of nanoceria and to exposure routes relevant for humans.
Methods:
Experimental biokinetic data on nanoceria (obtained from various exposure routes, sizes, coatings, doses, and tissues sampled) in rats were collected from the literature and also obtained from the researchers. The PBPK model was first calibrated and validated against IV data for 30 nm citrate coated ceria and then recalibrated for 5 nm ceria. Finally, the model was modified and tested against inhalation, intratracheal (IT) instillation, and oral nanoceria data.
Results:
The PBPK model adequately described nanoceria time courses in various tissues for 5 nm ceria given IV. The time courses of 30 nm ceria were reasonably well predicted for liver and spleen, whereas the biokinetics in other tissues were not well captured. For the inhalation, IT instillation, and oral exposure routes, re-optimization was difficult due to low absorption and, hence, low and variable nanoceria tissue levels. Moreover, the nanoceria properties and exposure conditions varied widely among the inhalation, IT instillation, and oral studies, making it difficult to assess the importance of different factors.
Conclusion:
Overall, our modeling efforts suggest that nanoceria biokinetics depend largely on the exposure route and dose.
Related Concept Videos
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:15
Model Approaches for Pharmacokinetic Data: Physiological Models
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 01:06
Model Approaches for Pharmacokinetic Data: Distributed Parameter Models
The distributed parameter models are specifically designed to account for variations and differences in some drug classes. This model is particularly useful for assessing regional concentrations of anticancer or...
 01:20
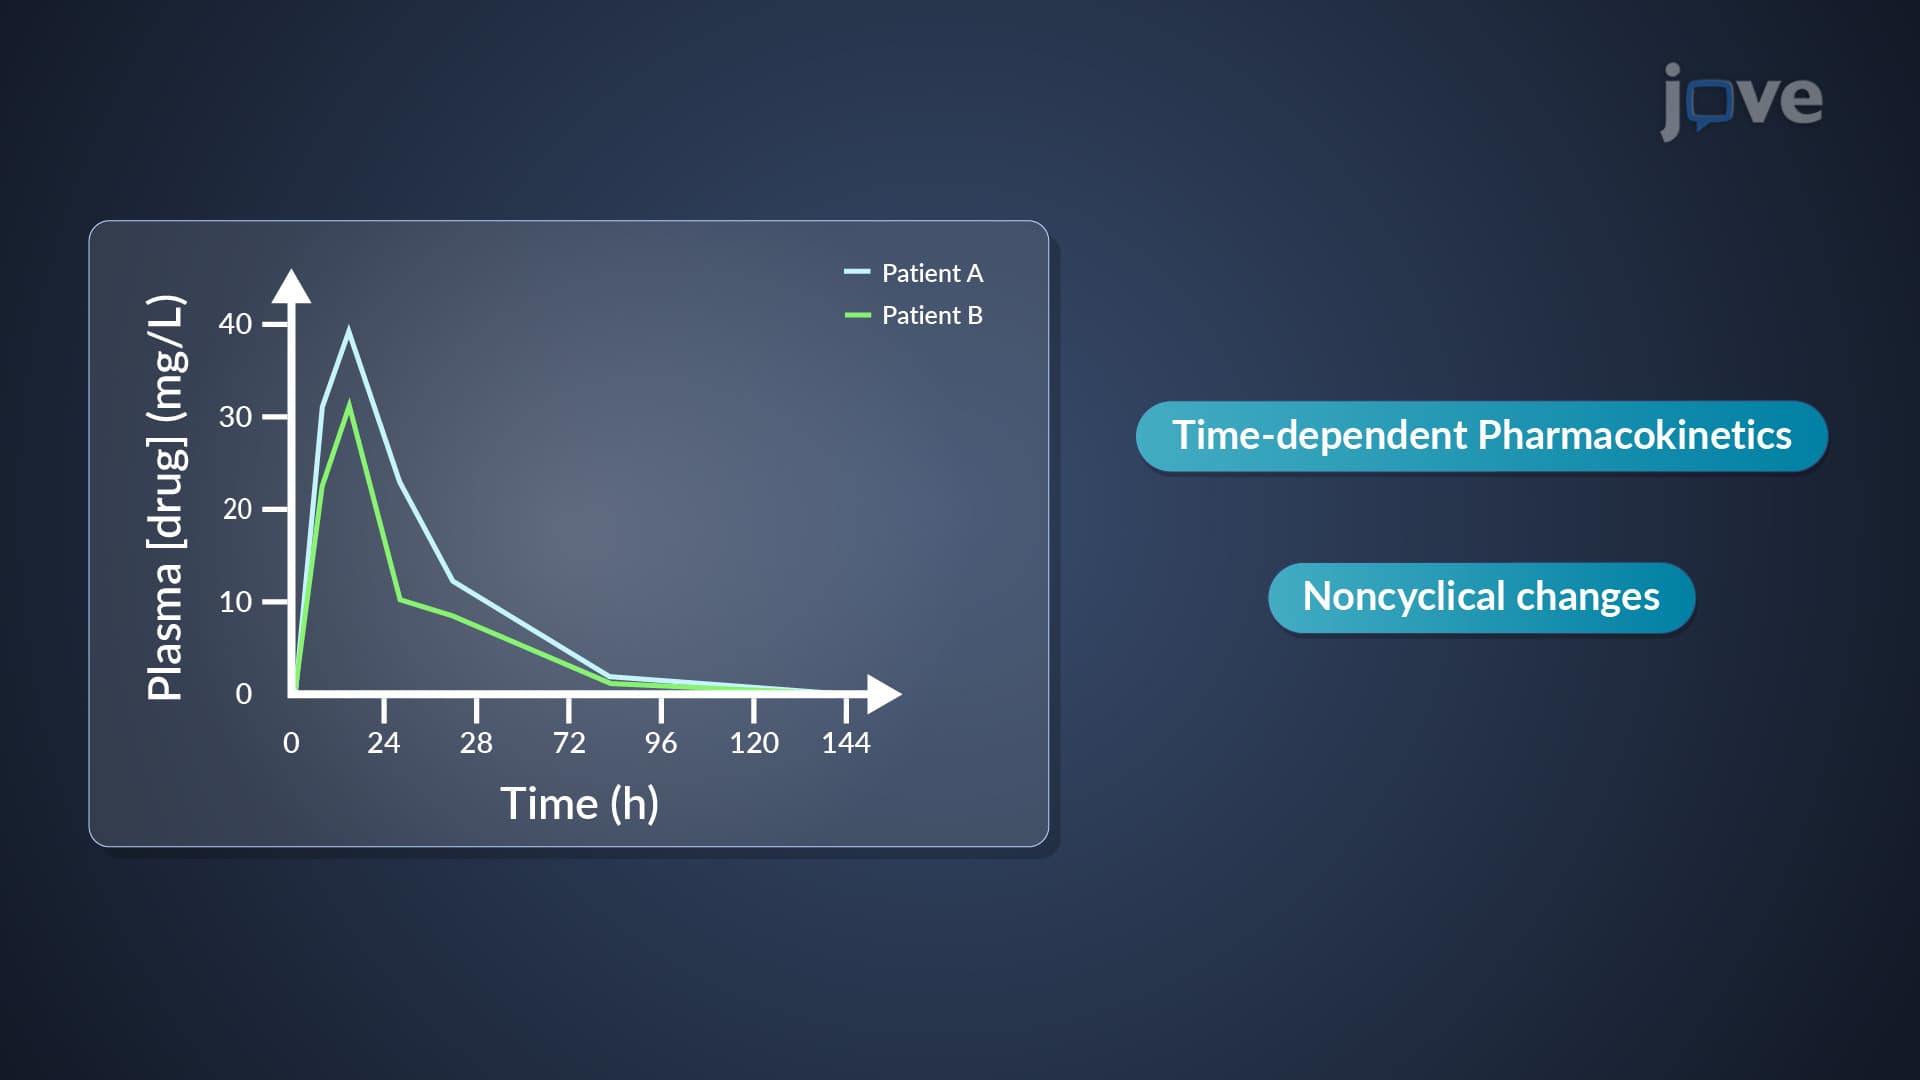
Chronopharmacokinetics: Time-Dependent Pharmacokinetics
Time-dependent pharmacokinetics refers to non-cyclical changes in drug rate processes over a period of time. It can lead to nonlinear pharmacokinetics, where the relationship between drug concentration and time is not proportional. Non-cyclical...
 00:57
Physiological Pharmacokinetic Models: Blood Flow-Limited Versus Diffusion-Limited Models