Related Experiment Video
Updated: Feb 10, 2026

Characterization of In Vitro Differentiation of Human Primary Keratinocytes by RNA-Seq Analysis
Published on: May 16, 2020
A comparative study of RNA-Seq and microarray data analysis on the two examples of rectal-cancer patients and Burkitt
Alexander Wolff1, Michaela Bayerlová1, Jochen Gaedcke2
1Dept. of Medical Statistics, University Medical Center Göttingen, Göttingen, Germany.

Background:
Pipeline comparisons for gene expression data are highly valuable for applied real data analyses, as they enable the selection of suitable analysis strategies for the dataset at hand. Such pipelines for RNA-Seq data should include mapping of reads, counting and differential gene expression analysis or preprocessing, normalization and differential gene expression in case of microarray analysis, in order to give a global insight into pipeline performances.
Methods:
Four commonly used RNA-Seq pipelines (STAR/HTSeq-Count/edgeR, STAR/RSEM/edgeR, Sailfish/edgeR, TopHat2/Cufflinks/CuffDiff)) were investigated on multiple levels (alignment and counting) and cross-compared with the microarray counterpart on the level of gene expression and gene ontology enrichment. For these comparisons we generated two matched microarray and RNA-Seq datasets: Burkitt Lymphoma cell line data and rectal cancer patient data.
Results:
The overall mapping rate of STAR was 98.98% for the cell line dataset and 98.49% for the patient dataset. Tophat's overall mapping rate was 97.02% and 96.73%, respectively, while Sailfish had only an overall mapping rate of 84.81% and 54.44%. The correlation of gene expression in microarray and RNA-Seq data was moderately worse for the patient dataset (ρ = 0.67-0.69) than for the cell line dataset (ρ = 0.87-0.88). An exception were the correlation results of Cufflinks, which were substantially lower (ρ = 0.21-0.29 and 0.34-0.53). For both datasets we identified very low numbers of differentially expressed genes using the microarray platform. For RNA-Seq we checked the agreement of differentially expressed genes identified in the different pipelines and of GO-term enrichment results.
Conclusion:
In conclusion the combination of STAR aligner with HTSeq-Count followed by STAR aligner with RSEM and Sailfish generated differentially expressed genes best suited for the dataset at hand and in agreement with most of the other transcriptomics pipelines.
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 01:20
Comparing the Survival Analysis of Two or More Groups
 01:08
Newton's Third Law: Examples
 01:07
Free Body Diagrams: Examples
 01:12
Analysis of Population Pharmacokinetic Data
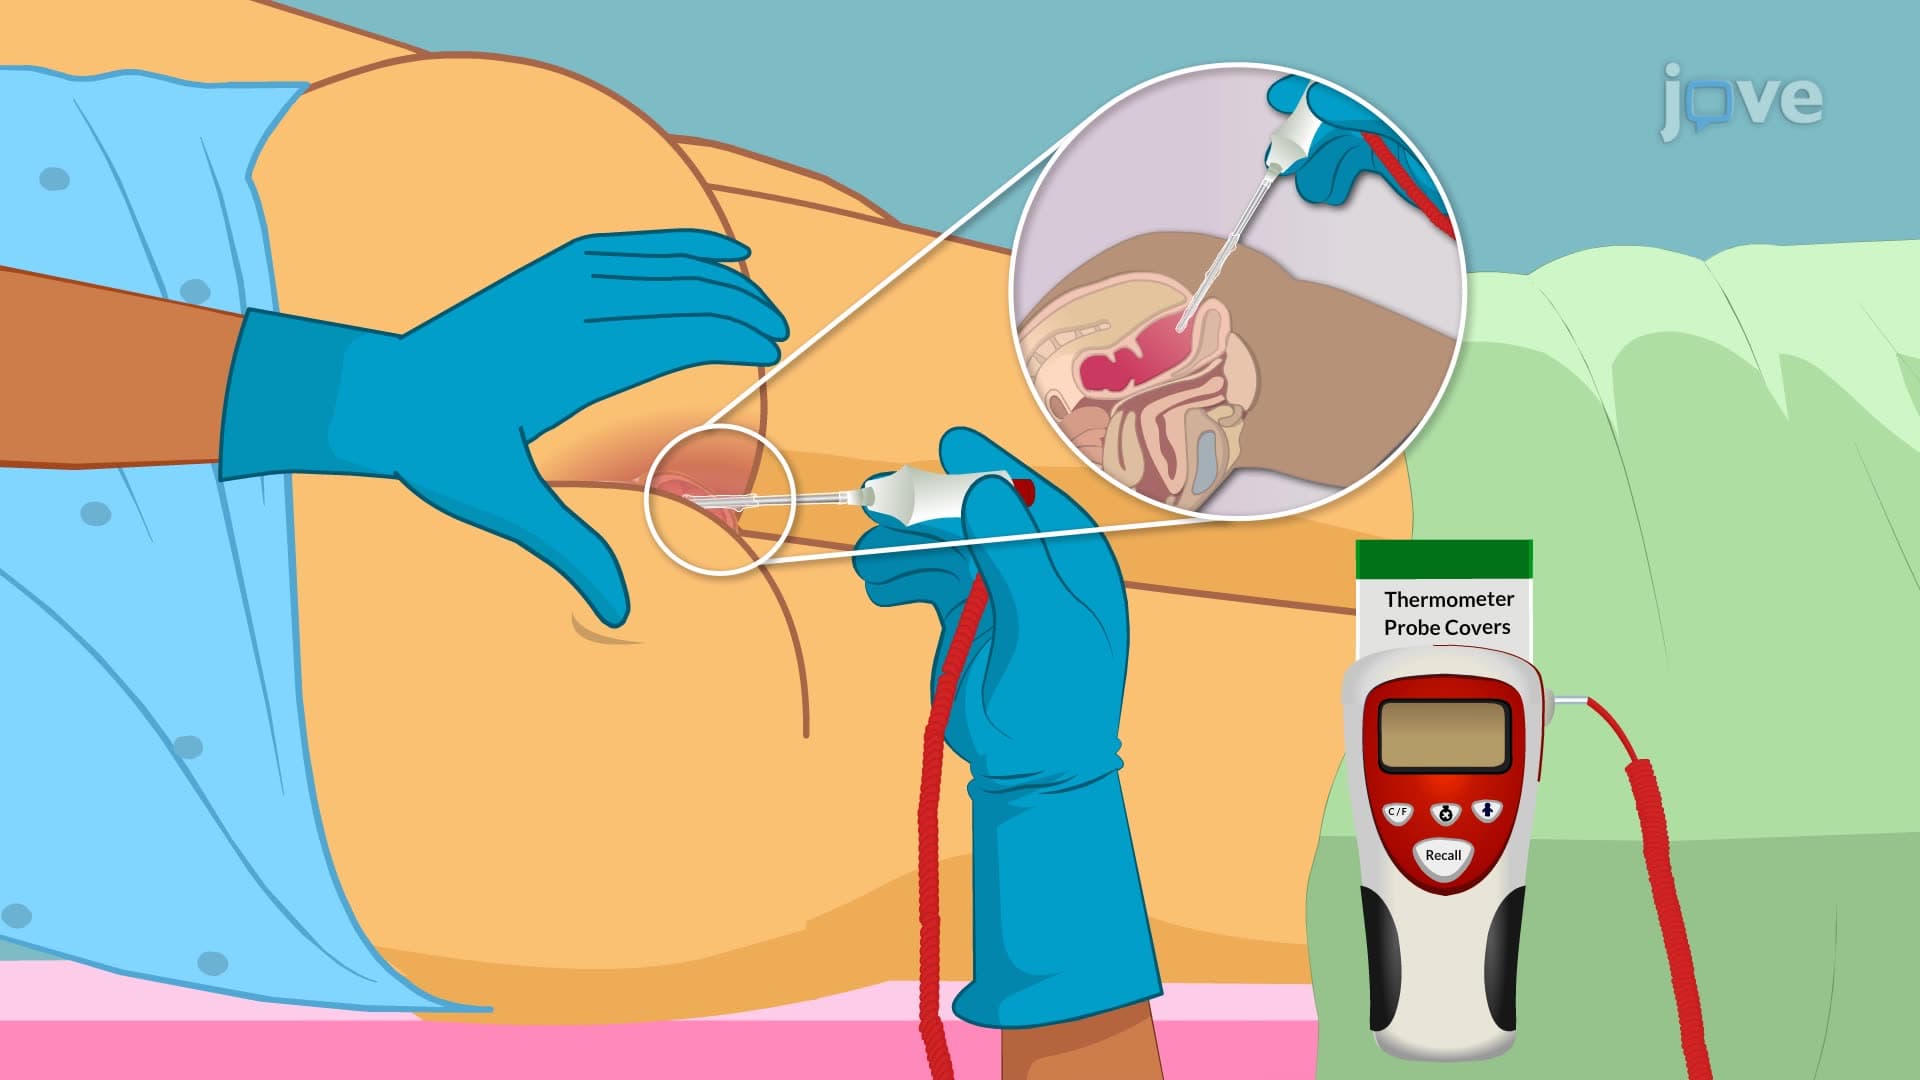 01:27
Assessing Body Temperature - Rectal
Follow these steps for rectal temperature assessment:
Step 1: Perform hand hygiene and don clean gloves to prevent cross-infection.
Step 2: Position the patient in a side-lying position to better visualize the rectal...