Related Experiment Video
Updated: Feb 8, 2026
Using Synchrotron Radiation Microtomography to Investigate Multi-scale Three-dimensional Microelectronic Packages
Published on: April 13, 2016
DMPy: a Python package for automated mathematical model construction of large-scale metabolic systems
Robert W Smith1,2, Rik P van Rosmalen1, Vitor A P Martins Dos Santos1,2
1Laboratory of Systems & Synthetic Biology, Wageningen UR, Stippeneng 4, Wageningen, 6708WE, The Netherlands.

Background:
Models of metabolism are often used in biotechnology and pharmaceutical research to identify drug targets or increase the direct production of valuable compounds. Due to the complexity of large metabolic systems, a number of conclusions have been drawn using mathematical methods with simplifying assumptions. For example, constraint-based models describe changes of internal concentrations that occur much quicker than alterations in cell physiology. Thus, metabolite concentrations and reaction fluxes are fixed to constant values. This greatly reduces the mathematical complexity, while providing a reasonably good description of the system in steady state. However, without a large number of constraints, many different flux sets can describe the optimal model and we obtain no information on how metabolite levels dynamically change. Thus, to accurately determine what is taking place within the cell, finer quality data and more detailed models need to be constructed.
Results:
In this paper we present a computational framework, DMPy, that uses a network scheme as input to automatically search for kinetic rates and produce a mathematical model that describes temporal changes of metabolite fluxes. The parameter search utilises several online databases to find measured reaction parameters. From this, we take advantage of previous modelling efforts, such as Parameter Balancing, to produce an initial mathematical model of a metabolic pathway. We analyse the effect of parameter uncertainty on model dynamics and test how recent flux-based model reduction techniques alter system properties. To our knowledge this is the first time such analysis has been performed on large models of metabolism. Our results highlight that good estimates of at least 80% of the reaction rates are required to accurately model metabolic systems. Furthermore, reducing the size of the model by grouping reactions together based on fluxes alters the resulting system dynamics.
Conclusion:
The presented pipeline automates the modelling process for large metabolic networks. From this, users can simulate their pathway of interest and obtain a better understanding of how altering conditions influences cellular dynamics. By testing the effects of different parameterisations we are also able to provide suggestions to help construct more accurate models of complete metabolic systems in the future.
Related Concept Videos
 01:29
Mathematical Modeling: Problem Solving
 00:58
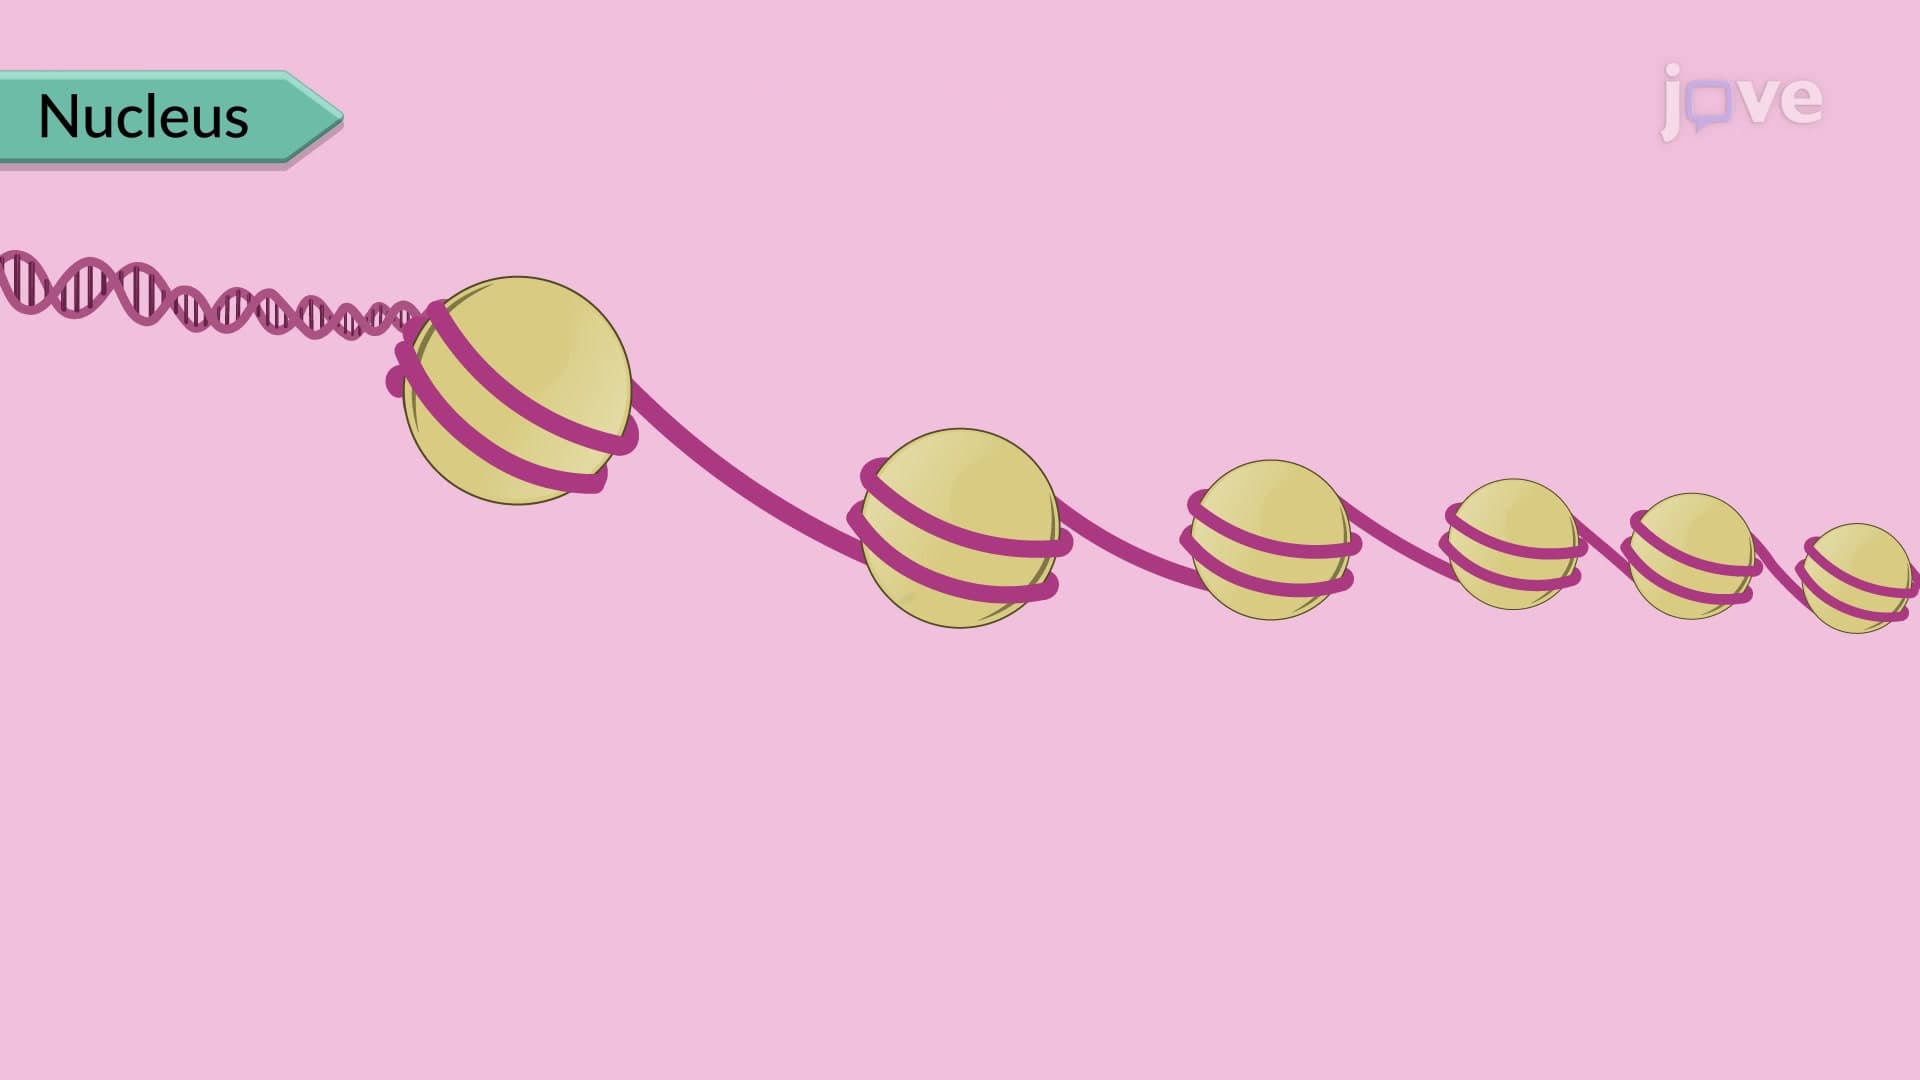
DNA Packaging
 01:32
Chromatin Packaging
 02:21
Chromatin Packaging
The chromatin
In combination with specialized DNA binding protein called Histones, the DNA double helix forms a compact DNA: protein complex called chromatin. The chromatin itself is further compacted into higher-order...
02:21Chromatin Packaging
 00:52
What is Metabolism?