Accurate genotyping across variant classes and lengths using variant graphs
Related Experiment Video
Updated: Feb 8, 2026

Detection of Rare Genomic Variants from Pooled Sequencing Using SPLINTER
Published on: June 23, 2012
Jonas Andreas Sibbesen1, Lasse Maretty1,
1The Bioinformatics Centre, Department of Biology, University of Copenhagen, Copenhagen, Denmark.

Abstract:
Genotype estimates from short-read sequencing data are typically based on the alignment of reads to a linear reference, but reads originating from more complex variants (for example, structural variants) often align poorly, resulting in biased genotype estimates. This bias can be mitigated by first collecting a set of candidate variants across discovery methods, individuals and databases, and then realigning the reads to the variants and reference simultaneously. However, this realignment problem has proved computationally difficult. Here, we present a new method (BayesTyper) that uses exact alignment of read k-mers to a graph representation of the reference and variants to efficiently perform unbiased, probabilistic genotyping across the variation spectrum. We demonstrate that BayesTyper generally provides superior variant sensitivity and genotyping accuracy relative to existing methods when used to integrate variants across discovery approaches and individuals. Finally, we demonstrate that including a 'variation-prior' database containing already known variants significantly improves sensitivity.
Related Concept Videos
 02:30
Histone Variants at the Centromere
 01:10
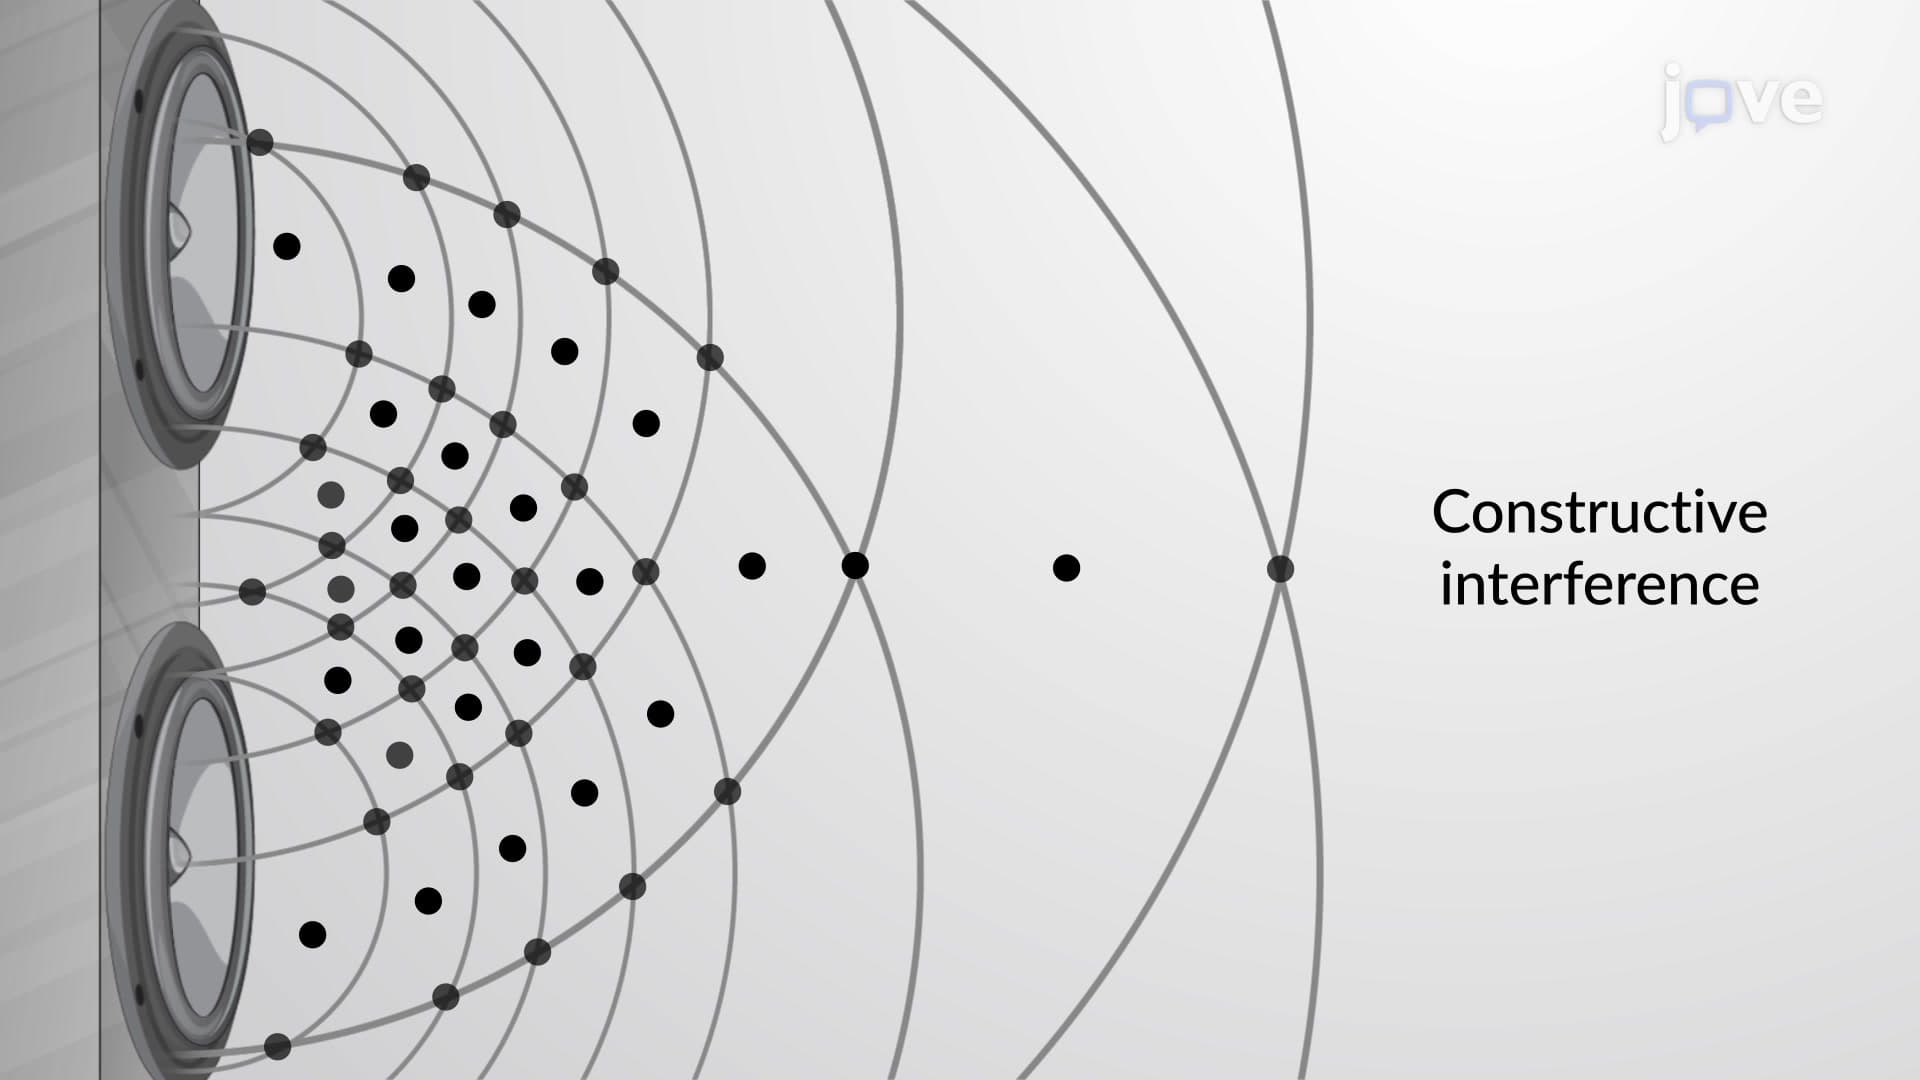
Interference: Path Lengths
Two special sources may be considered when they are in phase. This can be easily achieved by feeding the two sources from the same source. An example would be synchronizing the two speakers by feeding them with the same source, such as the sound waves produced by a tuning fork. This setup ensures that the two sources have the same frequency and are...
 02:49
Bond Energies and Bond Lengths
 01:07
Ogive Graph
 01:30
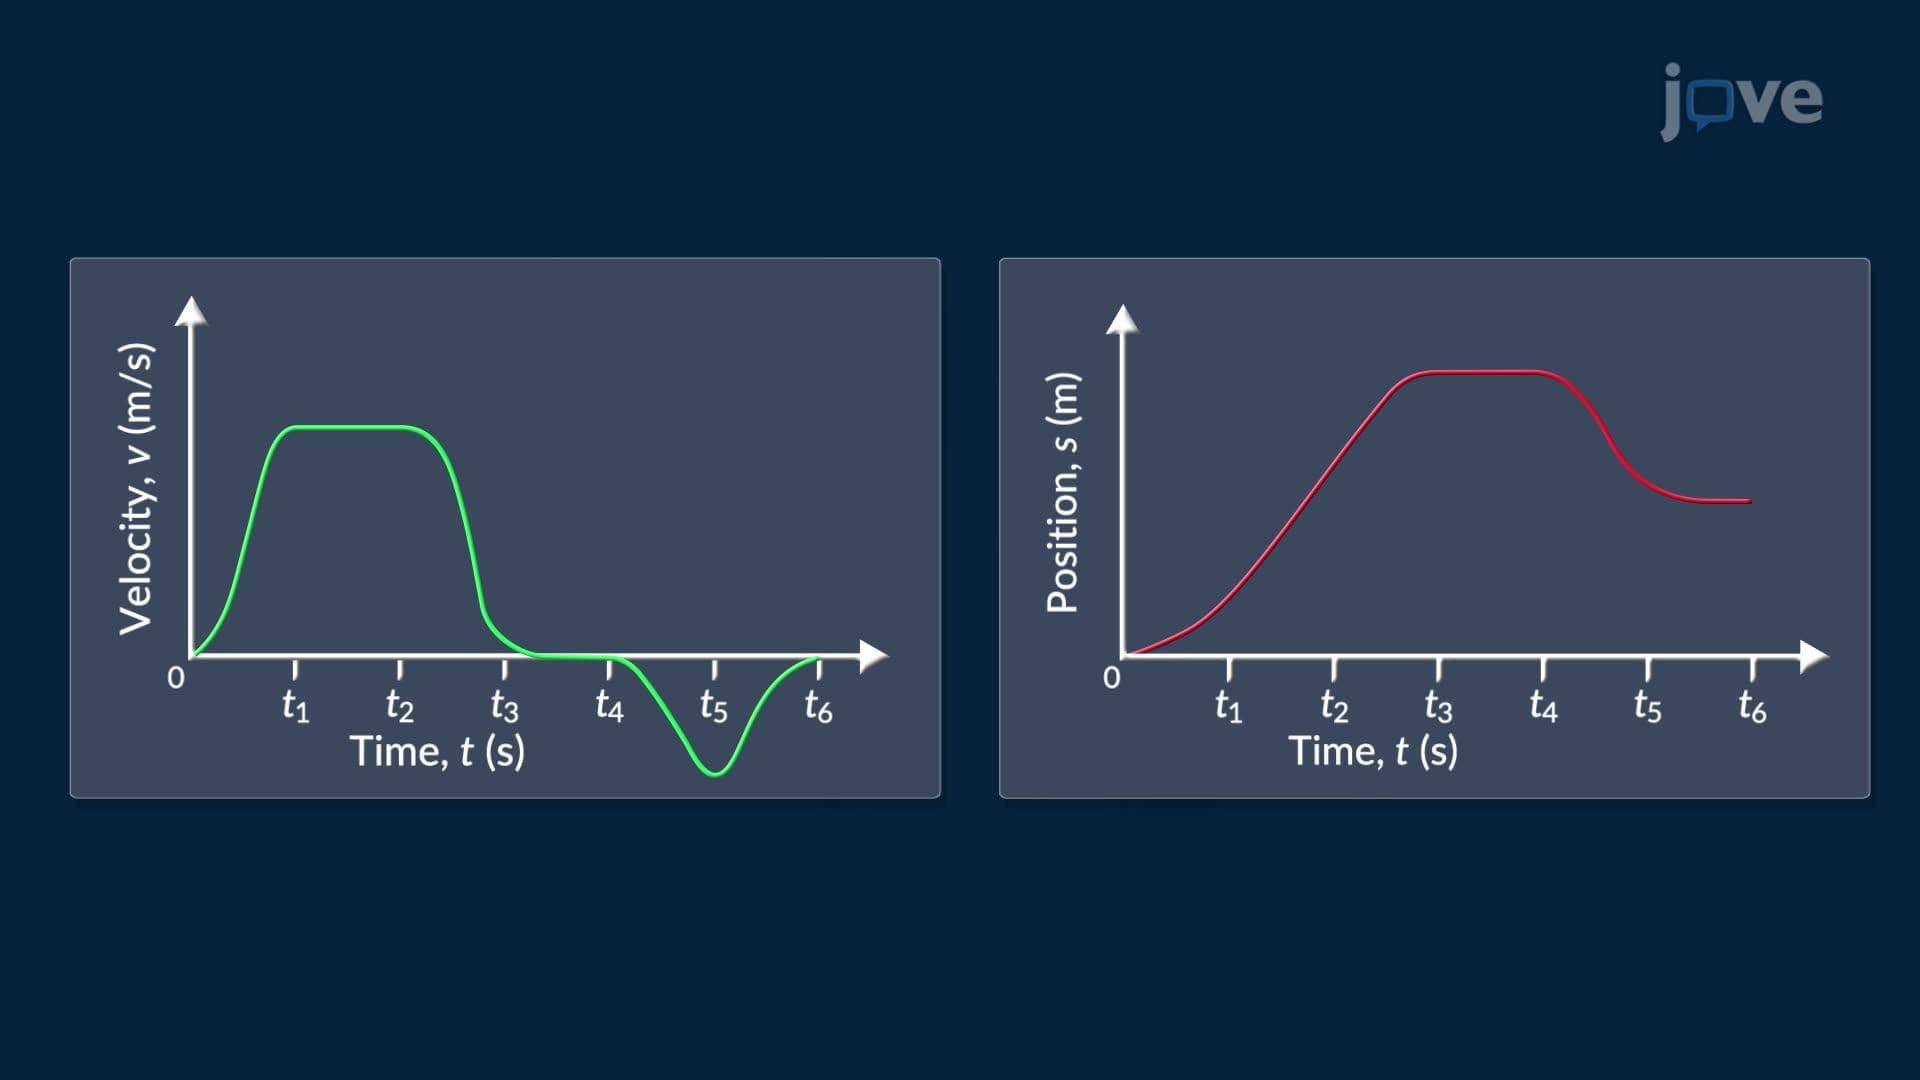
Graphing Antiderivatives
 01:07
Bar Graph