Related Experiment Video
Updated: Feb 8, 2026

Phage Therapy Application to Counteract Pseudomonas aeruginosa Infection in Cystic Fibrosis Zebrafish Embryos
Published on: May 12, 2020
A G542X cystic fibrosis mouse model for examining nonsense mutation directed therapies
Daniel R McHugh1,2, Miarasa S Steele2, Dana M Valerio2
1Department of Genetics and Genome Sciences, Case Western Reserve University, Cleveland, Ohio, United States of America.

Abstract:
Nonsense mutations are present in 10% of patients with CF, produce a premature termination codon in CFTR mRNA causing early termination of translation, and lead to lack of CFTR function. There are no currently available animal models which contain a nonsense mutation in the endogenous Cftr locus that can be utilized to test nonsense mutation therapies. In this study, we create a CF mouse model carrying the G542X nonsense mutation in Cftr using CRISPR/Cas9 gene editing. The G542X mouse model has reduced Cftr mRNA levels, demonstrates absence of CFTR function, and displays characteristic manifestations of CF mice such as reduced growth and intestinal obstruction. Importantly, CFTR restoration is observed in G542X intestinal organoids treated with G418, an aminoglycoside with translational readthrough capabilities. The G542X mouse model provides an invaluable resource for the identification of potential therapies of CF nonsense mutations as well as the assessment of in vivo effectiveness of these potential therapies targeting nonsense mutations.
Insights
Researchers developed a new mouse model for cystic fibrosis (CF) with a nonsense mutation. This model helps test therapies aimed at restoring CFTR function in CF patients with these specific mutations.
Area of Science:
- Biomedical Research
- Genetics
- Disease Modeling
Background:
- Nonsense mutations in the CFTR gene affect 10% of cystic fibrosis (CF) patients, leading to non-functional CFTR protein.
- Current animal models lack endogenous nonsense mutations, hindering the development of targeted therapies.
Purpose of the Study:
- To create a novel CF mouse model with a G542X nonsense mutation in the Cftr gene.
- To establish a preclinical tool for evaluating therapies for CF nonsense mutations.
Main Methods:
- CRISPR/Cas9 gene editing was employed to introduce the G542X nonsense mutation into the endogenous Cftr locus in mice.
- Phenotypic characterization of the G542X mouse model, including CFTR expression, function, and disease manifestations.
- In vitro assessment of CFTR restoration using G418, an aminoglycoside, in intestinal organoids derived from the G542X model.
Main Results:
- The G542X mouse model exhibits reduced Cftr mRNA levels and a complete absence of CFTR function.
- The model displays characteristic CF manifestations, including reduced growth and intestinal obstruction.
- Treatment with G418 demonstrated CFTR restoration in G542X intestinal organoids, indicating translational readthrough efficacy.
Conclusions:
- The G542X mouse model is a valuable preclinical tool for studying CF caused by nonsense mutations.
- This model will facilitate the identification and in vivo assessment of novel therapies targeting CFTR nonsense mutations.
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation,...
 01:24
Cystic Fibrosis: Management
Sinus disease and chronic...
 01:39
Mutations
 01:35
Mutations
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...
 02:27
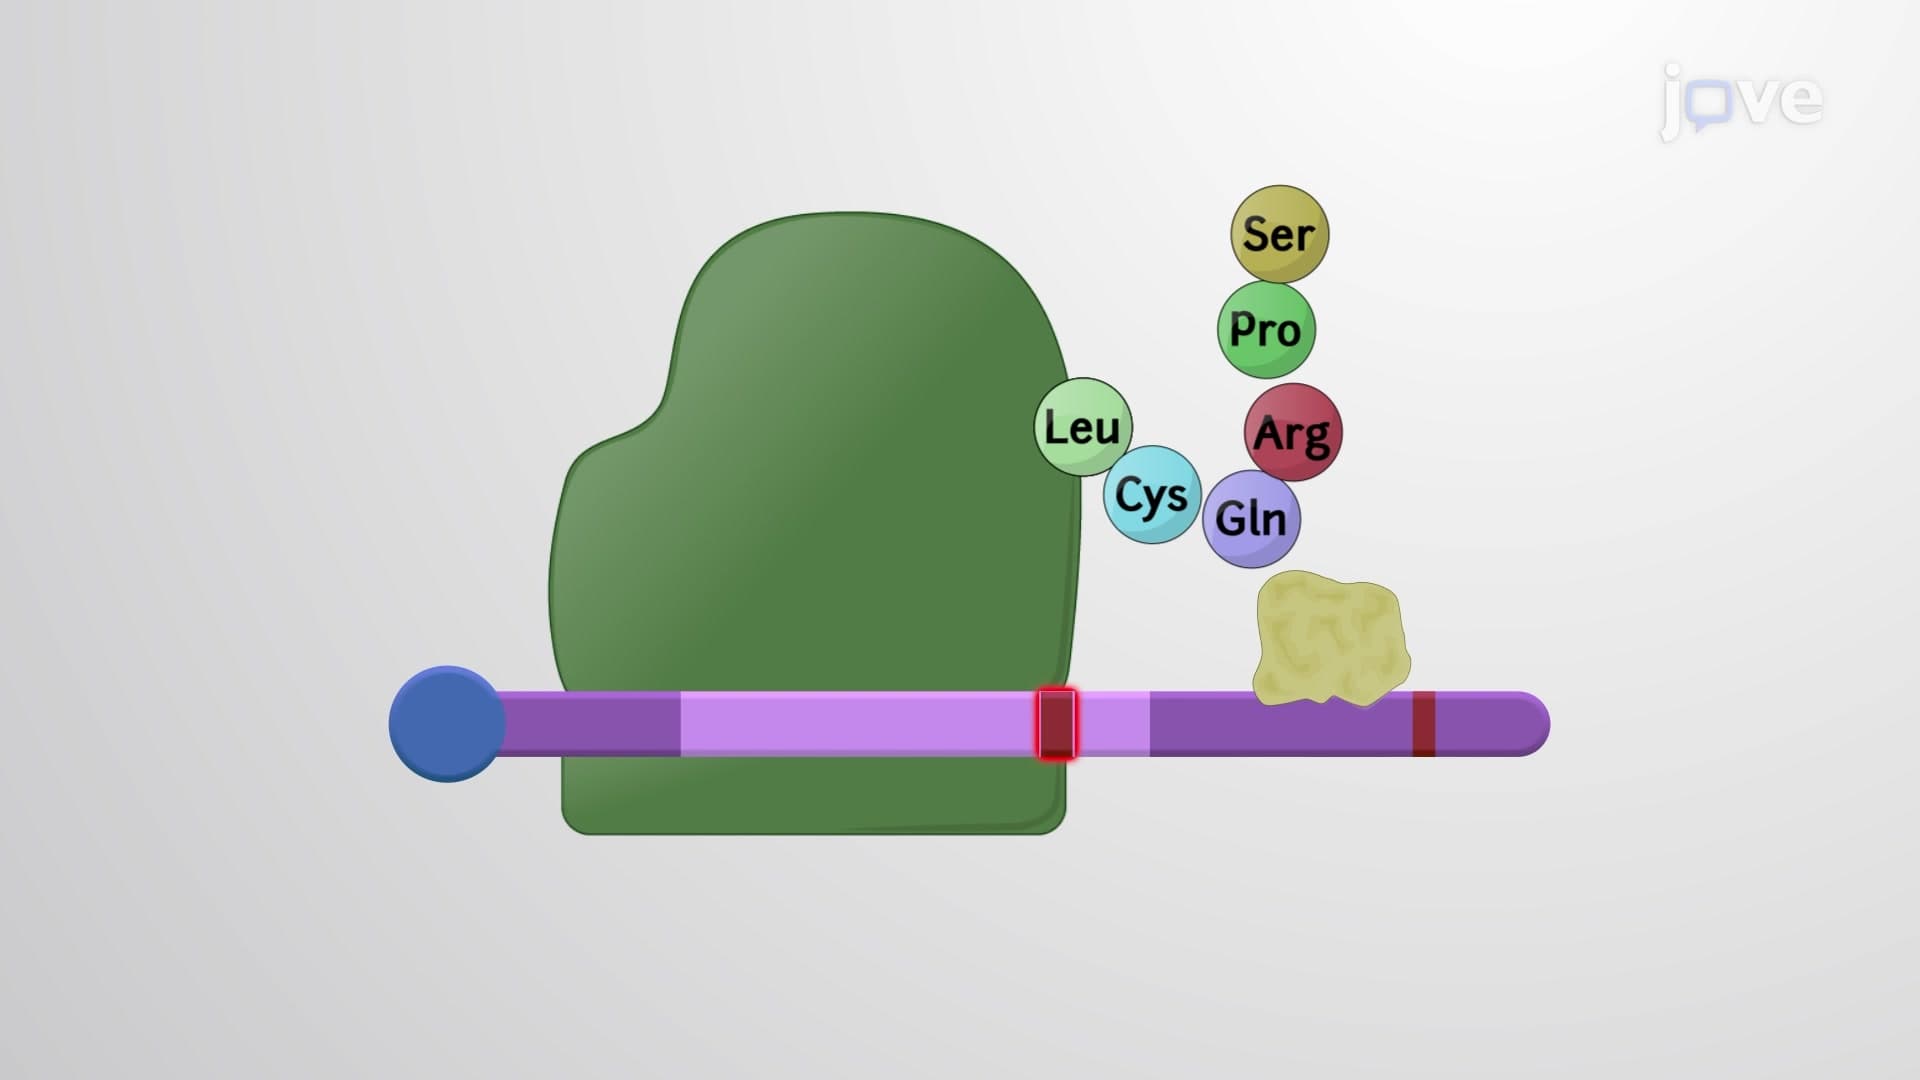
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
02:27Nonsense-mediated mRNA Decay