Related Experiment Video
Updated: Feb 8, 2026

Rapid Screening of HIV Reverse Transcriptase and Integrase Inhibitors
Published on: April 9, 2014
Improving Prediction Accuracy of Binding Free Energies and Poses of HIV Integrase Complexes Using the Binding Energy
Junchao Xia1, William Flynn2, Ronald M Levy2
1Center for Biophysics and Computational Biology and Department of Physics , Temple University , Philadelphia , Pennsylvania 19122 , United States.

Abstract:
To accelerate conformation sampling of slow dynamics from receptor or ligand, we introduced flattening potentials on selected bonded and nonbonded intramolecular interactions to the binding energy distribution analysis method (BEDAM) for calculating absolute binding free energies of protein-ligand complexes using an implicit solvent model and implemented flattening BEDAM using the asynchronous replica exchange (AsyncRE) framework for performing large scale replica exchange molecular dynamics (REMD) simulations. The advantage of using the flattening feature to reduce high energy barriers was exhibited first by the p-xylene-T4 lysozyme complex, where the intramolecular interactions of a protein side chain on the binding site were flattened to accelerate the conformational transition of the side chain from the trans to the gauche state when the p-xylene ligand is present in the binding site. Much more extensive flattening BEDAM simulations were performed for 53 experimental binders and 248 nonbinders of HIV-1 integrase which formed the SAMPL4 challenge, with the total simulation time of 24.3 μs. We demonstrated that the flattening BEDAM simulations not only substantially increase the number of true positives (and reduce false negatives) but also improve the prediction accuracy of binding poses of experimental binders. Furthermore, the values of area under the curve (AUC) of receiver operating characteristic (ROC) and the enrichment factors at 20% cutoff calculated from the flattening BEDAM simulations were improved significantly in comparison with that of simulations without flattening as we previously reported for the whole SAMPL4 database. Detailed analysis found that the improved ability to discriminate the binding free energies between the binders and nonbinders is due to the fact that the flattening simulations reduce the reorganization free energy penalties of binders and decrease the overlap of binding free energy distributions of binders relative to that of nonbinders. This happens because the conformational ensemble distributions for both the ligand and protein in solution match those at the fully coupled (complex) state more closely when the systems are more fully sampled after the flattening potentials are applied to the intermediate states.
Related Concept Videos
 02:13
Nuclear Binding Energy
 00:52
Potential Energy
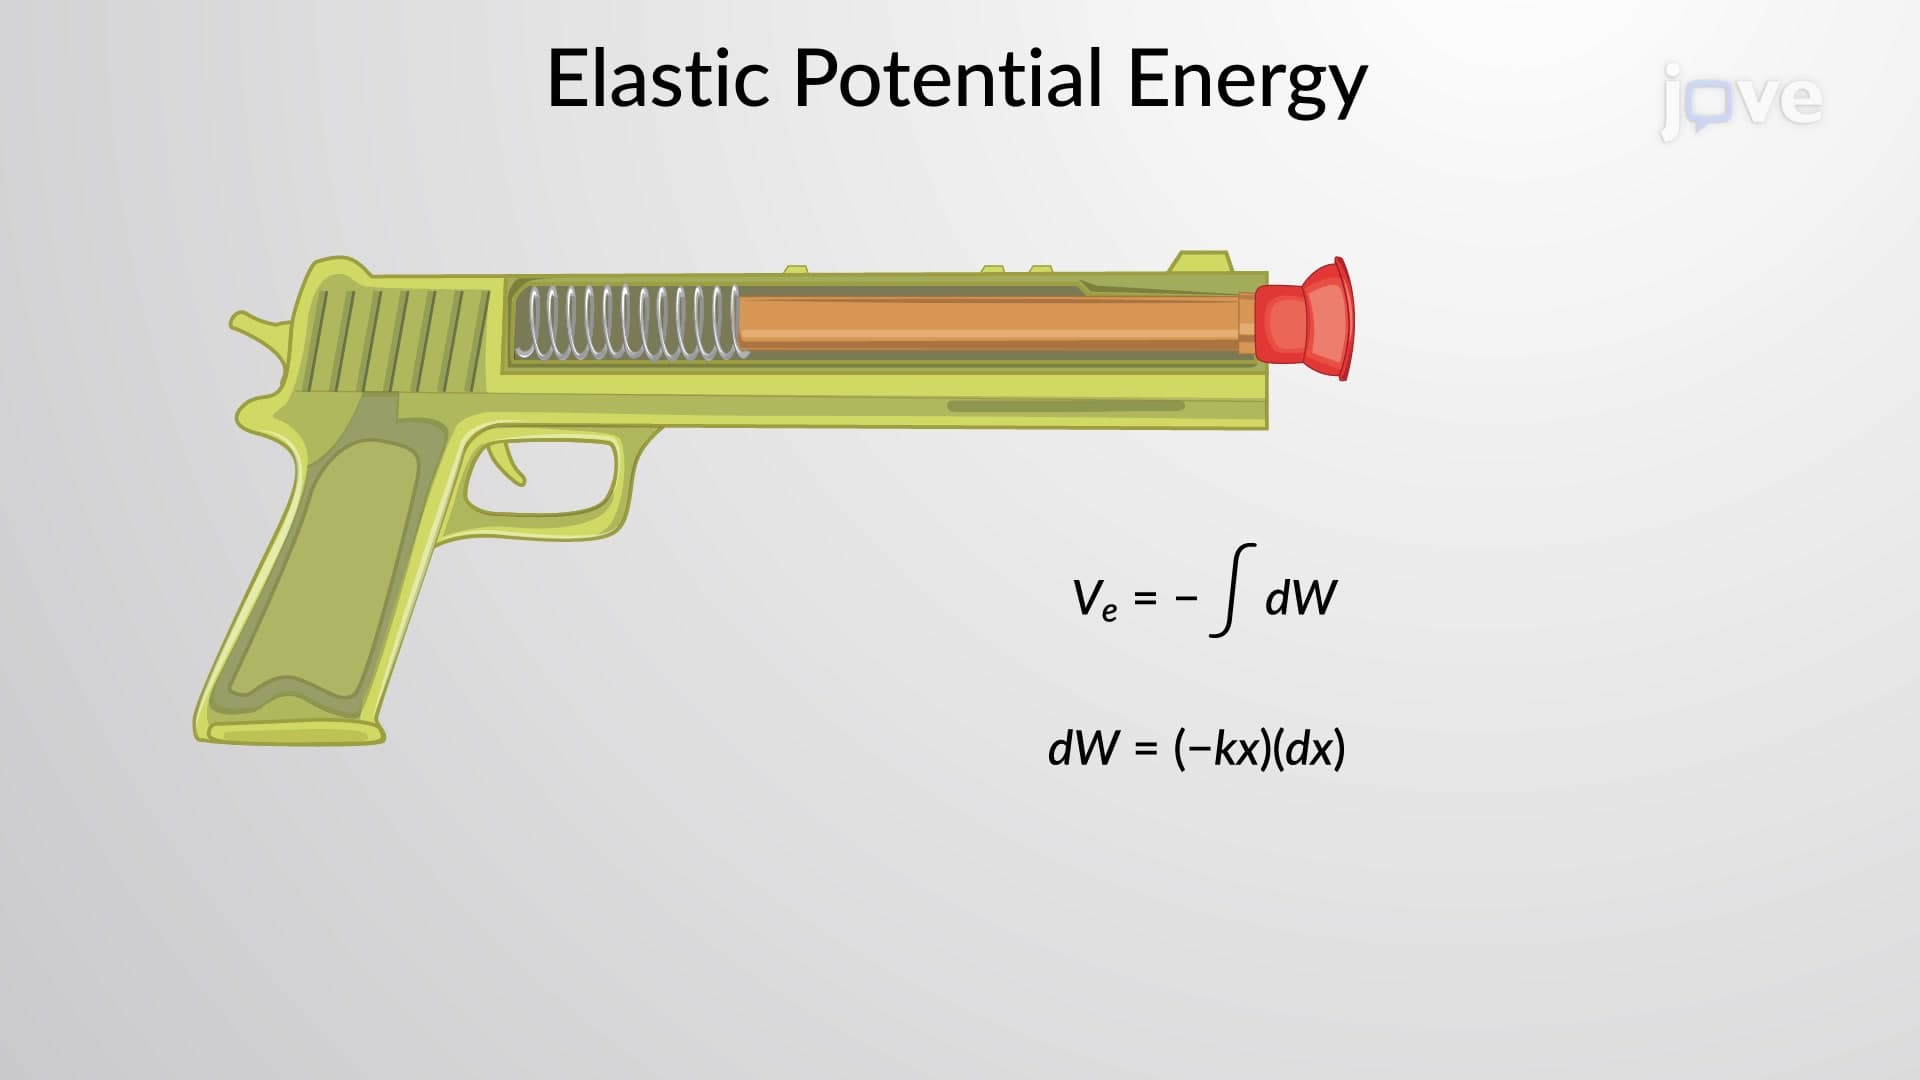
Chemical bonds that form attractive forces between atoms also contain potential energy, called chemical energy. When a chemical reaction...
 01:09
Potential Energy
 02:58
Cell Potential and Free Energy
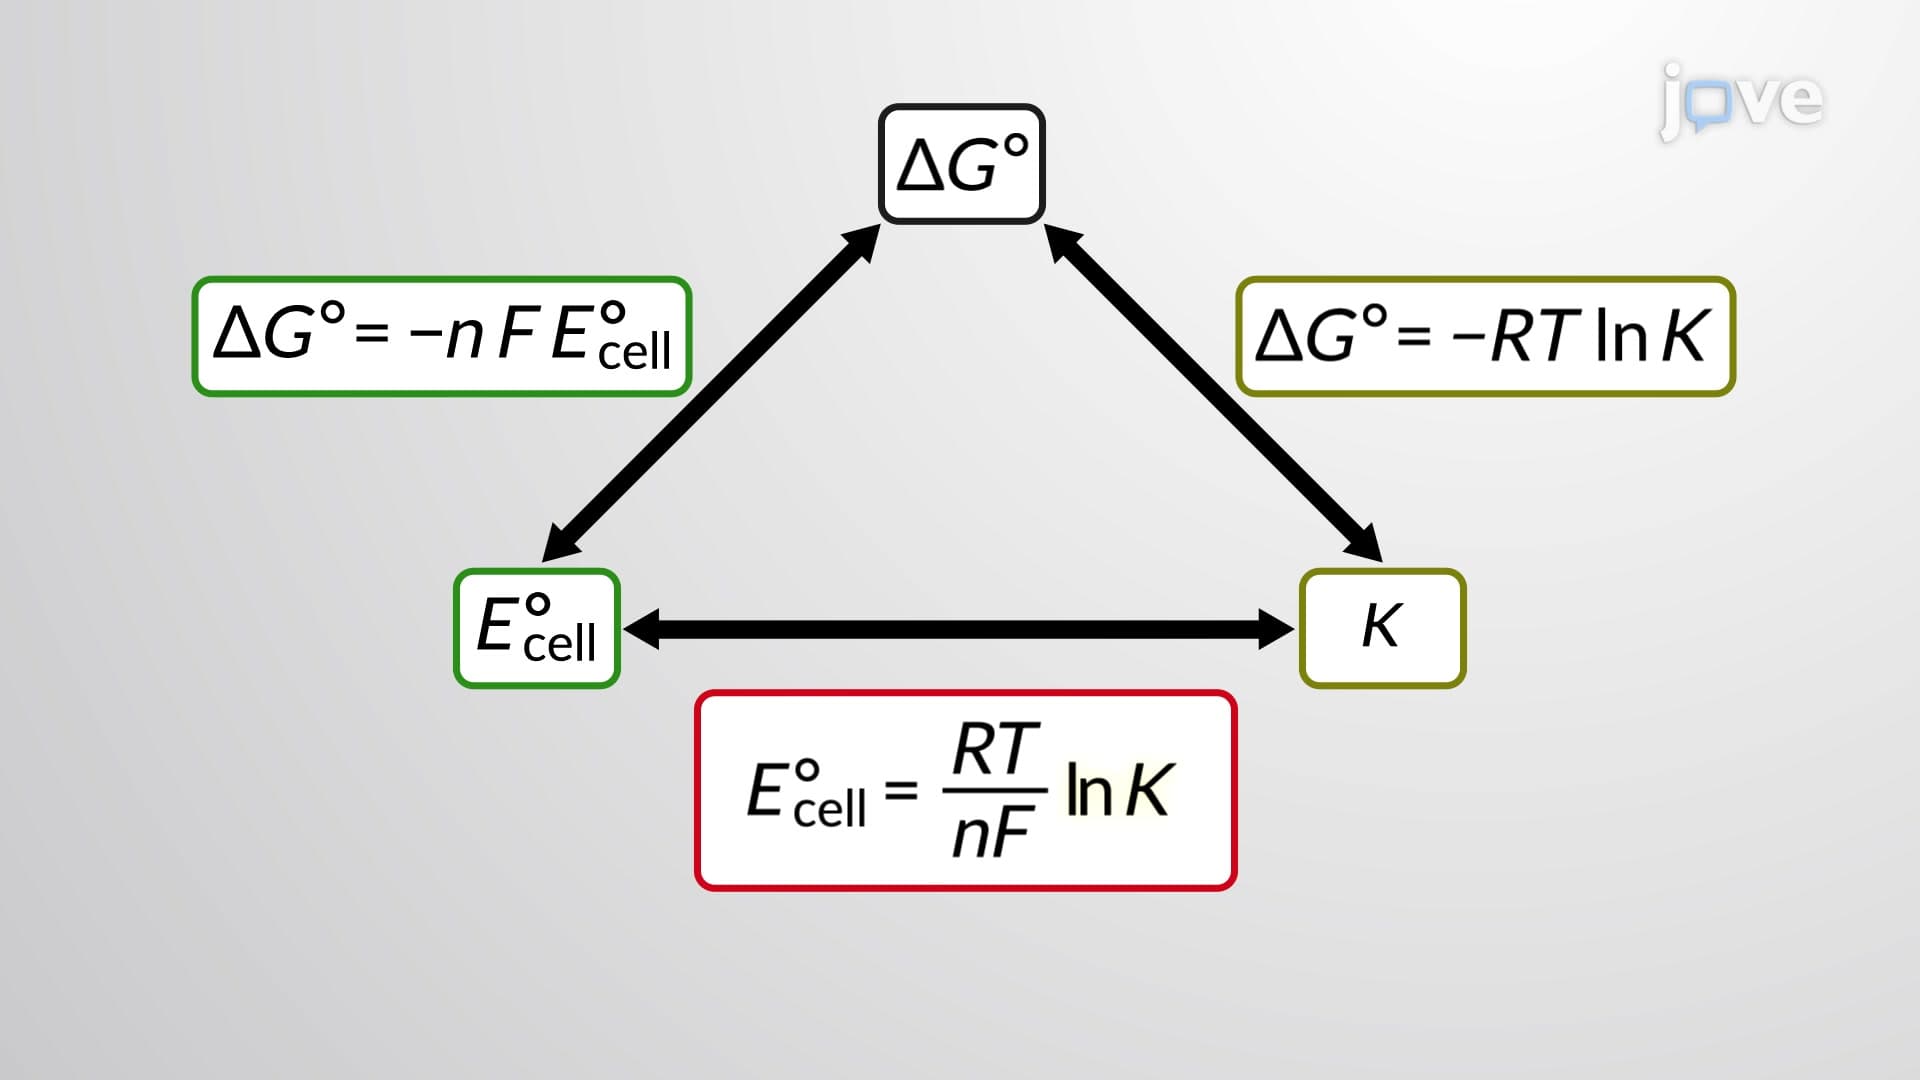
Thermodynamics is the branch of physics dealing with the relationship between heat and other forms of energy. In an electrochemical cell, chemical energy is converted into electrical energy.
Thus, a link can be predicted between cell potential, free energy change, and the equilibrium constant for the reaction. Cell potential can also be measured as the oxidant or the reducing strength, and similar acid-base strength measures are reflected in equilibrium...
 02:18
The Equilibrium Binding Constant and Binding Strength
 04:10
What is Energy?