Related Experiment Video
Updated: Feb 8, 2026
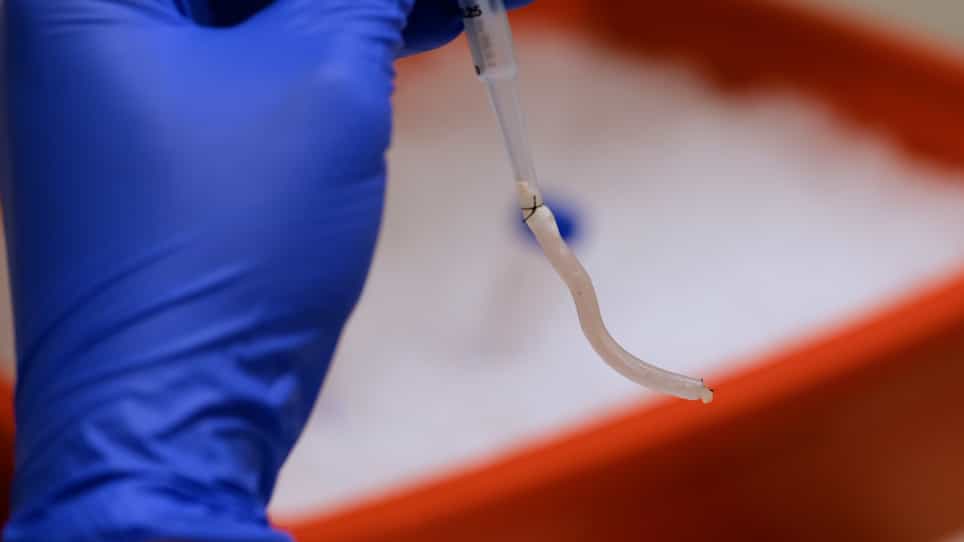

Generation of Murine Primary Colon Epithelial Monolayers from Intestinal Crypts
Published on: February 6, 2021
Insulin/IGF-1 enhances intestinal epithelial crypt proliferation through PI3K/Akt, and not ERK signaling in obese
Weinan Zhou1, Blair M Rowitz2, Megan J Dailey1
11 Department of Animal Sciences, University of Illinois at Urbana-Champaign, Urbana, IL 61801, USA.

Abstract:
The intestinal epithelium is continuously regenerated through proliferation and differentiation of stem cells located in the intestinal crypts. Obesity affects this process and results in greater stem cell proliferation and altered tissue growth and function. Obesity-induced high levels of insulin and insulin-like growth factor-1 in the stem cell niche are found to impact proliferation in rodents indicating that insulin and insulin-like growth factor-1 receptors may play a role in modulating intestinal epithelial stem cell proliferation. To determine whether insulin or insulin-like growth factor-1 can induce proliferation in human intestinal epithelial stem cells, and if two downstream insulin and insulin-like growth factor-1 receptor signaling pathways, PI3K/Akt and ERK, are involved, we used primary small intestinal epithelial crypts isolated from obese humans and investigated (1) the effect of insulin or insulin-like growth factor-1 on crypt proliferation, and (2) the effect of insulin and insulin-like growth factor-1 signaling inhibitors on insulin or insulin-like growth factor-1-induced proliferation. We found that insulin and insulin-like growth factor-1 enhanced the proliferation of crypt cells, including intestinal epithelial stem cells. Inhibition of the PI3K/Akt pathway attenuated insulin and insulin-like growth factor-1-induced proliferation, but inhibition of the ERK pathway had no effect. These results suggest that the classical metabolic PI3K pathway and not the canonical proliferation ERK pathway is involved in the insulin/insulin-like growth factor-1-induced increase in crypt proliferation in obese humans, which may contribute to abnormal tissue renewal and function. Impact statement This study investigates if insulin or insulin-like growth factor-1 (IGF-1) induces intestinal epithelial proliferation in humans, and if insulin and IGF-1 receptor signaling is involved in this process in obesity. Although obesity-induced high levels of insulin and IGF-1 in the stem cell niche are found to impact the proliferation of intestinal epithelial stem cells in rodents, we are the first to investigate this effect in humans. We found that insulin and IGF-1 enhanced the proliferation of intestinal crypts (including stem cells and other crypt cells) isolated from obese humans, and PI3K/Akt, and not ERK signaling was involved in insulin or IGF-1-induced proliferation. The imbalance in signaling between PI3K/Akt and ERK pathways may point to a pathway-specific impairment in insulin/IGF-1 receptor signaling. We propose that this may contribute to reciprocal relationships between insulin/IGF-1 receptor resistance and intestinal epithelial proliferation that leads to abnormal tissue renewal and function.
More Related Videos
 07:15
Mechanism of Kemeng Fang's Inhibition of Podocyte Apoptosis in Rats with Membranous Nephropathy through the PI3K/AKT Signaling Pathway
Published on: August 23, 2024
 08:19
Culture of Piglet Intestinal 3D Organoids from Cryopreserved Epithelial Crypts and Establishment of Cell Monolayers
Published on: February 10, 2023
Related Concept Videos
 01:22
PI3K/mTOR/AKT Signaling Pathway
 01:28
Insulin: The Receptor and Signaling Pathways
 01:23
Anatomy of the Intestines
Small Intestines
The small intestine is an ~7 meter-long tube with an inner diameter of just 2.5 cm. Since most nutrients are absorbed here, the inner lining of the...
 01:24
Obesity
 01:10
Interpreting ¹H NMR Signal Splitting: The (n + 1) Rule
 01:12
Role Of Notch Signalling In Intestinal Stem Cell Renewal
Direct cell-to-cell contact is needed for the activation of Notch signaling. The signal is initiated when a notch ligand binds to a receptor on an adjacent cell, also...