Related Experiment Video
Updated: Feb 7, 2026

Automated Analysis of Intracellular Phenotypes of Salmonella Using ImageJ
Published on: August 9, 2022
Proper conditional analysis in the presence of missing data: Application to large scale meta-analysis of tobacco use
Yu Jiang1, Sai Chen2, Daniel McGuire1
1Department of Public Health Sciences, Penn State College of Medicine, Hershey, Pennsylvania, United States of America.

Abstract:
Meta-analysis of genetic association studies increases sample size and the power for mapping complex traits. Existing methods are mostly developed for datasets without missing values, i.e. the summary association statistics are measured for all variants in contributing studies. In practice, genotype imputation is not always effective. This may be the case when targeted genotyping/sequencing assays are used or when the un-typed genetic variant is rare. Therefore, contributed summary statistics often contain missing values. Existing methods for imputing missing summary association statistics and using imputed values in meta-analysis, approximate conditional analysis, or simple strategies such as complete case analysis all have theoretical limitations. Applying these approaches can bias genetic effect estimates and lead to seriously inflated type-I or type-II errors in conditional analysis, which is a critical tool for identifying independently associated variants. To address this challenge and complement imputation methods, we developed a method to combine summary statistics across participating studies and consistently estimate joint effects, even when the contributed summary statistics contain large amounts of missing values. Based on this estimator, we proposed a score statistic called PCBS (partial correlation based score statistic) for conditional analysis of single-variant and gene-level associations. Through extensive analysis of simulated and real data, we showed that the new method produces well-calibrated type-I errors and is substantially more powerful than existing approaches. We applied the proposed approach to one of the largest meta-analyses to date for the cigarettes-per-day phenotype. Using the new method, we identified multiple novel independently associated variants at known loci for tobacco use, which were otherwise missed by alternative methods. Together, the phenotypic variance explained by these variants was 1.1%, improving that of previously reported associations by 71%. These findings illustrate the extent of locus allelic heterogeneity and can help pinpoint causal variants.
Related Concept Videos
 01:12
Analysis of Population Pharmacokinetic Data
 01:13
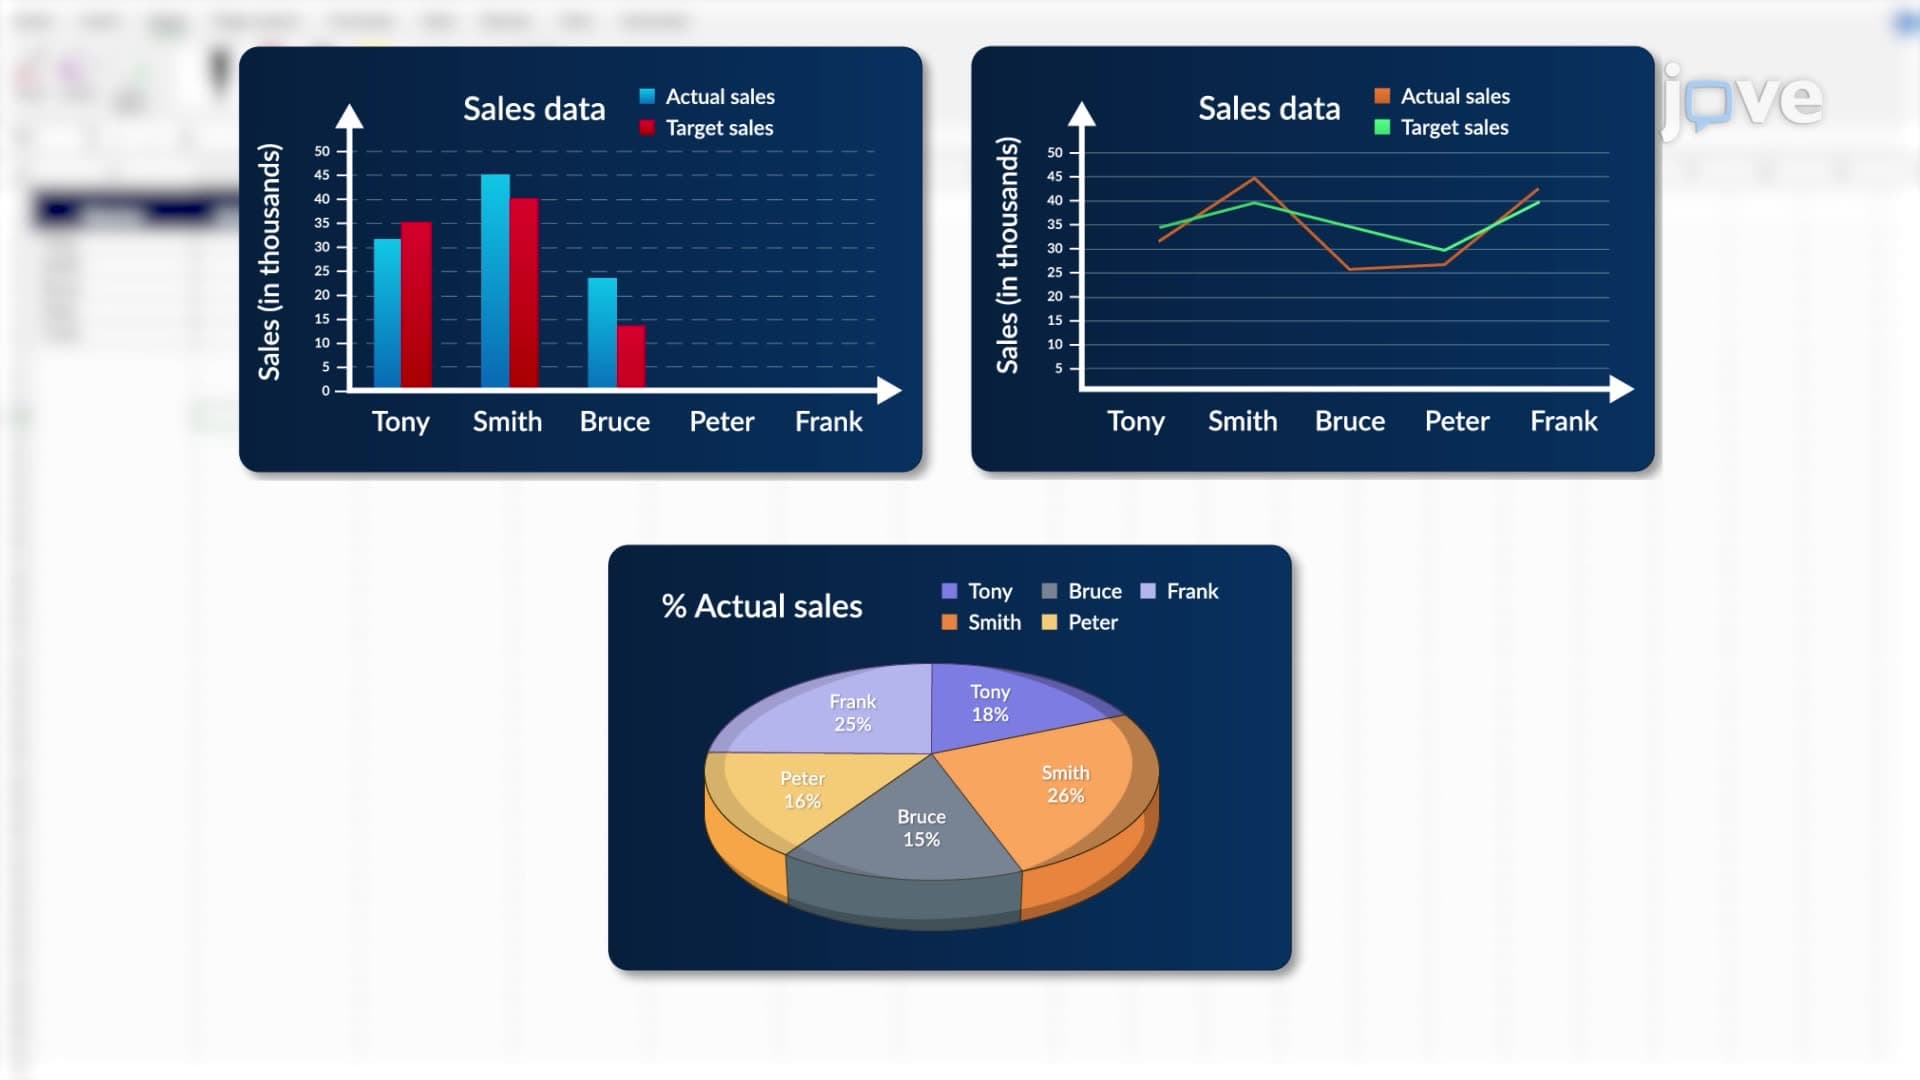
Overview of Microsoft Excel as a Data Analysis Tool
 01:17
Performing a Simple Data Analysis using MS-Excel Function
SUM: This function calculates the total sum of a range of values. It's the foundation for aggregating data, essential for determining overall trends and totals in datasets.
AVERAGE: It computes the mean value of a given set of numbers, providing a quick insight into the central...
 01:12
Statistical Software for Data Analysis and Clinical Trials
 01:35
Pedigree Analysis
 00:59
Model-Independent Approaches for Pharmacokinetic Data: Noncompartmental Analysis
One important characteristic of noncompartmental analyses is that drug exposure increases proportionally with increasing doses. This...