Related Experiment Video
Updated: Feb 7, 2026
Determination of Molecular Structures of HIV Envelope Glycoproteins using Cryo-Electron Tomography and Automated Sub-tomogram Averaging
Published on: December 1, 2011
Transferability in Machine Learning for Electronic Structure via the Molecular Orbital Basis
Matthew Welborn1, Lixue Cheng1, Thomas F Miller1
1Division of Chemistry and Chemical Engineering , California Institute of Technology , Pasadena , California 91125 , United States.

Abstract:
We present a machine learning (ML) method for predicting electronic structure correlation energies using Hartree-Fock input. The total correlation energy is expressed in terms of individual and pair contributions from occupied molecular orbitals, and Gaussian process regression is used to predict these contributions from a feature set that is based on molecular orbital properties, such as Fock, Coulomb, and exchange matrix elements. With the aim of maximizing transferability across chemical systems and compactness of the feature set, we avoid the usual specification of ML features in terms of atom- or geometry-specific information, such atom/element-types, bond-types, or local molecular structure. ML predictions of MP2 and CCSD energies are presented for a range of systems, demonstrating that the method maintains accuracy while providing transferability both within and across chemical families; this includes predictions for molecules with atom-types and elements that are not included in the training set. The method holds promise both in its current form and as a proof-of-principle for the use of ML in the design of generalized density-matrix functionals.
Related Concept Videos
 03:51
Molecular Orbital Theory II
 02:35
Molecular Orbital Theory I
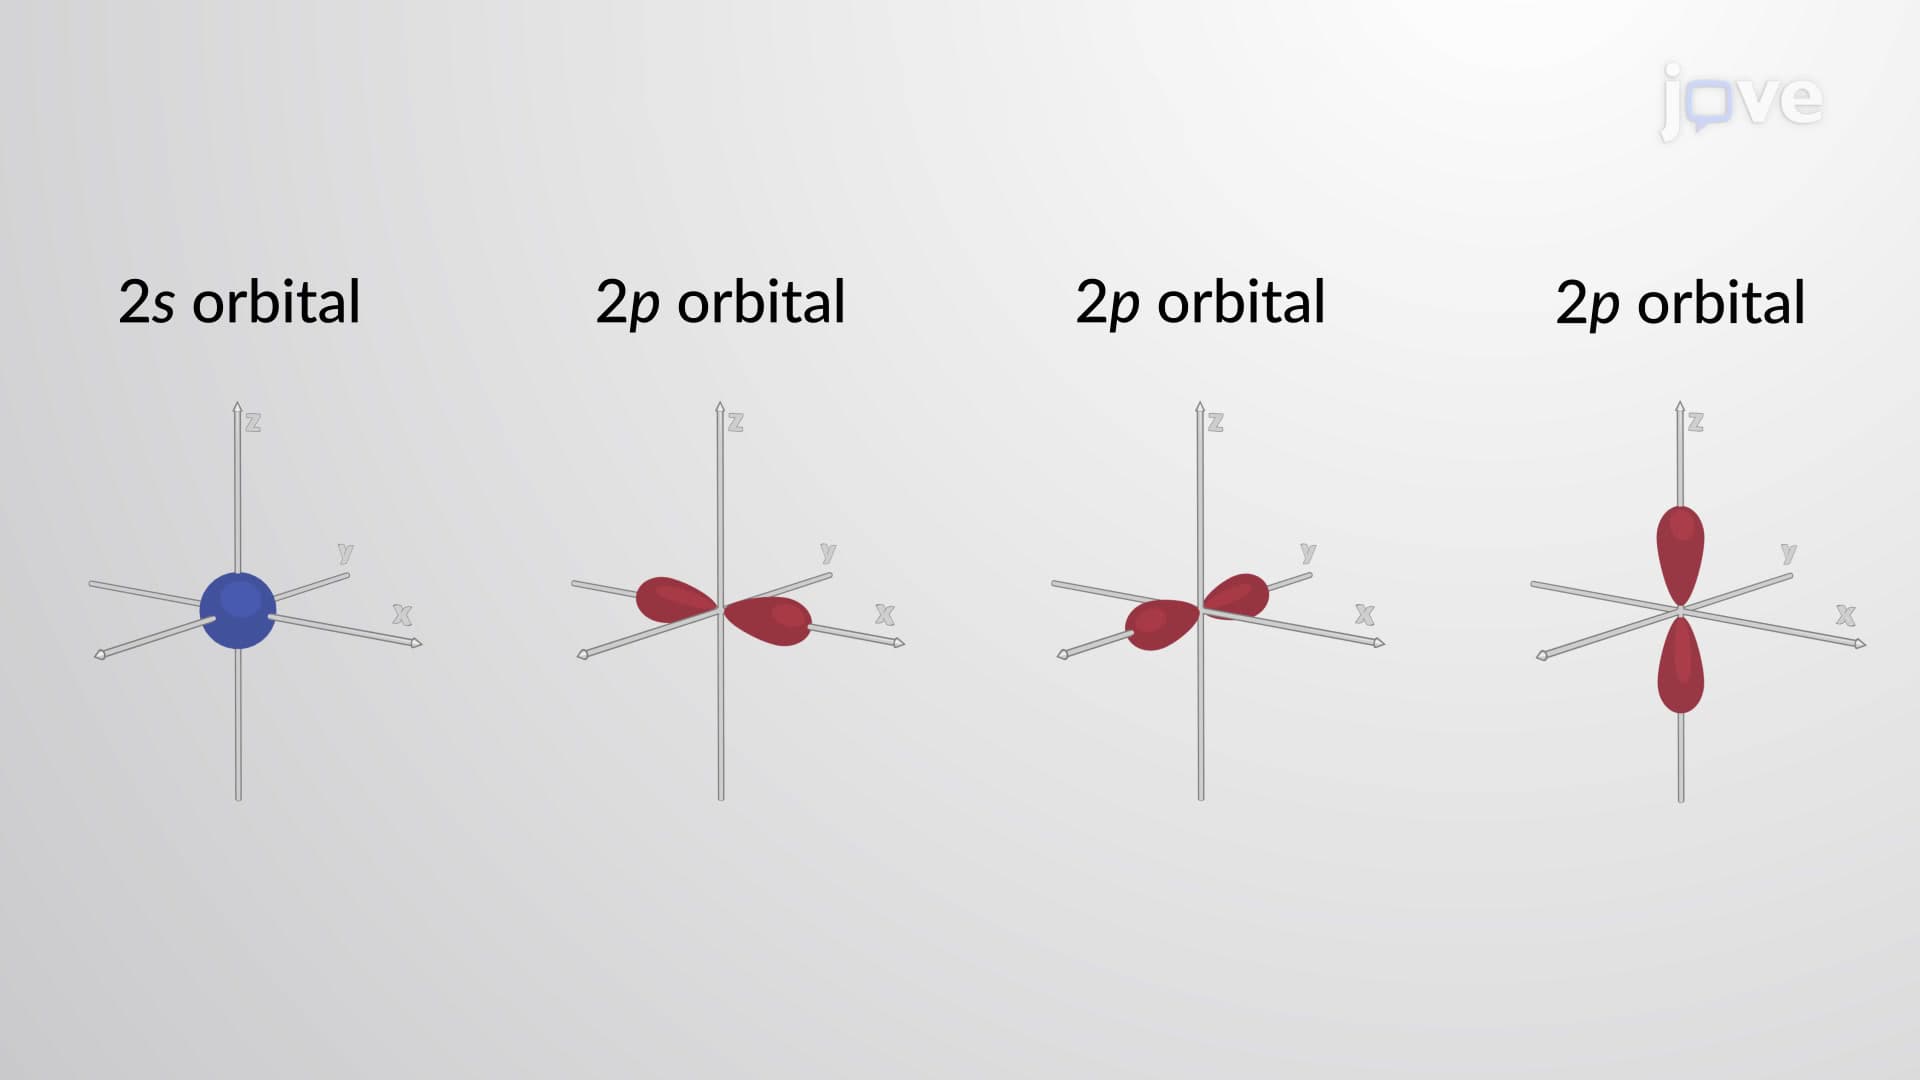 01:18
Electron Orbital Model
The first shell is closest to the nucleus, and it has only one subshell with a single spherical orbital called the...
 01:18
Structure of Benzene: Molecular Orbital Model
 02:48
Ionic Bonding and Electron Transfer
 03:35
Hybridization of Atomic Orbitals II