GAPDH with NAD+-binding site mutation competitively inhibits the wild-type and affects glucose metabolism in cancer
Related Experiment Video
Updated: Feb 7, 2026
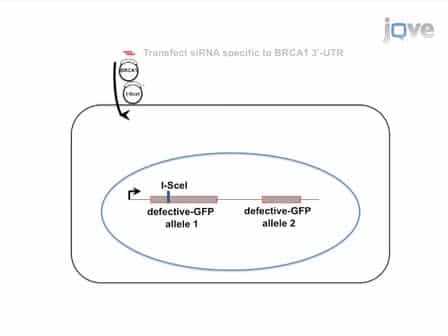
Identifying the Effects of BRCA1 Mutations on Homologous Recombination using Cells that Express Endogenous Wild-type BRCA1
Published on: February 17, 2011
Rani Kunjithapatham1, Shanmugasundaram Ganapathy-Kanniappan1
1The Division of Interventional Radiology, Russell H. Morgan Department of Radiology & Radiological Science, The Johns Hopkins University School of Medicine, Baltimore, MD, USA.

Background:
Rapid utilization of glucose is a metabolic signature of majority of cancers, hence enzymes of the glycolytic pathway remain attractive therapeutic targets. Recent reports have shown that targeting the glycolytic enzyme, glyceraldehyde-3-phosphate dehydrogenase (GAPDH), an abundant, ubiquitous multifunctional protein frequently upregulated in cancer, affects cancer progression. Here, we report that a catalytically-deficient mutant-GAPDH competitively inhibits the wild-type, and disrupts glucose metabolism in cancer cells.
Methods:
Using site-directed mutagenesis, the human GAPDH clone was mutated at one of the NAD+-binding sites, (i.e.) arginine (R13) and isoleucine (I14) to glutamine (Q13) and phenylalanine (F14), respectively. The inhibitory role of the mutant-GAPDH, and its effect on energy metabolism and cancer phenotype was determined using in vitro and in vivo models of cancer.
Results:
The enzymatically-dysfunctional mutant-GAPDH competitively inhibited the wild-type GAPDH in a cell-free system. In cancer cells, ectopic expression of the mutant-GAPDH, but not the wild-type, inhibited the glycolytic capacity of cellular-GAPDH, and led to the induction of metabolic stress accompanied by a sharp decline in glucose-uptake. Furthermore, expression of mutant-GAPDH affected cancer growth in vitro and in vivo. Mechanistically, structural analysis by bioinformatics revealed that the mutations at the NAD+-binding site altered the solvent-accessibility that perhaps affected the functionality of mutant-GAPDH.
Conclusion:
Mutant-GAPDH affects the enzymatic function of cellular-GAPDH and disrupts energy metabolism.
General Significance:
Our findings demonstrate that a minimal mutation at the NAD+-binding site is sufficient to generate a competitive but dysfunctional GAPDH, and its ectopic expression inhibits the wild-type to disrupt glycolysis.
Insights
A modified enzyme, glyceraldehyde-3-phosphate dehydrogenase (GAPDH), competitively inhibits the normal enzyme, disrupting glucose metabolism and cancer cell growth. This finding offers a new strategy for targeting cancer by interfering with energy production.
Area of Science:
- Biochemistry
- Cancer Biology
- Metabolic Pathways
Background:
- Rapid glucose utilization is a hallmark of many cancers, making glycolytic enzymes attractive therapeutic targets.
- Glyceraldehyde-3-phosphate dehydrogenase (GAPDH), a key glycolytic enzyme, is frequently upregulated in cancer and influences cancer progression.
- Targeting GAPDH offers a potential strategy for cancer therapy due to its critical role in cancer metabolism.
Purpose of the Study:
- To investigate the potential of a catalytically-deficient mutant GAPDH to inhibit wild-type GAPDH function.
- To determine the effect of mutant GAPDH on cancer cell glucose metabolism, energy production, and overall cancer phenotype.
- To explore the mechanistic basis of mutant GAPDH inhibition through structural analysis.
Main Methods:
- Site-directed mutagenesis was used to create a catalytically deficient human GAPDH mutant by altering key residues in the NAD+ binding site (R13Q, I14F).
- The inhibitory effect of the mutant GAPDH on wild-type GAPDH was assessed in cell-free systems and in cancer cells with ectopic expression.
- In vitro and in vivo cancer models were utilized to evaluate the impact of mutant GAPDH on cancer cell metabolism, growth, and phenotype.
Main Results:
- The enzymatically dysfunctional mutant GAPDH competitively inhibited wild-type GAPDH in vitro.
- Ectopic expression of mutant GAPDH, but not wild-type, significantly inhibited cancer cell glycolytic capacity, leading to metabolic stress and reduced glucose uptake.
- Mutant GAPDH expression impaired cancer growth in both in vitro and in vivo models, with bioinformatics suggesting altered NAD+ binding site functionality.
Conclusions:
- A minimally mutated GAPDH can act as a competitive inhibitor of the wild-type enzyme.
- Ectopic expression of this dysfunctional GAPDH disrupts cellular energy metabolism by inhibiting glycolysis.
- This strategy presents a novel approach to targeting cancer progression by disrupting essential metabolic pathways.
More Related Videos
Related Concept Videos
 02:34
Competition
 00:46
Feedback Inhibition
 01:39
Mutations
 02:21
Cancers Originate from Somatic Mutations in a Single Cell
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 00:36
Viral Mutations