Accurate Spin-State Energetics for Aryl Carbenes
Reza Ghafarian Shirazi1, Frank Neese1, Dimitrios A Pantazis1
1Max-Planck-Institut für Kohlenforschung, Kaiser-Wilhelm-Platz 1 , 45470 Mülheim an der Ruhr , Germany.

Abstract:
A test set of 12 aryl carbenes (AC12) is compiled with the purpose of establishing their adiabatic singlet-triplet energy splittings using correlated wave function based methods. The set covers both singlet and triplet ground state aryl carbenes, as well as a range of magnitudes for the ground state to excited state gap. The performance of coupled cluster methods is examined with respect to the reference wave function, the basis set, and a number of additional methodological parameters that enter the calculation. Inclusion of perturbative triples and basis set extrapolation with a combination of triple and quadruple-ζ basis sets are both required to ensure high accuracy. When canonical coupled cluster calculations become too expensive, the domain-based local pair natural orbital approach DLPNO-CCSD(T) can be used as a reliable method for larger systems, as it achieves a mean absolute error of only 0.2 kcal/mol for the singlet-triplet gaps in the present test set. Other first-principles wave function methods and selected density functional methods are also evaluated. Second-order Møller-Plesset perturbation theory approaches are only applicable in conjunction with orbital optimization (OO-MP2). Among the representative density functional methods tested, only double hybrid functionals perform sufficiently accurately to be considered useful for systems with small singlet-triplet gaps. On the basis of the reference coupled cluster results, projected gas-phase free energies are reported for all aryl carbenes. Finally, the treatment of singlet-triplet gaps in solution is discussed in terms of both implicit and explicit solvation.
Related Concept Videos
 02:35
Energetics of Solution Formation
When the strengths of the intermolecular forces of attraction between solute and solvent species in a solution are no different than those present in the separated components, the solution is formed with no accompanying energy change. Formation of the solution requires the solute–solute and solvent–solvent...
 01:10
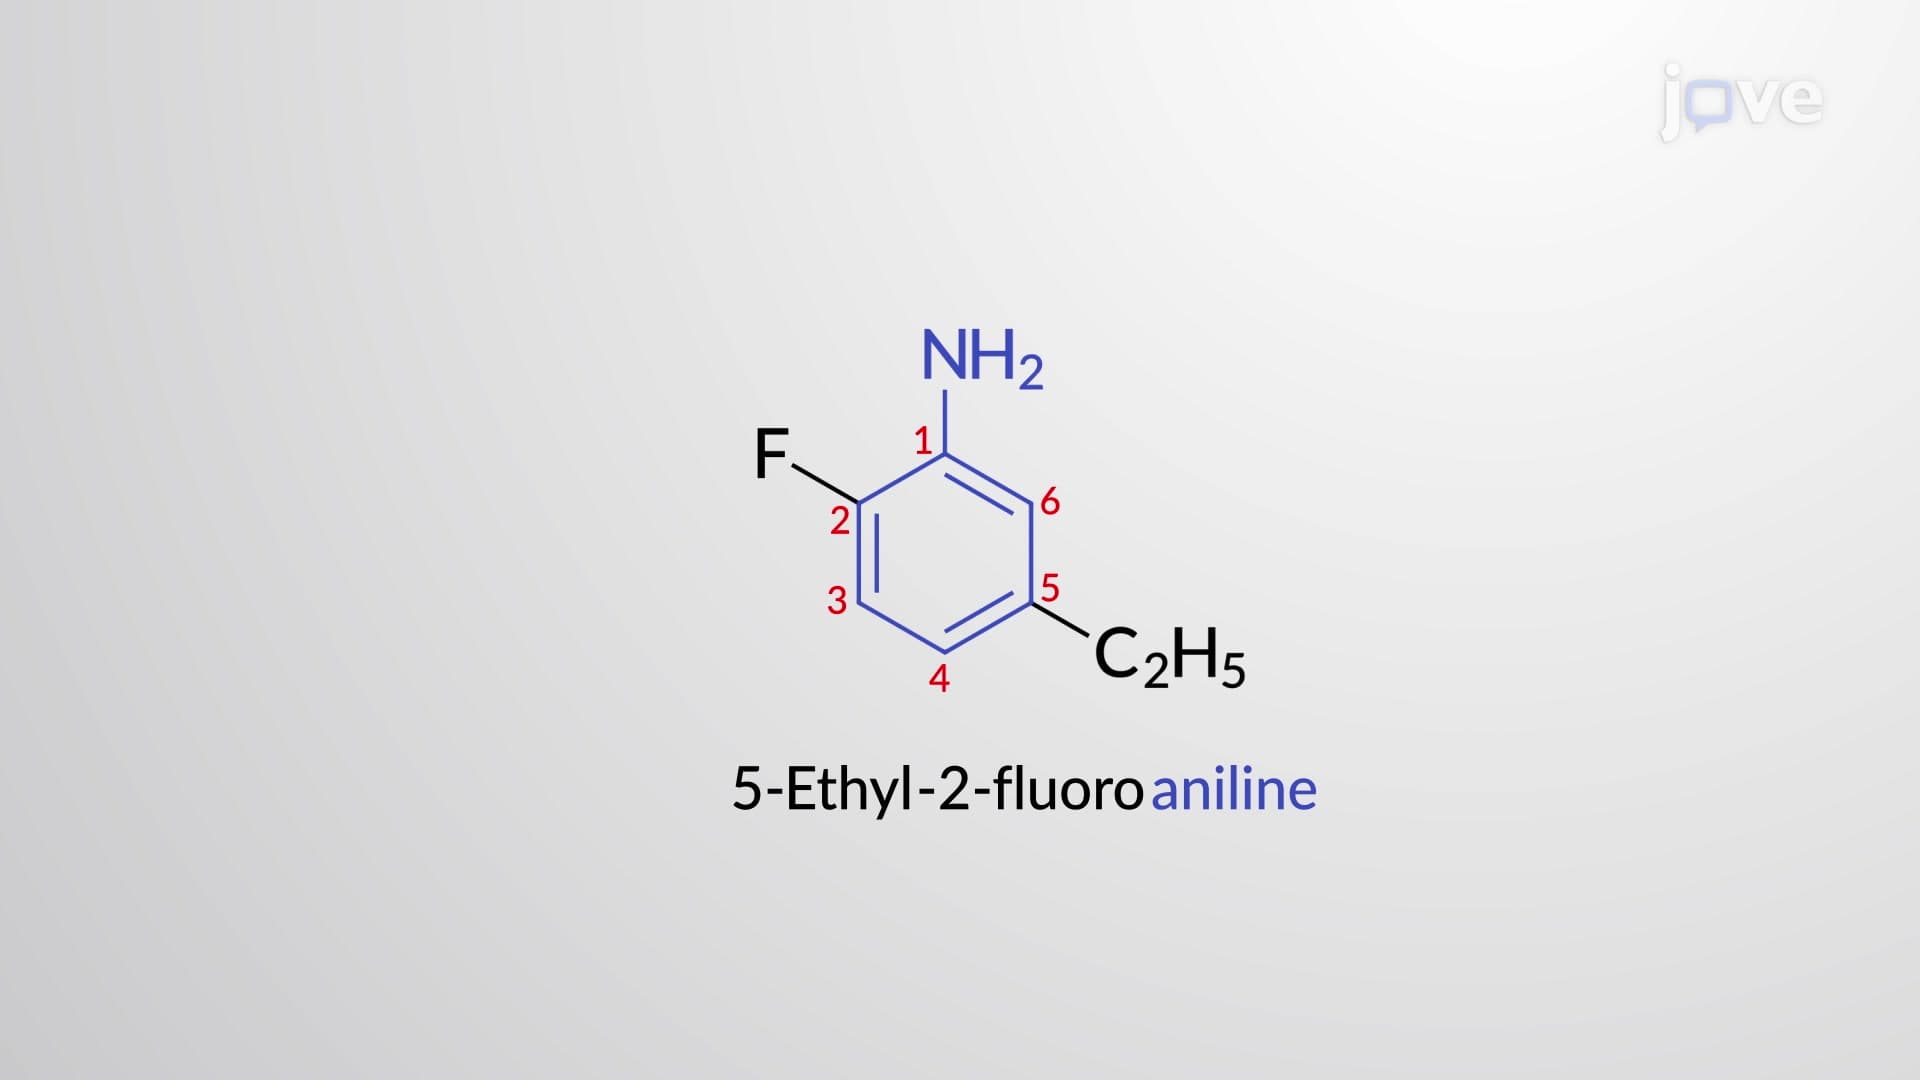
Nomenclature of Aryl and Heterocyclic Amines
 01:08
NMR Spectroscopy: Spin–Spin Coupling
 01:17
Spin–Spin Coupling: One-Bond Coupling
 01:08
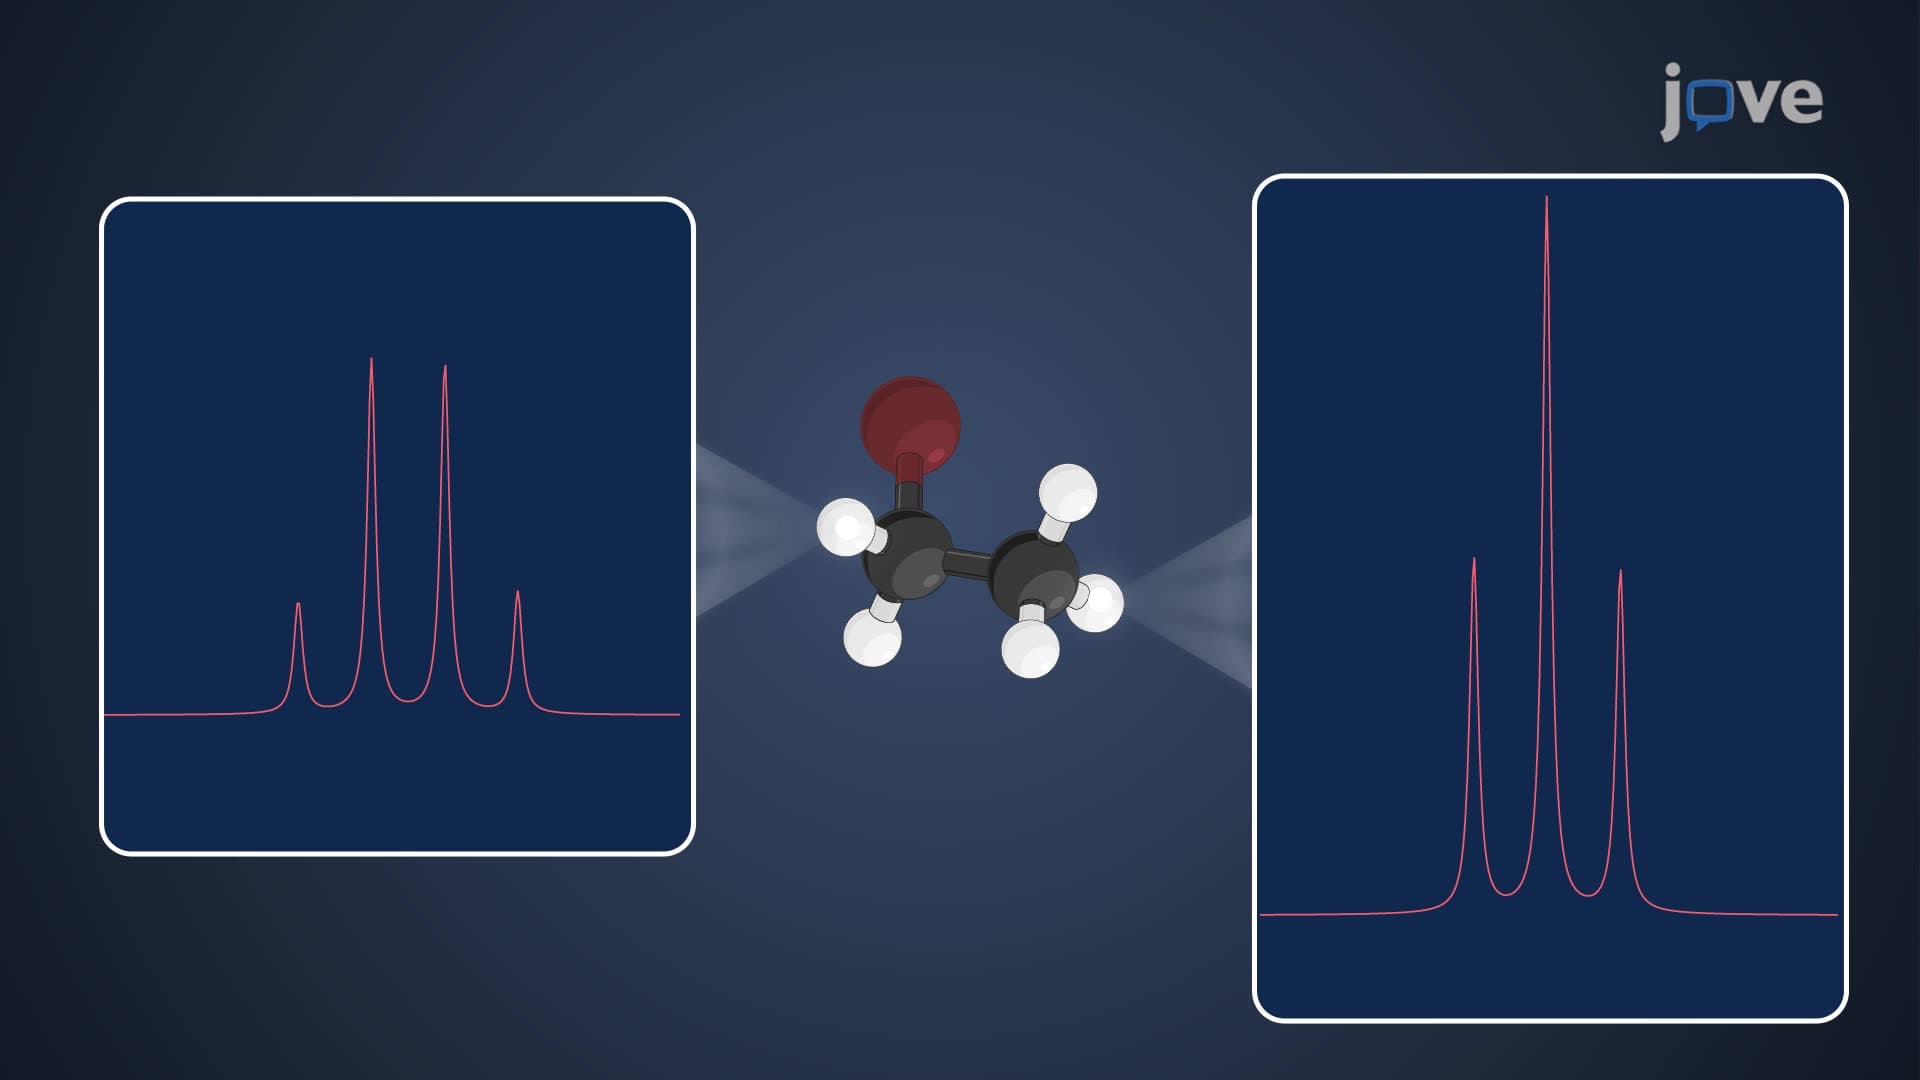
Spin–Spin Coupling Constant: Overview
Qualitatively, any spin plus-half nucleus polarizes the spins of its electrons to the minus-half state. Consequently, the paired electron in the hydrogen–carbon bond must...
 01:20
Spin–Spin Coupling: Two-Bond Coupling (Geminal Coupling)
The central atom need not be NMR-active because its electrons are affected by the electron polarization of the spin-active atoms. However, spin information is transmitted less effectively than in one-bond coupling, and 2J values are usually weaker than 1J values. The energy of...