
Mechanistic In Vitro Studies Indicate that the Clinical Drug-Drug Interaction between Telithromycin and Simvastatin
Robert Elsby1, Victoria Hare2, Hannah Neal2
1Department of ADME Sciences, Cyprotex Discovery Ltd. (an Evotec company), Macclesfield, Cheshire, United Kingdom (R.E., V.H., H.N., S.O., C.P., K.P., R.U.G., P.B.); and Drug Metabolism and Pharmacokinetics, Evotec, Abingdon, Oxfordshire, United Kingdom (R.J.R.) r.elsby@cyprotex.com.

Abstract:
A previous attempt to accurately quantify the increased simvastatin acid exposure due to drug-drug interaction (DDI) with coadministered telithromycin, using a mechanistic static model, substantially underpredicted the magnitude of the area under the plasma concentration-time curve ratio (AUCR) based on reversible inhibition of CYP3A4 and organic anion transporting polypeptide 1B1 (OATP1B1). To reconcile this disconnect between predicted and clinically observed AUCR, telithromycin was evaluated as a time-dependent inhibitor of CYP3A4 in vitro, as well as an inhibitor of OATP1B1. Telithromycin inhibited OATP1B1-mediated [3H]-estradiol 17β-d-glucuronide (0.02 µM) transport with a mean IC50 of 12.0 ± 1.45 µM and was determined by IC50 shift and kinetic analyses to be a competitive reversible inhibitor of CYP3A4-mediated midazolam1- hydroxylation with a mean absolute inhibition constant (Ki) value of 3.65 ± 0.531 µM. The 2.83-fold shift in IC50 (10.4-3.68 µM) after a 30-minute metabolic preincubation confirmed telithromycin as a time-dependent inhibitor of CYP3A4; the mean inhibitor concentration that causes half-maximal inactivation of enzyme (KI) and maximal rate of inactivation of enzyme (kinact) values determined for inactivation were 1.05 ± 0.226 µM and 0.02772 ± 0.00272 min-1, respectively. After the integration of an enzyme time-dependent inhibition component into the previous mechanistic static model using the in vitro inhibitory kinetic parameters determined above, the newly predicted simvastatin acid AUCR (10.8 or 5.4) resulting from perturbation of its critical disposition pathways matched the clinically observed AUCR (10.8 or 4.3) after coadministration, or staggered administration, with telithromycin, respectively. These results indicate the time-dependent inhibition of CYP3A4 by telithromycin as the primary driver underlying its clinical DDI with simvastatin acid.
Related Concept Videos
 01:17
Drug Dependence
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:14
Time Course of Drug Effect
 01:16
Methods for Studying Drug Absorption: In vitro
The diffusion cell method uses a two-compartment cell, including a donor compartment with the drug solution, which simulates the environment where the drug is applied, and a receptor compartment with a buffer solution, which simulates the environment...
 01:29
Drug-Receptor Interactions
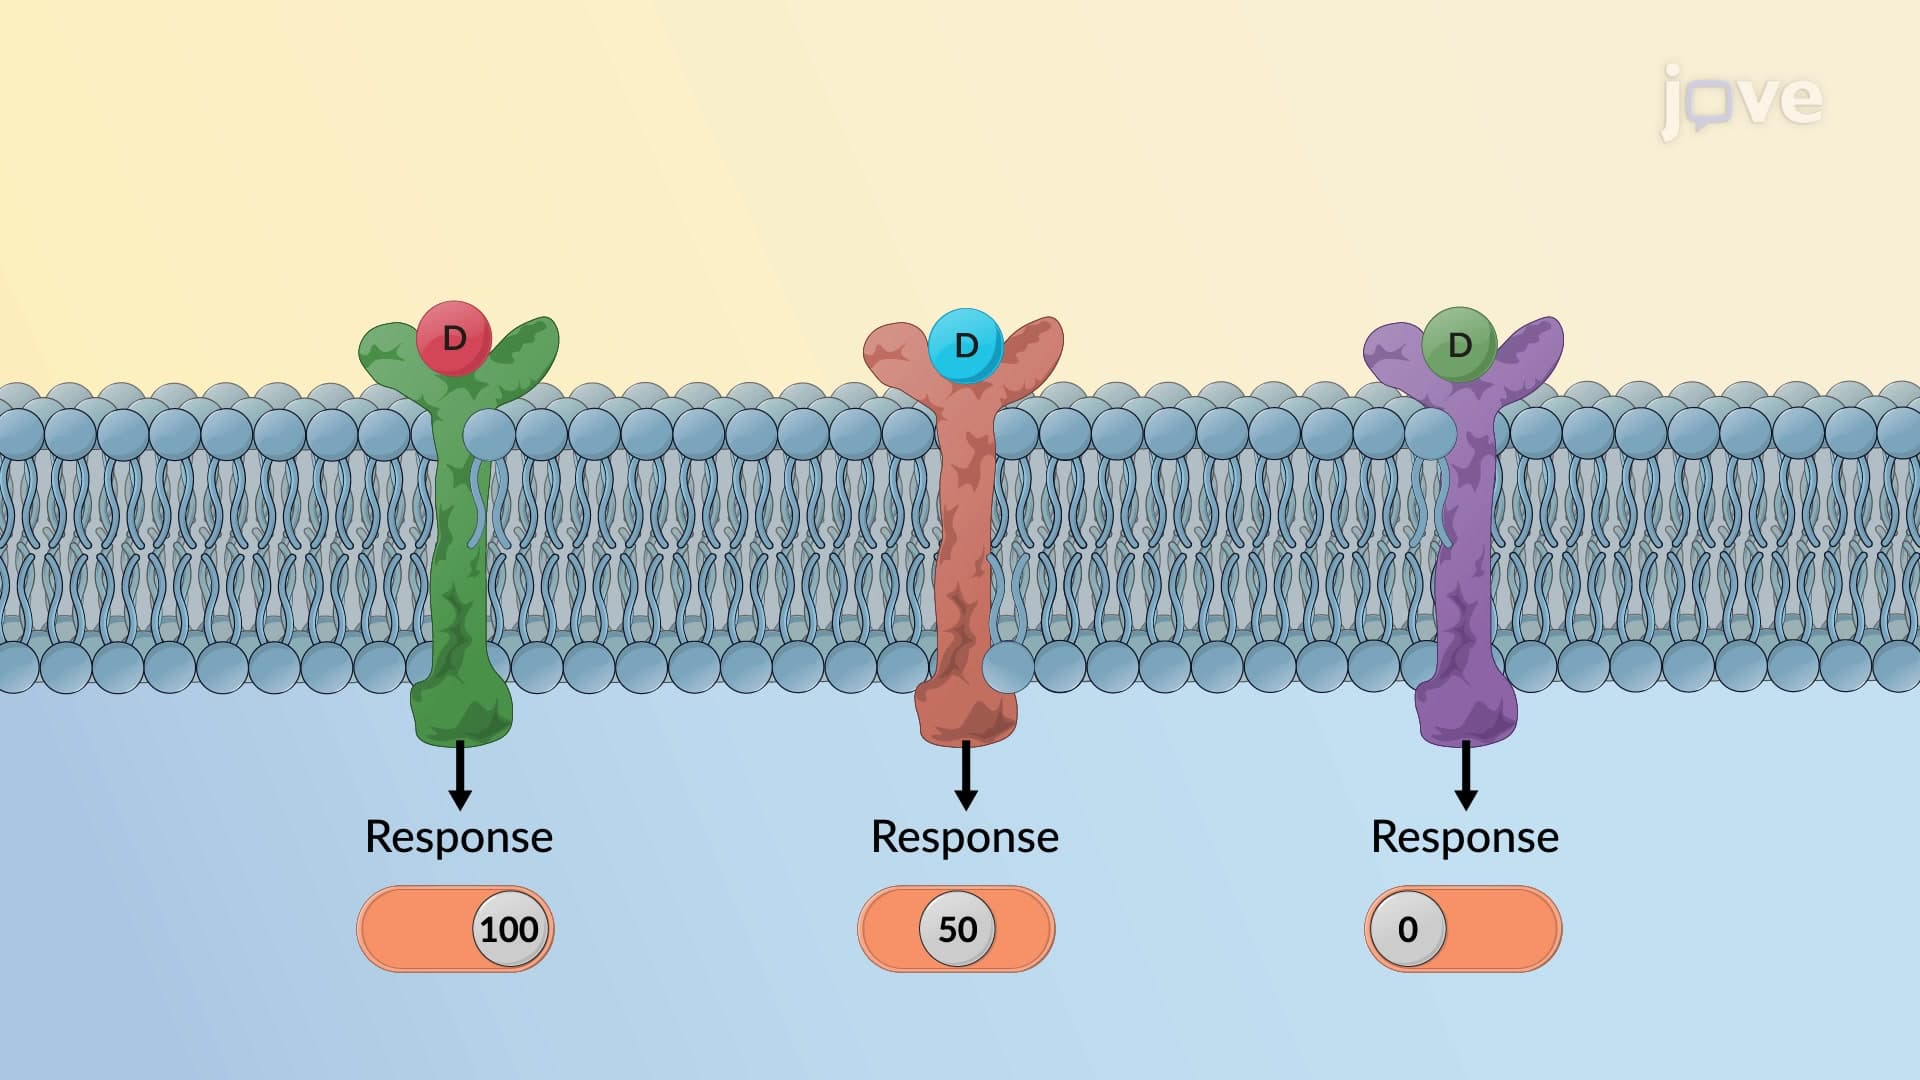
Several parameters, such as the drug's affinity for its receptor and its efficacy, which is its ability to activate the receptor, determine the drug's effect on the tissue....
 01:26
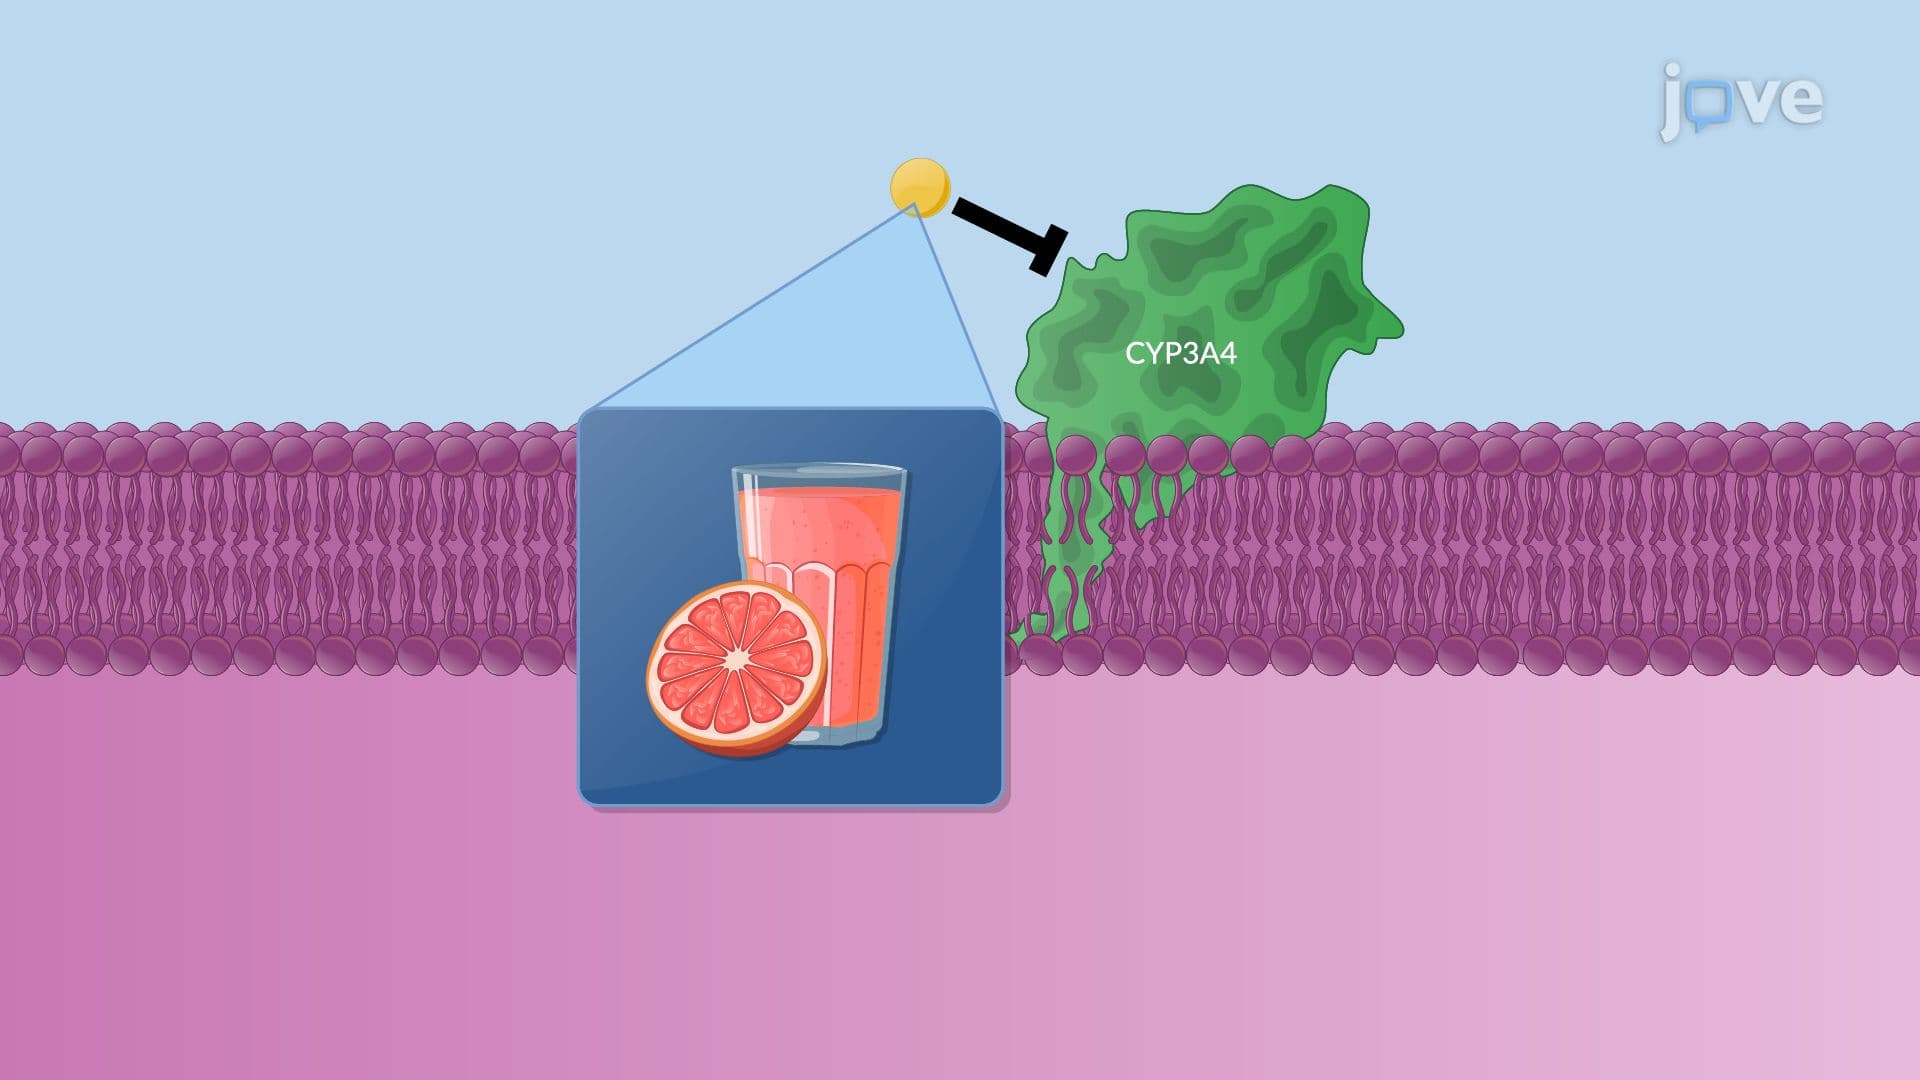
Pharmacokinetics: Drug–Food and Drug–Viral Interactions