Related Experiment Video
Updated: Feb 3, 2026
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
Predicting Relative Binding Affinity Using Nonequilibrium QM/MM Simulations
Meiting Wang1,2, Ye Mei1,3, Ulf Ryde2
1State Key Laboratory of Precision Spectroscopy, School of Physics and Materials Science , East China Normal University , Shanghai 200062 , China.

Abstract:
Calculating binding free energies with quantum-mechanical (QM) methods is notoriously time-consuming. In this work, we studied whether such calculations can be accelerated by using nonequilibrium (NE) molecular dynamics simulations employing Jarzynski's equality. We studied the binding of nine cyclic carboxylate ligands to the octa-acid deep-cavity host from the SAMPL4 challenge with the reference potential approach. The binding free energies were first calculated at the molecular mechanics (MM) level with free energy perturbation using the generalized Amber force field with restrained electrostatic potential charges for the host and the ligands. Then the free energy corrections for going from the MM Hamiltonian to a hybrid QM/MM Hamiltonian were estimated by averaging over many short NE molecular dynamics simulations. In the QM/MM calculations, the ligand was described at the semiempirical PM6-DH+ level. We show that this approach yields MM → QM/MM free energy corrections that agree with those from other approaches within statistical uncertainties. The desired precision can be obtained by running a proper number of independent NE simulations. For the systems studied in this work, a total simulation length of 20 ps was appropriate for most of the ligands, and 36-324 simulations were necessary in order to reach a precision of 0.3 kJ/mol.
Related Concept Videos
 02:18
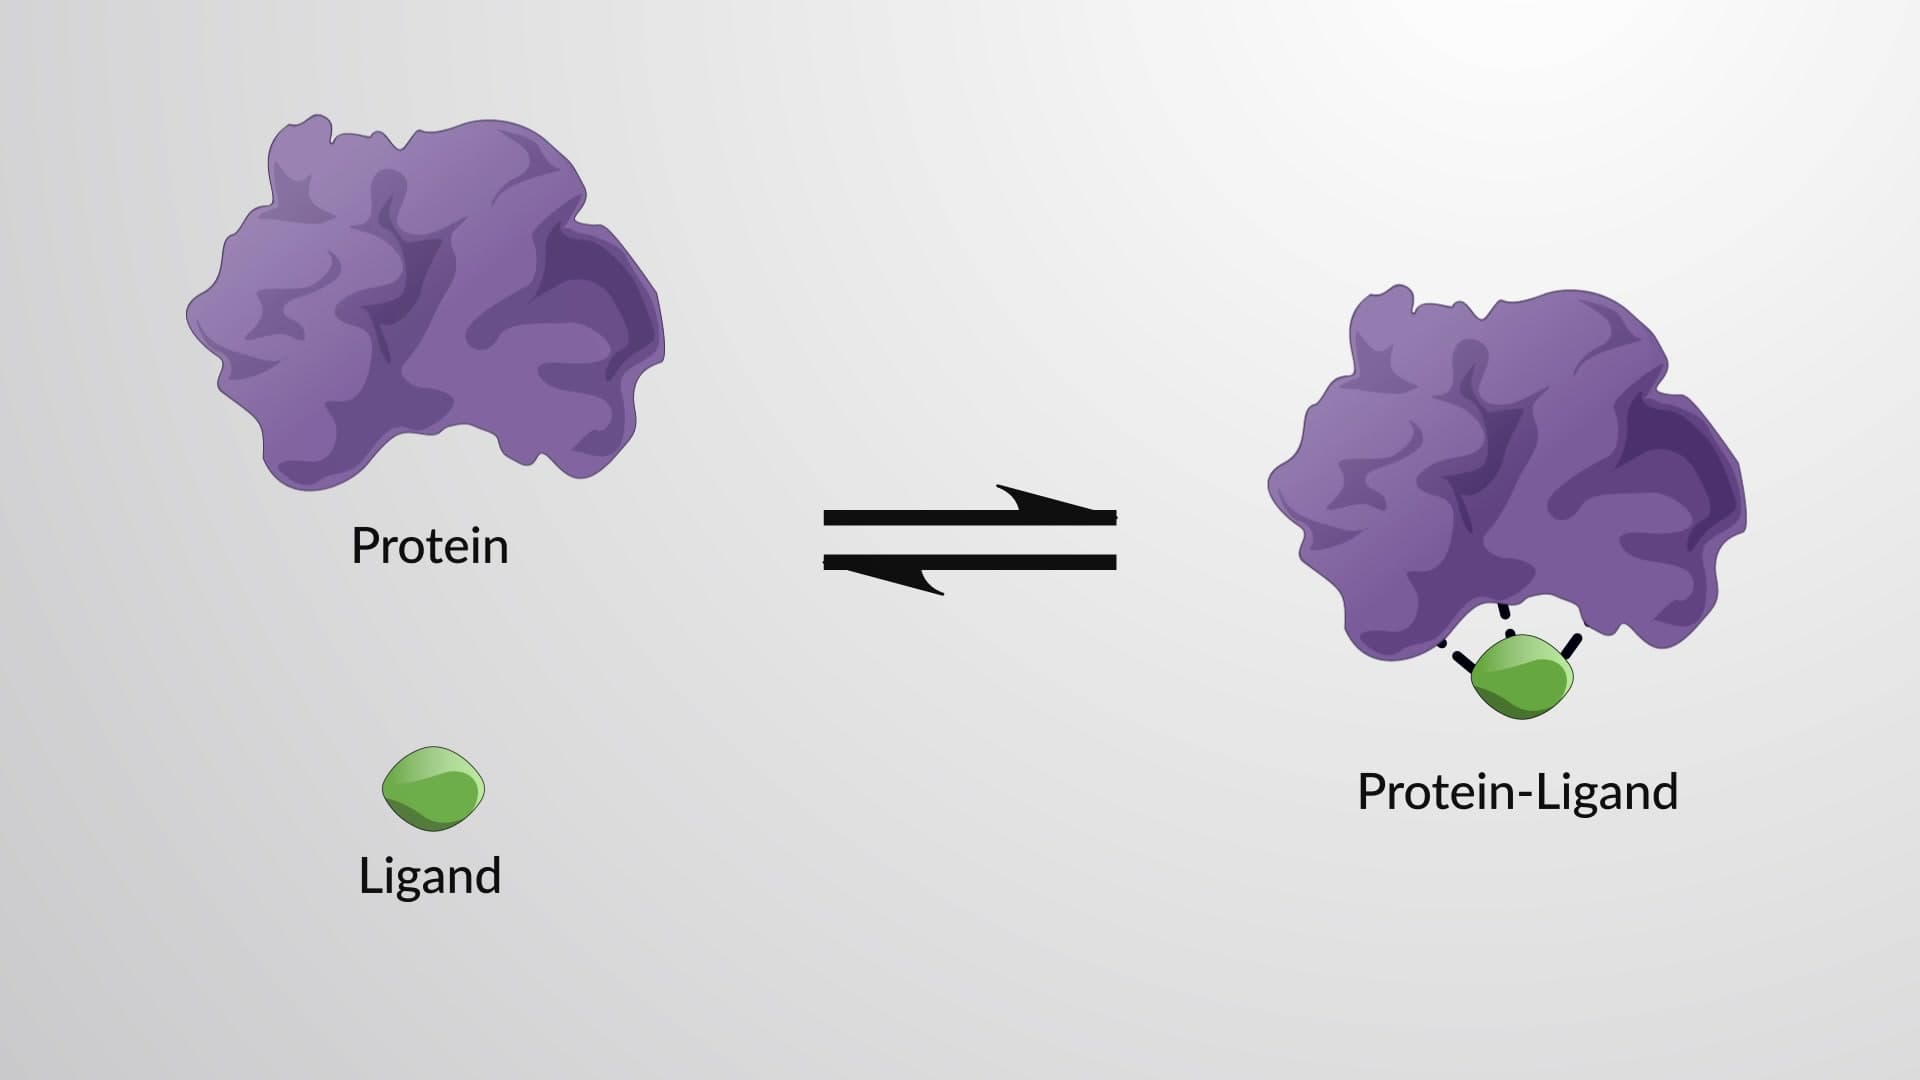
The Equilibrium Binding Constant and Binding Strength
 01:41
Affinity and Avidity
 03:07
Electron Affinity
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:27
Predicting Molecular Geometry
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...