Related Experiment Video
Updated: Feb 3, 2026
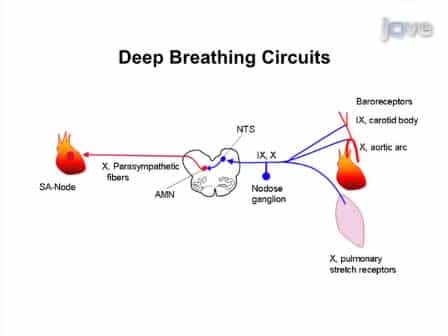

Quantitative Autonomic Testing
Published on: July 19, 2011
IGHMBP2 mutation associated with organ-specific autonomic dysfunction
Pedro J Tomaselli1, Alejandro Horga2, Alexander M Rossor2
1MRC Centre for Neuromuscular Diseases, National Hospital for Neurology and Neurosurgery and UCL Institute of Neurology, Queen Square, London WC1N 3AR, UK; Department of Neuromuscular Disorders, Clinical Hospital of Ribeirão Preto, University of São Paulo, Ribeirão Preto, SP 14640-900, Brazil.

Insights
IGHMBP2 gene mutations cause severe neuromuscular disorders. This case highlights a novel mutation leading to progressive muscle weakness, respiratory failure, and gastrointestinal autonomic dysfunction.
Area of Science:
- Genetics and Molecular Biology
- Neurology
- Rare Diseases
Background:
- Biallelic mutations in the IGHMBP2 gene are linked to distinct neuromuscular disorders, including spinal muscular atrophy with respiratory distress type 1 (SMARD1) and Charcot-Marie-Tooth disease type 2S (CMT2S).
- Understanding the full spectrum of IGHMBP2-related phenotypes is crucial for accurate diagnosis and management.
Observation:
- A patient presented with infantile-onset progressive muscle weakness and wasting in all limbs.
- Respiratory involvement emerged at age 9, necessitating continuous non-invasive ventilation, alongside severe gastrointestinal autonomic dysfunction.
- Late-stage neurophysiological studies showed absent sensory/motor responses and severe denervation in proximal upper limb muscles.
Findings:
- Targeted sequencing identified a novel homozygous missense variant (c.1325A>G; p.Tyr442Cys) in the IGHMBP2 gene.
- Sanger sequencing and co-segregation analysis confirmed the variant's novelty and parental carrier status.
Implications:
- This case expands the known clinical spectrum of IGHMBP2-related disorders.
- It underscores the potential for severe peripheral neuropathy and significant gastrointestinal autonomic dysfunction, possibly requiring parenteral nutrition, in patients with IGHMBP2 mutations.
Abstract:
Biallelic mutations in the IGHMBP2 have been associated with two distinct phenotypes: spinal muscular atrophy with respiratory distress type 1 (SMARD1) and CMT2S. We describe a patient who developed progressive muscle weakness and wasting in her upper and lower limbs from infancy. She developed respiratory involvement at age 9, eventually requiring 24-h non-invasive ventilation, and severe autonomic dysfunction restricted to the gastrointestinal tract. Neurophysiological studies at age 27 years revealed absent sensory and motor responses and severe chronic denervation changes in proximal muscles of the upper limbs. Targeted multigene panel sequencing detected a novel homozygous missense variant in the IGHMBP2 gene (c.1325A > G; p.Tyr442Cys). This variant was validated by Sanger sequencing and co-segregation analysis confirmed that both parents were asymptomatic heterozygous carriers. This case report confirms that IGHMBP2 related disorders can result in a severe peripheral neuropathy with gastrointestinal autonomic dysfunction requiring parenteral nutrition.
Related Concept Videos
 01:39
Mutations
 01:35
Mutations
Chromosomal Alterations Are Large-Scale Mutations
While point mutations are changes in a single nucleotide in...
 00:36
Viral Mutations
 01:09
Mutation, Gene Flow, and Genetic Drift
 01:22
Autonomic Nervous System
The ANS comprises two main divisions: the sympathetic and parasympathetic divisions. These divisions function antagonistically to maintain a dynamic...
 01:26
Autonomic Nervous System: Overview