Related Experiment Video
Updated: Feb 2, 2026

Next Generation Sequencing for the Detection of Actionable Mutations in Solid and Liquid Tumors
Published on: September 20, 2016
Gene characteristics predicting missense, nonsense and frameshift mutations in tumor samples
Ivan P Gorlov1, Claudio W Pikielny2, Hildreth R Frost2
1The Geisel School of Medicine, Department of Biomedical Data Science, Dartmouth College, HB7936, One Medical Center Dr., Dartmouth-Hitchcock Medical Center, Beirut, NH, 03756, Lebanon. ivan.p.gorlov@dartmouth.edu.

Background:
Because driver mutations provide selective advantage to the mutant clone, they tend to occur at a higher frequency in tumor samples compared to selectively neutral (passenger) mutations. However, mutation frequency alone is insufficient to identify cancer genes because mutability is influenced by many gene characteristics, such as size, nucleotide composition, etc. The goal of this study was to identify gene characteristics associated with the frequency of somatic mutations in the gene in tumor samples.
Results:
We used data on somatic mutations detected by genome wide screens from the Catalog of Somatic Mutations in Cancer (COSMIC). Gene size, nucleotide composition, expression level of the gene, relative replication time in the cell cycle, level of evolutionary conservation and other gene characteristics (totaling 11) were used as predictors of the number of somatic mutations. We applied stepwise multiple linear regression to predict the number of mutations per gene. Because missense, nonsense, and frameshift mutations are associated with different sets of gene characteristics, they were modeled separately. Gene characteristics explain 88% of the variation in the number of missense, 40% of nonsense, and 23% of frameshift mutations. Comparisons of the observed and expected numbers of mutations identified genes with a higher than expected number of mutations- positive outliers. Many of these are known driver genes. A number of novel candidate driver genes was also identified.
Conclusions:
By comparing the observed and predicted number of mutations in a gene, we have identified known cancer-associated genes as well as 111 novel cancer associated genes. We also showed that adding the number of silent mutations per gene reported by genome/exome wide screens across all cancer type (COSMIC data) as a predictor substantially exceeds predicting accuracy of the most popular cancer gene predicting tool - MutsigCV.
More Related Videos
 05:51
A Strategy to Identify de Novo Mutations in Common Disorders such as Autism and Schizophrenia
Published on: June 15, 2011
 13:34
A Combined 3D Tissue Engineered In Vitro/In Silico Lung Tumor Model for Predicting Drug Effectiveness in Specific Mutational Backgrounds
Published on: April 6, 2016
Related Concept Videos
 01:39
Mutations
 01:30
Point and Frameshift Mutations
 02:27
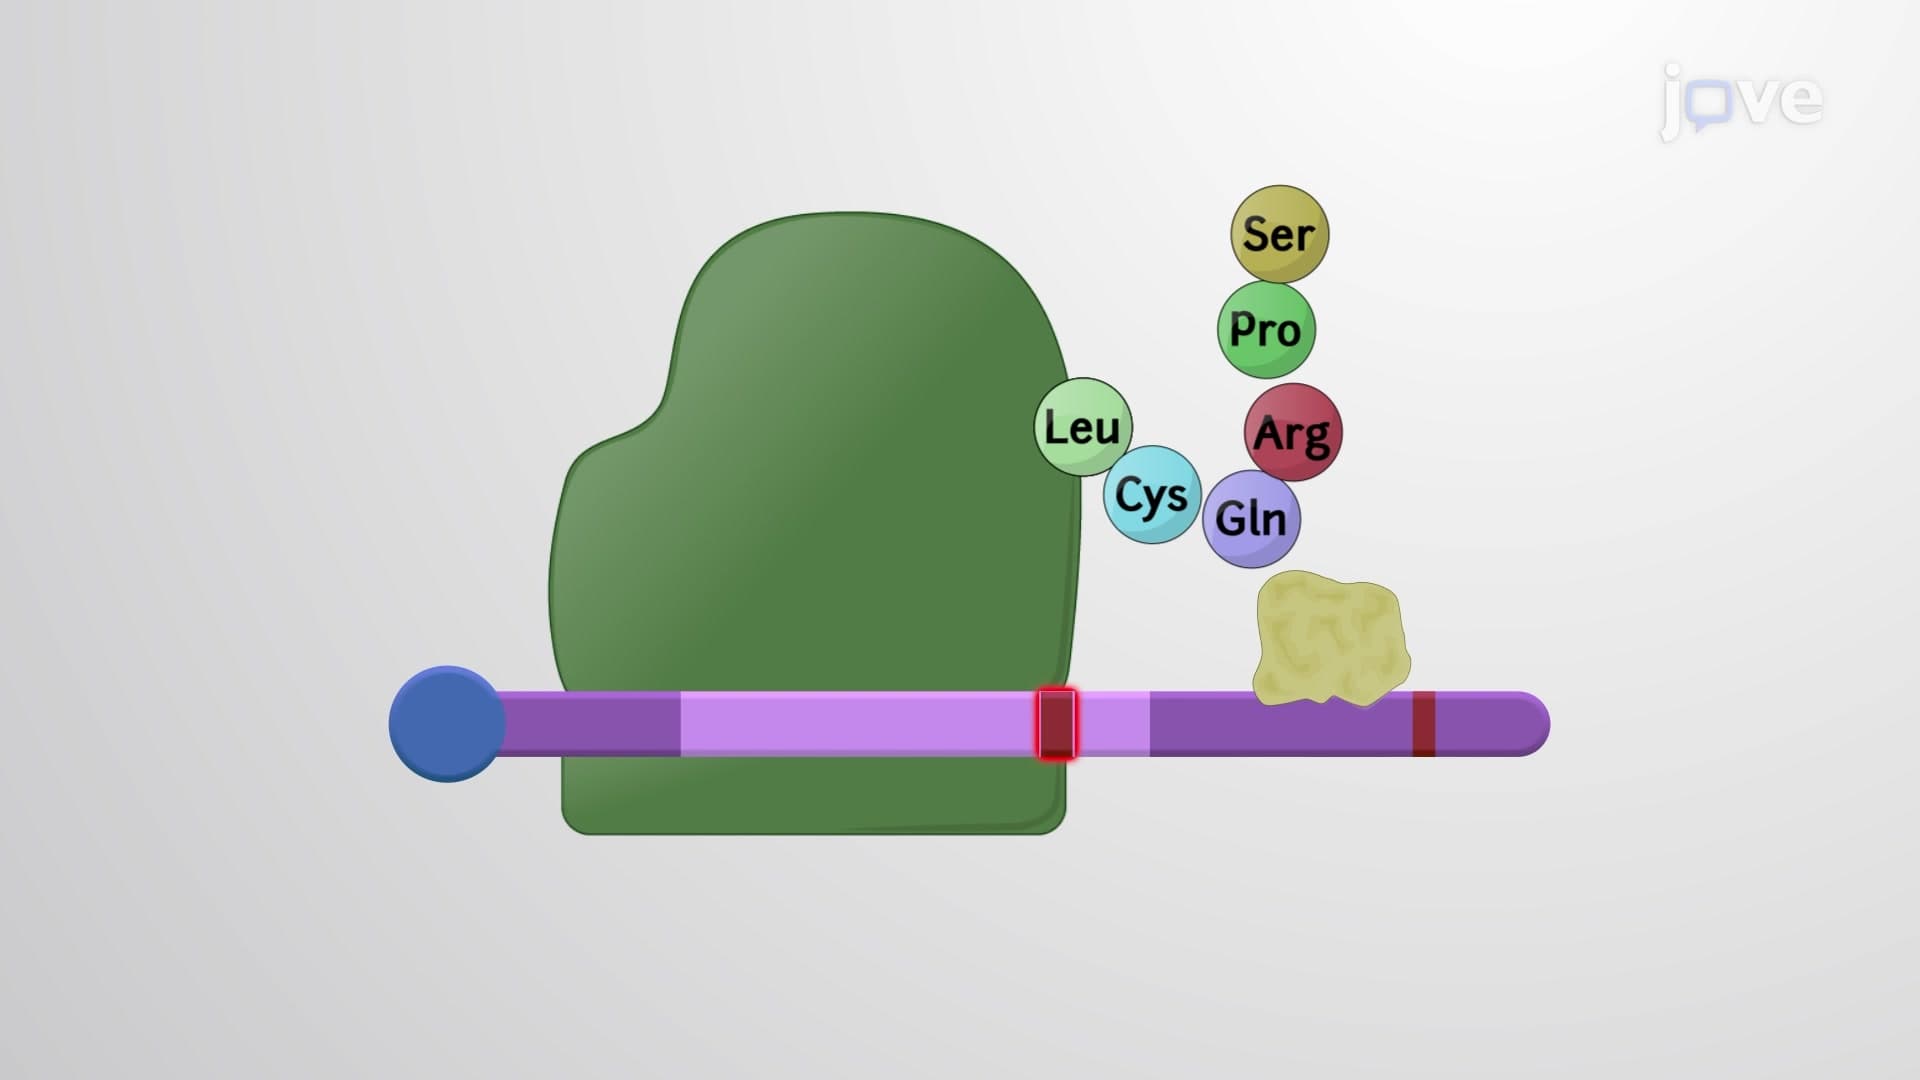
Nonsense-mediated mRNA Decay
Usually, Upf3 binds to an Exon Junction Complex (EJC) at mRNA splice sites. If a ribosome fully translates the mRNA,...
02:27Nonsense-mediated mRNA Decay
 01:09
Mutation, Gene Flow, and Genetic Drift
 00:36
Viral Mutations