Related Experiment Video
Updated: Feb 1, 2026

A Flow Cytometry-based Assay for Measuring Mitochondrial Membrane Potential in Cardiac Myocytes After Hypoxia/Reoxygenation
Published on: July 13, 2018
Restricting mitochondrial GRK2 post-ischemia confers cardioprotection by reducing myocyte death and maintaining
Priscila Y Sato1,2,3, J Kurt Chuprun1,2, Laurel A Grisanti1,2,4
1Center for Translational Medicine, Lewis Katz School of Medicine, Temple University, Philadelphia, PA 19140, USA.

Abstract:
Increased abundance of GRK2 [G protein-coupled receptor (GPCR) kinase 2] is associated with poor cardiac function in heart failure patients. In animal models, GRK2 contributes to the pathogenesis of heart failure after ischemia-reperfusion (IR) injury. In addition to its role in down-regulating activated GPCRs, GRK2 also localizes to mitochondria both basally and post-IR injury, where it regulates cellular metabolism. We previously showed that phosphorylation of GRK2 at Ser670 is essential for the translocation of GRK2 to the mitochondria of cardiomyocytes post-IR injury in vitro and that this localization promotes cell death. Here, we showed that mice with a S670A knock-in mutation in endogenous GRK2 showed reduced cardiomyocyte death and better cardiac function post-IR injury. Cultured GRK2-S670A knock-in cardiomyocytes subjected to IR in vitro showed enhanced glucose-mediated mitochondrial respiratory function that was partially due to maintenance of pyruvate dehydrogenase activity and improved glucose oxidation. Thus, we propose that mitochondrial GRK2 plays a detrimental role in cardiac glucose oxidation post-injury.
Insights
Mitochondrial G protein-coupled receptor kinase 2 (GRK2) worsens heart function after injury. Blocking its mitochondrial localization improves cardiac function and glucose metabolism post-ischemia-reperfusion injury.
Area of Science:
- Cardiology
- Molecular Biology
- Mitochondrial Biology
Background:
- Increased G protein-coupled receptor kinase 2 (GRK2) abundance correlates with poor cardiac function in heart failure.
- GRK2 contributes to heart failure pathogenesis following ischemia-reperfusion (IR) injury and regulates mitochondrial metabolism.
- GRK2's mitochondrial localization post-IR injury is crucial for cardiomyocyte death.
Purpose of the Study:
- To investigate the role of GRK2 mitochondrial localization in cardiac function and metabolism post-IR injury.
- To determine if inhibiting GRK2's mitochondrial translocation improves outcomes after cardiac injury.
Main Methods:
- Utilized a mouse model with a S670A knock-in mutation in GRK2, preventing phosphorylation and mitochondrial translocation.
- Assessed cardiac function and cardiomyocyte death in GRK2-S670A mutant mice post-IR injury.
- Examined mitochondrial respiratory function and glucose oxidation in cultured GRK2-S670A cardiomyocytes subjected to IR.
Main Results:
- Mice with the GRK2 S670A mutation exhibited reduced cardiomyocyte death and improved cardiac function after IR injury.
- GRK2-S670A cardiomyocytes showed enhanced glucose-mediated mitochondrial respiration and improved glucose oxidation post-IR.
- Maintenance of pyruvate dehydrogenase activity contributed to improved glucose oxidation in GRK2-S670A cardiomyocytes.
Conclusions:
- Mitochondrial GRK2 plays a detrimental role in cardiac glucose oxidation following injury.
- Inhibiting GRK2's mitochondrial localization represents a potential therapeutic strategy for heart failure post-IR injury.
- Targeting GRK2's role in mitochondrial metabolism may improve cardiac recovery after ischemic events.
Related Concept Videos
 01:26
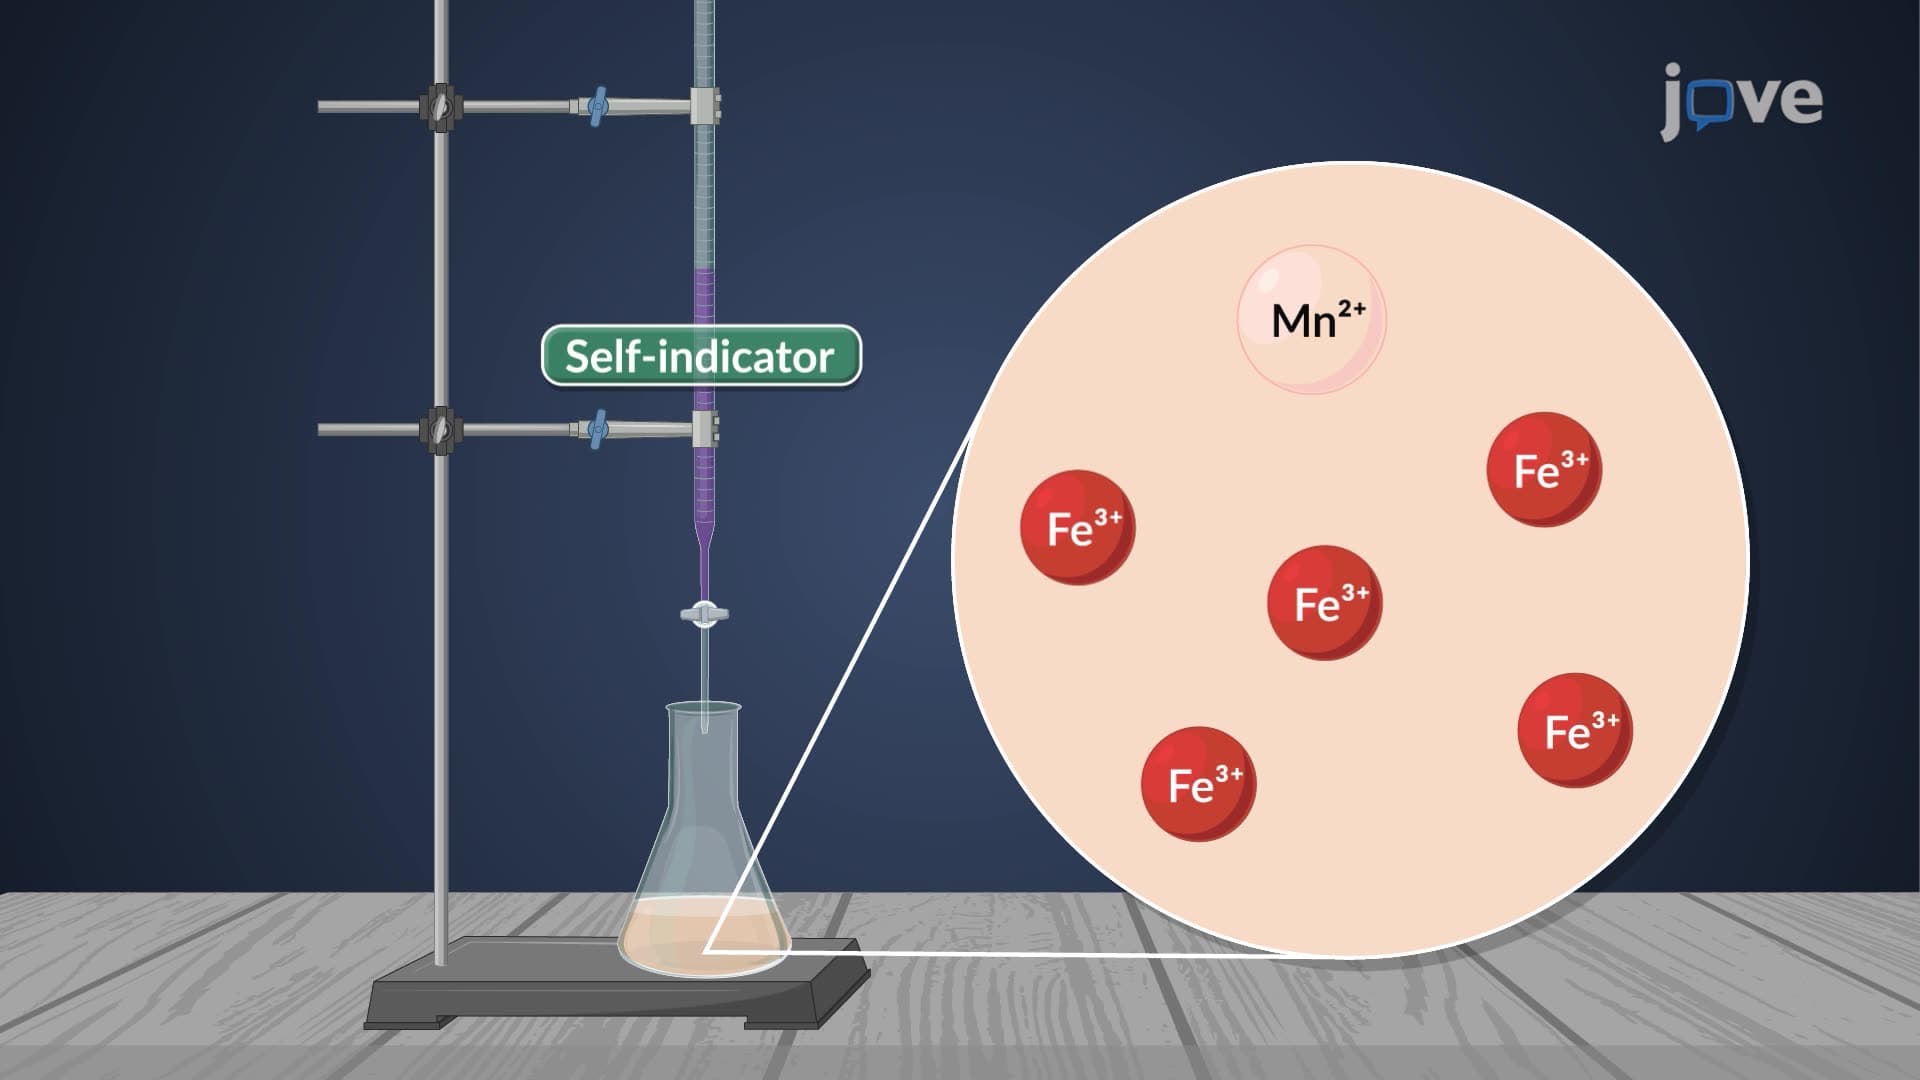
Redox Titration: Other Oxidizing and Reducing Agents
 03:14
Oxidation Numbers
 02:59
Animal Mitochondrial Genetics
 03:11
Oxidation-Reduction Reactions
 01:11
Restriction Enzymes
The host bacteria protect their own genomic DNA from these enzymes by methylating these sites. Some...
 02:19
Export of Mitochondrial and Chloroplast Genes