Related Experiment Video
Updated: Jan 31, 2026

A Method for Screening and Validation of Resistant Mutations Against Kinase Inhibitors
Published on: December 7, 2014
Ensemble-Based Replica Exchange Alchemical Free Energy Methods: The Effect of Protein Mutations on Inhibitor Binding
Agastya P Bhati1, Shunzhou Wan1, Peter V Coveney1
1Centre for Computational Science, Department of Chemistry , University College London , 20 Gordon Street , London , WC1H 0AJ , United Kingdom.

Abstract:
The accurate prediction of the binding affinity changes of drugs caused by protein mutations is a major goal in clinical personalized medicine. We have developed an ensemble-based free energy approach called thermodynamic integration with enhanced sampling (TIES), which yields accurate, precise, and reproducible binding affinities. TIES has been shown to perform well for predictions of free energy differences of congeneric ligands to a wide range of target proteins. We have recently introduced variants of TIES, which incorporate the enhanced sampling technique REST2 (replica exchange with solute tempering) and the free energy estimator MBAR (Bennett acceptance ratio). Here we further extend the TIES methodology to study relative binding affinities caused by protein mutations when bound to a ligand, a variant which we call TIES-PM. We apply TIES-PM to fibroblast growth factor receptor 3 (FGFR3) to investigate binding free energy changes upon protein mutations. The results show that TIES-PM with REST2 successfully captures a large conformational change and generates correct free energy differences caused by a gatekeeper mutation located in the binding pocket. Simulations without REST2 fail to overcome the energy barrier between the conformations, and hence the results are highly sensitive to the initial structures. We also discuss situations where REST2 does not improve the accuracy of predictions.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 10:33
Development of Inhibitors of Protein-protein Interactions through REPLACE: Application to the Design and Development Non-ATP Competitive CDK Inhibitors
Published on: October 26, 2015
Related Concept Videos
 02:13
Nuclear Binding Energy
 01:22
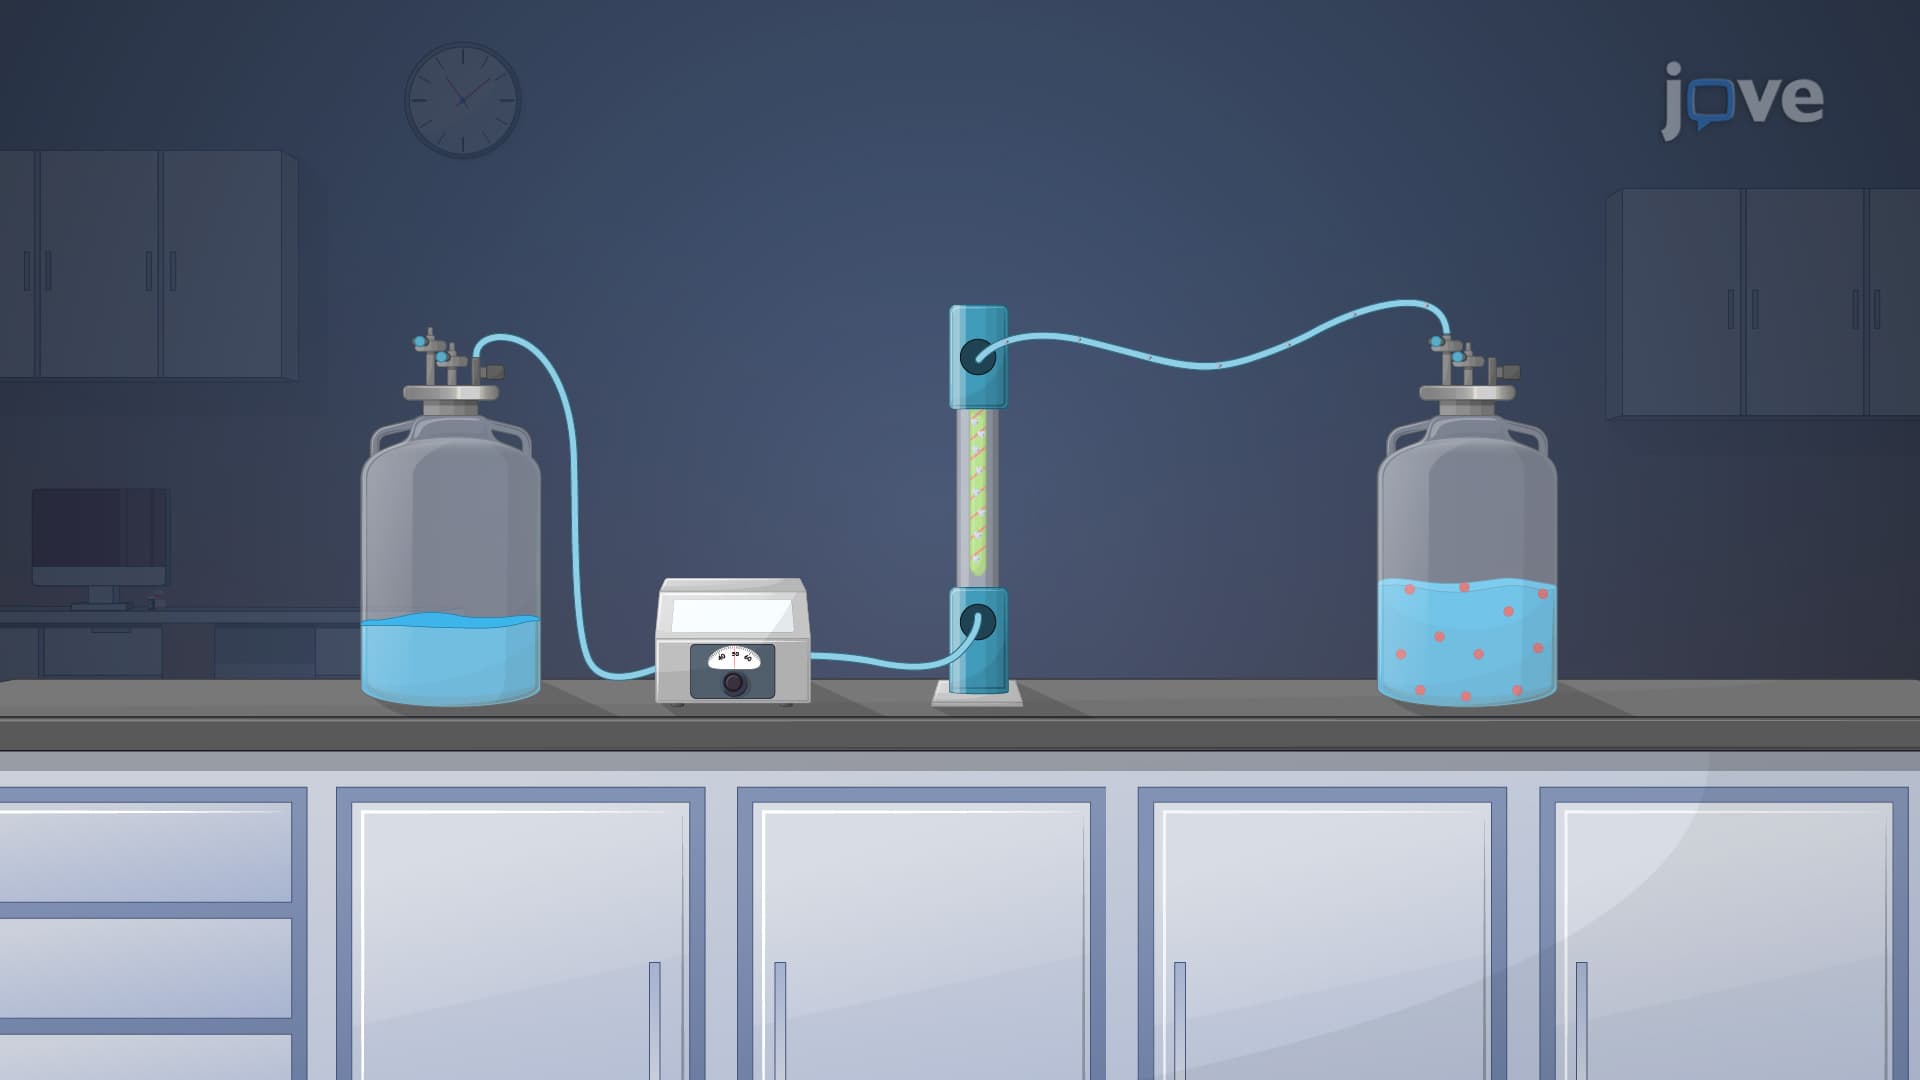
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:39
Mutations
 01:52
Eukaryotic Transcription Inhibitors
Eukaryotic transcription inhibitors usually contain two distinct domains, a...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:20
Gas Exchange and Transport