Related Experiment Video
Updated: Jan 31, 2026

Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Gas-Phase Ozone Reactions with a Structurally Diverse Set of Molecules: Barrier Heights and Reaction Energies
Daniela Trogolo1, J Samuel Arey1,2, Peter R Tentscher2
1School of Architecture, Civil and Environmental Engineering , École Polytechnique Fédérale de Lausanne , 1015 Lausanne , Switzerland.

Abstract:
Reactions with ozone transform organic and inorganic molecules in water treatment systems as well as in atmospheric chemistry, either in the aqueous phase, at gas/particle interfaces, or in the gas phase. Computed thermokinetic data can be used to estimate the reactivities of molecules toward ozone in cases where no experimental data are available. Although the gas-phase reactivity of olefins with ozone has been characterized extensively in the literature, this is not the case for the richer chemistry of ozone with polar molecules, which occurs in the aqueous phase or in microhydrated environments. Here, we selected a number of model reactions with small molecules (ethene, ethyne, hydrogen cyanide, hydrogen chloride, ammonia, bromide, and trimethylamine) to study the accuracy of different quantum chemical methods for describing the reactivities of these molecules with ozone. We calculated benchmark electronic energies of gas-phase reactions of these systems with single-reference coupled cluster (CC) theory. These benchmark results for the binding energy in the van der Waals complex, the energy of the transition structure, and the reaction energy were estimated to be accurate within 1-2 kcal mol-1. Singlet oxygen (1O2) is a common product of ozone reactions. Coupled cluster calculations with up to perturbative quadruples (CCSDT( Q)) were needed to obtain reaction energies accurate within 1 kcal mol-1 when this species was involved. In (micro)hydrated environments or at interfaces, coupled cluster methods are prohibitively expensive in most cases. We tested the suitability of some contemporary density functional theory (DFT) methods to reproduce the benchmark electronic energy differences. Range-separated functionals were found to be promising candidates to estimate forward barrier heights, with LC-ωPBE rivaling the accuracy of CCSD( T). For energies of reaction, however, DFT methods exhibited large systematic errors, depending on their fraction of orbital exchange. This was found to worsen when 1O2 is a product, and no safe recommendation can be given for DFT reaction energies in such cases.
More Related Videos
Related Concept Videos
 02:34
Kinetic Molecular Theory and Gas Laws Explain Properties of Gas Molecules
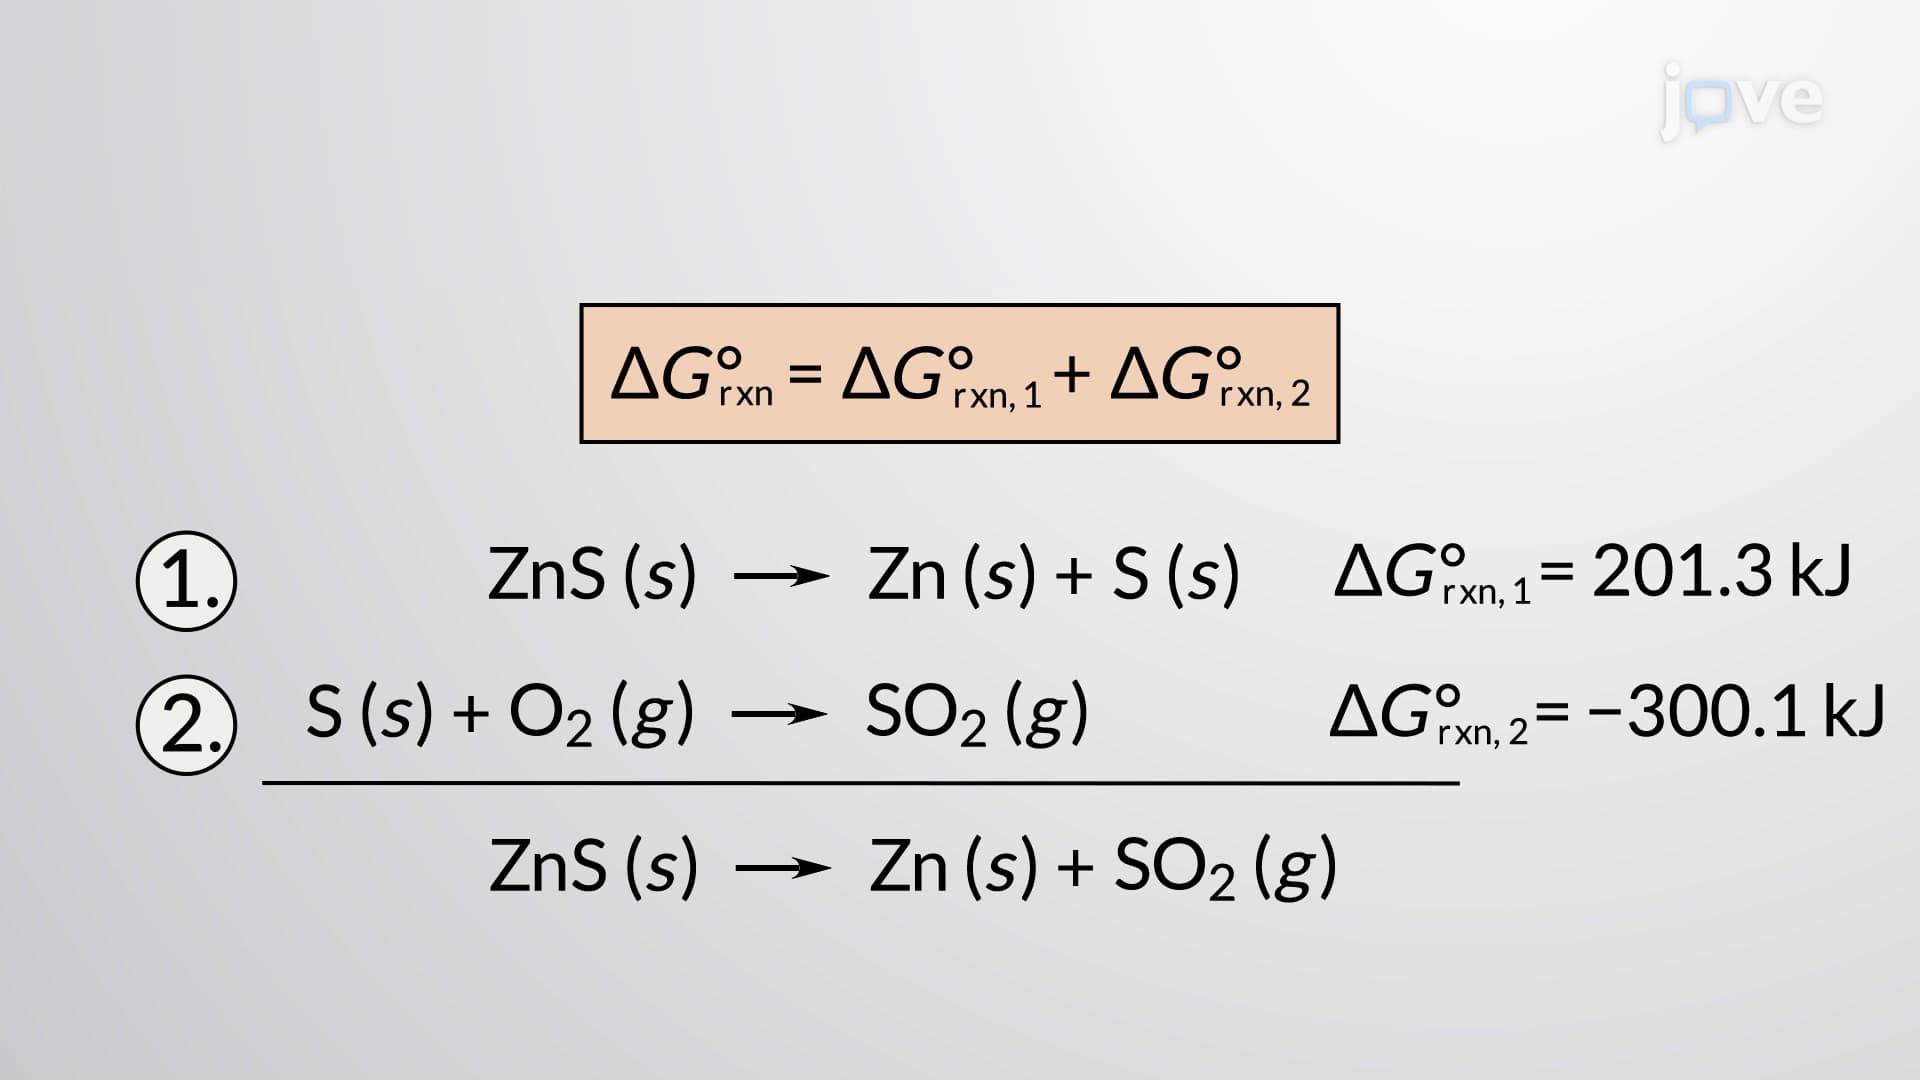 02:49
Calculating Standard Free Energy Changes
 01:15
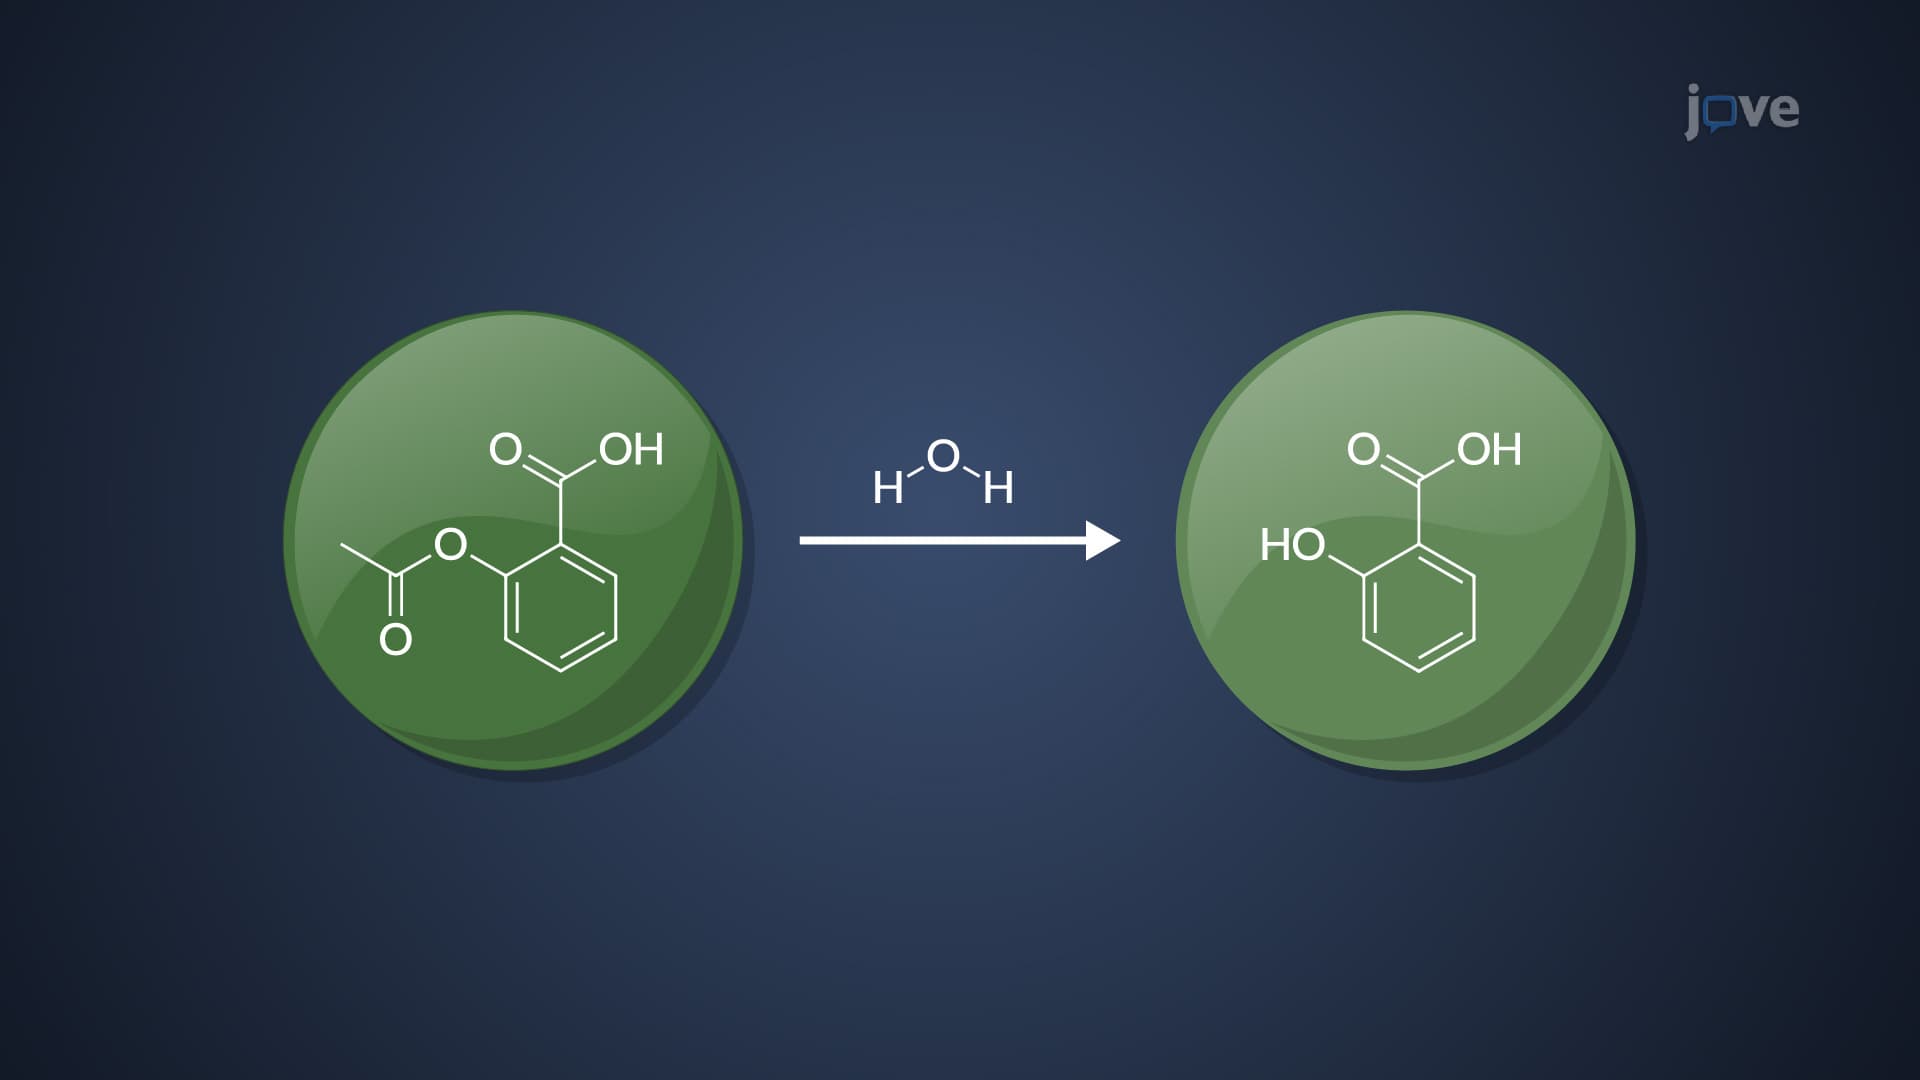
Phase I Reactions: Hydrolytic Reactions
An important hydrolytic reaction is ester hydrolysis. Ester bonds, often found in prodrugs, are broken down, increasing the solubility of drugs like aspirin and lidocaine for more straightforward elimination. Amide hydrolysis is another critical reaction, targeting amide bonds prevalent...
 01:24
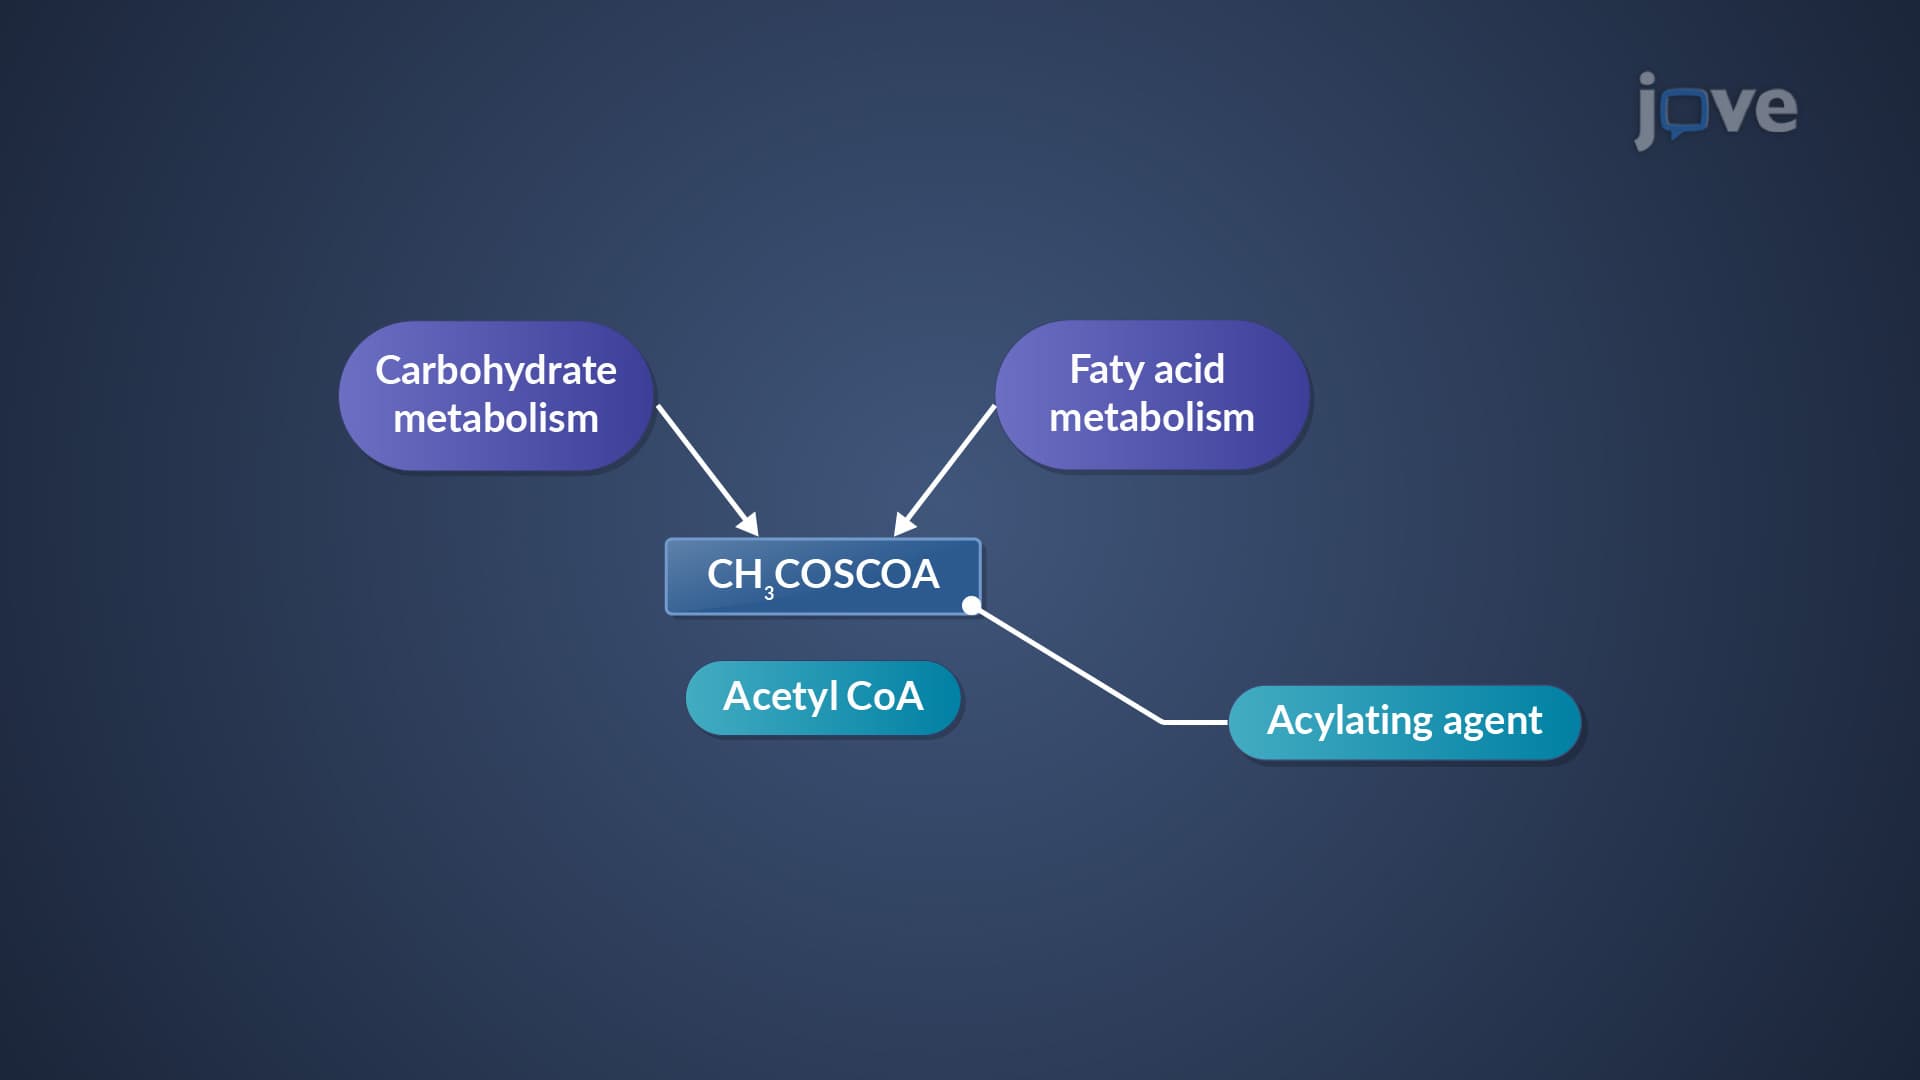
Phase II Reactions: Acetylation Reactions
The substrates for acetylation are typically drugs or their metabolites with an amino, sulfonamide, or hydrazine functional group. Acetylation can occur at several points in the drug molecule, including primary, secondary, and...
 01:17
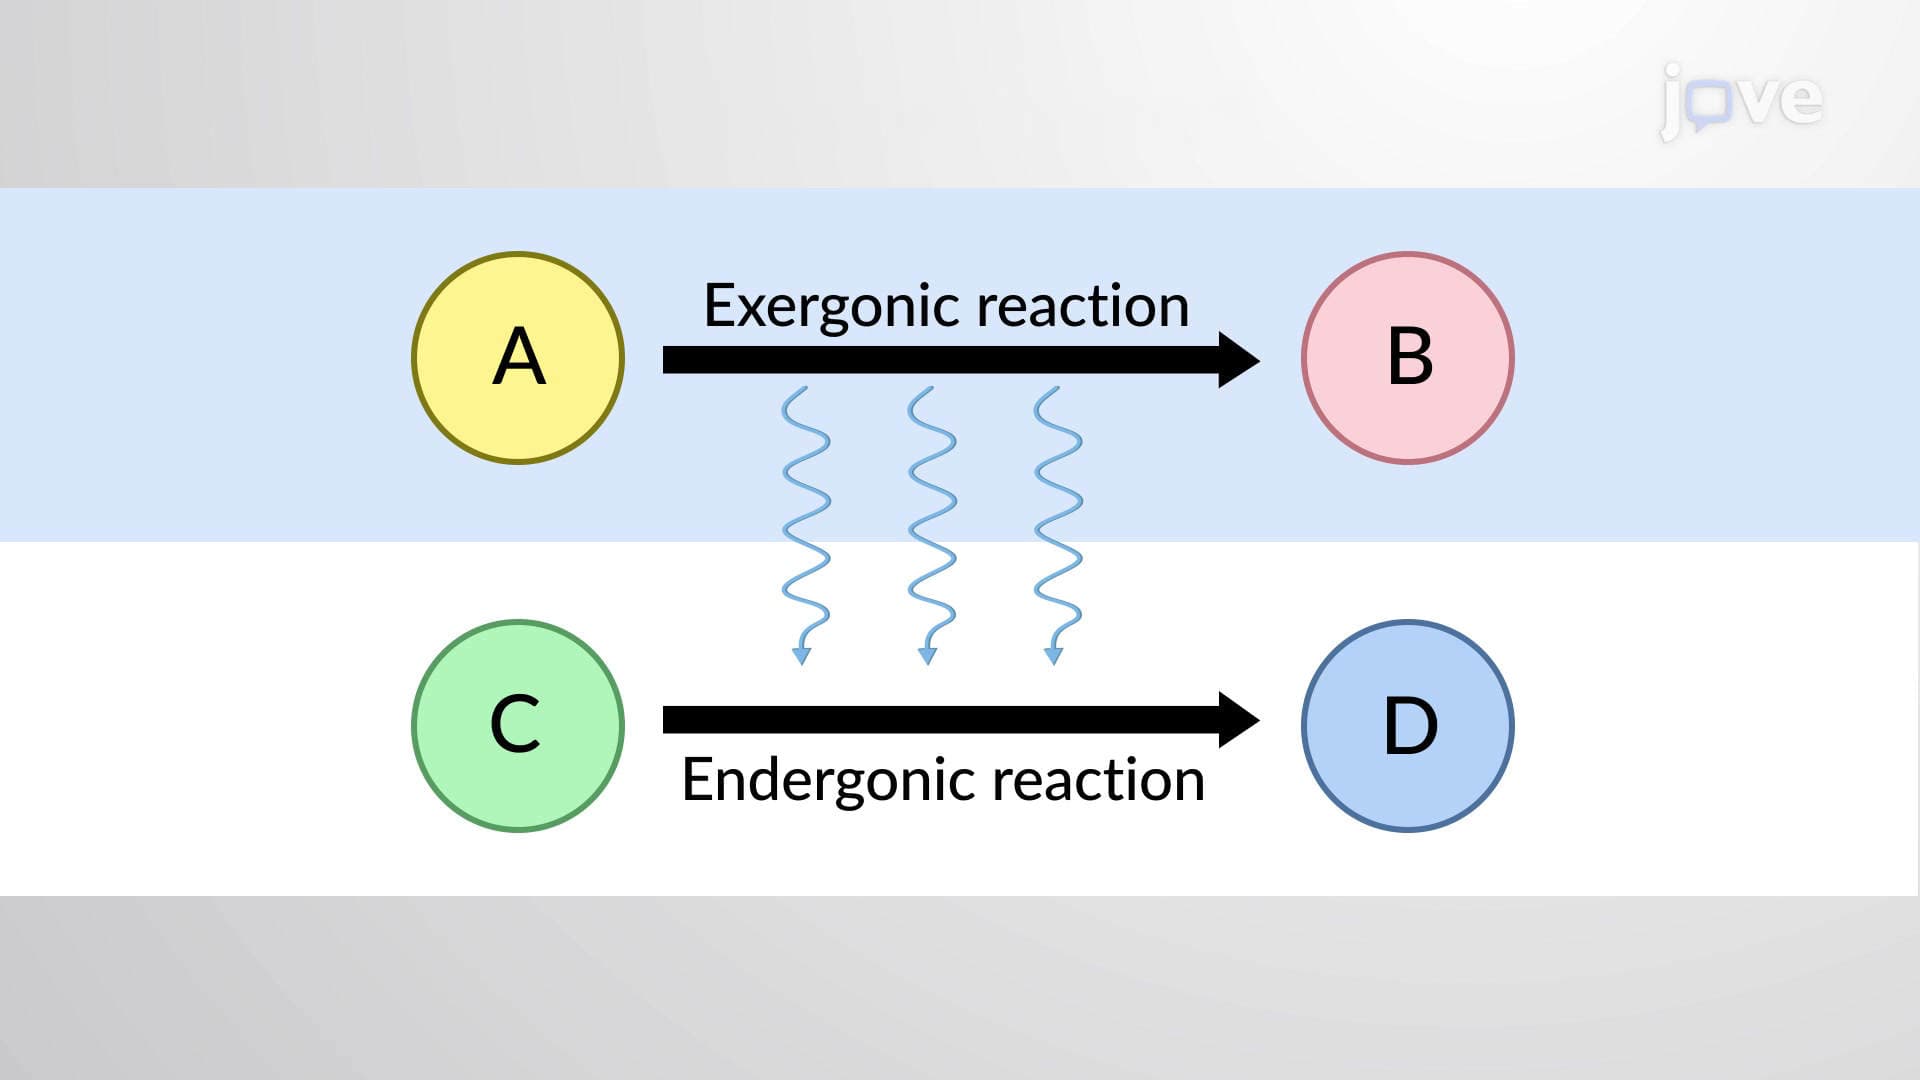
Coupled Reactions
Energy in adenosine triphosphate or ATP molecules is easily accessible to do work. ATP powers the majority of energy-requiring cellular reactions....
 01:27
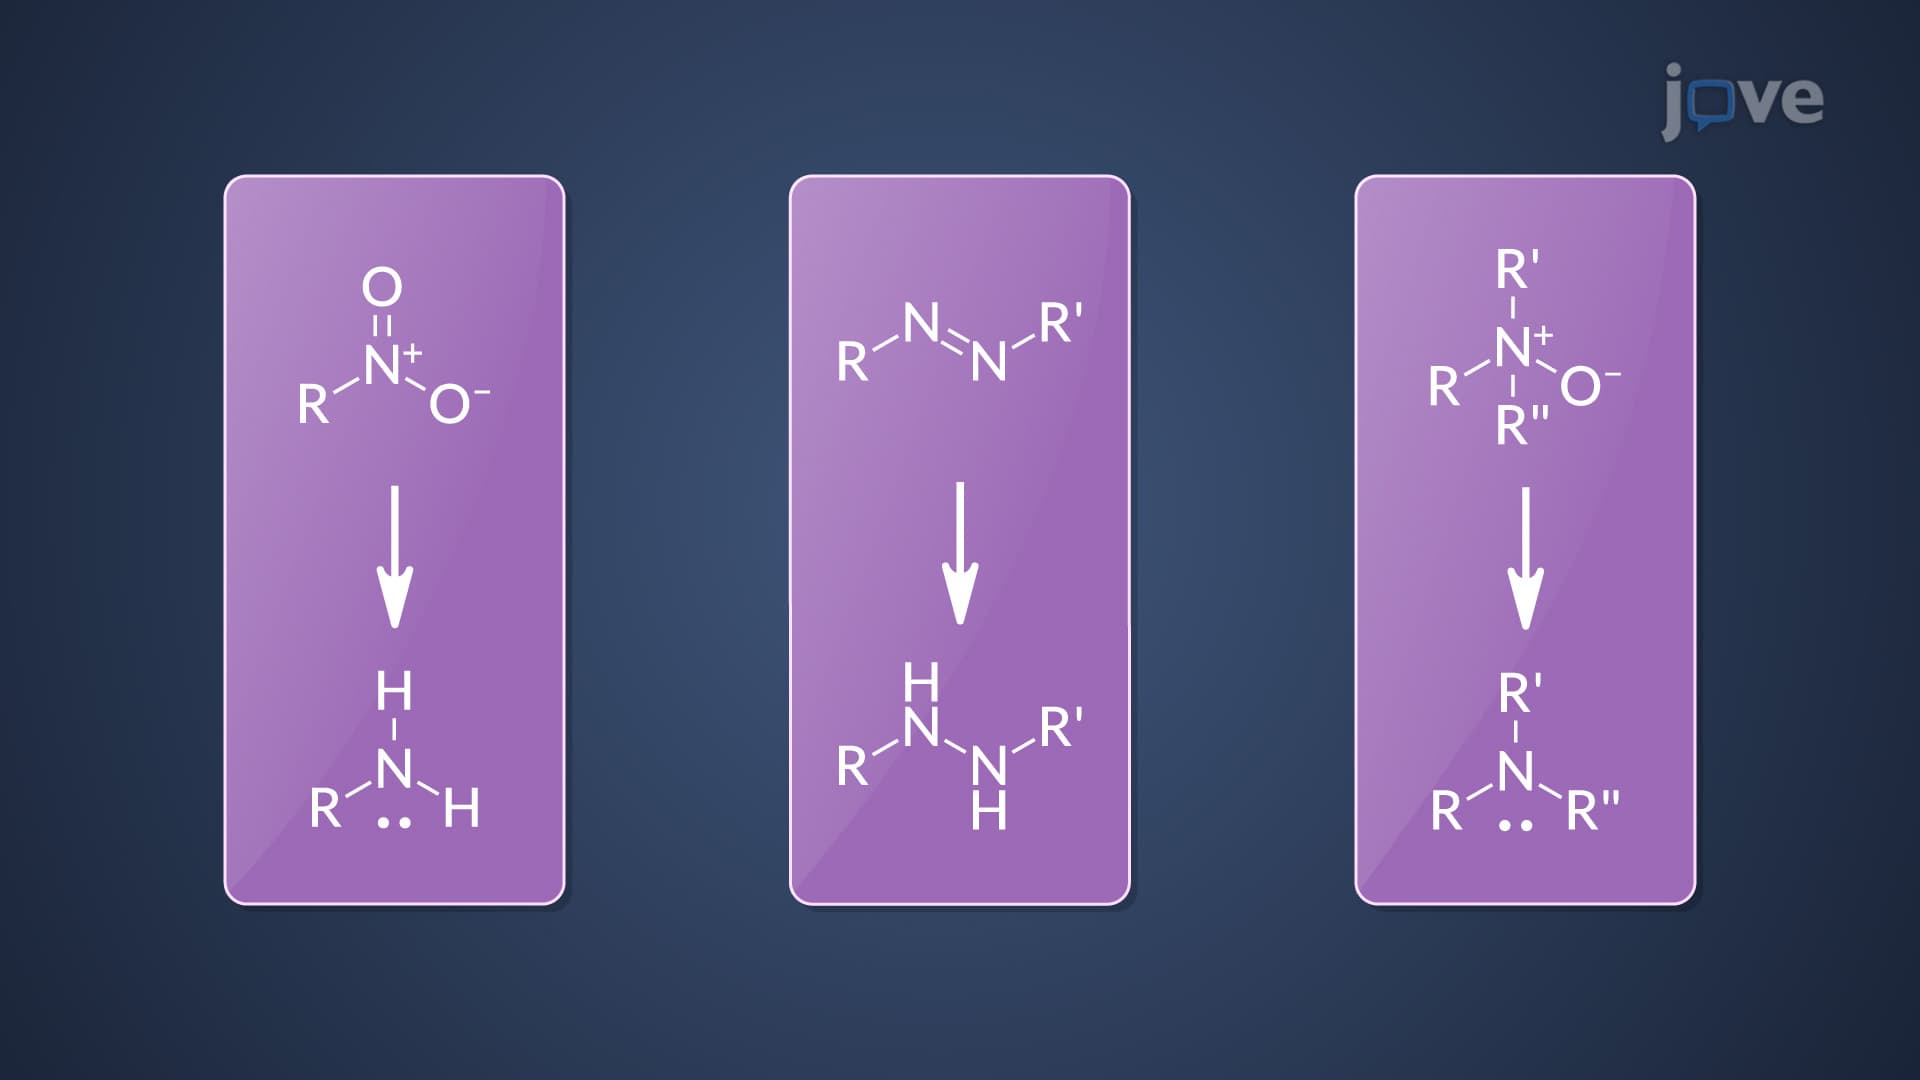
Phase I Reactions: Reductive Reactions