
Role of Conformational Flexibility in Monte Carlo Simulations of Many-Protein Systems
Bibhab Bandhu Majumdar1, Vera Prytkova2, Eric K Wong2
1Max-Planck-Institut für Kohlenforschung , Kaiser-Wilhelm-Platz 1 , D-45470 Mülheim an der Ruhr , Germany.

Abstract:
Efficient computational modeling of biological systems characterized by high concentrations of macromolecules often relies on rigid-body Brownian Dynamics or Monte Carlo (MC) simulations. However, the accuracy of rigid-body models is limited by the fixed conformation of the simulated biomolecules. Multi-conformation Monte Carlo (mcMC) simulations of protein solutions incorporate conformational flexibility via a conformational swap trial move within a predetermined library of discrete protein structures, thereby alleviating artifacts arising from the use of a single protein conformation. Here, we investigate the impact of the number of distinct protein structures in the conformational library and the extent of conformational sampling used in its generation on structural observables computed from simulations of hen egg white lysozyme (HEWL), human γD-Crystallin, and bovine γB-Crystallin solutions. We find that the importance of specific protocols for the construction of the protein structure library is strongly dependent on the nature of the simulated system.
More Related Videos

Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:20
Conformity
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 02:18
Intrinsically Disordered Proteins
 01:28
Role of Proteins in the Human Body
 02:20
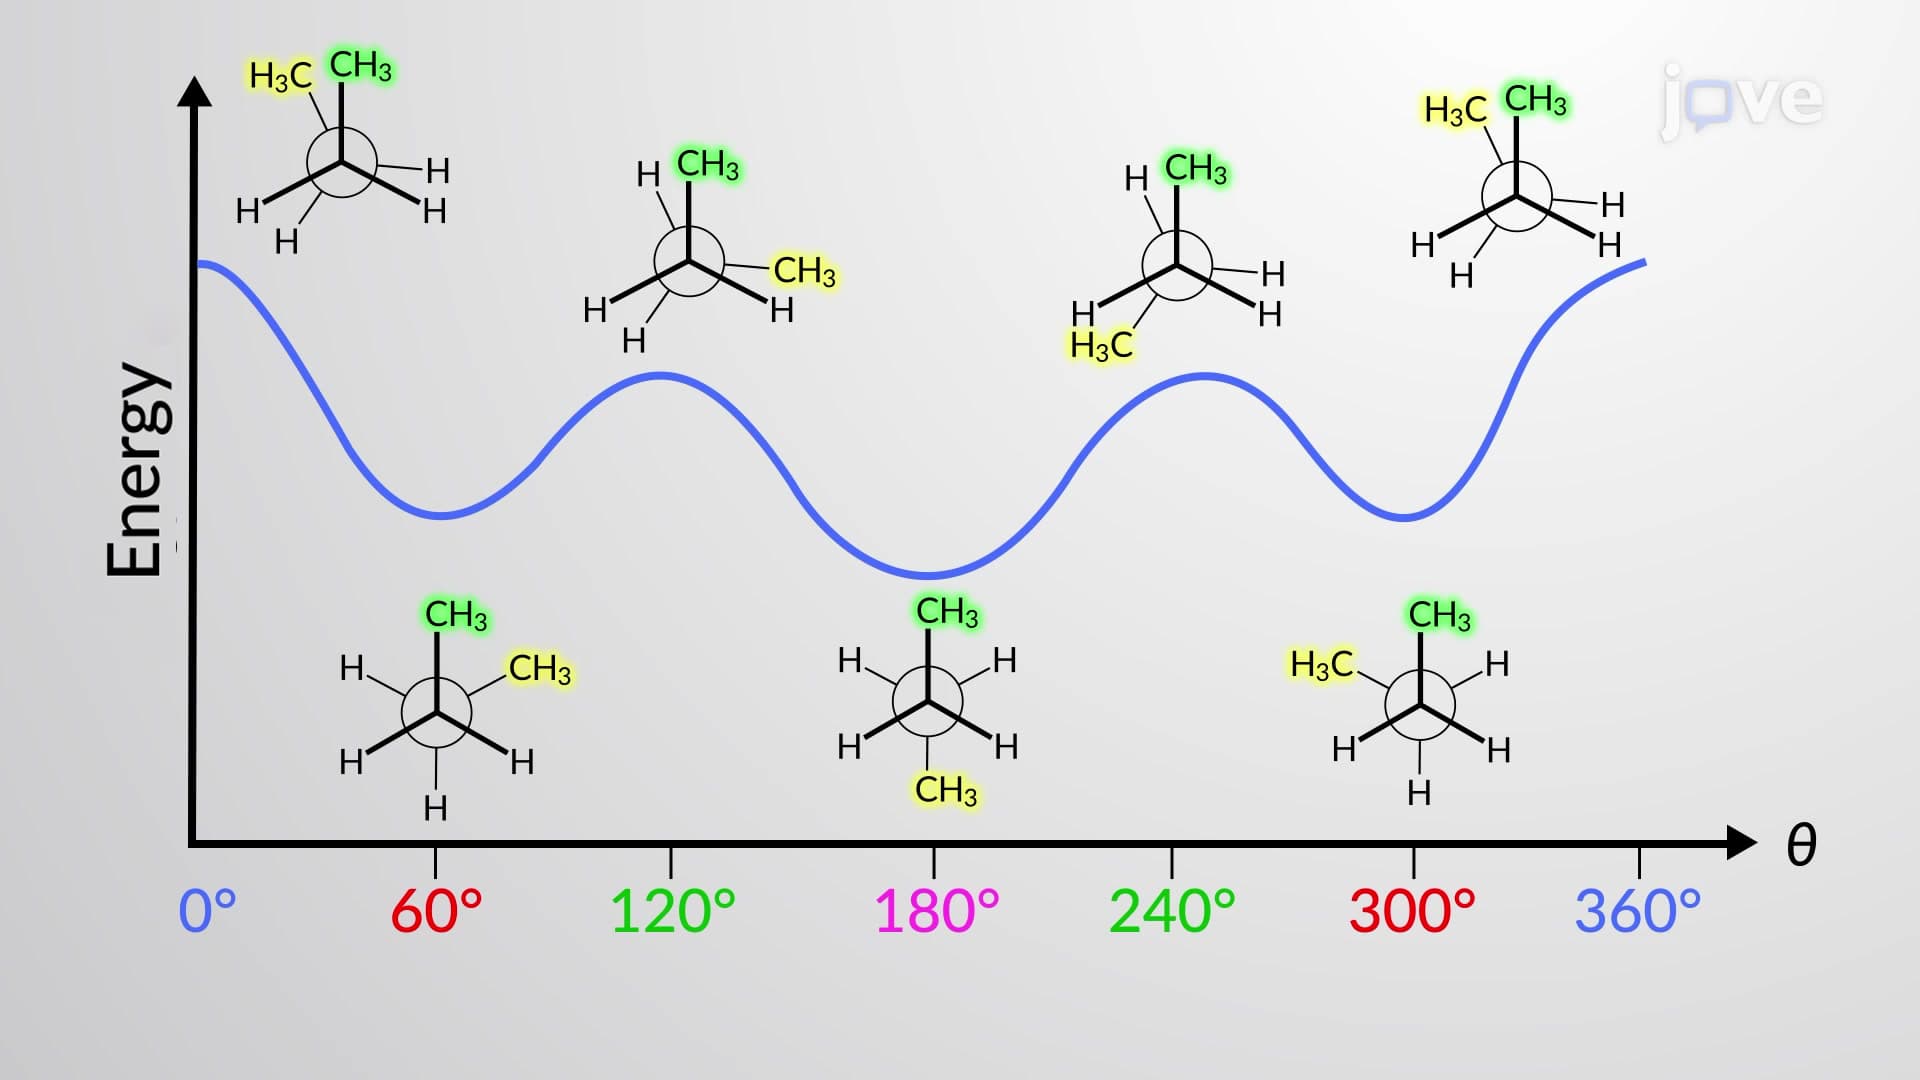
Conformations of Butane