Related Experiment Video
Updated: Jan 30, 2026
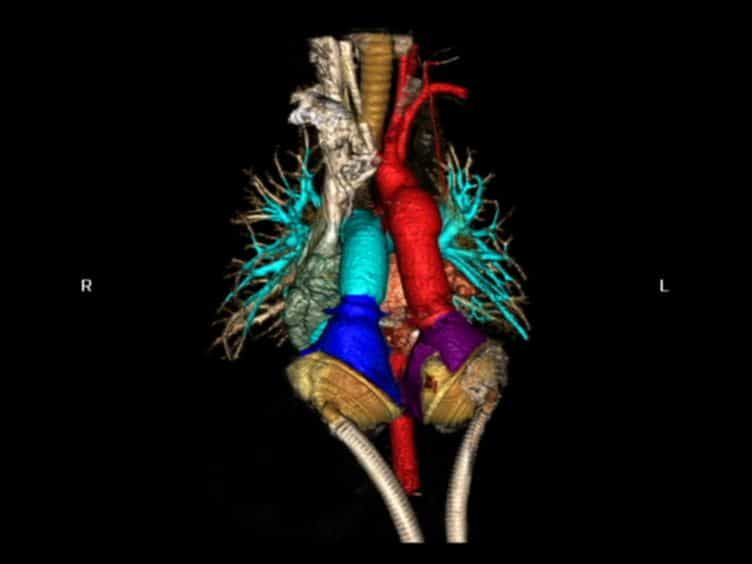

Implantation of Total Artificial Heart in Congenital Heart Disease
Published on: July 18, 2014
Leukoencephalopathy due to variants in GFPT1-associated congenital myasthenic syndrome
Guy Helman1, Suvasini Sharma1, Joanna Crawford1
1From the Murdoch Children's Research Institute (G.H., C.S.), Parkville, Melbourne; Institute for Molecular Bioscience (G.H., J.C., C.S.), the University of Queensland, Brisbane, Australia; Neurology Division (S.S., B.P., P.J.), Department of Pediatrics, Lady Hardinge Medical College, New Delhi, India; Division of Neurology (P.J.), Department of Pediatrics, the Hospital for Sick Children, Toronto, Canada; Data61 (S.J.B.), Commonwealth Scientific and Industrial Research Organisation, Brisbane, Australia; Hôpital Marin (J.A.U.), Centre Neuromusculaire, Filnemus, Hendaye, France; Department of Pathology (R.K.S.), G.B. Pant Hospital, New Delhi, India; Illumina, Inc. (R.J.T.), San Diego, CA; Department of Child Neurology (M.S.v.d.K.), Emma Children's Hospital, Amsterdam University Medical Centers, Vrije Universiteit Amsterdam and Amsterdam Neuroscience; and Department of Functional Genomics (M.S.v.d.K.), Neuroscience Campus Amsterdam, the Netherlands.

Objective:
To determine the molecular etiology of disease in 4 individuals from 2 unrelated families who presented with proximal muscle weakness and features suggestive of mitochondrial disease.
Methods:
Clinical information and neuroimaging were reviewed. Genome sequencing was performed on affected individuals and biological parents.
Results:
All affected individuals presented with muscle weakness and difficulty walking. In one family, both children had neonatal respiratory distress while the other family had 2 children with episodic deteriorations. In each family, muscle biopsy demonstrated ragged red fibers. MRI was suggestive of a mitochondrial leukoencephalopathy, with extensive deep cerebral white matter T2 hyperintense signal and selective involvement of the middle blade of the corpus callosum. Through genome sequencing, homozygous GFPT1 missense variants were identified in the affected individuals of each family. The variants detected (p.Arg14Leu and p.Thr151Lys) are absent from population databases and predicted to be damaging by in silico prediction tools. Following the genetic diagnosis, nerve conduction studies were performed and demonstrated a decremental response to repetitive nerve stimulation, confirming the diagnosis of myasthenia. Treatment with pyridostigmine was started in one family with favorable response.
Conclusions:
GFPT1 encodes a widely expressed protein that controls the flux of glucose into the hexosamine-biosynthesis pathway that produces precursors for glycosylation of proteins. GFPT1 variants and defects in other enzymes of this pathway have previously been associated with congenital myasthenia. These findings identify leukoencephalopathy as a previously unrecognized phenotype in GFPT1-related disease and suggest that mitochondrial dysfunction could contribute to this disorder.
Related Concept Videos
 02:30
Histone Variants at the Centromere
 01:24
Nephrotic Syndrome I : Introduction
 01:30
Acute Coronary Syndrome I: Introduction
 01:17
Irritable Bowel Syndrome I: Introduction
IBS is a chronic condition that can persist over a long period or recur frequently.
The pathogenesis of IBS involves a complex interplay of the following factors:
Altered...
 01:27
Restless Leg Syndrome and Night Terrors
The exact cause of RLS is not fully understood, but it is believed to involve dopamine, a neurotransmitter that helps regulate muscle movement. Imbalances in dopamine levels...
 01:26
Acute Coronary Syndrome V: Nursing Management