Related Experiment Video
Updated: Jan 30, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
A combined drug discovery strategy based on machine learning and molecular docking
Yanmin Zhang1, Yuchen Wang1, Weineng Zhou1
1Laboratory of Molecular Design and Drug Discovery, School of Science, China Pharmaceutical University, Nanjing, China.

Abstract:
Data mining methods based on machine learning play an increasingly important role in drug design and discovery. In the current work, eight machine learning methods including decision trees, k-Nearest neighbor, support vector machines, random forests, extremely randomized trees, AdaBoost, gradient boosting trees, and XGBoost were evaluated comprehensively through a case study of ACC inhibitor data sets. Internal and external data sets were employed for cross-validation of the eight machine learning methods. Results showed that the extremely randomized trees model performed best and was adopted as the first step of virtual screening. Together with structure-based virtual screening in the second step, this combined strategy obtained desirable results. This work indicates that the combination of machine learning methods with traditional structure-based virtual screening can effectively strengthen the ability in finding potential hits from large compound database for a given target.
Related Concept Videos
 01:26
Drug Discovery: Overview
 01:27
Combined Effects of Drugs: Synergism
Such synergistic combinations...
 01:30
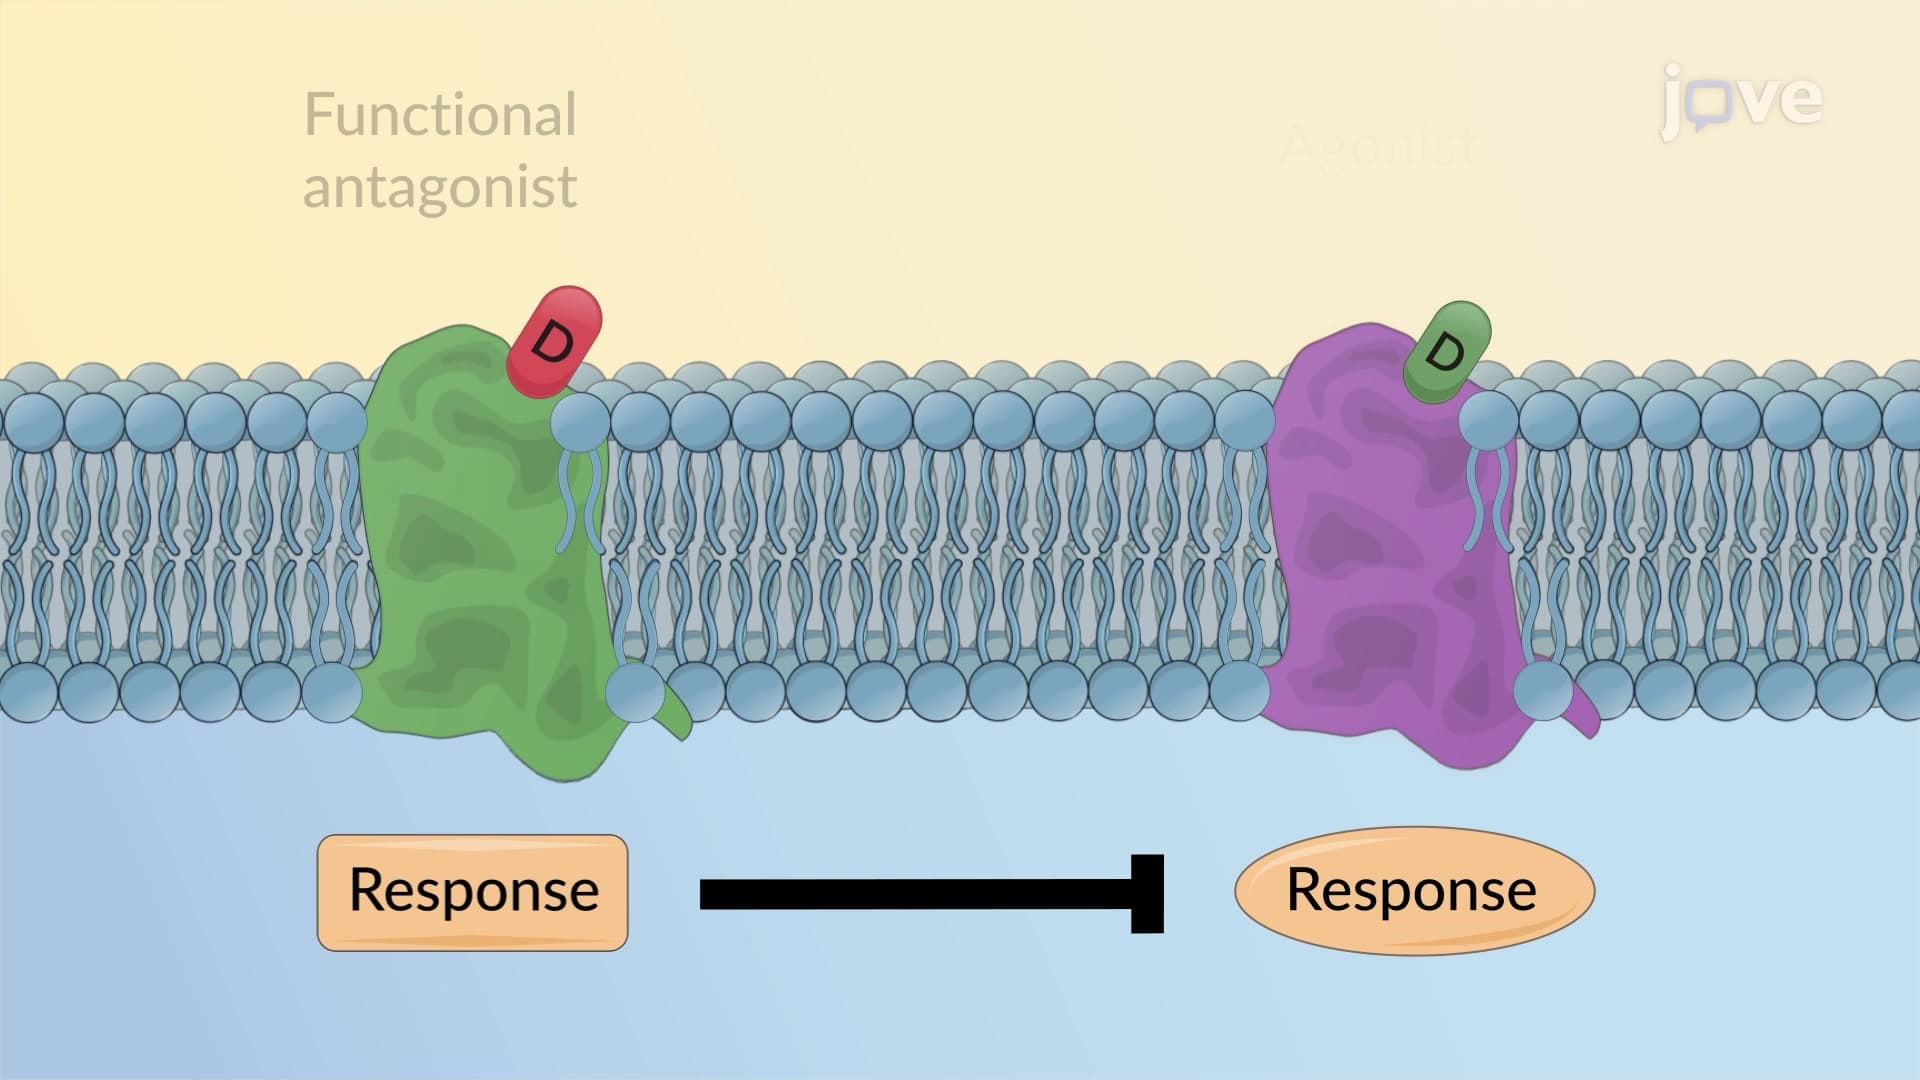
Combined Effects of Drugs: Antagonism
The most common type is receptor antagonism, where one drug acts as an antagonist to block the effects of another drug by...
 01:30
Anticoagulant Drugs: Low-Molecular-Weight Heparins
 01:19
Machines
A free-body diagram of the...
 01:30
Machines: Problem Solving II