Related Experiment Video
Updated: Jan 29, 2026

Methods for Quantitative Detection of Antibody-induced Complement Activation on Red Blood Cells
Published on: January 29, 2014
Hyperfunctional complement C3 promotes C5-dependent atypical hemolytic uremic syndrome in mice
Kate Smith-Jackson1,2, Yi Yang1, Harriet Denton1
1Institute of Cellular Medicine, Newcastle University, Newcastle upon Tyne, United Kingdom.

Atypical hemolytic uremic syndrome (aHUS) is frequently associated in humans with loss-of-function mutations in complement-regulating proteins or gain-of-function mutations in complement-activating proteins. Thus, aHUS provides an archetypal complement-mediated disease with which to model new therapeutic strategies and treatments. Herein, we show that, when transferred to mice, an aHUS-associated gain-of-function change (D1115N) to the complement-activation protein C3 results in aHUS. Homozygous C3 p.D1115N (C3KI) mice developed spontaneous chronic thrombotic microangiopathy together with hematuria, thrombocytopenia, elevated creatinine, and evidence of hemolysis. Mice with active disease had reduced plasma C3 with C3 fragment and C9 deposition within the kidney. Therapeutic blockade or genetic deletion of C5, a protein downstream of C3 in the complement cascade, protected homozygous C3KI mice from thrombotic microangiopathy and aHUS. Thus, our data provide in vivo modeling evidence that gain-of-function changes in complement C3 drive aHUS. They also show that long-term C5 deficiency is not accompanied by development of other renal complications (such as C3 glomerulopathy) despite sustained dysregulation of C3. Our results suggest that this preclinical model will allow testing of novel complement inhibitors with the aim of developing precisely targeted therapeutics that could have application in many complement-mediated diseases.
Atypical hemolytic uremic syndrome (aHUS) is frequently associated in humans with loss-of-function mutations in complement-regulating proteins or gain-of-function mutations in complement-activating proteins. Thus, aHUS provides an archetypal complement-mediated disease with which to model new therapeutic strategies and treatments. Herein, we show that, when transferred to mice, an aHUS-associated gain-of-function change (D1115N) to the complement-activation protein C3 results in aHUS. Homozygous C3 p.D1115N (C3KI) mice developed spontaneous chronic thrombotic microangiopathy together with hematuria, thrombocytopenia, elevated creatinine, and evidence of hemolysis. Mice with active disease had reduced plasma C3 with C3 fragment and C9 deposition within the kidney. Therapeutic blockade or genetic deletion of C5, a protein downstream of C3 in the complement cascade, protected homozygous C3KI mice from thrombotic microangiopathy and aHUS. Thus, our data provide in vivo modeling evidence that gain-of-function changes in complement C3 drive aHUS. They also show that long-term C5 deficiency is not accompanied by development of other renal complications (such as C3 glomerulopathy) despite sustained dysregulation of C3. Our results suggest that this preclinical model will allow testing of novel complement inhibitors with the aim of developing precisely targeted therapeutics that could have application in many complement-mediated diseases.
Related Concept Videos
 01:27
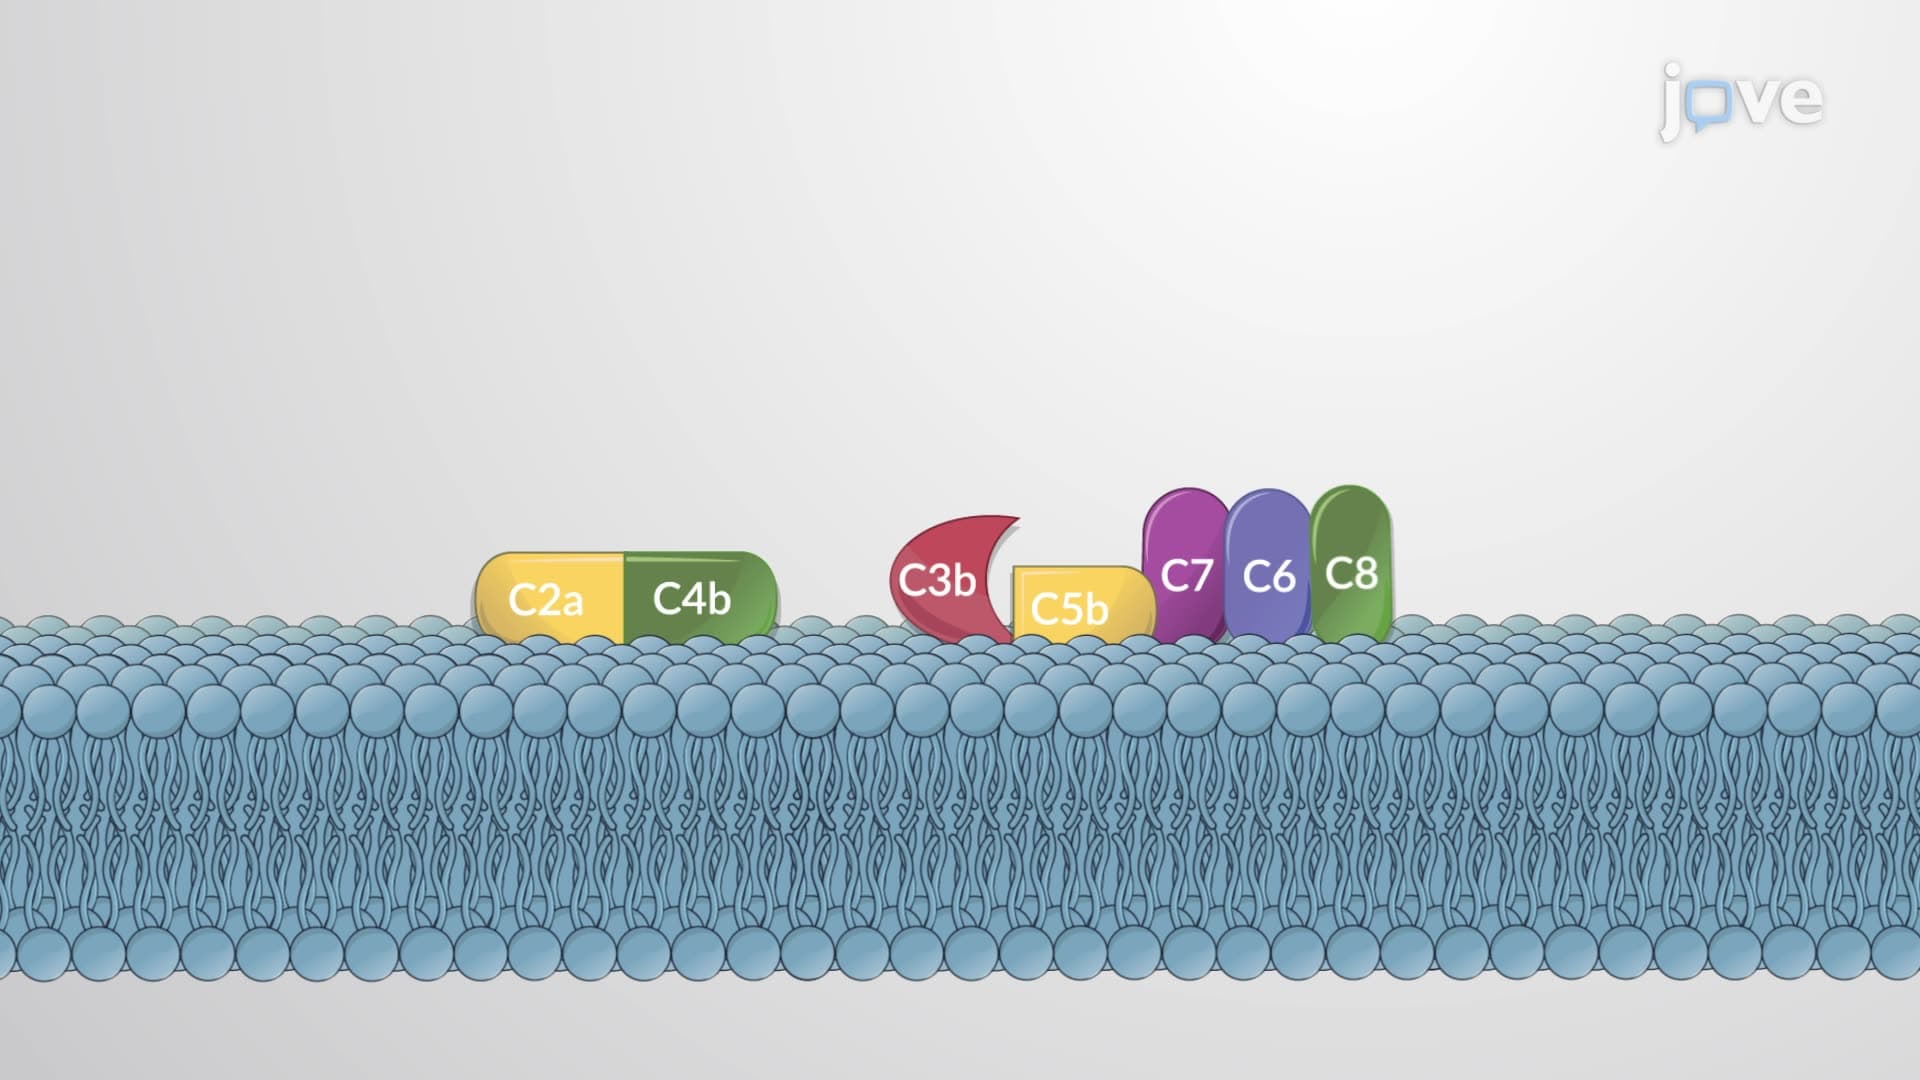
Complement System
 00:49
Complementation Tests
Organisms heterozygous for different mutations are crossed pairwise in all combinations. If present on different genes, the mutations can complement each other by providing the missing...
 01:21
Antipsychotic Drugs: Typical and Atypical Agents
 02:40
The Eukaryotic Promoter Region
02:40The Eukaryotic Promoter Region
 01:21
Frequency-dependent Selection