Related Experiment Video
Updated: Jan 29, 2026
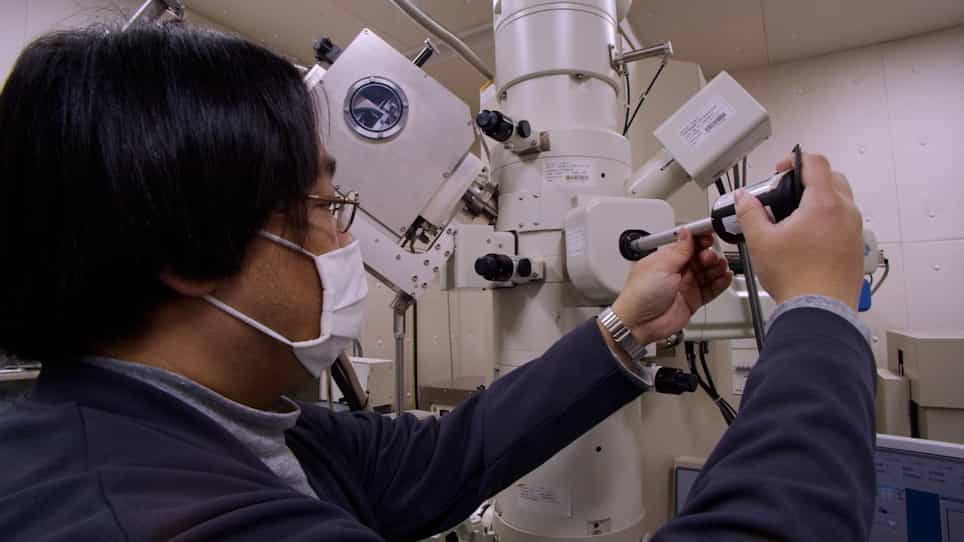

Quantitative Atomic-Site Analysis of Functional Dopants/Point Defects in Crystalline Materials by Electron-Channeling-Enhanced Microanalysis
Published on: May 10, 2021
Interaction between Bi Dopants and Intrinsic Defects in LiNbO3 from Local and Hybrid Density Functional Theory
Lili Li1, Yanlu Li1, Xian Zhao1
1State Key Lab of Crystal Materials and Institute of Crystal Materials , Shandong University , Jinan 250100 , China.

Abstract:
The interactions between Bi dopants including Bi-substituting Li (BiLi) and Bi-substituting Nb (BiNb) and the intrinsic antisite defects (NbLi) and Li vacancies (VLi) in LiNbO3 are investigated using local and hybrid density functional theories. Three charge-compensated defect clusters, BiLi4+ + NbLi4+ + 8VLi-, BiLi4+ + 4VLi-, and BiLi4+ + BiNb0 + 4VLi-, are modeled in this work to investigate the effects of the Bi concentration. The most stable cluster configurations, the Bi-doping stability in the clusters, and the electronic state interaction between Bi and intrinsic defects have been studied in detail. It is found that BiLi4+ has a stronger electron-capturing ability than NbLi4+ in Bi-doped congruent LiNbO3. The BiLi-doping-induced local lattice distortion and the electron-trapping behavior remain unchanged with increasing Bi-doping concentration. However, the position of the Bi defect states in the band gap is found to be shifted in congruent LiNbO3. This is mainly attributed to the large lattice relaxation induced by the large number of Li vacancies instead of the ionic level redistribution caused by the direct interaction between Bi and intrinsic defects.
Related Concept Videos
 02:18
Intrinsically Disordered Proteins
 02:35
Molecular Orbital Theory I
 03:24
Hybridization of Atomic Orbitals I
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory