Related Experiment Video
Updated: Jan 28, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Structure Based Drug Design and Molecular Docking Studies of Anticancer Molecules Paclitaxel, Etoposide and Topotecan
Manisha Yadav1, Swasti Dhagat1, Jujjavarapu S Eswari1
1Department of Biotechnology, National Institute of Technology, Raipur, India.

Background:
Tubulin is the biochemical target for several clinically used anticancer drugs as it helps in the formation of mitotic spindle during mitosis stage of cell division. Many of the anti-cancer drugs are known to interact with tubulin and microtubules including some plant alkaloids, such as paclitaxel, etoposide and topotecan. In silico drug design of these molecules were performed prior to testing these drugs in vitro. In silico drug design of these anti-cancer drugs becomes a challenge due to the complex structure of target protein. This challenge was overcome by predicting the structure of the target protein (tubulin) by homology modeling.
Methods:
In this study, computer aided drug designing approach was applied to predict the suitable docking site in target protein and the interaction of tubulin protein with paclitaxel, etoposide and topotecan was explored by molecular docking using Schrödinger software. Docking score and glide energy were determined with ligands to validate their anticancer properties.
Results:
The results indicate that etoposide is the best drug for tubulin with a docking score of - 4.916 and glide energy of -46.470 kcal/mol compared to paclitaxel and topotecan.
Conclusion:
The testing of these drugs in silico provides an alternate to in vitro testing of these molecules on cancer cell lines which is a time and cost intensive process. The in silico study of parameters, such as docking score and glide energy, will help pharmacists in developing new molecules as targets for cancers in a time and cost-effective manner.
More Related Videos
 06:03
Use of Viral Entry Assays and Molecular Docking Analysis for the Identification of Antiviral Candidates against Coxsackievirus A16
Published on: July 15, 2019
 10:25
Screening Traditional Chinese Medicine Compounds for Inhibiting UCHL3 Activity Based on Molecular Docking and Deubiquitinating Enzyme Probe Technology
Published on: November 22, 2024
Related Concept Videos
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence...
 02:51
Metal-Ligand Bonds
In these complexes, transition metals form coordinate covalent bonds, a kind of Lewis acid-base interaction in which both of the electrons in the bond are contributed by a donor (Lewis base) to an electron acceptor (Lewis acid). The Lewis acid in...
 03:05
Acid Strength and Molecular Structure
In the absence of any leveling effect, the acid strength of binary compounds of hydrogen with nonmetals (A) increases as the H-A bond strength decreases down a group in the periodic table. For group 17, the order of increasing acidity is HF < HCl < HBr < HI. Likewise, for group 16, the order of increasing acid strength is H2O < H2S < H2Se < H2Te. Across a row in the periodic table, the acid strength of binary hydrogen compounds increases with increasing...
 03:48
Behavior of Gas Molecules: Molecular Diffusion, Mean Free Path, and Effusion
 02:54
Lewis Structures of Molecular Compounds and Polyatomic Ions
 02:34
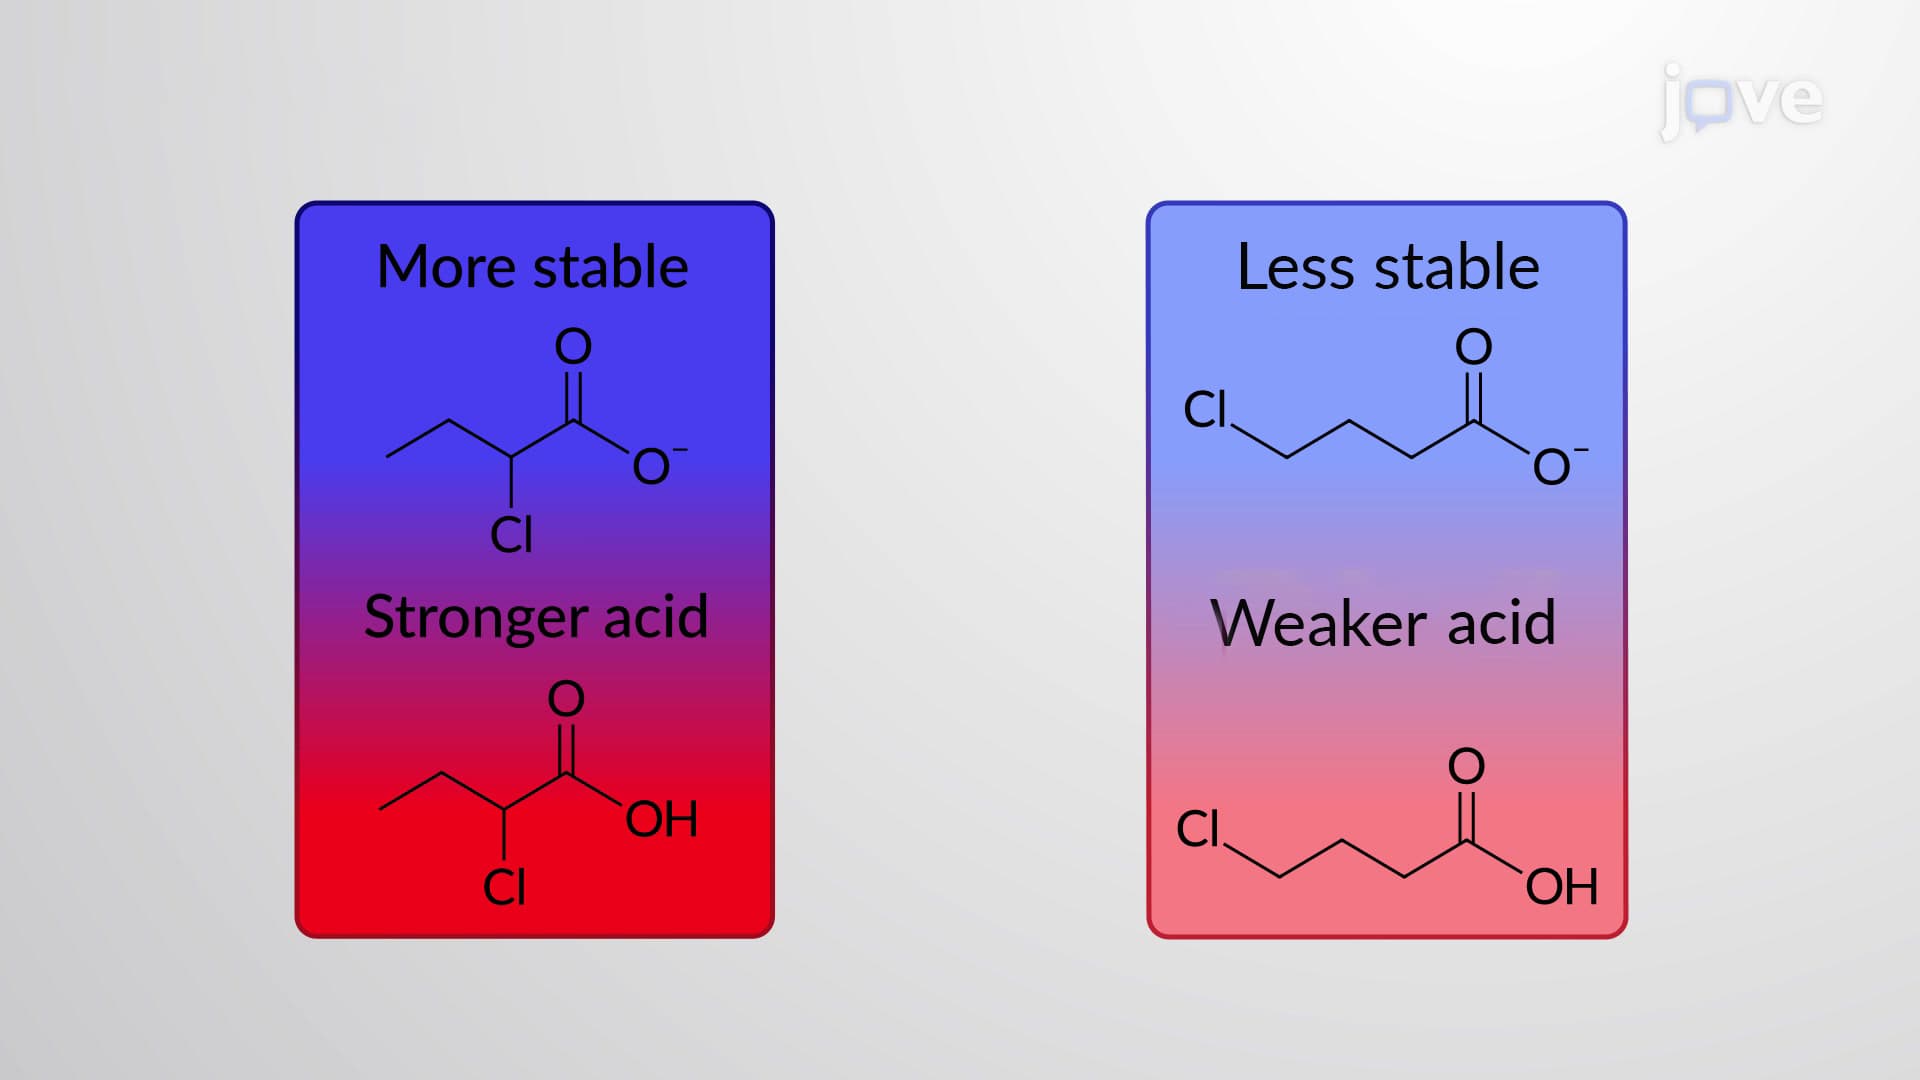
Molecular Structure and Acidity
The size effect explains the change in atomic size on acidity. When comparing the acids formed from elements that belong to the same column in the periodic table, their atomic sizes...