Related Experiment Video
Updated: Jan 27, 2026
Microsurgical Obstruction of Testes Fusion in Spodoptera litura
Published on: July 16, 2021
RNA-Seq analysis reveals transcript diversity and active genes after common cutworm (Spodoptera litura Fabricius)
Haiping Du1, Xiao Li1, Lihua Ning2
1National Center for Soybean Improvement, National Key Laboratory of Crop Genetics and Germplasm Enhancement, Jiangsu Collaborative Innovation Center for Modern Crop Production, Nanjing Agricultural University, Nanjing, 210095, China.

Background:
Common cutworm (CCW) is highly responsible for destabilizing soybean productivity. Wild soybean is a resource used by breeders to discover elite defensive genes.
Results:
The transcriptomes of two wild accessions (W11 and W99) with different resistance to CCW were analyzed at early- and late-induction time points. After induction, the susceptible accession W11 differentially expressed 1268 and 508 genes at the early and late time points, respectively. Compared with W11, the resistant accession W99 differentially expressed 1270 genes at the early time point and many more genes (2308) at the late time point. In total, 3836 non-redundant genes were identified in both lines. Gene Ontology (GO) and Kyoto Encyclopedia of Genes and Genomes (KEGG) analyses revealed that the differentially expressed genes (DEGs) in W99 at the late time point were mostly associated with specific processes and pathways. Among the non-redundant genes, 146 genes were commonly up-regulated in the treatment condition compared with the control condition at the early- and late-induction time points in both accessions used in this experiment. Approximately 40% of the common DEGs were related to secondary metabolism, disease resistance, and signal transduction based on their putative function. Excluding the common DEGs, W99 expressed more unique DEGs than W11. Further analysis of the 3836 DEGs revealed that the induction of CCW not only up-regulated defense-related genes, including 37 jasmonic acid (JA)-related genes, 171 plant-pathogen-related genes, and 17 genes encoding protease inhibitors, but also down-regulated growth-related genes, including 35 photosynthesis-related genes, 48 nutrition metabolism genes, and 28 auxin metabolism genes. Therefore, representative defense-related and growth-related genes were chosen for binding site prediction via co-expression of transcription factors (TFs) and spatial expression pattern analyses. In total, 53 binding sites of 28 TFs were identified based on 3 defense-related genes and 3 growth-related genes. Phosphate transporter PT1, which is a representative growth-related gene, was transformed into soybean, and the transgenic soybean plants were susceptible to CCW.
Conclusions:
In summary, we described transcriptome reprograming after herbivore induction in wild soybean, identified the susceptibility of growth-related genes, and provided new resources for the breeding of herbivore-resistant cultivated soybeans.
Related Concept Videos
 02:42
Eukaryotic Transcription Activators
The binding domains are capable of recognizing and interacting with regulatory sequences on the DNA. These...
 01:58
Prokaryotic Transcriptional Activators and Repressors
Transcription of prokaryotic...
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 01:10
Transcription
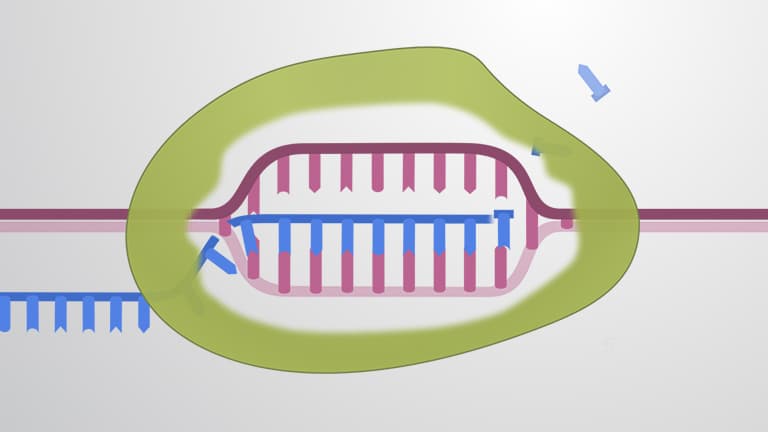
Transcription is the process of synthesizing RNA from a DNA sequence by RNA polymerase. It is the first step in producing a protein from a gene sequence. Additionally, many other proteins and regulatory sequences are involved in the proper synthesis of messenger RNA (mRNA). Regulation of transcription is responsible for the differentiation of all the different types of cells and often for the proper cellular response to environmental signals.
Transcription Can Produce Different Kinds...
 00:58
Eukaryotic RNA Polymerases
All three eukaryotic RNAPs require specific transcription factors, of which the...
 02:16
Transcription Factors