Related Experiment Video
Updated: Jan 26, 2026

Purifying the Impure: Sequencing Metagenomes and Metatranscriptomes from Complex Animal-associated Samples
Published on: December 22, 2014
Towards precision quantification of contamination in metagenomic sequencing experiments
M S Zinter1, M Y Mayday1, K K Ryckman2
1Department of Pediatrics, Division of Critical Care, University of California, San Francisco School of Medicine, Benioff Children's Hospital, San Francisco, CA, USA.

Abstract:
Metagenomic next-generation sequencing (mNGS) experiments involving small amounts of nucleic acid input are highly susceptible to erroneous conclusions resulting from unintentional sequencing of occult contaminants, especially those derived from molecular biology reagents. Recent work suggests that, for any given microbe detected by mNGS, an inverse linear relationship between microbial sequencing reads and sample mass implicates that microbe as a contaminant. By associating sequencing read output with the mass of a spike-in control, we demonstrate that contaminant nucleic acid can be quantified in order to identify the mass contributions of each constituent. In an experiment using a high-resolution (n = 96) dilution series of HeLa RNA spanning 3-logs of RNA mass input, we identified a complex set of contaminants totaling 9.1 ± 2.0 attograms. Given the competition between contamination and the true microbiome in ultra-low biomass samples such as respiratory fluid, quantification of the contamination within a given batch of biological samples can be used to determine a minimum mass input below which sequencing results may be distorted. Rather than completely censoring contaminant taxa from downstream analyses, we propose here a statistical approach that allows separation of the true microbial components from the actual contribution due to contamination. We demonstrate this approach using a batch of n = 97 human serum samples and note that despite E. coli contamination throughout the dataset, we are able to identify a patient sample with significantly more E. coli than expected from contamination alone. Importantly, our method assumes no prior understanding of possible contaminants, does not rely on any prior collection of environmental or reagent-only sequencing samples, and does not censor potentially clinically relevant taxa, thus making it a generalized approach to any kind of metagenomic sequencing, for any purpose, clinical or otherwise.
Related Concept Videos
 01:16
Contaminants and Errors
Another key consideration is determining the appropriate number of samples required to...
 03:37
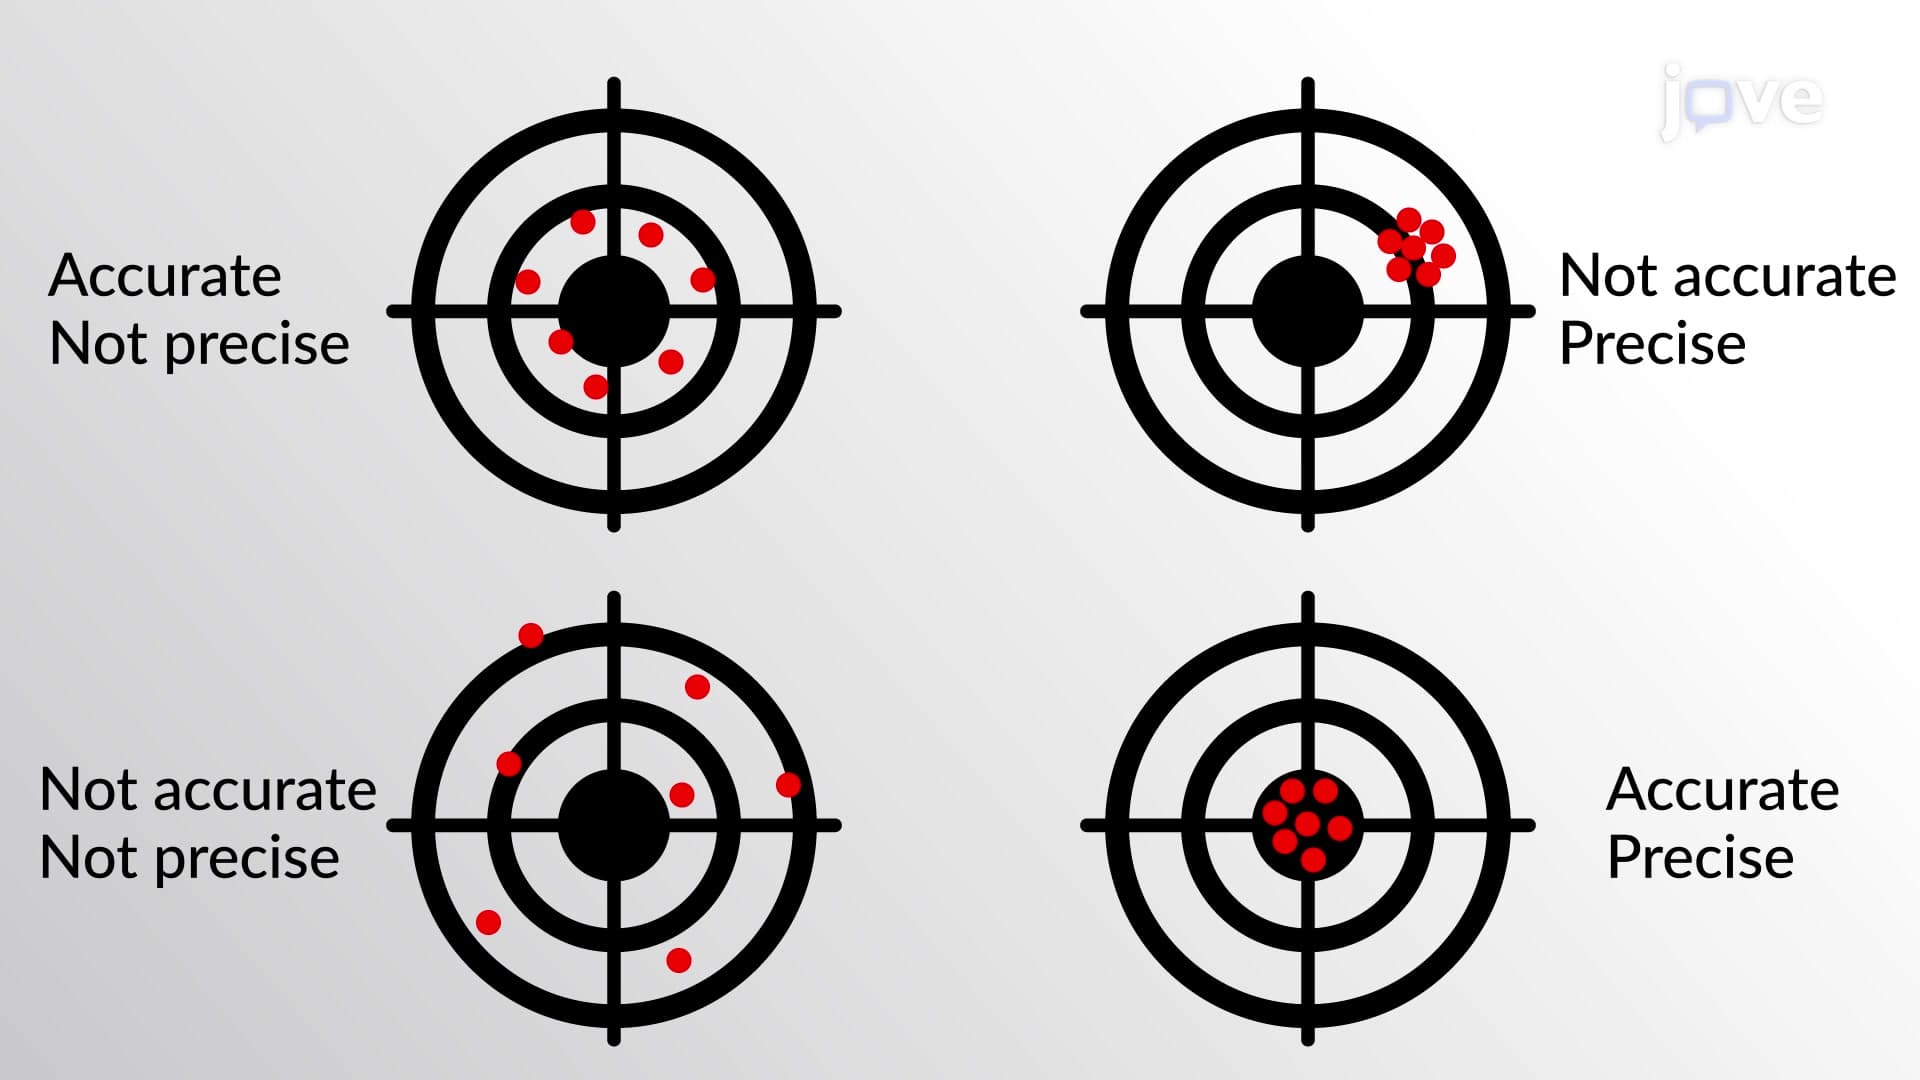
Uncertainty in Measurement: Accuracy and Precision
 01:52
Accuracy and Precision
 01:12
What is an Experiment?
 02:02
Cis-regulatory Sequences
 01:19
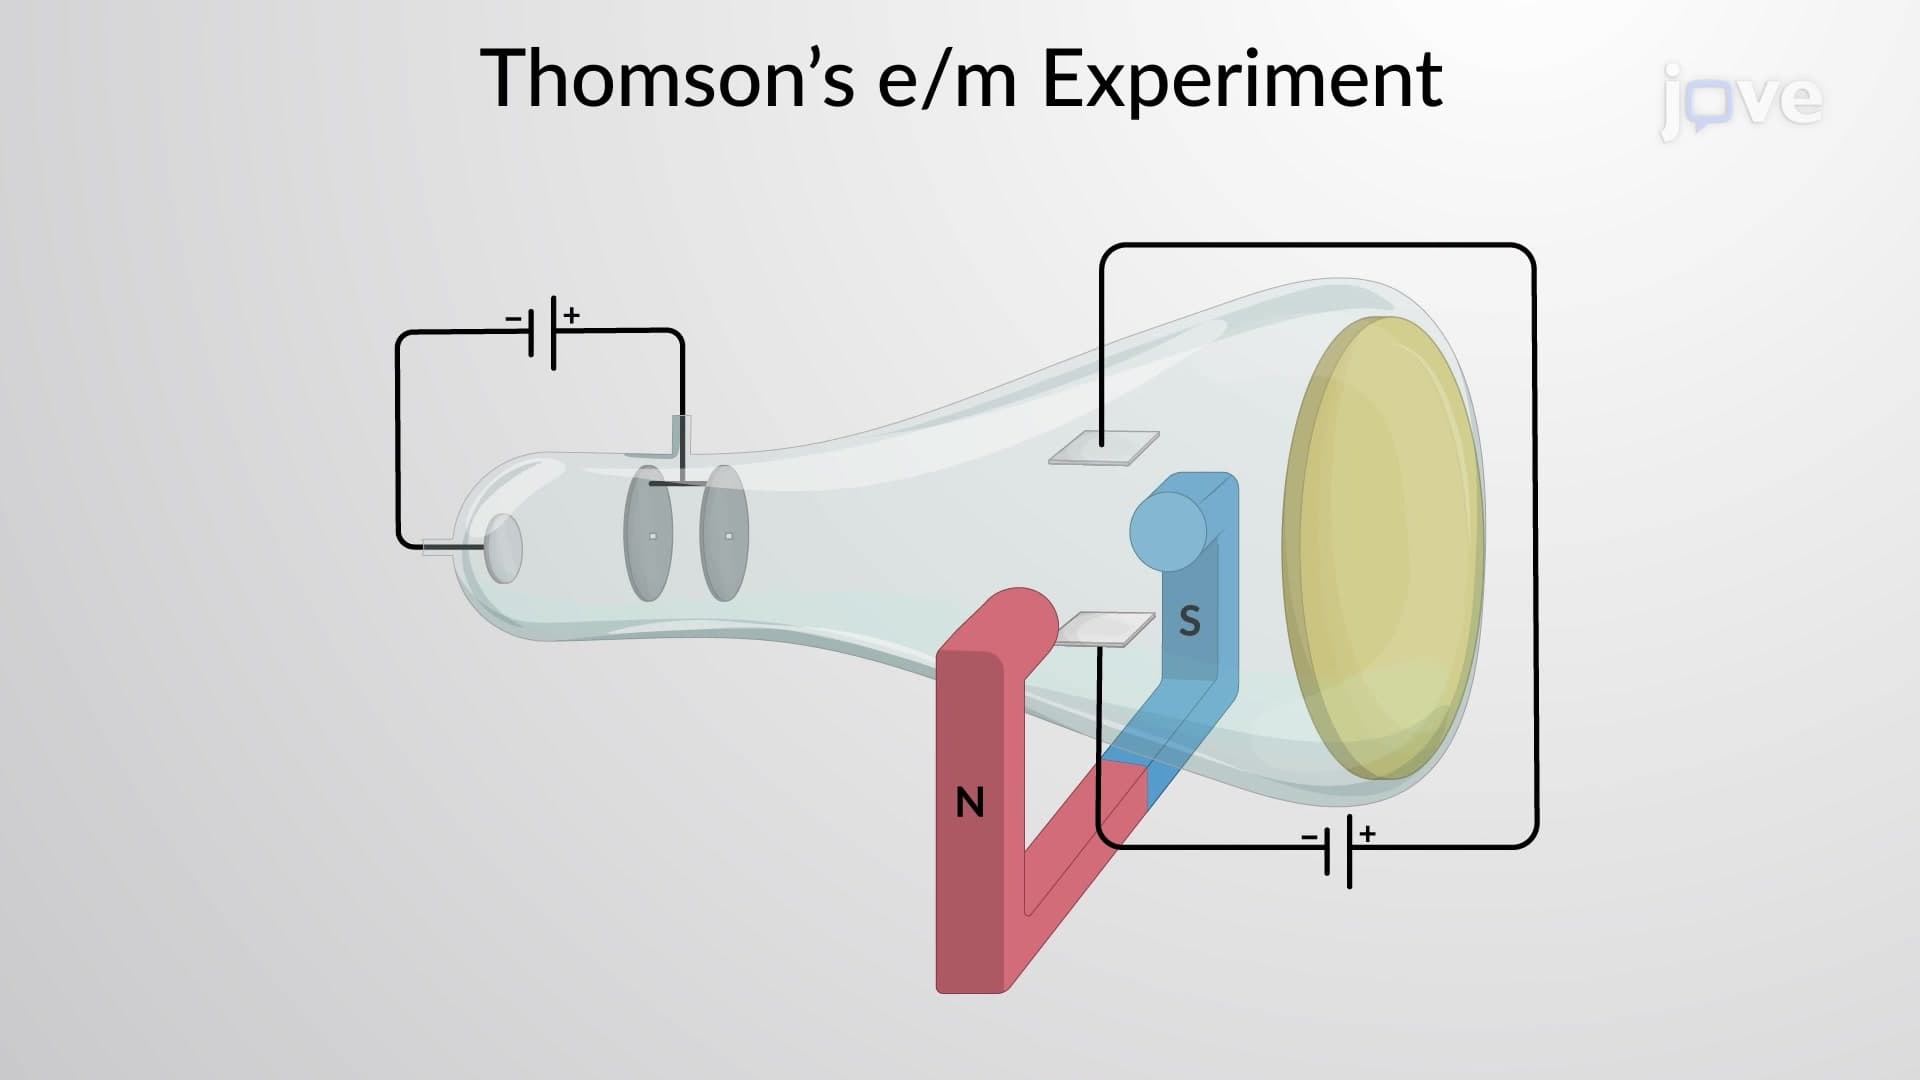
Thomson's e/m Experiment
A particle with charge q, speed v, and mass m enters an area from the top, where the magnetic and electric fields are perpendicular both to the particle's motion and to one another. The magnetic...