Related Experiment Video
Updated: Jan 25, 2026
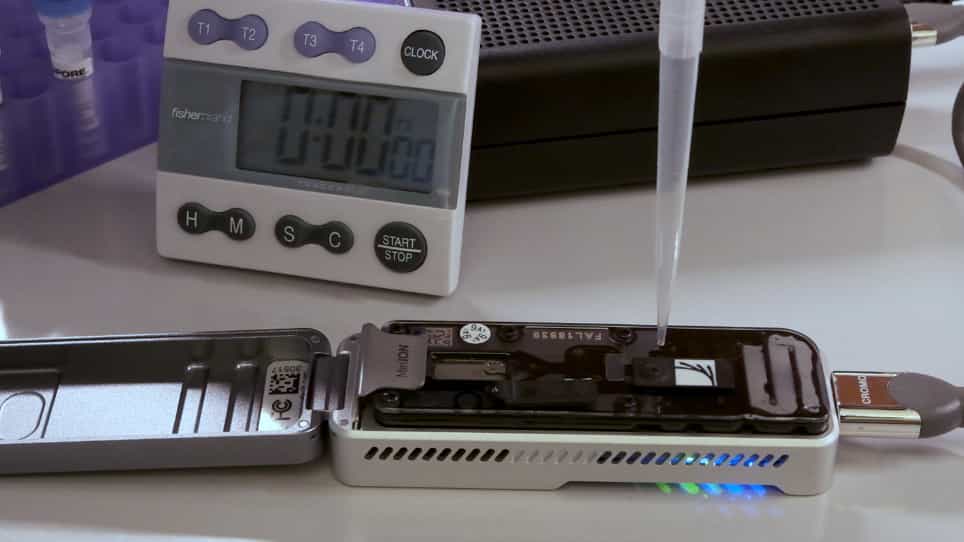

Hybrid De Novo Genome Assembly for the Generation of Complete Genomes of Urinary Bacteria using Short- and Long-read Sequencing Technologies
Published on: August 20, 2021
A comparative analysis of methods for de novo assembly of hymenopteran genomes using either haploid or diploid
1Department of Evolutionary and Environmental Biology, Institute of Evolution, University of Haifa, Haifa, 3498838, Israel.

Abstract:
Diverse invertebrate taxa including all 200,000 species of Hymenoptera (ants, bees, wasps and sawflies) have a haplodiploid sex determination system, where females are diploid and males are haploid. Thus, hymenopteran genome projects can make use of DNA from a single haploid male sample, which is assumed advantageous for genome assembly. For the purpose of gene annotation, transcriptome sequencing is usually conducted using RNA from a pool of individuals. We conducted a comparative analysis of genome and transcriptome assembly and annotation methods, using genetic sources of different ploidy: (1) DNA from a haploid male or a diploid female (2) RNA from the same haploid male or a pool of individuals. We predicted that the use of a haploid male as opposed to a diploid female will simplify the genome assembly and gene annotation thanks to the lack of heterozygosity. Using DNA and RNA from the same haploid individual is expected to provide better confidence in transcript-to-genome alignment, and improve the annotation of gene structure in terms of the exon/intron boundaries. The haploid genome assemblies proved to be more contiguous, with both contig and scaffold N50 size at least threefold greater than their diploid counterparts. Completeness evaluation showed mixed results. The SOAPdenovo2 diploid assembly was missing more genes than the haploid assembly. The SPAdes diploid assembly had more complete genes, but a higher level of duplicates, and a greatly overestimated genome size. When aligning the two transcriptomes against the male genome, the male transcriptome gave 2-3% more complete transcripts than the pool transcriptome for genes with comparable expression levels in both transcriptomes. However, this advantage disappears in the final results of the gene annotation pipeline that incorporates evidence from homologous proteins. The RNA pool is still required to obtain the full transcriptome with genes that are expressed in other life stages and castes. In conclusion, the use of a haploid source material for a de novo genome project provides a substantial advantage to the quality of the genome draft and the use of RNA from the same haploid individual for transcriptome to genome alignment provides a minor advantage for genes that are expressed in the adult male.
Related Concept Videos
 02:16
Comparing Mitochondrial, Chloroplast, and Prokaryotic Genomes
 03:36
Genome Annotation and Assembly
 01:20
Comparing the Survival Analysis of Two or More Groups
 02:30
Genomic Imprinting and Inheritance
The expression of some genes depends on which parent passed the gene to the offspring, through a phenomenon known as...
 02:02
Genomics
 01:18
Sampling Methods: Sample Types
Solid samples include a variety of substances, such as sediments from water bodies, soil, metals, and biological tissues. Two standard methods for extracting sediments from water bodies are grab sampling and piston coring. Grab sampling involves using a device to collect a discrete sediment sample from the bottom of a water body with minimal disturbance. Grab samples do not always represent the entire area due to...