Related Experiment Video
Updated: Jan 25, 2026
Protein Purification-free Method of Binding Affinity Determination by Microscale Thermophoresis
Published on: August 15, 2013
Assessing the performance of MM/PBSA and MM/GBSA methods. 9. Prediction reliability of binding affinities and binding
Gaoqi Weng1, Ercheng Wang, Fu Chen
1College of Pharmaceutical Sciences, Zhejiang University, Hangzhou, Zhejiang 310058, China. tingjunhou@zju.edu.cn.

Abstract:
A significant number of protein-protein interactions (PPIs) are mediated through the interactions between proteins and peptide segments, and therefore determination of protein-peptide interactions (PpIs) is critical to gain an in-depth understanding of the PPI network and even design peptides or small molecules capable of modulating PPIs. Computational approaches, especially molecular docking, provide an efficient way to model PpIs, and a reliable scoring function that can recognize the correct binding conformations for protein-peptide complexes is one of the most important components in protein-peptide docking. The end-point binding free energy calculation methods, such as MM/GBSA and MM/PBSA, are theoretically more rigorous than most empirical and semi-empirical scoring functions designed for protein-peptide docking, but their performance in predicting binding affinities and binding poses for protein-peptide systems has not been systematically assessed. In this study, we first evaluated the capability of MM/GBSA and MM/PBSA with different solvation models, interior dielectric constants (εin) and force fields to predict the binding affinities for 53 protein-peptide complexes. For the 19 short peptides with 5-12 residues, MM/PBSA based on the minimized structures in explicit solvent with the ff99 force field and εin = 2 yields the best correlation between the predicted binding affinities and the experimental data (rp = 0.748), while for the 34 medium-size peptides with 20-25 residues, MM/GBSA based on 1 ns of molecular dynamics (MD) simulations in implicit solvent with the ff03 force field, the GBOBC1 model and a low interior dielectric constant (εin = 1) yields the best accuracy (rp = 0.735). Then, we assessed the rescoring capability of MM/PBSA and MM/GBSA to distinguish the correct binding conformations from the decoys for 112 protein-peptide systems. The results illustrate that MM/PBSA based on the minimized structures with the ff99 or ff14SB force field and MM/GBSA based on the minimized structures with the ff03 force field show excellent capability to recognize the near-native binding poses for the short and medium-size peptides, respectively, and they outperform the predictions given by two popular protein-peptide docking algorithms (pepATTRACT and HPEPDOCK). Therefore, MM/PBSA and MM/GBSA are powerful tools to predict the binding affinities and identify the correct binding poses for protein-peptide systems.

Related Concept Videos
 02:18
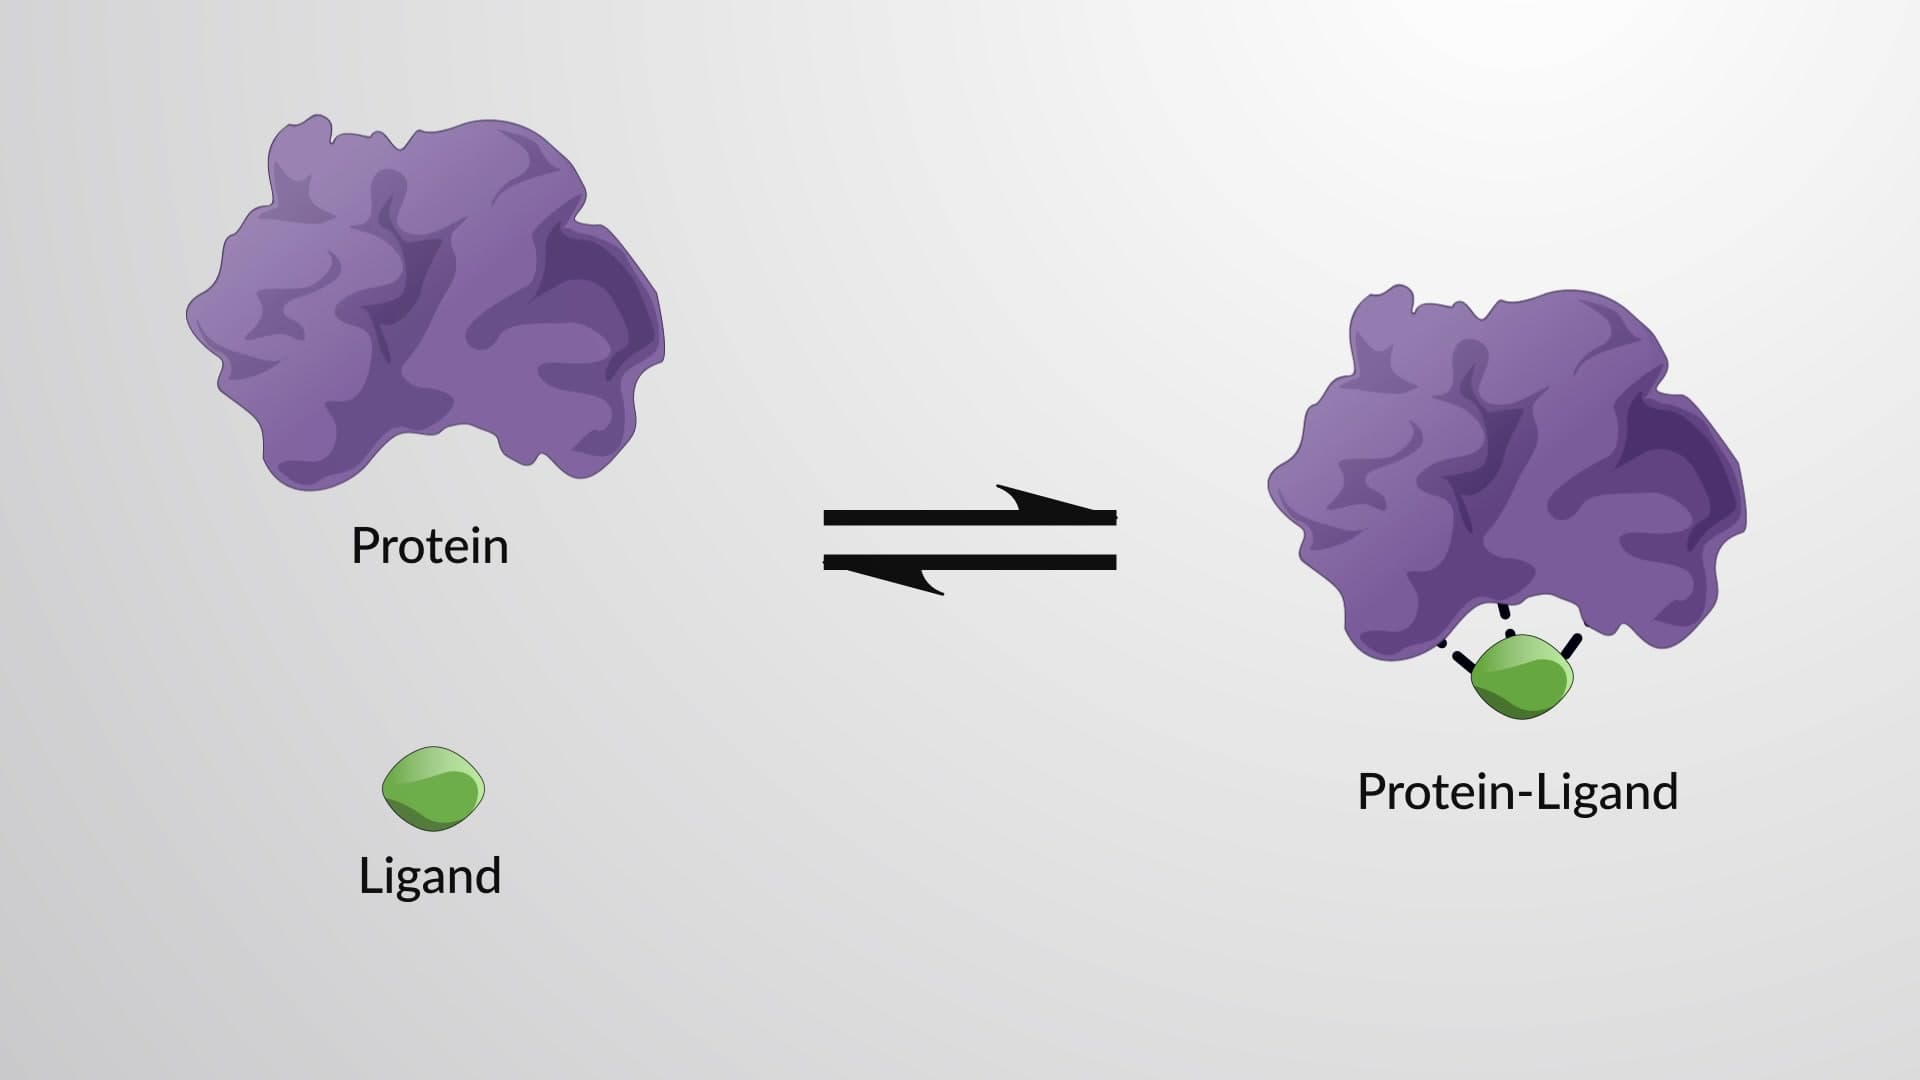
The Equilibrium Binding Constant and Binding Strength
02:18The Equilibrium Binding Constant and Binding Strength
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 00:49
Ligand Binding and Linkage