Related Experiment Video
Updated: Jan 24, 2026

Sedimentation Equilibrium of a Small Oligomer-forming Membrane Protein: Effect of Histidine Protonation on Pentameric Stability
Published on: April 2, 2015
A Hierarchical Approach to Predict Conformation-Dependent Histidine Protonation States in Stable and Flexible
Serzhan N Sakipov1, Jose C Flores-Canales1, Maria G Kurnikova1
1Chemistry Department , Carnegie Mellon University , Pittsburgh , Pennsylvania 15213 , United States.

Abstract:
Solution acidity measured by pH is an important environmental factor that affects protein structure. It influences the protonation state of protein residues, which in turn may be coupled to protein conformational changes, unfolding, and ligand binding. It remains difficult to compute and measure the p Ka of individual residues, as well as to relate them to pH-dependent protein transitions. This paper presents a hierarchical approach to compute the p Ka of individual protonatable residues, specifically histidines, coupled with underlying structural changes of a protein. A fast and efficient free energy perturbation (FEP) algorithm has also been developed utilizing a fast implementation of standard molecular dynamics (MD) algorithms. Specifically, a CUDA version of the AMBER MD engine is used in this paper. Eight histidine p Ka's are computed in a diverse set of pH stable proteins to demonstrate the proposed approach's utility and assess the predictive quality of the AMBER FF99SB force field. A reference molecule is carefully selected and tested for convergence. A hierarchical approach is used to model p Ka's of the six histidine residues of the diphtheria toxin translocation domain (DTT), which exhibits a diverse ensemble of individual conformations and pH-dependent unfolding. The hierarchical approach consists of first sampling equilibrium conformational ensembles of a protein with protonated and neutral histidine residues via long equilibrium MD simulations (Flores-Canales, J. C.; et al. bioRxiv, 2019, 572040). A clustering method is then used to identify sampled protein conformations, and p Ka's of histidines in each protein conformation are computed. Finally, an ensemble averaging formalism is developed to compute weighted average histidine p Ka's. These can be compared with an apparent experimentally measured p Ka of the DTT protein and thus allows us to propose a mechanism of pH-dependent unfolding of the DTT protein.
More Related Videos
 10:03
Proton Transfer and Protein Conformation Dynamics in Photosensitive Proteins by Time-resolved Step-scan Fourier-transform Infrared Spectroscopy
Published on: June 27, 2014
 12:52
IridiumIII Luminescent Probe for Detection of the Malarial Protein Biomarker Histidine Rich Protein-II
Published on: July 7, 2015
Related Concept Videos
 00:52
¹H NMR of Conformationally Flexible Molecules: Temporal Resolution
 01:15
¹H NMR of Conformationally Flexible Molecules: Variable-Temperature NMR
 01:20
Conformity
 02:18
Intrinsically Disordered Proteins
 01:25
cAMP-dependent Protein Kinase Pathways
 02:20
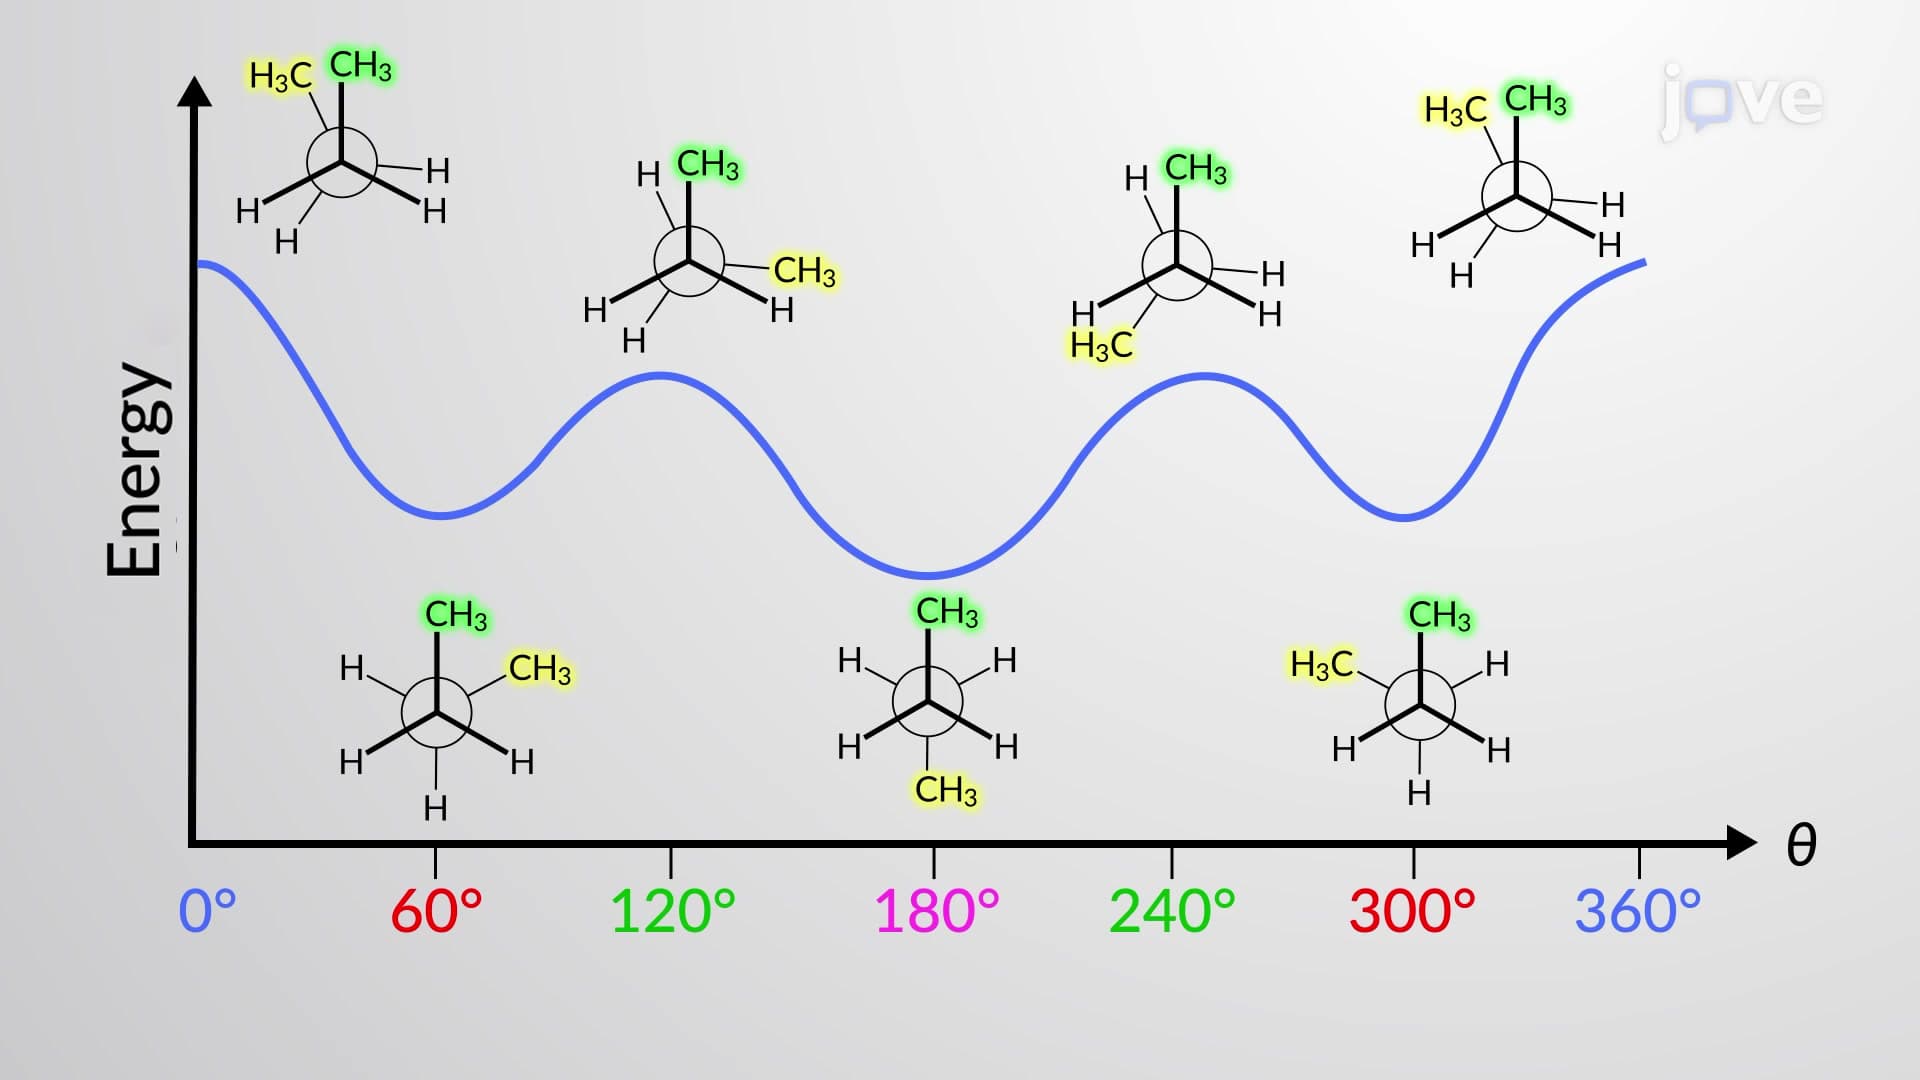
Conformations of Butane