Related Experiment Video
Updated: Jan 23, 2026


Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Molecular dimers of methane clathrates: ab initio potential energy surfaces and variational vibrational states
Michael P Metz1, Krzysztof Szalewicz, János Sarka
1Department of Physics and Astronomy, University of Delaware, Newark, Delaware 19716, USA. szalewic@udel.edu.

Abstract:
Motivated by the energetic and environmental relevance of methane clathrates, highly accurate ab initio potential energy surfaces (PESs) have been developed for the three possible dimers of the methane and water molecules: (H2O)2, CH4·H2O, and (CH4)2. While only a single monomer geometry was used for each monomer in the ab initio calculations, the PES parameterization makes it possible to produce distinct surfaces for all isotopologues within the rigid-monomer approximation. The PESs were fitted to computations at the frozen-core coupled-cluster level with single, double, and non-iterative triple excitations, employing basis sets of augmented triple- and quadruple-zeta quality plus bond functions, followed by extrapolations to the complete basis set limit. The long-range parts of the PESs are computed using the asymptotic version of symmetry-adapted perturbation theory based on a density-functional description of the monomers. All PESs are polarizable, i.e., in cluster or condensed-phase applications they approximate many-body effects by the induced dipole polarization model. The PESs were developed in a fully automated procedure applying the autoPES method, which is used for the first time to generate near-spectroscopic quality surfaces. The stationary points (SPs) on the PESs have been determined and compared with literature data. For CH4·H2O, previously unknown SPs have been identified and the first detailed study of the (CH4)2 potential energy landscape has been carried out. The PESs were used in variational quantum nuclear motion computations. For the water dimer, the resulting vibrational transitions are in excellent agreement with available high-resolution spectroscopic data. For (CH4)2, the intermonomer vibrational states are reported for the first time.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Related Concept Videos
 00:52
Potential Energy
Chemical bonds that form attractive forces between atoms also contain potential energy, called chemical energy. When a chemical reaction...
 01:09
Potential Energy
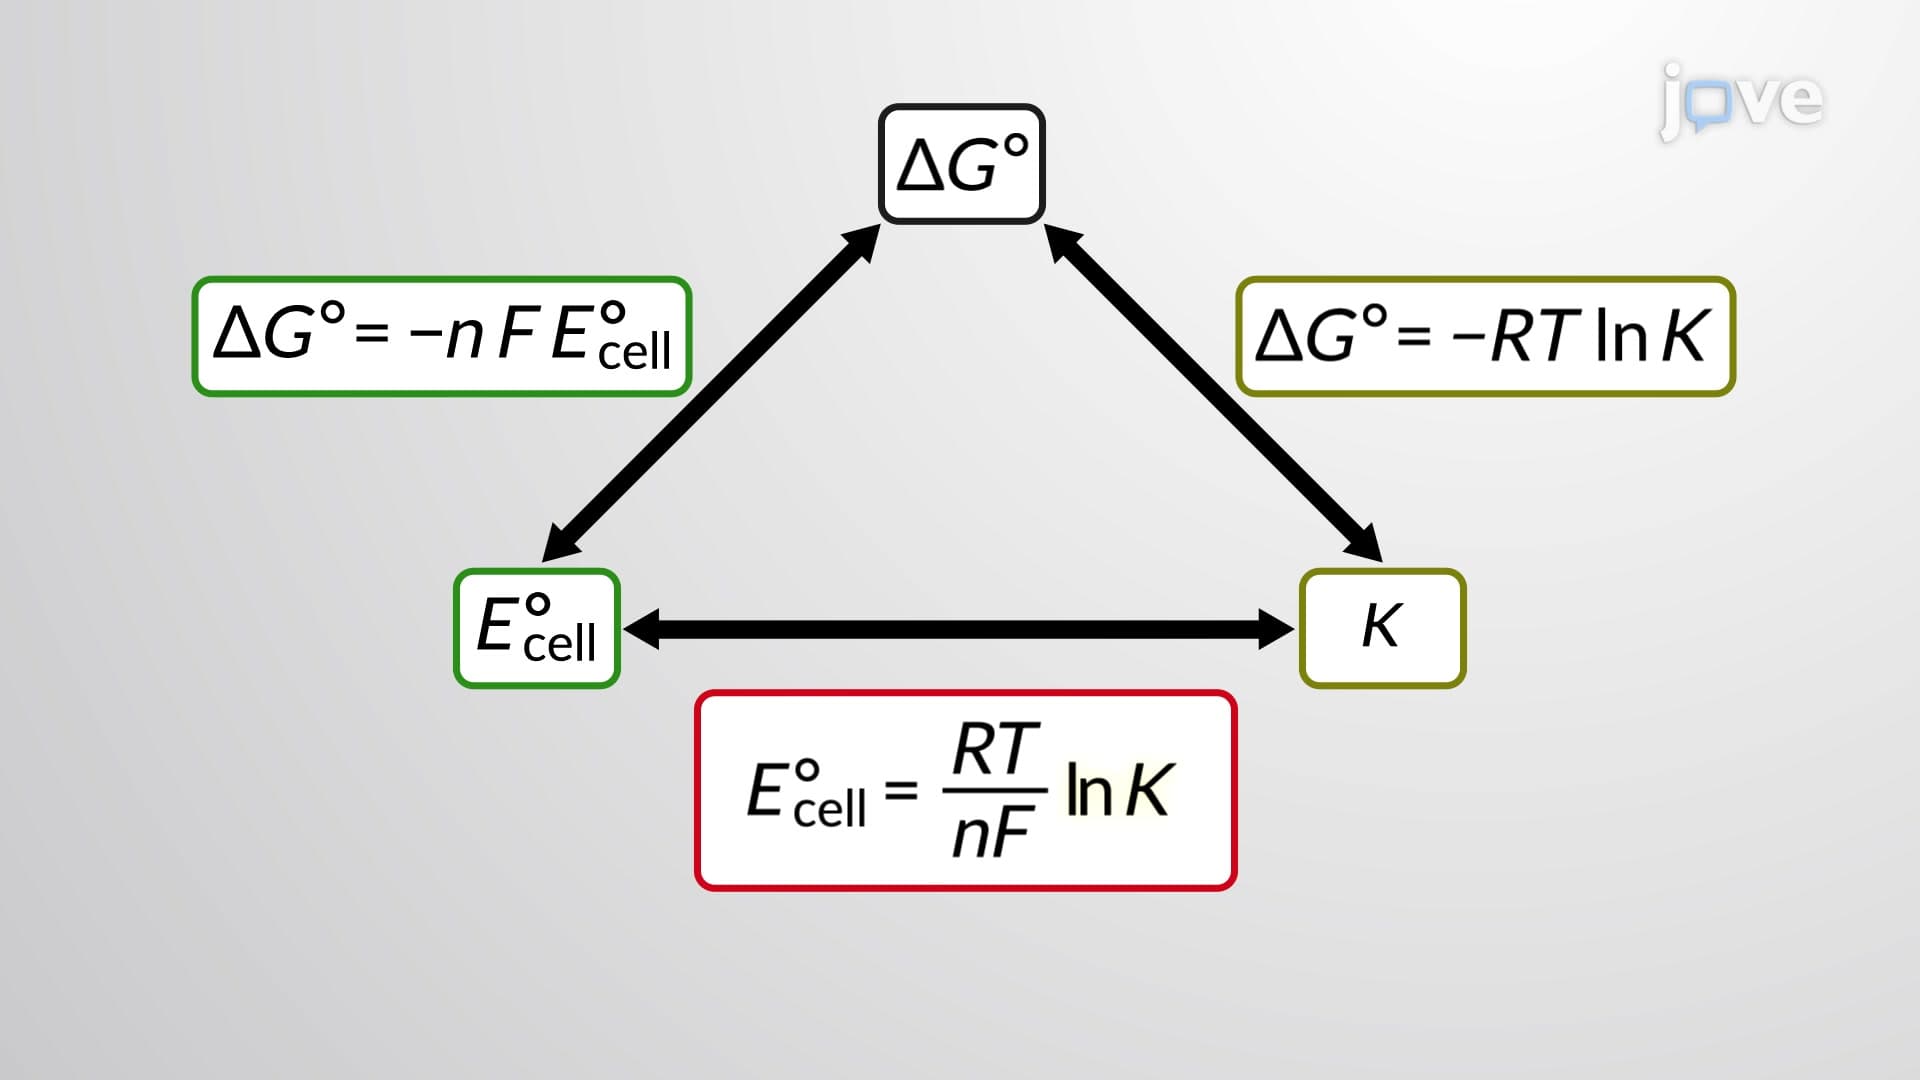 02:58
Cell Potential and Free Energy
Thermodynamics is the branch of physics dealing with the relationship between heat and other forms of energy. In an electrochemical cell, chemical energy is converted into electrical energy.
Thus, a link can be predicted between cell potential, free energy change, and the equilibrium constant for the reaction. Cell potential can also be measured as the oxidant or the reducing strength, and similar acid-base strength measures are reflected in equilibrium...
 01:24
IR Spectroscopy: Molecular Vibration Overview
Stretching vibrations are vibrational motions that occur along the bond line, changing the bond length or distance between two bonded atoms. They are further distinguished as symmetric or asymmetric. In symmetric stretching, the...
 03:07
Kinetic Molecular Theory: Molecular Velocities, Temperature, and Kinetic Energy
 01:16
Surface Tension and Surface Energy
Consider a beaker filled with liquid. The bulk molecules in the liquid experience equal attractive forces on all sides with the surrounding molecules. However, the surface molecules experience a net attractive force downward due to the bulk molecules. The surface of the liquid behaves like a stretched membrane,...