Related Experiment Video
Updated: Jan 21, 2026

Characterization of In Vitro Differentiation of Human Primary Keratinocytes by RNA-Seq Analysis
Published on: May 16, 2020
Next Generation DNA-Seq and Differential RNA-Seq Allow Re-annotation of the Pyrococcus furiosus DSM 3638 Genome and
Felix Grünberger1, Robert Reichelt1, Boyke Bunk2
1Institute of Microbiology and Archaea Center, University of Regensburg, Regensburg, Germany.

Abstract:
Pyrococcus furiosus DSM 3638 is a model organism for hyperthermophilic archaea with an optimal growth temperature near 100°C. The genome was sequenced about 18 years ago. However, some publications suggest that in contrast to other Pyrococcus species, the genome of P. furiosus DSM 3638 is prone to genomic rearrangements. Therefore, we re-sequenced the genome using third generation sequencing techniques. The new de novo assembled genome is 1,889,914 bp in size and exhibits high sequence identity to the published sequence. However, two major deviations were detected: (1) The genome is 18,342 bp smaller than the NCBI reference genome due to a recently described deletion. (2) The region between PF0349 and PF0388 is inverted most likely due an assembly problem for the original sequence. In addition, numerous minor variations, ranging from single nucleotide exchanges, deletions or insertions were identified. The total number of insertion sequence (IS) elements is also reduced from 30 to 24 in the new sequence. Re-sequencing of a 2-year-old "lab culture" using Nanopore sequencing confirmed the overall stability of the P. furiosus DSM 3638 genome even under normal lab conditions without taking any special care. To improve genome annotation, the updated DNA sequence was combined with an RNA sequencing approach. Here, RNAs from eight different growth conditions were pooled to increase the number of detected transcripts. Furthermore, a differential RNA-Seq approach was employed for the identification of transcription start sites (TSSs). In total, 2515 TSSs were detected and classified into 834 primary (pTSS), 797 antisense (aTSS), 739 internal and 145 secondary TSSs. Our analysis of the upstream regions revealed a well conserved archaeal promoter structure. Interrogation of the distances between pTSSs and aTSSs revealed a significant number of antisense transcripts, which are a result of bidirectional transcription from the same TATA box. This mechanism of antisense transcript production could be further confirmed by in vitro transcription experiments. We assume that bidirectional transcription gives rise to non-functional antisense RNAs and that this is a widespread phenomenon in archaea due to the architecture of the TATA element and the symmetric structure of the TATA-binding protein.
Related Concept Videos
 03:21
RNA-seq
Before the discovery of RNA-seq, microarray-based methods and Sanger sequencing were used for transcriptome analysis. However, while...
 03:36
Genome Annotation and Assembly
 02:35
Transcription Elongation Factors
The transcription elongation is regulated via pausing of RNA polymerase on several occasions during transcription. In bacteria, these halts are necessary because the transcription of DNA into mRNA is coupled to the translation of that mRNA...
 01:10
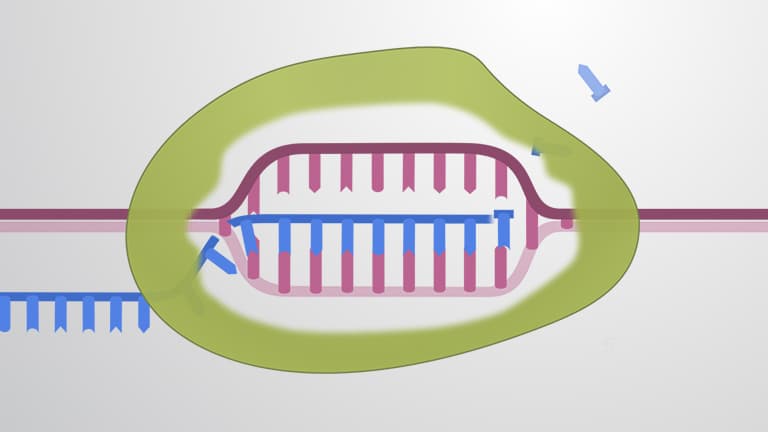
Transcription
Transcription is the process of synthesizing RNA from a DNA sequence by RNA polymerase. It is the first step in producing a protein from a gene sequence. Additionally, many other proteins and regulatory sequences are involved in the proper synthesis of messenger RNA (mRNA). Regulation of transcription is responsible for the differentiation of all the different types of cells and often for the proper cellular response to environmental signals.
Transcription Can Produce Different Kinds...
 02:16
Transcription Factors
 02:02
Genomics