Effect of food on the serum concentration profile of enteric-coated valproic acid

J H Fischer1, A N Barr, F P Paloucek

  • 1Clinical Pharmacokinetics Laboratory, College of Pharmacy, University of Illinois, Chicago 60612.

Neurology
|August 1, 1988
PubMed

Related Concept Videos

Drug Absorption: Factors Affecting GI Absorption01:19

Drug Absorption: Factors Affecting GI Absorption

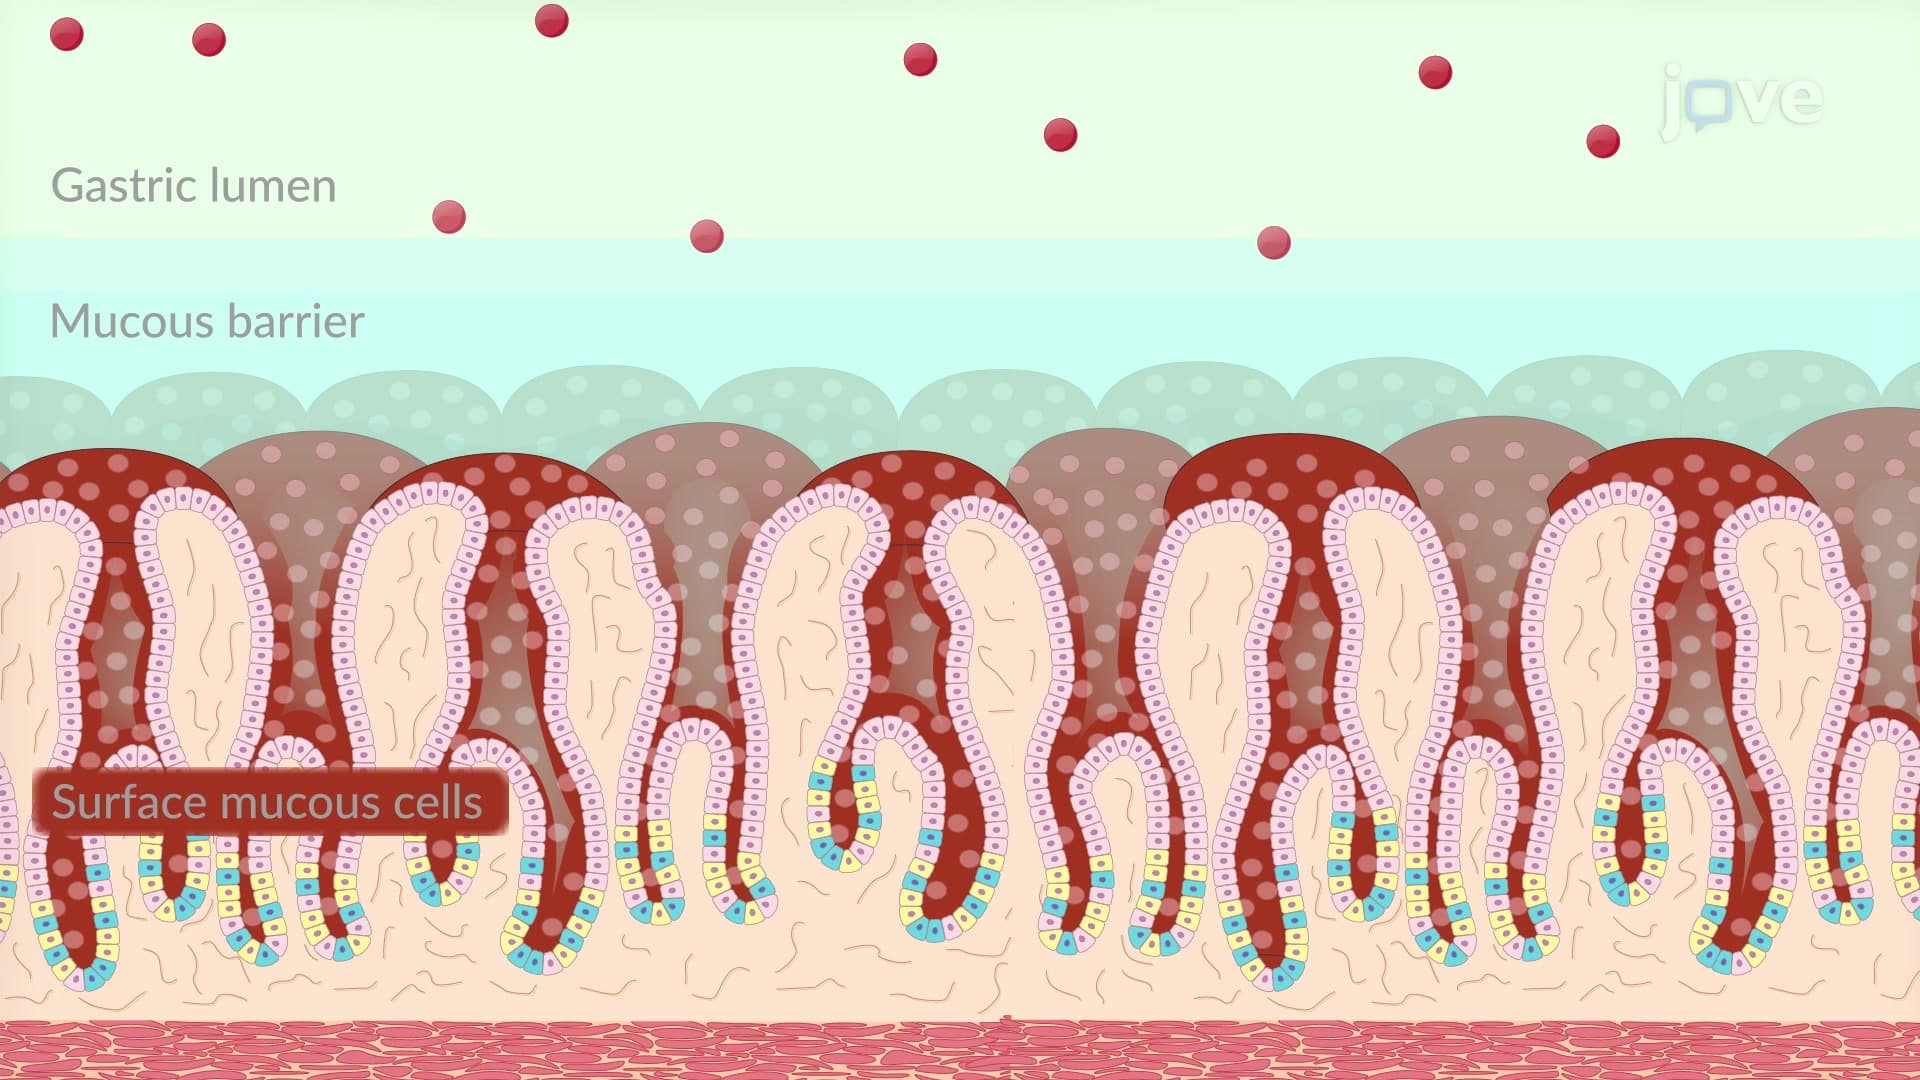
The process of oral drug absorption can be influenced by several factors. Weakly acidic drugs tend to be absorbed more readily from the stomach due to their nonionized state. However, absorption may be less efficient in the upper intestine, where drugs are often ionized. Interestingly, despite the stomach's apparent advantage for drug absorption, its mucous layer can hinder diffusion. Its surface area is also smaller than the intestine's, which can further slow down the absorption rate.
In...
Time Course of Drug Effect01:14

Time Course of Drug Effect

The progression of a drug's impact can be analyzed by examining both the concentration-time course and the effect-time course. The concentration-time course is determined by the drug's half-life and is influenced by factors such as its pharmacokinetics, including absorption, distribution, metabolism, and elimination. The effect of the drug is often related to its concentration in the plasma and is calculated using the maximum drug effect and the plasma concentration that generates 50 percent of...
Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH01:21

Factors Affecting Dissolution: Drug pKa, Lipophilicity and GI pH

Drug absorption within the gastrointestinal (GI) tract is a complex process influenced by several critical factors, including the site pH, the drug's dissociation constant (pKa), and the drug's lipophilicity. The GI tract exhibits a pH gradient, with an acidic environment in the stomach and a more alkaline environment in the small intestine. This pH variation directly affects the ionization state of drugs.
A drug's pKa and the pH of the gastrointestinal (GI) tract play crucial roles in drug...
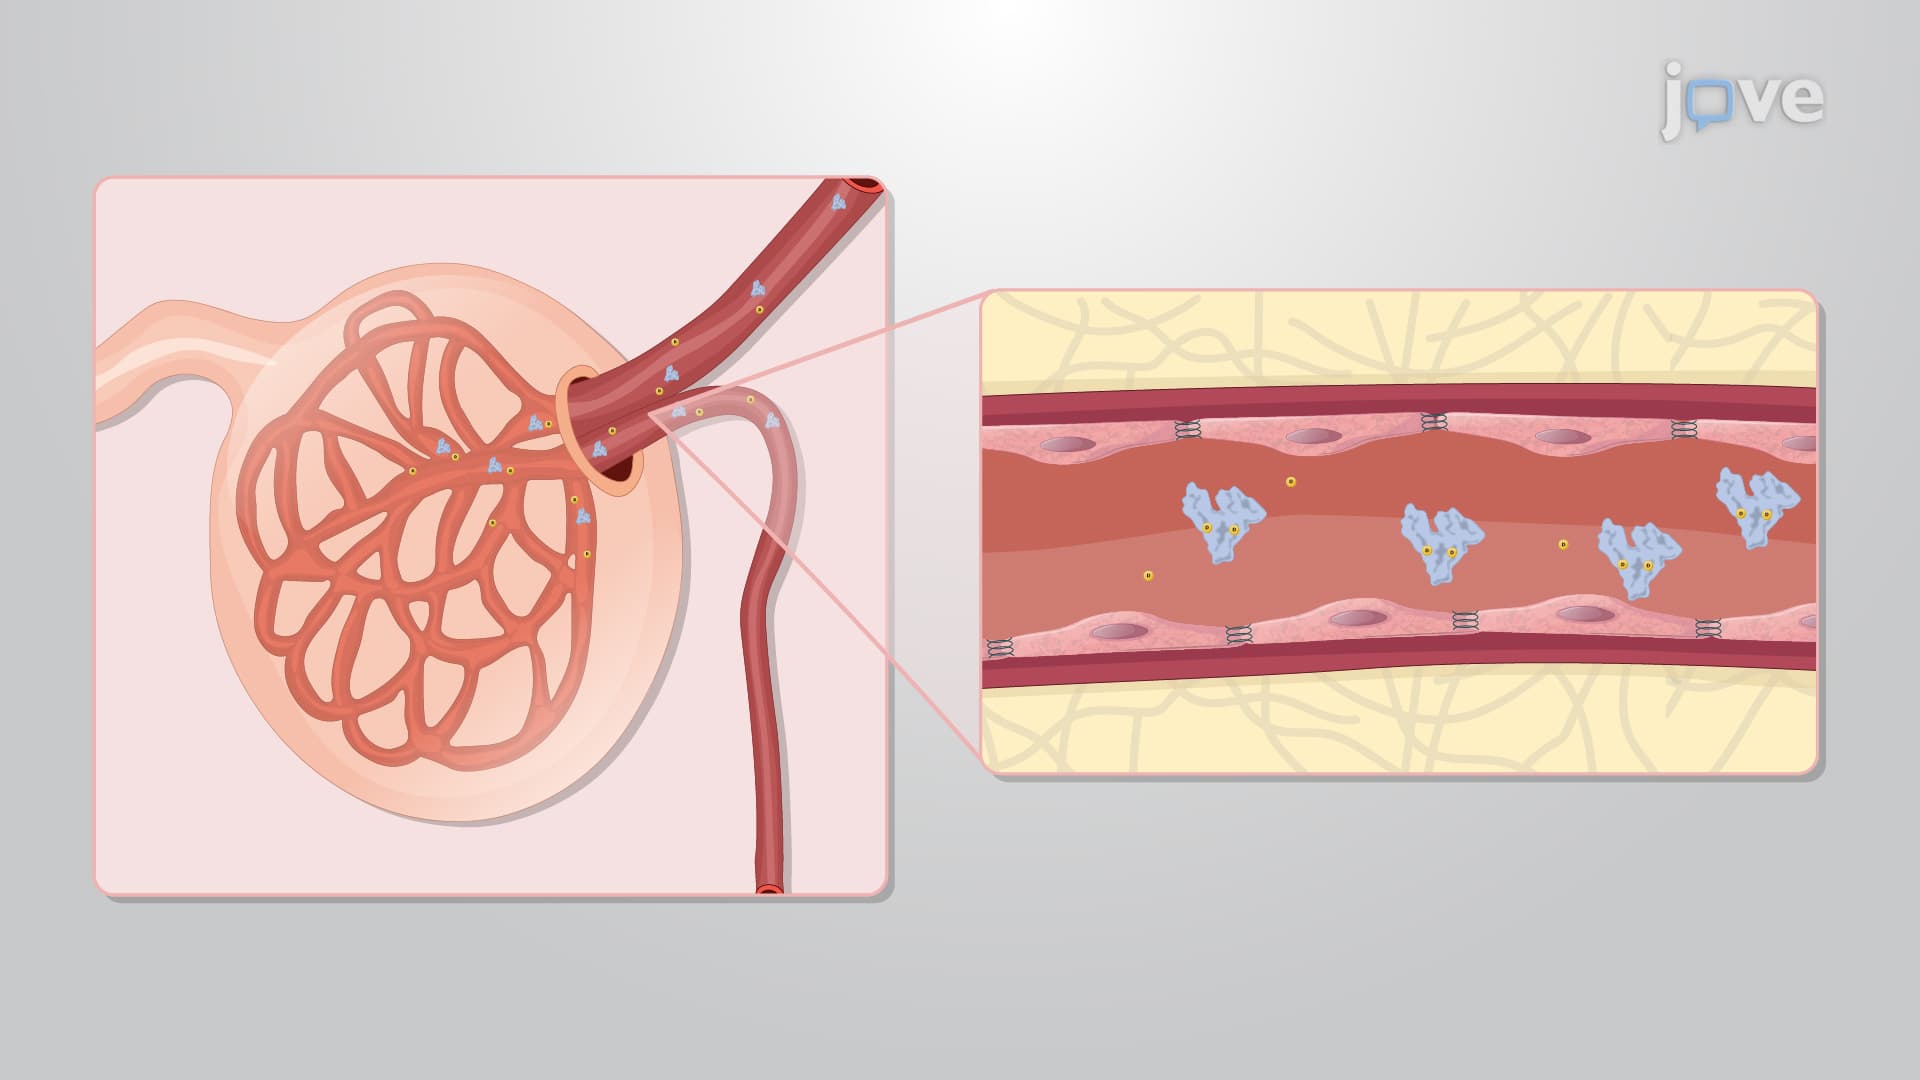
Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding01:22

Nonlinear Pharmacokinetics: Bioavailability and Protein-Drug Binding

When a drug follows nonlinear pharmacokinetics, its bioavailability, the amount of the drug that reaches the systemic circulation, can change with different doses. This is due to the presence of a saturable pathway. The pathway becomes saturated as the drug concentration increases, decreasing the absorption rate. Consequently, the drug's bioavailability may be lower than expected at higher doses.
To quantify the extent of bioavailability, pharmacologists often use a parameter called .
Bioavailability: Influencing Factors01:22

Bioavailability: Influencing Factors
Bioavailability refers to the extent and rate at which a drug reaches systemic circulation in its active form. Extent refers to the amount of the drug that makes it into circulation, while rate is the speed at which it enters circulation. It is influenced by several factors critical for optimizing drug formulations, dosing regimens, and therapeutic outcomes.Physicochemical properties of drugs and formulationsThe solubility, stability, and dissolution rate of a drug significantly impact its...
Pharmacokinetics: Drug–Food and Drug–Viral Interactions01:26

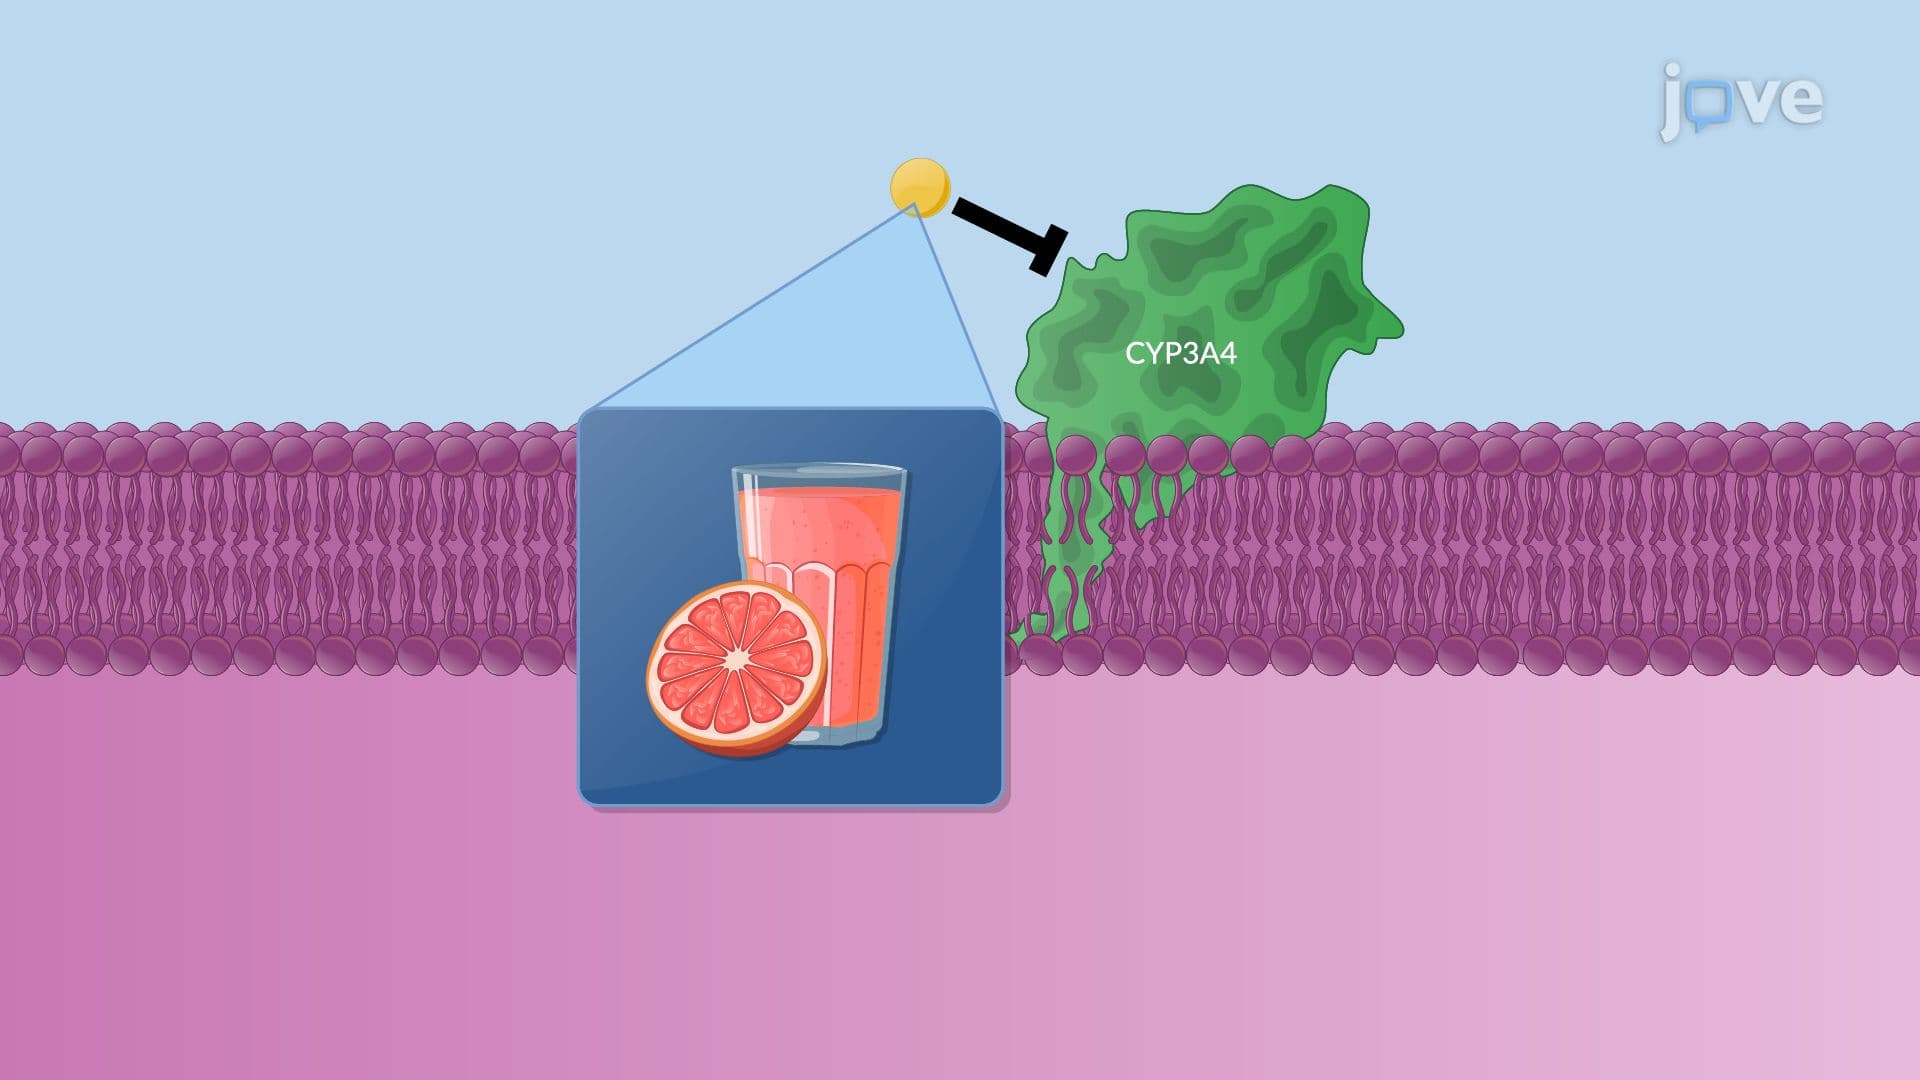
Pharmacokinetics: Drug–Food and Drug–Viral Interactions

A drug interaction occurs when the concurrent use of another drug, food, or an external substance alters the pharmacological activity of a drug. This interaction can modify the action of the original drug, affecting its effectiveness and safety.Drug–food interactions are significant as they impact drug absorption, metabolism, and excretion. For example, grapefruit juice is a well-known disruptor of drug metabolism. It inhibits the cytochrome P450 3A4 enzyme, crucial for the metabolism of many...