Related Experiment Video
Updated: Jan 20, 2026

Using Scaffold Liposomes to Reconstitute Lipid-proximal Protein-protein Interactions In Vitro
Published on: January 11, 2017
Assessing the performance of the MM/PBSA and MM/GBSA methods. 10. Impacts of enhanced sampling and variable
Ercheng Wang1, Gaoqi Weng1, Huiyong Sun1
1Hangzhou Institute of Innovative Medicine, College of Pharmaceutical Sciences, Zhejiang University, Hangzhou, Zhejiang 310058, China. tingjunhou@zju.edu.cn.

Abstract:
Enhanced sampling has been extensively used to capture the conformational transitions in protein folding, but it attracts much less attention in the studies of protein-protein recognition. In this study, we evaluated the impact of enhanced sampling methods and solute dielectric constants on the overall accuracy of the molecular mechanics/Poisson-Boltzmann surface area (MM/PBSA) and molecular mechanics/generalized Born surface area (MM/GBSA) approaches for the protein-protein binding free energy calculations. Here, two widely used enhanced sampling methods, including aMD and GaMD, and conventional molecular dynamics (cMD) simulations with two AMBER force fields (ff03 and ff14SB) were used to sample the conformations for 21 protein-protein complexes. The MM/PBSA and MM/GBSA calculation results illustrate that the standard MM/GBSA based on the cMD simulations yields the best Pearson correlation (rp = -0.523) between the predicted binding affinities and the experimental data, which is much higher than that given by MM/PBSA (rp = -0.212). Two enhanced sampling methods (aMD and GaMD) are indeed more efficient for conformational sampling, but they did not improve the binding affinity predictions for protein-protein systems, suggesting that the aMD or GaMD sampling (at least in short timescale simulations) may not be a good choice for the MM/PBSA and MM/GBSA predictions of protein-protein complexes. The solute dielectric constant of 1.0 is recommended to MM/GBSA, but a higher solute dielectric constant is recommended to MM/PBSA, especially for the systems with higher polarity on the protein-protein binding interfaces. Then, a preliminary assessment of the MM/GBSA calculations based on a variable dielectric generalized Born (VDGB) model was conducted. The results highlight the potential power of VDGB in the free energy predictions for protein-protein systems, but more thorough studies should be done in the future.
More Related Videos
 05:43
A Protein Preparation Method for the High-throughput Identification of Proteins Interacting with a Nuclear Cofactor Using LC-MS/MS Analysis
Published on: January 24, 2017
 10:20
Method for Identifying Small Molecule Inhibitors of the Protein-protein Interaction Between HCN1 and TRIP8b
Published on: November 11, 2016
Related Concept Videos
 02:04
Protein-protein Interfaces
 02:15
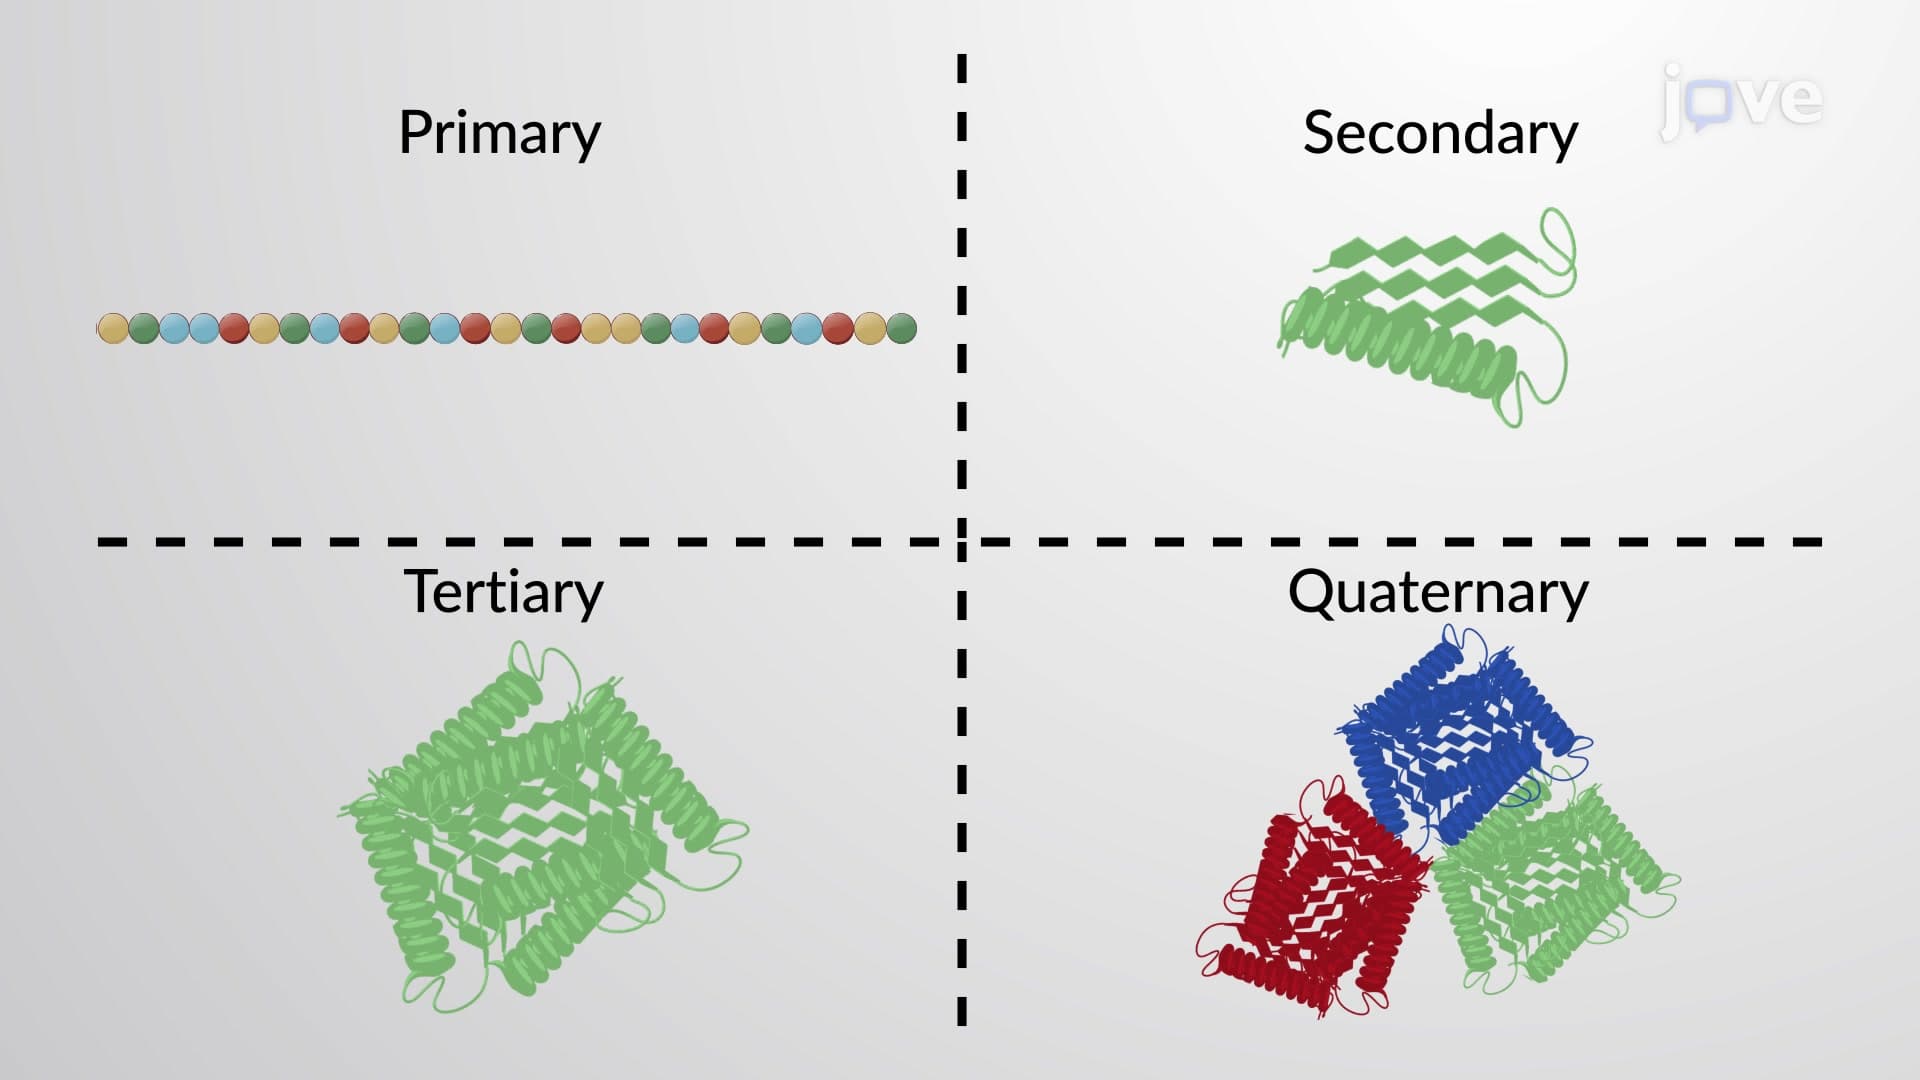
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to...
 02:26
Protein Networks
These interactions can be represented through maps depicting protein-protein interaction networks, represented as nodes and edges. Nodes are circles that are representative of a protein,...
 01:55
What are Proteins?
 02:36
RNA Polymerase II Accessory Proteins