Related Experiment Video
Updated: Jan 20, 2026

Robotics and Dynamic Image Analysis for Studies of Gene Expression in Plant Tissues
Published on: May 5, 2010
fagin: synteny-based phylostratigraphy and finer classification of young genes
Zebulun Arendsee1,2,3, Jing Li1,2, Urminder Singh1,2,3
1Department of Genetics Development and Cell Biology, Iowa State University, Ames, IA, 50010, USA.

Background:
With every new genome that is sequenced, thousands of species-specific genes (orphans) are found, some originating from ultra-rapid mutations of existing genes, many others originating de novo from non-genic regions of the genome. If some of these genes survive across speciations, then extant organisms will contain a patchwork of genes whose ancestors first appeared at different times. Standard phylostratigraphy, the technique of partitioning genes by their age, is based solely on protein similarity algorithms. However, this approach relies on negative evidence ─ a failure to detect a homolog of a query gene. An alternative approach is to limit the search for homologs to syntenic regions. Then, genes can be positively identified as de novo orphans by tracing them to non-coding sequences in related species.
Results:
We have developed a synteny-based pipeline in the R framework. Fagin determines the genomic context of each query gene in a focal species compared to homologous sequence in target species. We tested the fagin pipeline on two focal species, Arabidopsis thaliana (plus four target species in Brassicaseae) and Saccharomyces cerevisiae (plus six target species in Saccharomyces). Using microsynteny maps, fagin classified the homology relationship of each query gene against each target genome into three main classes, and further subclasses: AAic (has a coding syntenic homolog), NTic (has a non-coding syntenic homolog), and Unknown (has no detected syntenic homolog). fagin inferred over half the "Unknown" A. thaliana query genes, and about 20% for S. cerevisiae, as lacking a syntenic homolog because of local indels or scrambled synteny.
Conclusions:
fagin augments standard phylostratigraphy, and extends synteny-based phylostratigraphy with an automated, customizable, and detailed contextual analysis. By comparing synteny-based phylostrata to standard phylostrata, fagin systematically identifies those orphans and lineage-specific genes that are well-supported to have originated de novo. Analyzing within-species genomes should distinguish orphan genes that may have originated through rapid divergence from de novo orphans. Fagin also delineates whether a gene has no syntenic homolog because of technical or biological reasons. These analyses indicate that some orphans may be associated with regions of high genomic perturbation.
More Related Videos
 07:35
Selecting Multiple Biomarker Subsets with Similarly Effective Binary Classification Performances
Published on: October 11, 2018
 06:22
Machine Learning-Based Cough Tone Classification: Diagnostic Exploration of Chronic Obstructive Pulmonary Disease and Respiratory Tract Infections
Published on: September 19, 2025
Related Concept Videos
 02:31
Synteny and Evolution
Around 80 million years ago, the human and mice lineages diverged from the common ancestor. During the course of evolution, the ancestral...
 01:01
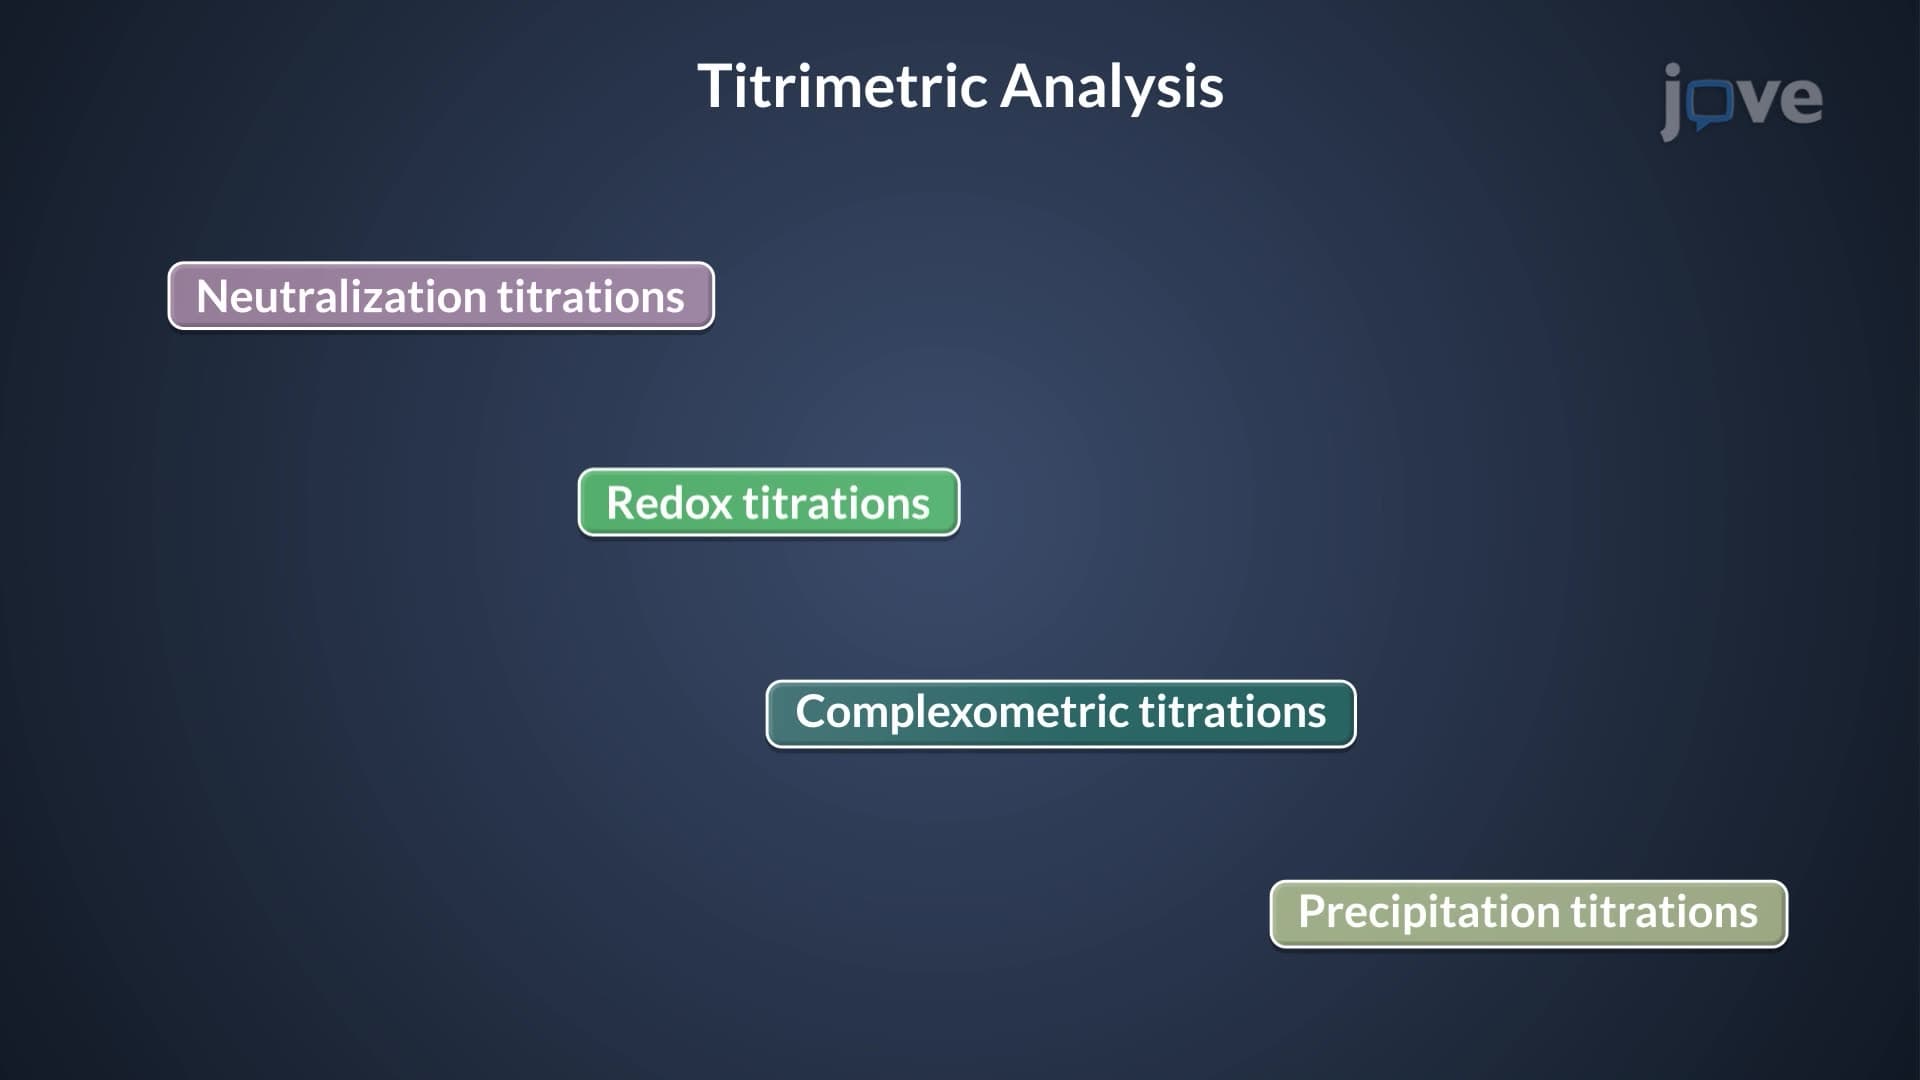
Classification of Titrimetric Analysis Based on Reaction Types
Titrations between an acid and a base lead to neutralization reactions that form...
 01:18
Cardiovascular Drugs: Classification based on Therapeutic Indications
 01:30
Classification of Neurotransmitters
 01:30
Classification of Leukocytes
Neutrophils are the most abundant type of granular leukocytes, comprising 50-70% of all leukocytes. They feature small, evenly distributed granules and a...
 01:18
Classification of Bones
Long and Short Bones
The appendicular skeleton, particularly the upper and lower limbs, is primarily made of long and short bones. The...