Related Experiment Video
Updated: Jan 20, 2026

Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
Performance of Density-Functional Tight-Binding in Comparison to Ab Initio and First-Principles Methods for Isomer
Ka Hung Lee1,2, Udo Schnupf3, Bobby G Sumpter4,4
1Department of Chemistry, Graduate School of Science, Nagoya University, Nagoya 464-8602, Japan.

Abstract:
Density functional theory (DFT) is a widely used methodology for the computation of molecular and electronic structure, and we confirm that B3LYP and the high-level ab initio G3B3 method are in excellent agreement for the lowest-energy isomers of the 16 glucose epimers. Density-functional tight-binding (DFTB) is an approximate version of DFT with typically comparable accuracy that is 2 to 3 orders of magnitude faster, therefore generally very suitable for processing large numbers of complex structures. Conformational isomerism in sugars is well known to give rise to a large number of isomer structures. On the basis of a comprehensive study of glucose epimers in vacuo and aqueous solution, we found that the performance of DFTB is on par to B3LYP in terms of geometrical parameters excluding hydrogen bonds and isomer energies. However, DFTB underestimates both hydrogen bonding interactions as well as torsional barriers associated with rotations of the hydroxy groups, resulting in a counterintuitive overemphasis of hydrogen bonding in both gas phase as well as in water. Although the associated root mean squared deviation from B3LYP within epimer isomer groups is only on the order of 1 kcal/mol, this deviation affects the correct assignment of major isomer ordering, which span less than 10 kcal/mol. Both second- as well as third-order DFTB methods are exhibiting similar deviations from B3LYP. Even after the inclusion of empirical dispersion corrections in vacuum, these deviations remain for a large majority of isomer energies and geometries when compared to dispersion-corrected B3LYP.
More Related Videos
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
 10:27
Contrast-Matching Detergent in Small-Angle Neutron Scattering Experiments for Membrane Protein Structural Analysis and Ab Initio Modeling
Published on: October 21, 2018
Related Concept Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
 10:27
Contrast-Matching Detergent in Small-Angle Neutron Scattering Experiments for Membrane Protein Structural Analysis and Ab Initio Modeling
 02:13
Nuclear Binding Energy
 01:05
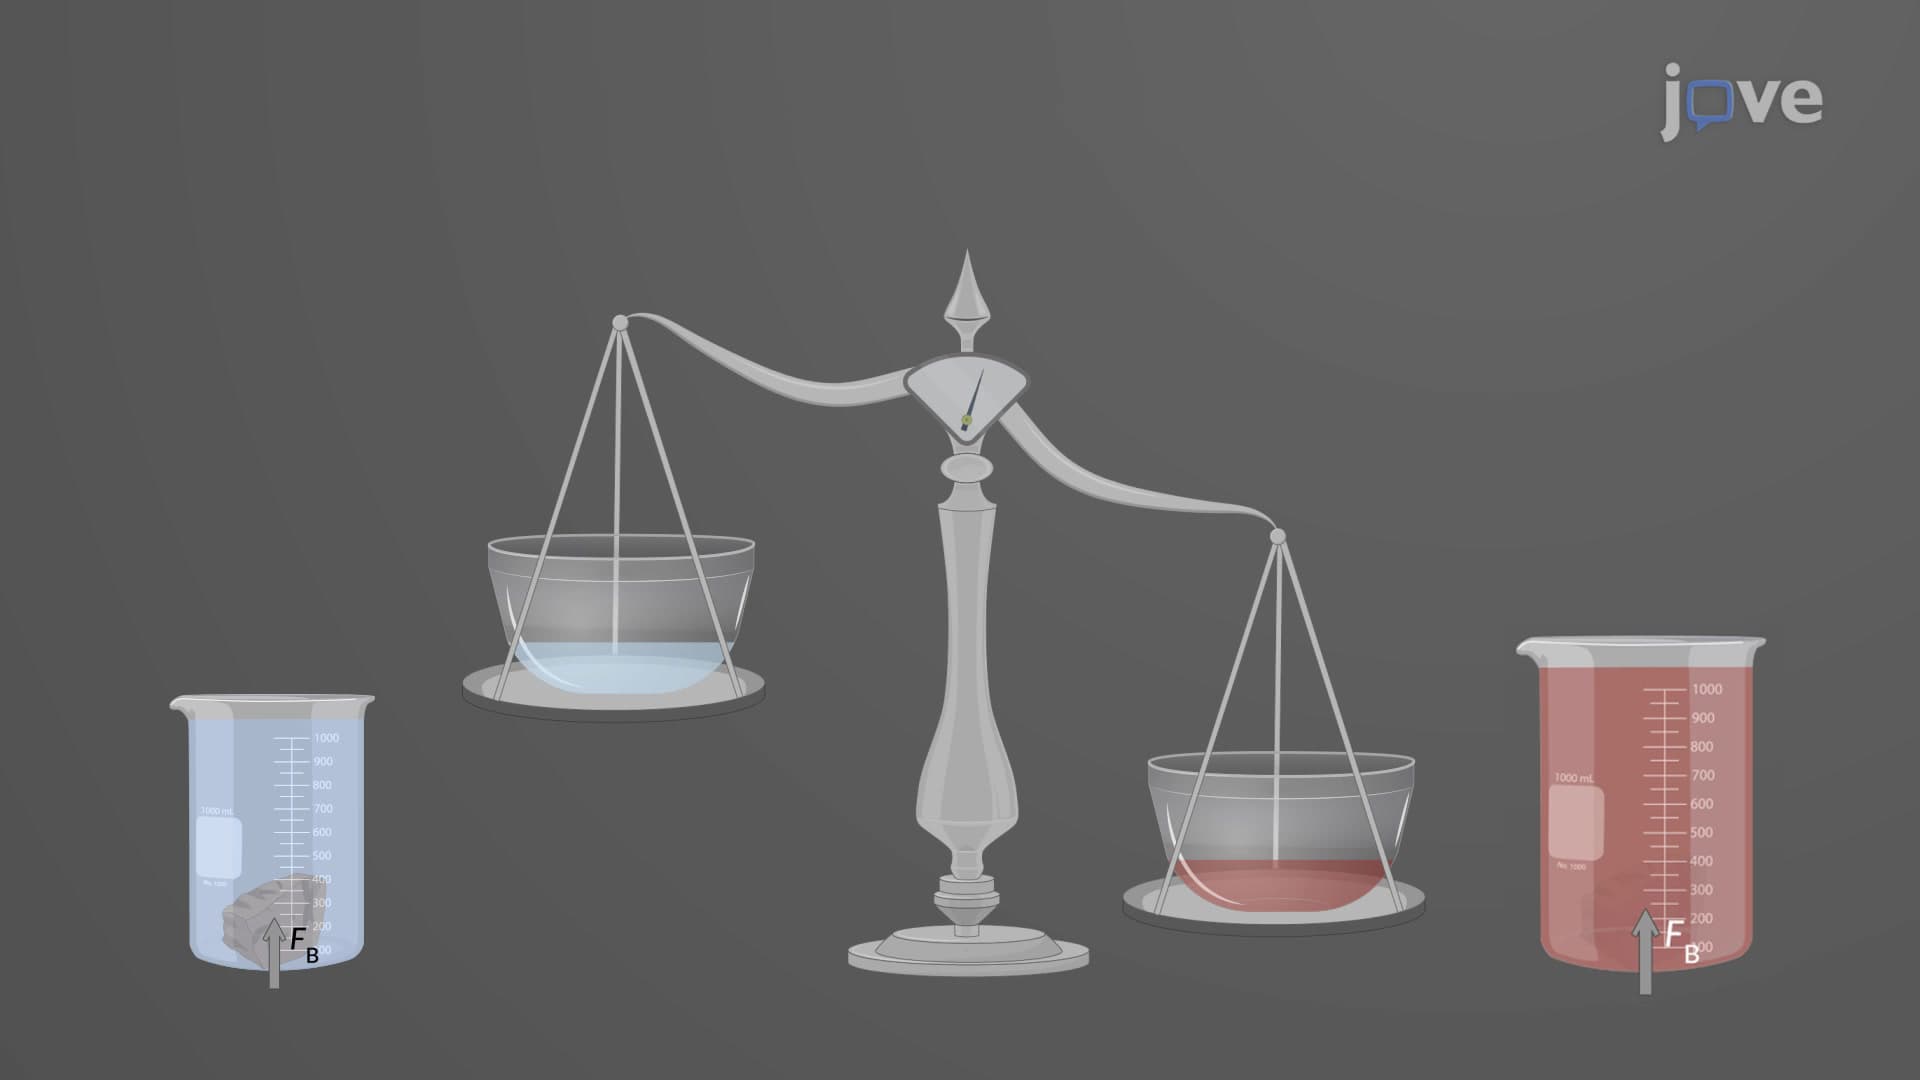
Density and Archimedes' Principle
 01:20
Strain-Energy Density
In the elastic region of a material, the relationship between the stress and the strain is linear and follows Hooke's Law. The strain energy density in this region...