Related Experiment Video
Updated: Jan 20, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Prediction of Accurate Binding Modes Using Combination of Classical and Accelerated Molecular Dynamics and
Filip Fratev1,2, Thomas Steinbrecher3, Svava Ósk Jónsdóttir4
1Department of Pharmaceutical Sciences, School of Pharmacy, The University of Texas at El Paso, 1101 N Campbell Street, El Paso, Texas 79902, United States.

Abstract:
Estimating the correct binding modes of ligands in protein-ligand complexes is crucial not only in the drug discovery process but also for elucidating potential toxicity mechanisms. In the current paper, we propose a computational modeling workflow using the combination of docking, classical molecular dynamics (cMD), accelerated molecular dynamics (aMD) and free-energy perturbation (FEP+ protocol) for identification of possible ligand binding modes. It was applied for investigation of selected perfluorocarboxyl acids (PFCAs) in the PPARγ nuclear receptor. Although both regular and induced fit docking failed to reproduce the experimentally determined binding mode of the ligands when docked into a non-native X-ray structure, cMD and aMD simulations successfully identified the most probable binding conformations. Moreover, multiple binding modes were identified for all of these compounds and the shorter-chain PFCAs continuously moved between a few energetically favorable binding conformations. On the basis of MD predictions of binding conformations, we applied the default and also redesigned FEP+ sampling protocols, which accurately reproduced experimental differences in the binding energies. Thus, the preliminary MD simulations can also provide helpful information about correct setup of the FEP+ calculations. These results show that the PFCA binding modes were accurately predicted and that the FEP+ protocol can be used to estimate free energies of binding of flexible ligands that are not typical druglike compounds. Our in silico workflow revealed the specific ligand-residue interactions within the ligand binding domain and the main characteristics of the PFCAs, and it was concluded that these compounds are week PPARγ partial agonists. This work also suggests a common pipeline for identification of ligand binding modes, ligand-protein dynamics description, and relative free-energy calculations.

Related Concept Videos
 10:29
Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
 02:13
Nuclear Binding Energy
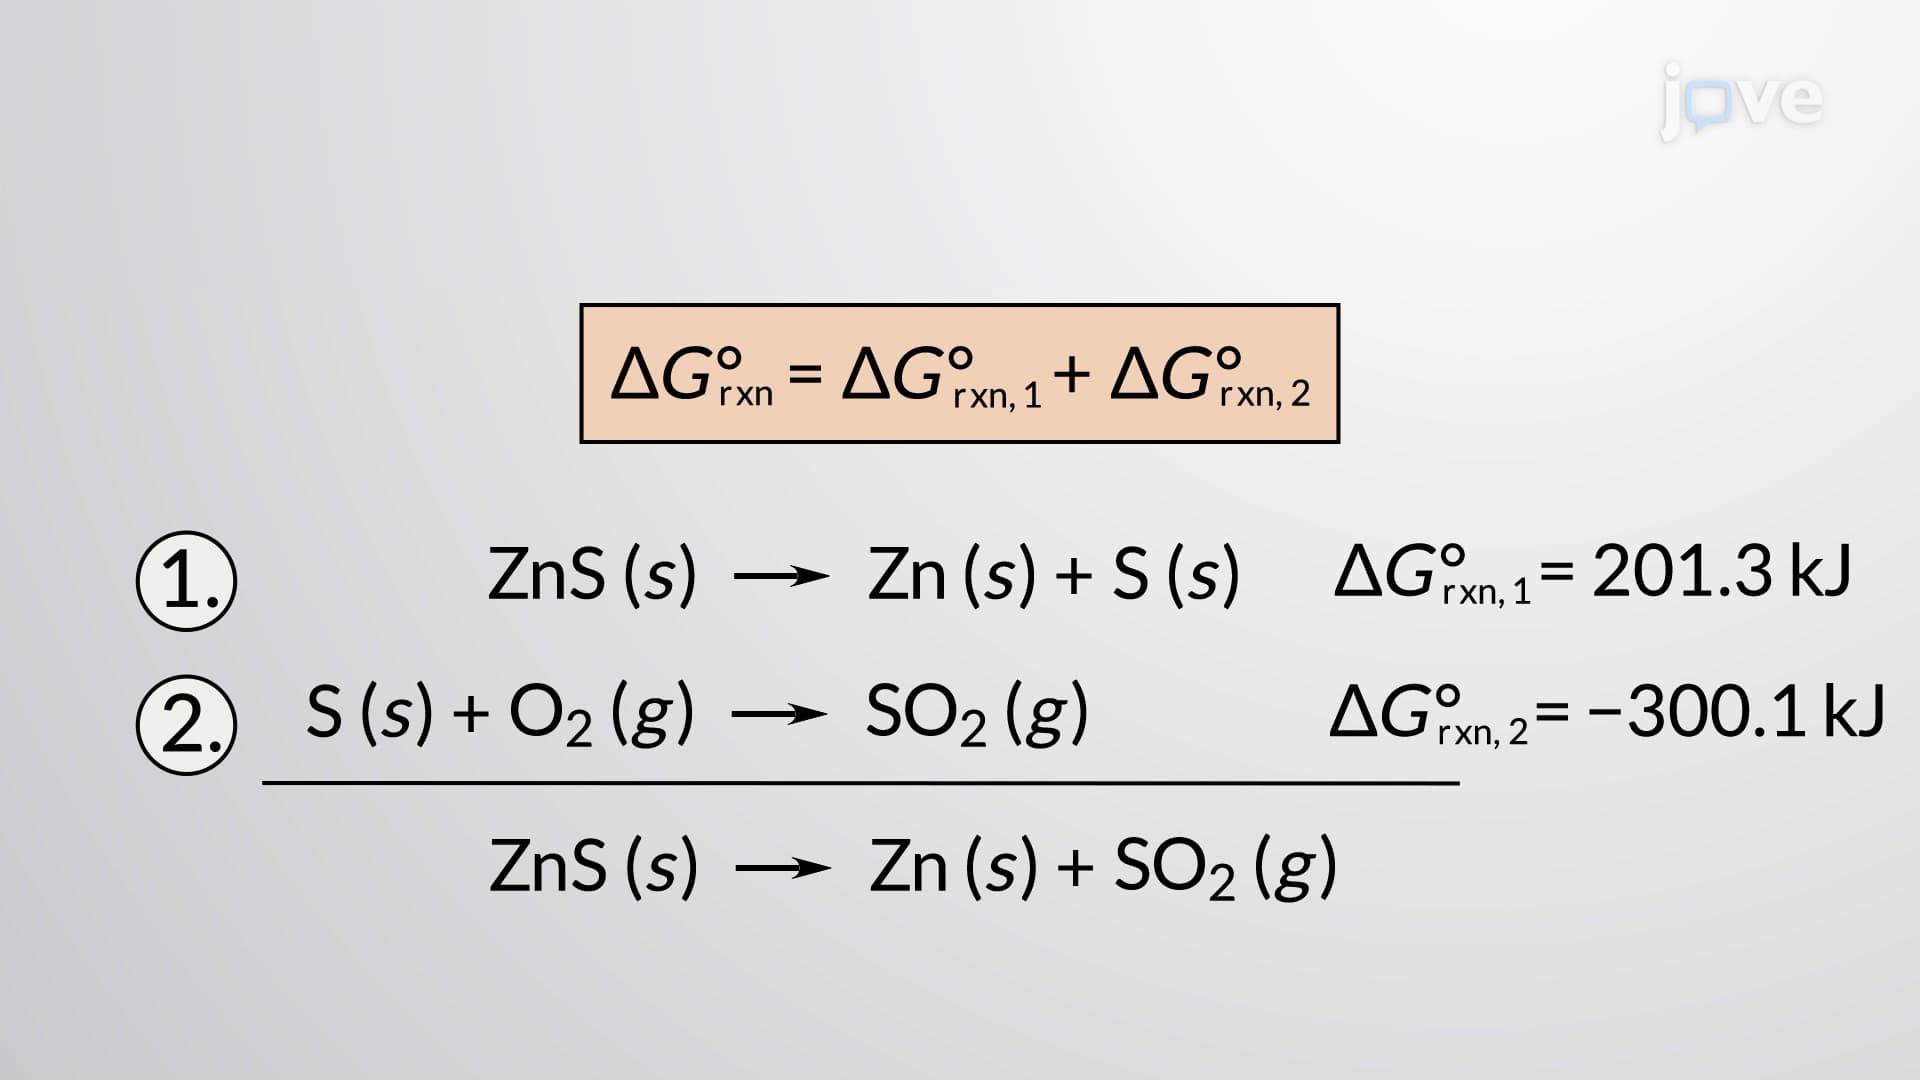 02:49
Calculating Standard Free Energy Changes
 02:27
Predicting Molecular Geometry
 03:15
Energy Dynamics
Energy is one of the most important abiotic factors in an ecosystem and organisms in an ecosystem are connected by the flow of energy and matter among one another. Since energy can be neither created nor destroyed, it can only change form or be transferred to the next organism in a food chain. For example, every time a cow grazes on grass or an osprey hunts and consumes fish, energy is transferred from the consumed organism to the consumer. Each of these interactions in a food...
 05:16
Energy Dynamics
ExpandNOTE: In this lab you will grow caterpillars in conditions with and without food to examine the transfer of biomass in a typical trophic system.
Weigh and record the wet mass of enough leaves of cabbage to line the bottom of a plastic tub.
Then, collect two batches of five caterpillars from their cabbage habitat, and weigh and record the wet mass of each batch separately in individual weigh boats. Make sure to tare the balance before each measurement....