Related Experiment Video
Updated: Jan 19, 2026

Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Evaluating the strengths of salt bridges in the CutA1 protein using molecular dynamic simulations: a comparison of
Yoshinori Matsuura1, Yasumasa Joti1,2, Bagautdin Bagautdinov2
1RIKEN SPring-8 Center, Sayo, Hyogo, Japan.

Abstract:
Ion-ion interactions (salt bridges) between favorable pairs of charged residues are important for the conformational stability of proteins. Molecular dynamic (MD) simulations are useful for elucidating the interactions among charged residues fluctuating in solution. However, the quality of MD results depends strongly on the force fields used. In this study, we compared the strengths of salt bridges among force fields by performing MD simulations using the CutA1 protein (trimer) from the hyperthermophile Pyrococcus horikoshii (PhCutA1), which has an unusually large proportion of charged residues. The force fields Chemistry at HARvard Macromolecular Mechanics (Charmm)27, Assisted Model Building and Energy Refinement (Amber)99sb, Amber14sb, GROningen Molecular Simulation (Gromos)43a1, and Gromos53a6 were used in combination with two different water models, tip3p (for Charmm27, Amber99sb, and Amber14sb) and simple point charge/extended (for Amber99sb, Gromos43a1, and Gromos53a6), yielding a total of six combinations. The RMSDs of all Cα atoms of PhCutA1 were similar among force fields, except for Charmm27, during 400-ns MD simulations at 300 K; however, the radius of gyration (Rg ) was greater for Amber99sb and shorter for Gromos43a1. The average strengths of salt bridges for each positively charged residue did not differ greatly among force fields, but the strengths at specific sites within the structure depended sensitively on the force field used. In the case of the Gromos group, positively charged residues could engage in favorable interactions with many more charged residues than in the other force fields, especially in loop regions; consequently, the apparent strength at each site was lower.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 08:09
A Micro-agar Salt Bridge Electrode for Analyzing the Proton Turnover Rate of Recombinant Membrane Proteins
Published on: January 7, 2019
Related Concept Videos
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
 08:09
A Micro-agar Salt Bridge Electrode for Analyzing the Proton Turnover Rate of Recombinant Membrane Proteins
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
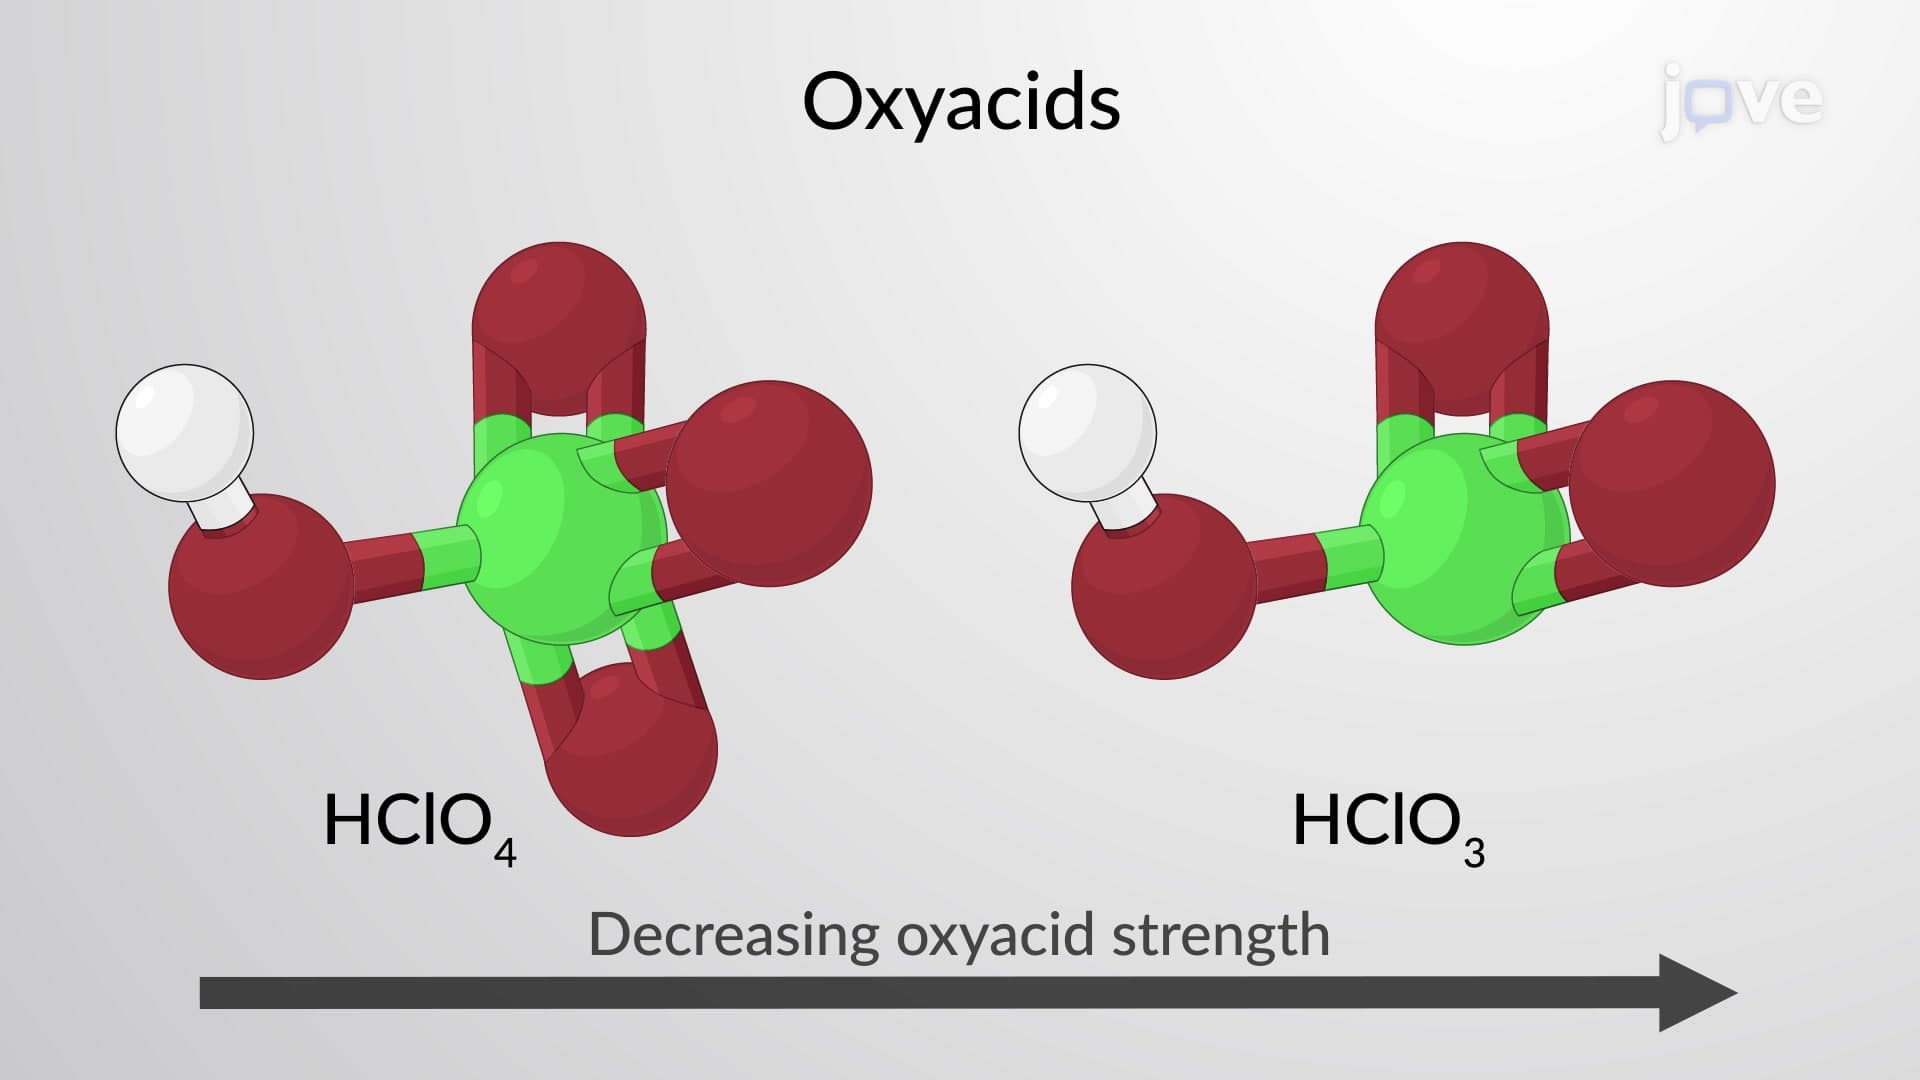 03:05
Acid Strength and Molecular Structure
In the absence of any leveling effect, the acid strength of binary compounds of hydrogen with nonmetals (A) increases as the H-A bond strength decreases down a group in the periodic table. For group 17, the order of increasing acidity is HF < HCl < HBr < HI. Likewise, for group 16, the order of increasing acid strength is H2O < H2S < H2Se < H2Te. Across a row in the periodic table, the acid strength of binary hydrogen compounds increases with increasing...