Related Experiment Video
Updated: Jan 19, 2026

Automated Sample Preparation for the Multiplexed Analysis of Single-Cell Histone Post-Translational Modifications (sc-hPTM2)
Published on: December 19, 2025
Quantitation of Single and Combinatorial Histone Modifications by Integrated Chromatography of Bottom-up Peptides and
Kevin A Janssen1,2, Mariel Coradin1,2, Congcong Lu2
1Biochemistry and Molecular Biophysics Graduate Group, Department of Biochemistry and Biophysics, Perelman School of Medicine, University of Pennsylvania, Philadelphia, PA, USA.

Abstract:
The analysis of histone post-translational modifications (PTMs) by mass spectrometry (MS) has been critical to the advancement of the field of epigenetics. The most sensitive and accurate workflow is similar to the canonical proteomics analysis workflow (bottom-up MS), where histones are digested into short peptides (4-20 aa) and quantitated in extracted ion chromatograms. However, this limits the ability to detect even very common co-occurrences of modifications on histone proteins, preventing biological interpretation of PTM crosstalk. By digesting with GluC rather than trypsin, it is possible to produce long polypeptides corresponding to intact histone N-terminal tails (50-60 aa), where most modifications reside. This middle-down MS approach is used to study distant PTM co-existence. However, the most sensitive middle-down workflow uses weak cation exchange-hydrophilic interaction chromatography (WCX-HILIC), which is less robust than conventional reversed-phase chromatography. Additionally, since the buffer systems for middle-down and bottom-up proteomics differ substantially, it is cumbersome to toggle back and forth between both experimental setups on the same LC system. Here, we present a new workflow using porous graphitic carbon (PGC) as a stationary phase for histone analysis where bottom-up and middle-down sized histone peptides can be analyzed simultaneously using the same reversed-phase buffer setup. By using this protocol for middle-down sized peptides, we identified 406 uniquely modified intact histone tails and achieved a correlation of 0.85 between PGC and WCX-HILIC LC methods. Together, our method facilitates the analysis of single and combinatorial histone PTMs with much simpler applicability for conventional proteomics labs than the state-of-the-art middle-down MS.
Related Concept Videos
 02:32
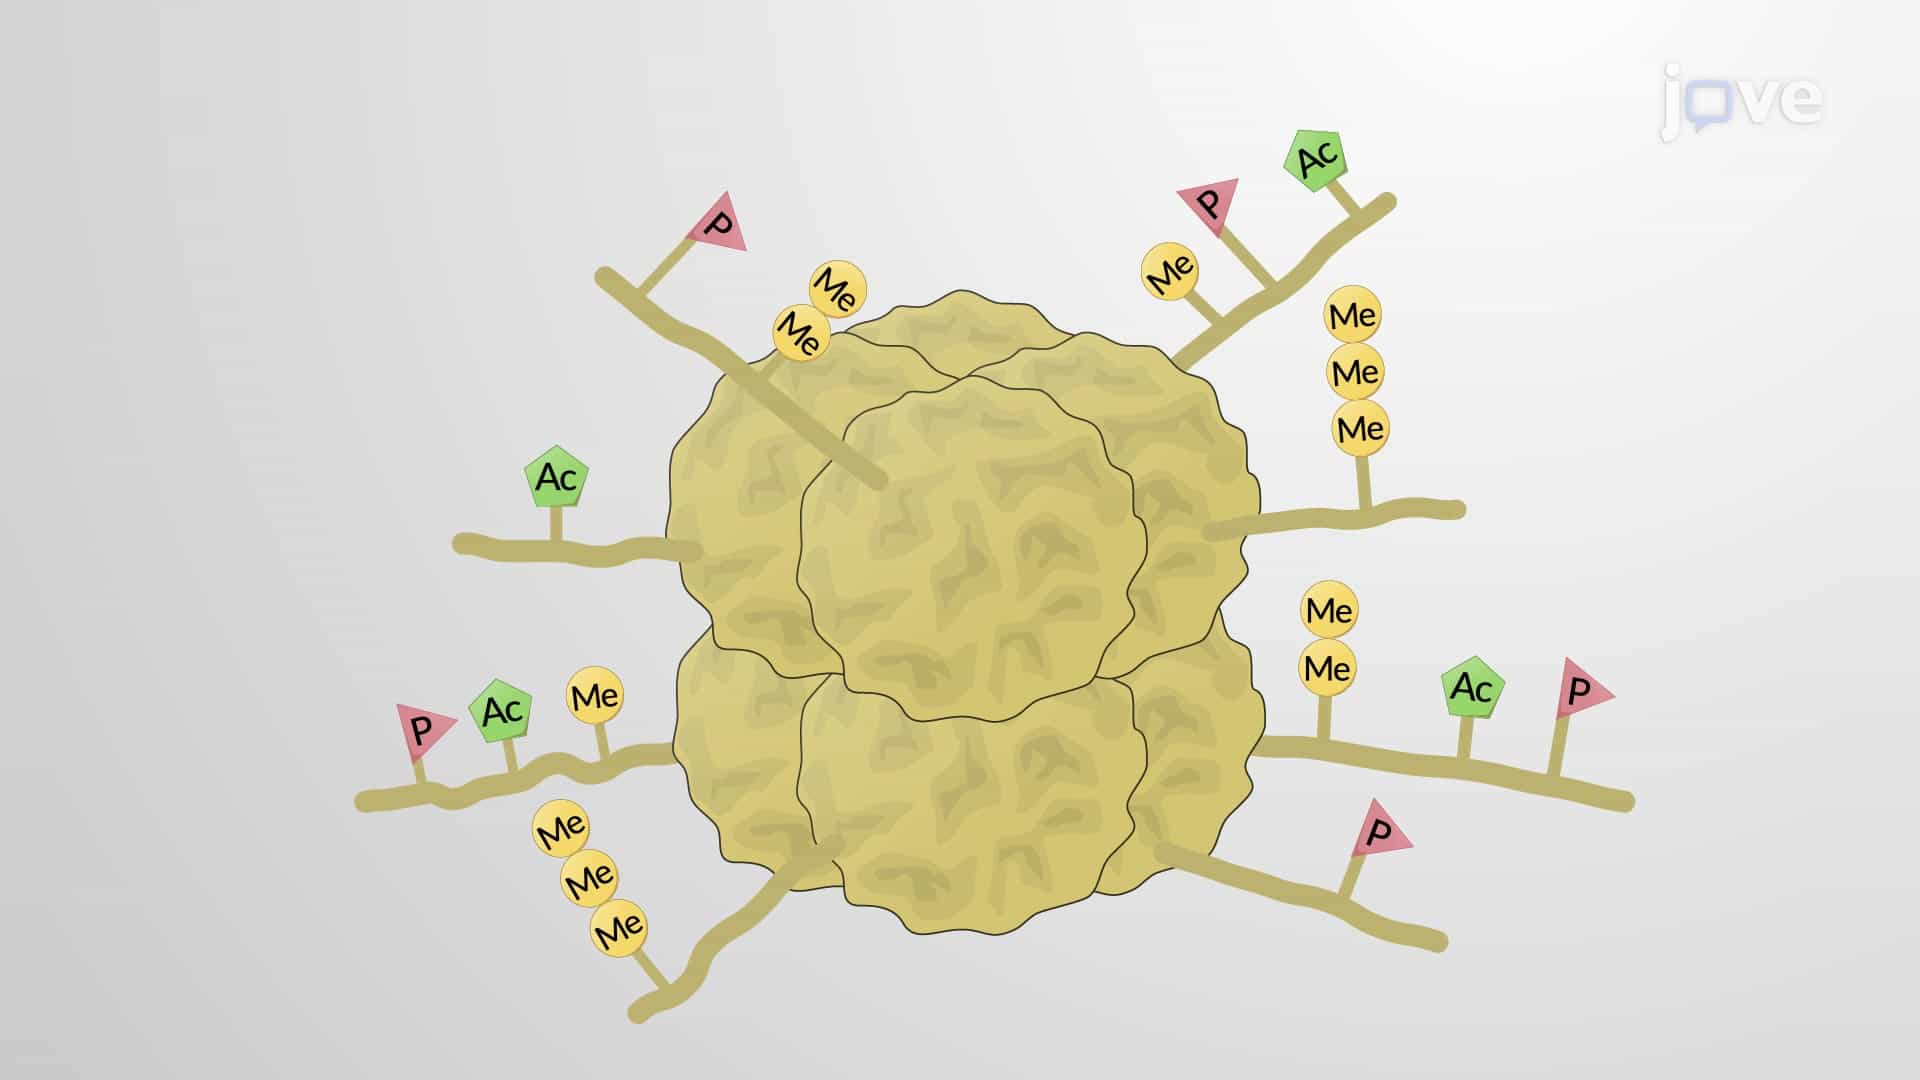
Histone Modification
Acetylation
The enzyme histone acetyltransferase adds acetyl group to the histones. Another enzyme, histone...
02:32Histone Modification
 07:21
Automated Sample Preparation for the Multiplexed Analysis of Single-Cell Histone Post-Translational Modifications (sc-hPTM2)
 11:02
Complete Workflow for Analysis of Histone Post-translational Modifications Using Bottom-up Mass Spectrometry: From Histone Extraction to Data Analysis
 09:47
Analysis of Histone Antibody Specificity with Peptide Microarrays
 05:52
Histone Modification Screening using Liquid Chromatography, Trapped Ion Mobility Spectrometry, and Time-Of-Flight Mass Spectrometry