Related Experiment Video
Updated: Jan 19, 2026
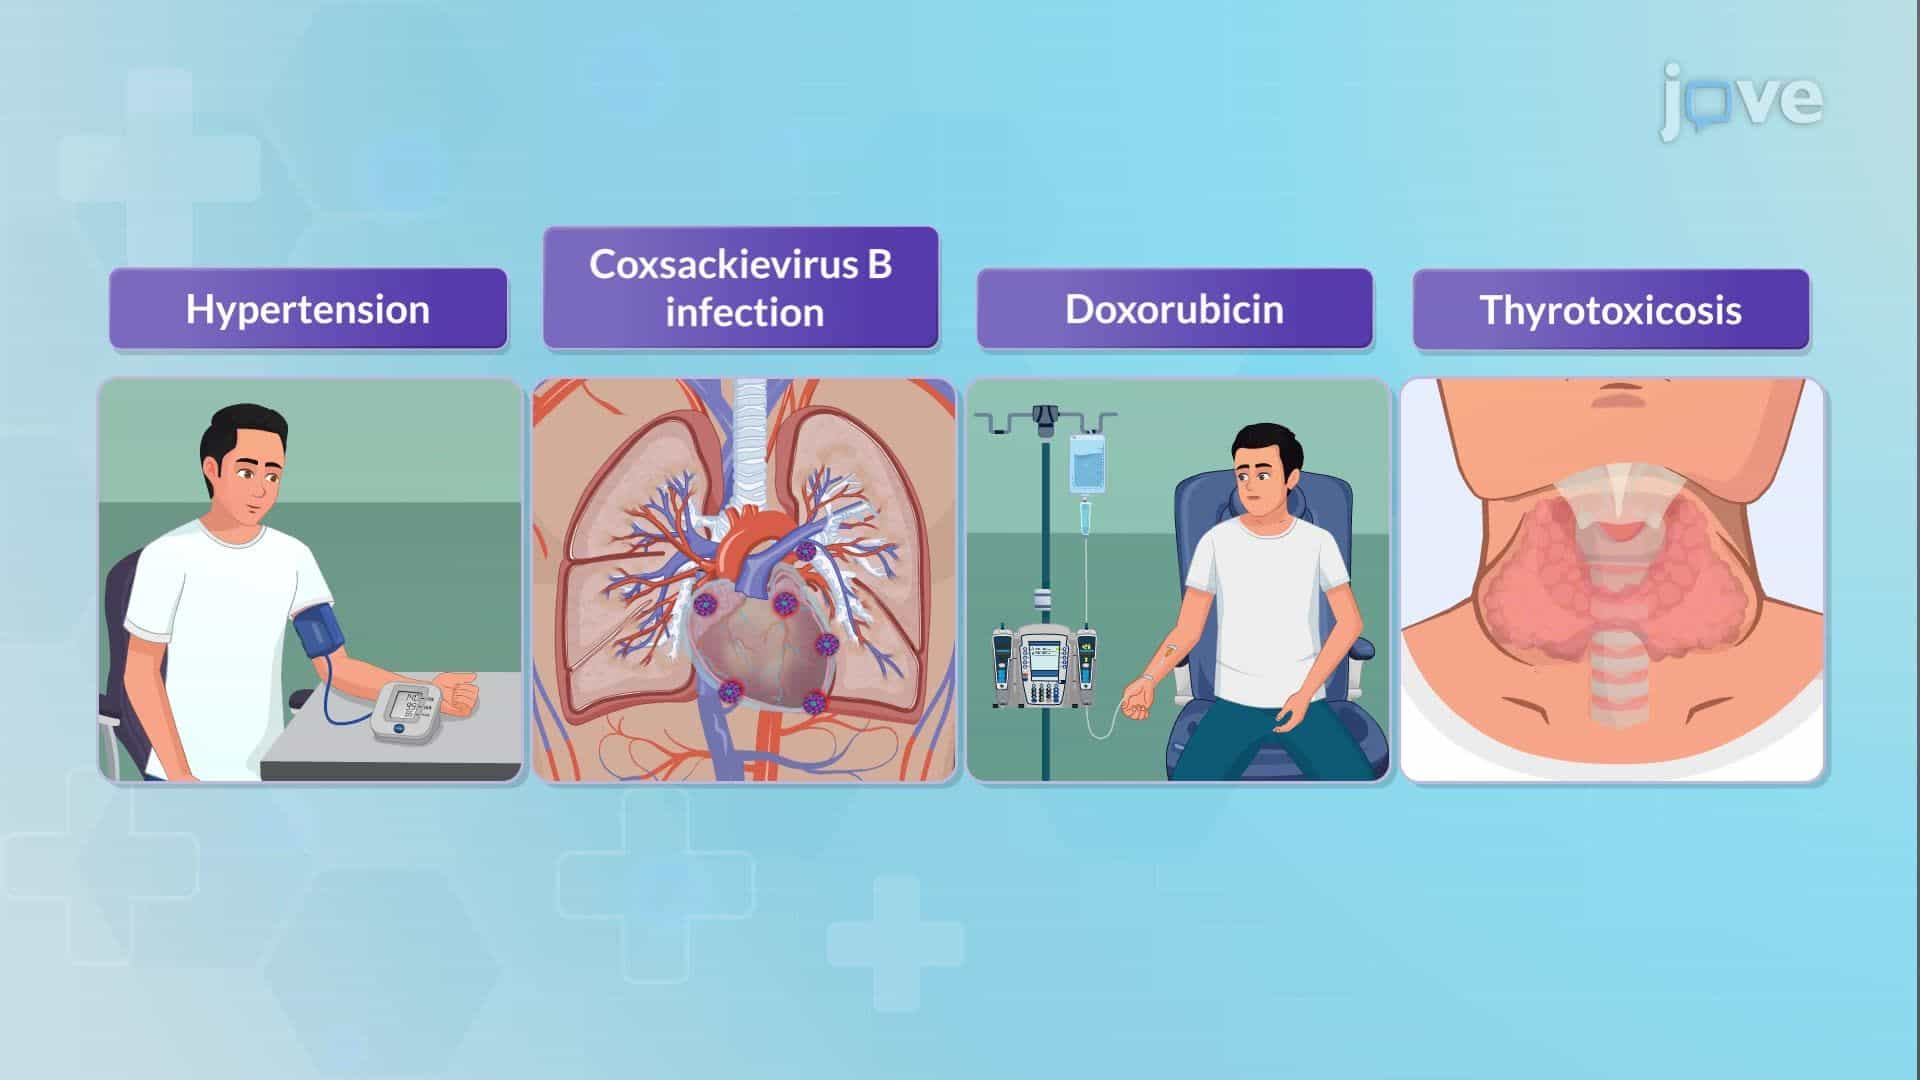

Cardiomyopathy II: Dilated Cardiomyopathy
Published on: June 19, 2025
Genetic Risk of Arrhythmic Phenotypes in Patients With Dilated Cardiomyopathy
Marta Gigli1, Marco Merlo2, Sharon L Graw3
1Cardiovascular Department, Azienda Sanitaria-Universitaria Integrata Trieste "ASUITS," Trieste, Italy; Cardiovascular Institute and Adult Medical Genetics Program, University of Colorado Anschutz Medical Campus, Aurora, Colorado.

Insights
Genetic variants in dilated cardiomyopathy (DCM) impact outcomes. Desmosomal and LMNA gene variants in DCM patients identify those at highest risk for sudden cardiac death and ventricular arrhythmias, irrespective of ejection fraction.
Area of Science:
- Cardiology
- Genetics
- Molecular Biology
Background:
- Genotype-phenotype correlations in dilated cardiomyopathy (DCM) and the impact of gene variants on clinical outcomes are not well understood.
- Understanding these relationships is crucial for predicting disease progression and patient prognosis.
Purpose of the Study:
- To investigate the prognostic significance of genetic variant carrier status in a large cohort of DCM patients.
- To identify specific gene variants associated with adverse clinical outcomes in DCM.
Main Methods:
- Next-generation sequencing was used to analyze 487 DCM patients.
- Patients were categorized by functional gene groups, and outcomes including all-cause mortality, heart failure events, and sudden cardiac death were assessed.
- Composite outcomes included heart failure-related death, heart transplantation, or ventricular assist device implantation (DHF/HTx/VAD), and sudden cardiac death/sustained ventricular tachycardia/ventricular fibrillation (SCD/VT/VF).
Main Results:
- Pathogenic/likely pathogenic variants were identified in 37% of patients, with Titin, sarcomeric genes, LMNA, and desmosomal genes being frequently implicated.
- No significant difference in all-cause mortality was observed between variant carriers and noncarriers.
- A trend towards worse outcomes for SCD/VT/VF and DHF/HTx/VAD was noted in carriers, with desmosomal and LMNA variants showing the highest rates of SCD/VT/VF, independent of left ventricular ejection fraction.
Conclusions:
- Desmosomal and LMNA gene variants are key indicators for identifying DCM patients at elevated risk for sudden cardiac death and life-threatening ventricular arrhythmias.
- These findings are significant regardless of the patient's left ventricular ejection fraction, offering a critical prognostic marker.
Background:
Genotype-phenotype correlations in dilated cardiomyopathy (DCM) and, in particular, the effects of gene variants on clinical outcomes remain poorly understood.
Objectives:
The purpose of this study was to investigate the prognostic role of genetic variant carrier status in a large cohort of DCM patients.
Methods:
A total of 487 DCM patients were analyzed by next-generation sequencing and categorized the disease genes into functional gene groups. The following composite outcome measures were assessed: 1) all-cause mortality; 2) heart failure-related death, heart transplantation, or destination left ventricular assist device implantation (DHF/HTx/VAD); and 3) sudden cardiac death/sustained ventricular tachycardia/ventricular fibrillation (SCD/VT/VF).
Results:
A total of 183 pathogenic/likely pathogenic variants were found in 178 patients (37%): 54 (11%) Titin; 19 (4%) Lamin A/C (LMNA); 24 (5%) structural cytoskeleton-Z disk genes; 16 (3.5%) desmosomal genes; 46 (9.5%) sarcomeric genes; 8 (1.6%) ion channel genes; and 11 (2.5%) other genes. All-cause mortality was no different between variant carriers and noncarriers (p = 0.99). A trend toward worse SCD/VT/VF (p = 0.062) and DHF/HTx/VAD (p = 0.061) was found in carriers. Carriers of desmosomal and LMNA variants experienced the highest rate of SCD/VT/VF, which was independent of the left ventricular ejection fraction.
Conclusions:
Desmosomal and LMNA gene variants identify the subset of DCM patients who are at greatest risk for SCD and life-threatening ventricular arrhythmias, regardless of the left ventricular ejection fraction.
Related Concept Videos
 01:30
Cardiomyopathy II: Dilated Cardiomyopathy
 05:14
A Doxorubicin-Induced Murine Model of Dilated Cardiomyopathy In Vivo
 06:04
Pulse-Wave Velocity, Flow-Mediated Dilation, and Carotid Intima-Media Thickness to Assess Cardiovascular Risk in Population with Metabolic Syndrome
 09:16
Isolation and Characterization of Cardiac Mesenchymal Stromal Cells from Endomyocardial Bioptic Samples of Arrhythmogenic Cardiomyopathy Patients
 01:29
Cardiomyopathy IV: Restrictive Cardiomyopathy
 01:29
Cardiomyopathy III: Hypertrophic Cardiomyopathy