Related Experiment Video
Updated: Jan 19, 2026

Author Spotlight: Unveiling the Pathway Linking Obesity to Autoimmune Inflammation in Multiple Sclerosis
Published on: February 23, 2024
Nrf2 deficiency increases oligodendrocyte loss, demyelination, neuroinflammation and axonal damage in an MS animal
Anna Nellessen1, Stella Nyamoya1,2,3, Adib Zendedel1
1Institute of Neuroanatomy, Uniklinik RWTH Aachen, Wendlingweg 2, 52074, Aachen, Germany.

Abstract:
Oxidative stress is a pathophysiological hallmark of many CNS diseases, among multiple sclerosis (MS). Accordingly, boosting the astrocytic transcription factor nuclear factor E2-related factor 2 (Nrf2) system in an MS mouse model efficiently ameliorates oligodendrocyte loss, neuroinflammation and axonal damage. Moreover, Dimethylfumarate, an efficient activator of Nrf2, has recently been approved as therapeutic option in MS treatment. Here, we use the cuprizone mouse model of MS to induce oxidative stress, selective oligodendrocyte loss, microglia and astrocyte activation as well as axonal damage in both wild type and Nrf2-deficient mice. We found increased oligodendrocyte apoptosis and loss, pronounced neuroinflammation and higher levels of axonal damage in cuprizone-fed Nrf2-deficient animals when compared to wild type controls. In addition, Nrf2-deficient animals showed a higher susceptibility towards cuprizone within the commissura anterior white matter tract, a structure that is relatively insensitive to cuprizone in wild type animals. Our data highlight the cuprizone model as a suitable tool to study the complex interplay of oxidative stress, neuroinflammation and axonal damage. Further studies will have to show whether distinct expression patterns of Nrf2 are involved in the variable susceptibility towards cuprizone in the mouse.
Insights
Nuclear factor E2-related factor 2 (Nrf2) deficiency exacerbates oxidative stress and damage in a mouse model of multiple sclerosis (MS). This highlights Nrf2
Area of Science:
- Neuroscience
- Immunology
- Pathology
Background:
- Oxidative stress is central to central nervous system (CNS) diseases like multiple sclerosis (MS).
- The nuclear factor E2-related factor 2 (Nrf2) pathway, particularly in astrocytes, shows promise in mitigating MS pathology.
- Dimethylfumarate, an Nrf2 activator, is an approved MS therapeutic.
Purpose of the Study:
- To investigate the role of Nrf2 in the cuprizone mouse model of MS.
- To assess the impact of Nrf2 deficiency on oxidative stress, oligodendrocyte loss, neuroinflammation, and axonal damage.
Main Methods:
- Utilized the cuprizone mouse model to induce MS-like pathology.
- Compared wild-type and Nrf2-deficient mice under cuprizone challenge.
- Assessed oligodendrocyte apoptosis, neuroinflammation markers, and axonal damage.
Main Results:
- Nrf2-deficient mice exhibited exacerbated oligodendrocyte apoptosis and loss compared to wild-type controls.
- Neuroinflammation and axonal damage were significantly higher in Nrf2-deficient mice.
- Nrf2-deficient mice showed increased susceptibility to cuprizone in the commissura anterior white matter tract.
Conclusions:
- Nrf2 plays a critical protective role against oxidative stress and MS-like pathology in the CNS.
- The cuprizone model is valuable for studying the interactions between oxidative stress, neuroinflammation, and axonal damage in MS.
- Further research is needed to explore Nrf2 expression patterns and variable susceptibility to cuprizone.
Related Concept Videos
 08:17
Scoring Central Nervous System Inflammation, Demyelination, and Axon Injury in Experimental Autoimmune Encephalomyelitis
 04:55
A Stably Established Two-Point Injection of Lysophosphatidylcholine-Induced Focal Demyelination Model in Mice
 01:30
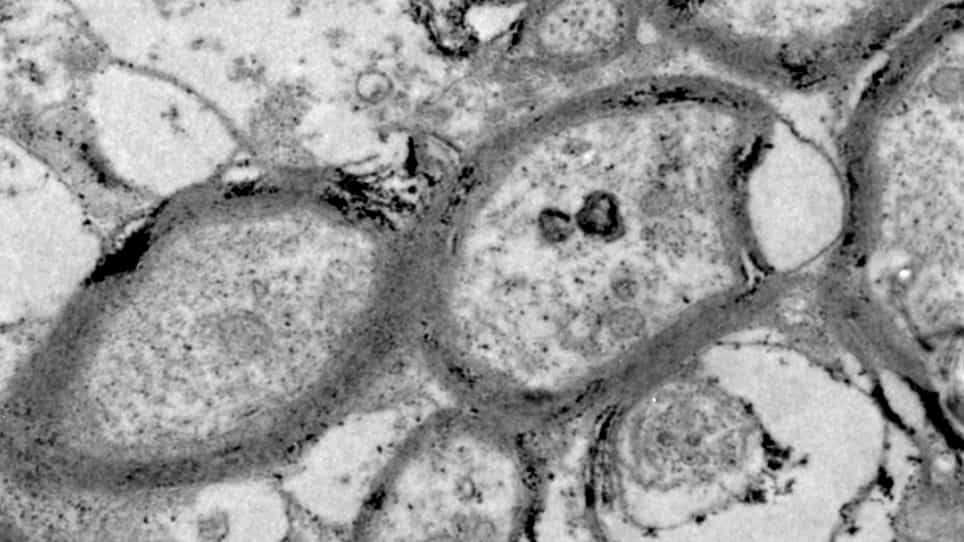
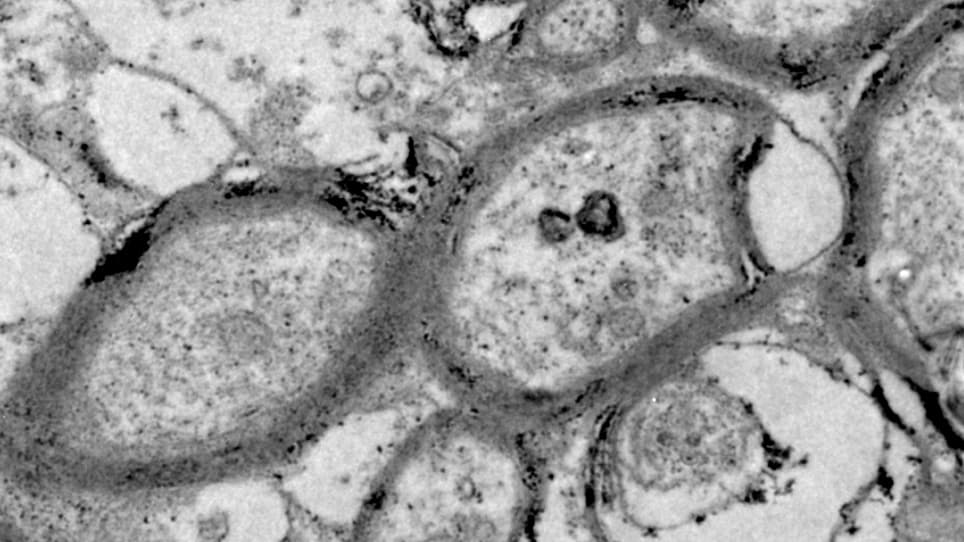
Ultrastructural Analysis of Demyelination in Mouse Spinal Cord Cross-Sections
 05:08
Demyelination and Remyelination of Mouse Spinal Cord Neurons
 09:46
Rat Model of Widespread Cerebral Cortical Demyelination Induced by an Intracerebral Injection of Pro-Inflammatory Cytokines
 04:53
Isolation of Mouse Primary Microglia by Magnetic-Activated Cell Sorting in Animal Models of Demyelination