Related Experiment Video
Updated: Jan 19, 2026

X-Ray Crystallography to Study the Oligomeric State Transition of the Thermotoga maritima M42 Aminopeptidase TmPep1050
Published on: May 13, 2020
The modular structure of α/β-hydrolases
Tabea L Bauer1, Patrick C F Buchholz1, Jürgen Pleiss1
1Institute of Biochemistry and Technical Biochemistry, University of Stuttgart, Germany.

Abstract:
The α/β-hydrolase fold family is highly diverse in sequence, structure and biochemical function. To investigate the sequence-structure-function relationships, the Lipase Engineering Database (https://led.biocatnet.de) was updated. Overall, 280 638 protein sequences and 1557 protein structures were analysed. All α/β-hydrolases consist of the catalytically active core domain, but they might also contain additional structural modules, resulting in 12 different architectures: core domain only, additional lids at three different positions, three different caps, additional N- or C-terminal domains and combinations of N- and C-terminal domains with caps and lids respectively. In addition, the α/β-hydrolases were distinguished by their oxyanion hole signature (GX-, GGGX- and Y-types). The N-terminal domains show two different folds, the Rossmann fold or the β-propeller fold. The C-terminal domains show a β-sandwich fold. The N-terminal β-propeller domain and the C-terminal β-sandwich domain are structurally similar to carbohydrate-binding proteins such as lectins. The classification was applied to the newly discovered polyethylene terephthalate (PET)-degrading PETases and MHETases, which are core domain α/β-hydrolases of the GX- and the GGGX-type respectively. To investigate evolutionary relationships, sequence networks were analysed. The degree distribution followed a power law with a scaling exponent γ = 1.4, indicating a highly inhomogeneous network which consists of a few hubs and a large number of less connected sequences. The hub sequences have many functional neighbours and therefore are expected to be robust toward possible deleterious effects of mutations. The cluster size distribution followed a power law with an extrapolated scaling exponent τ = 2.6, which strongly supports the connectedness of the sequence space of α/β-hydrolases. DATABASE: Supporting data about domains from other proteins with structural similarity to the N- or C-terminal domains of α/β-hydrolases are available in Data Repository of the University of Stuttgart (DaRUS) under doi: https://doi.org/10.18419/darus-458.
Related Concept Videos
 01:09
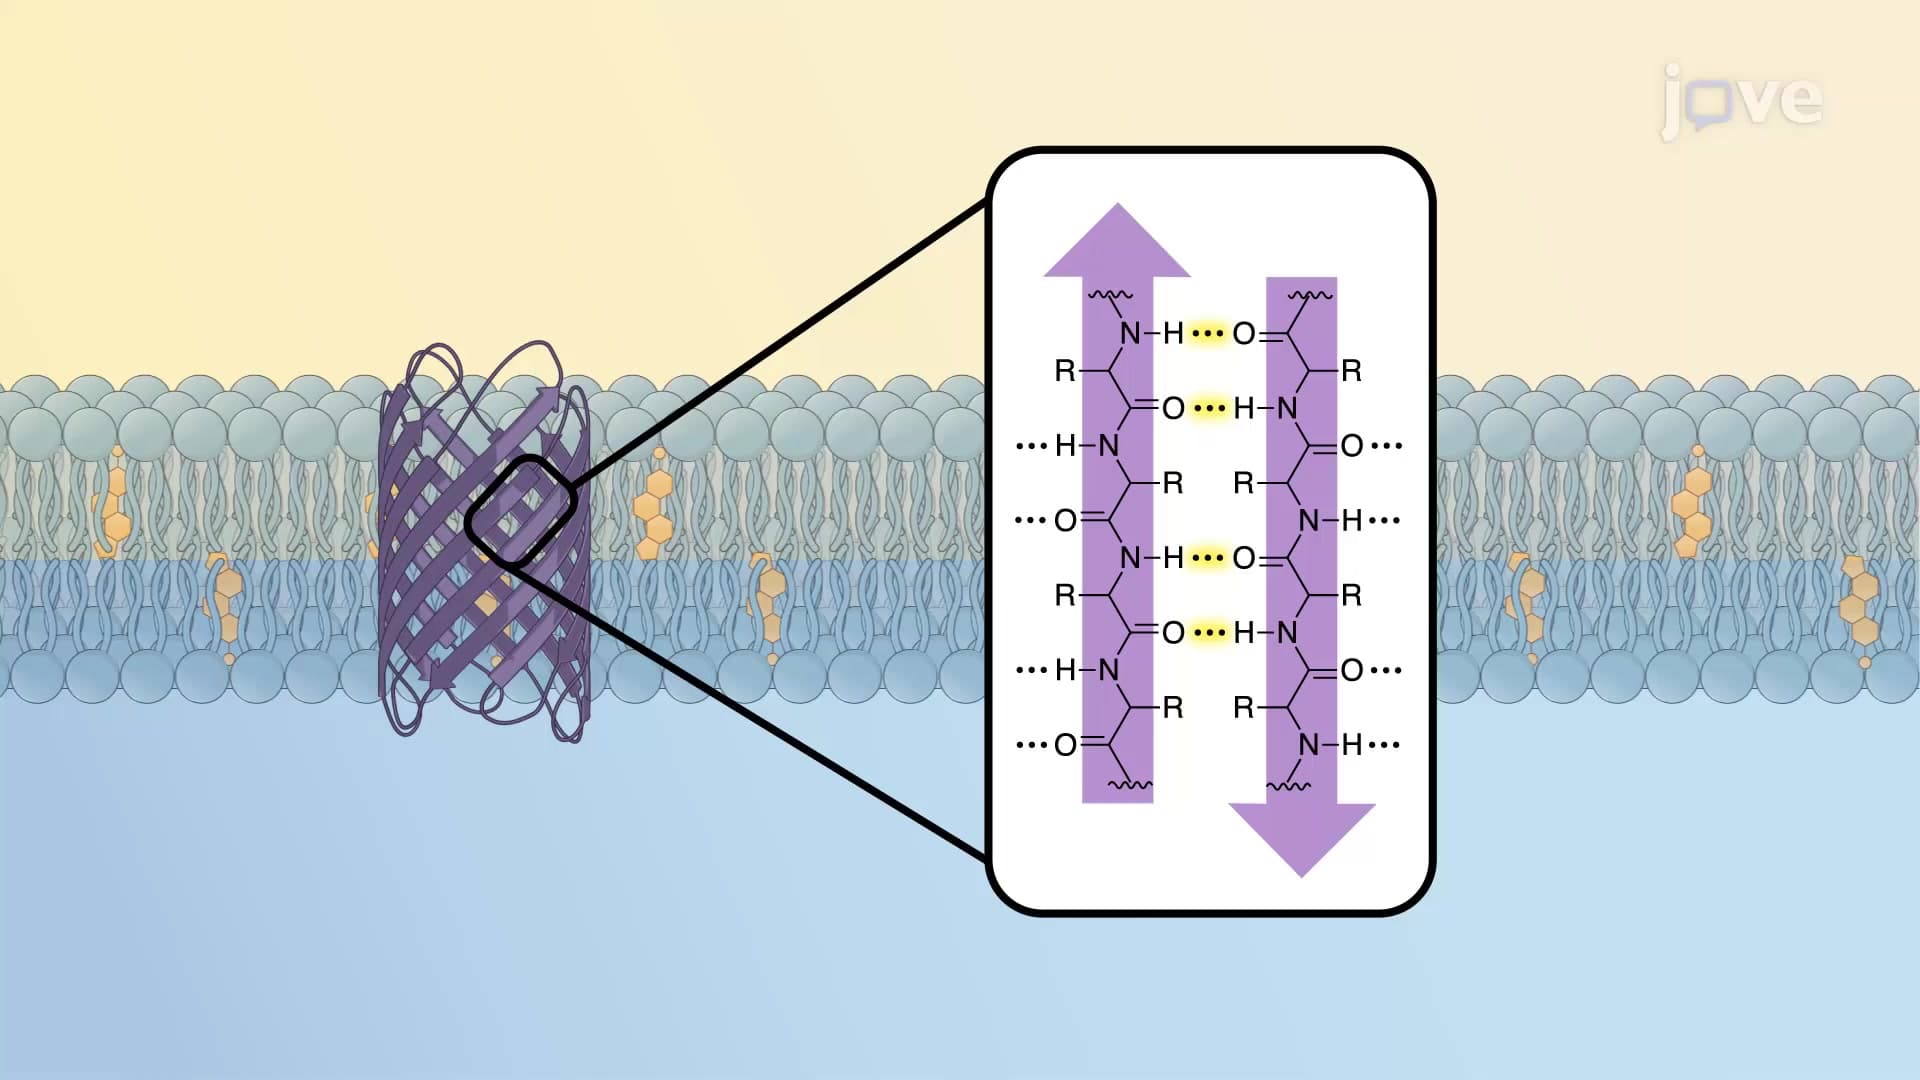
Multi-pass Transmembrane Proteins and β-barrels
α-Helix containing multi-pass transmembrane proteins
Multi-pass transmembrane proteins such as...
 01:22
Lysosomal Hydrolases
 02:15
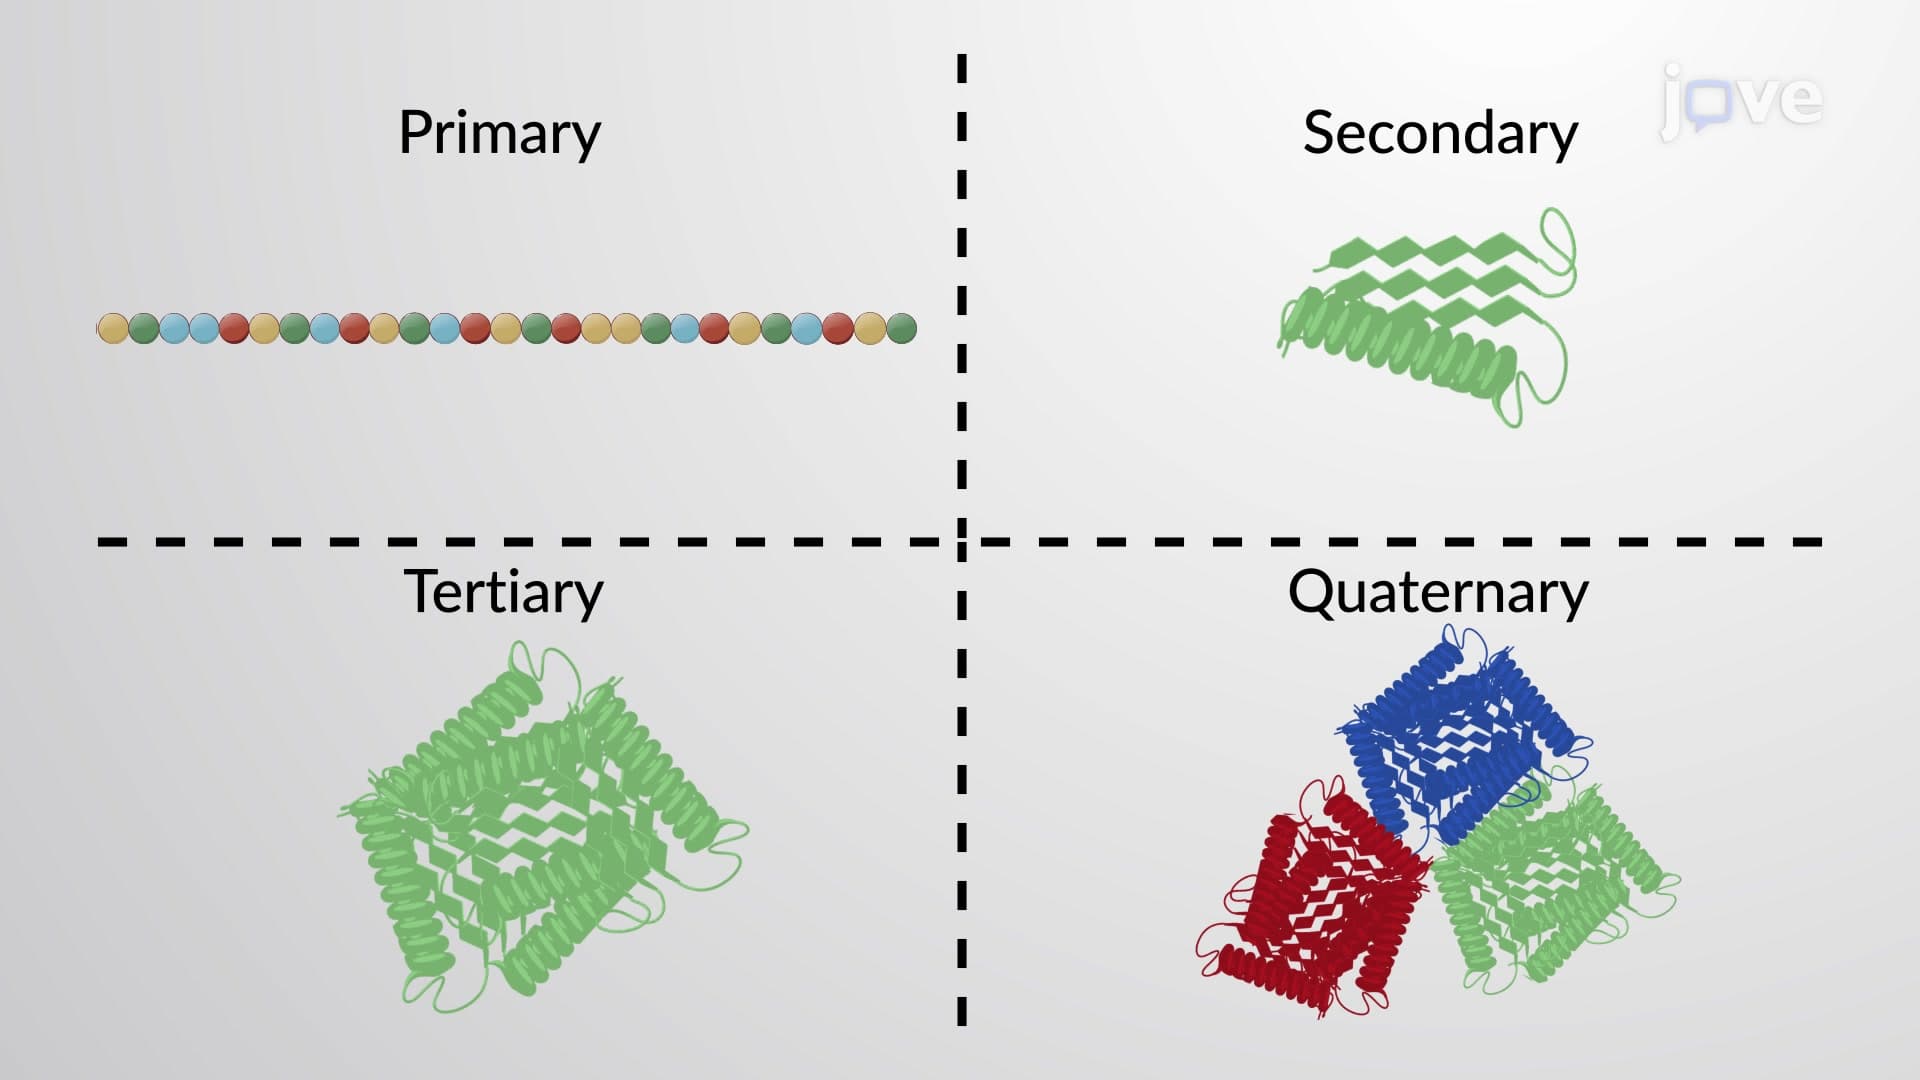
Protein and Protein Structure
A protein's shape is critical to its function. For example, an enzyme...
 01:22
Protein Folding
 01:25
Protein Folding
Protein Structure Is Critical to Its Biological Function
Proteins perform a wide range of biological functions such as catalyzing chemical reactions, providing...
 02:41
Protein Complex Assembly
Many viruses self-assemble into a fully functional unit using the infected host cell to...