Related Experiment Video
Updated: Jan 18, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quantum Dynamics of Dissociative Chemisorption of H2 on the Stepped Cu(211) Surface
Egidius W F Smeets1, Gernot Füchsel2, Geert-Jan Kroes1
1Leiden Institute of Chemistry, Gorlaeus Laboratories, Leiden University, P.O. Box 9502, 2300 RA Leiden, The Netherlands.

Abstract:
Reactions on stepped surfaces are relevant to heterogeneous catalysis, in which a reaction often takes place at the edges of nanoparticles where the edges resemble steps on single-crystal stepped surfaces. Previous results on H2 + Cu(211) showed that, in this system, steps do not enhance the reactivity and raised the question of whether this effect could be, in any way, related to the neglect of quantum dynamical effects in the theory. To investigate this, we present full quantum dynamical molecular beam simulations of sticking of H2 on Cu(211), in which all important rovibrational states populated in a molecular beam experiment are taken into account. We find that the reaction of H2 with Cu(211) is very well described with quasi-classical dynamics when simulating molecular beam sticking experiments, in which averaging takes place over a large number of rovibrational states and over translational energy distributions. Our results show that the stepped Cu(211) surface is distinct from its component Cu(111) terraces and Cu(100) steps and cannot be described as a combination of its component parts with respect to the reaction dynamics when considering the orientational dependence. Specifically, we present evidence that, at translational energies close to the reaction threshold, vibrationally excited molecules show a negative rotational quadrupole alignment parameter on Cu(211), which is not found on Cu(111) and Cu(100). The effect arises because these molecules react with a site-specific reaction mechanism at the step, that is, inelastic rotational enhancement, which is only effective for molecules with a small absolute value of the magnetic rotation quantum number. From a comparison to recent associative desorption experiments as well as Born-Oppenheimer molecular dynamics calculations, it follows that the effects of surface atom motion and electron-hole pair excitation on the reactivity fall within chemical accuracy, that is, modeling these effect shifts extracted reaction probability curves by less than 1 kcal/mol translational energy. We found no evidence in our fully state-resolved calculations for the "slow" reaction channel that was recently reported for associative desorption of H2 from Cu(111) and Cu(211), but our results for the fast channel are in good agreement with the experiments on H2 + Cu(211).
More Related Videos
 07:53
Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
 14:11
Quantification of Hydrogen Concentrations in Surface and Interface Layers and Bulk Materials through Depth Profiling with Nuclear Reaction Analysis
Published on: March 29, 2016
Related Concept Videos
 02:50
Catalysis
 03:51
Molecular Orbital Theory II
 03:35
Hybridization of Atomic Orbitals II
 01:19
2D NMR: Heteronuclear Single-Quantum Correlation Spectroscopy (HSQC)
 02:13
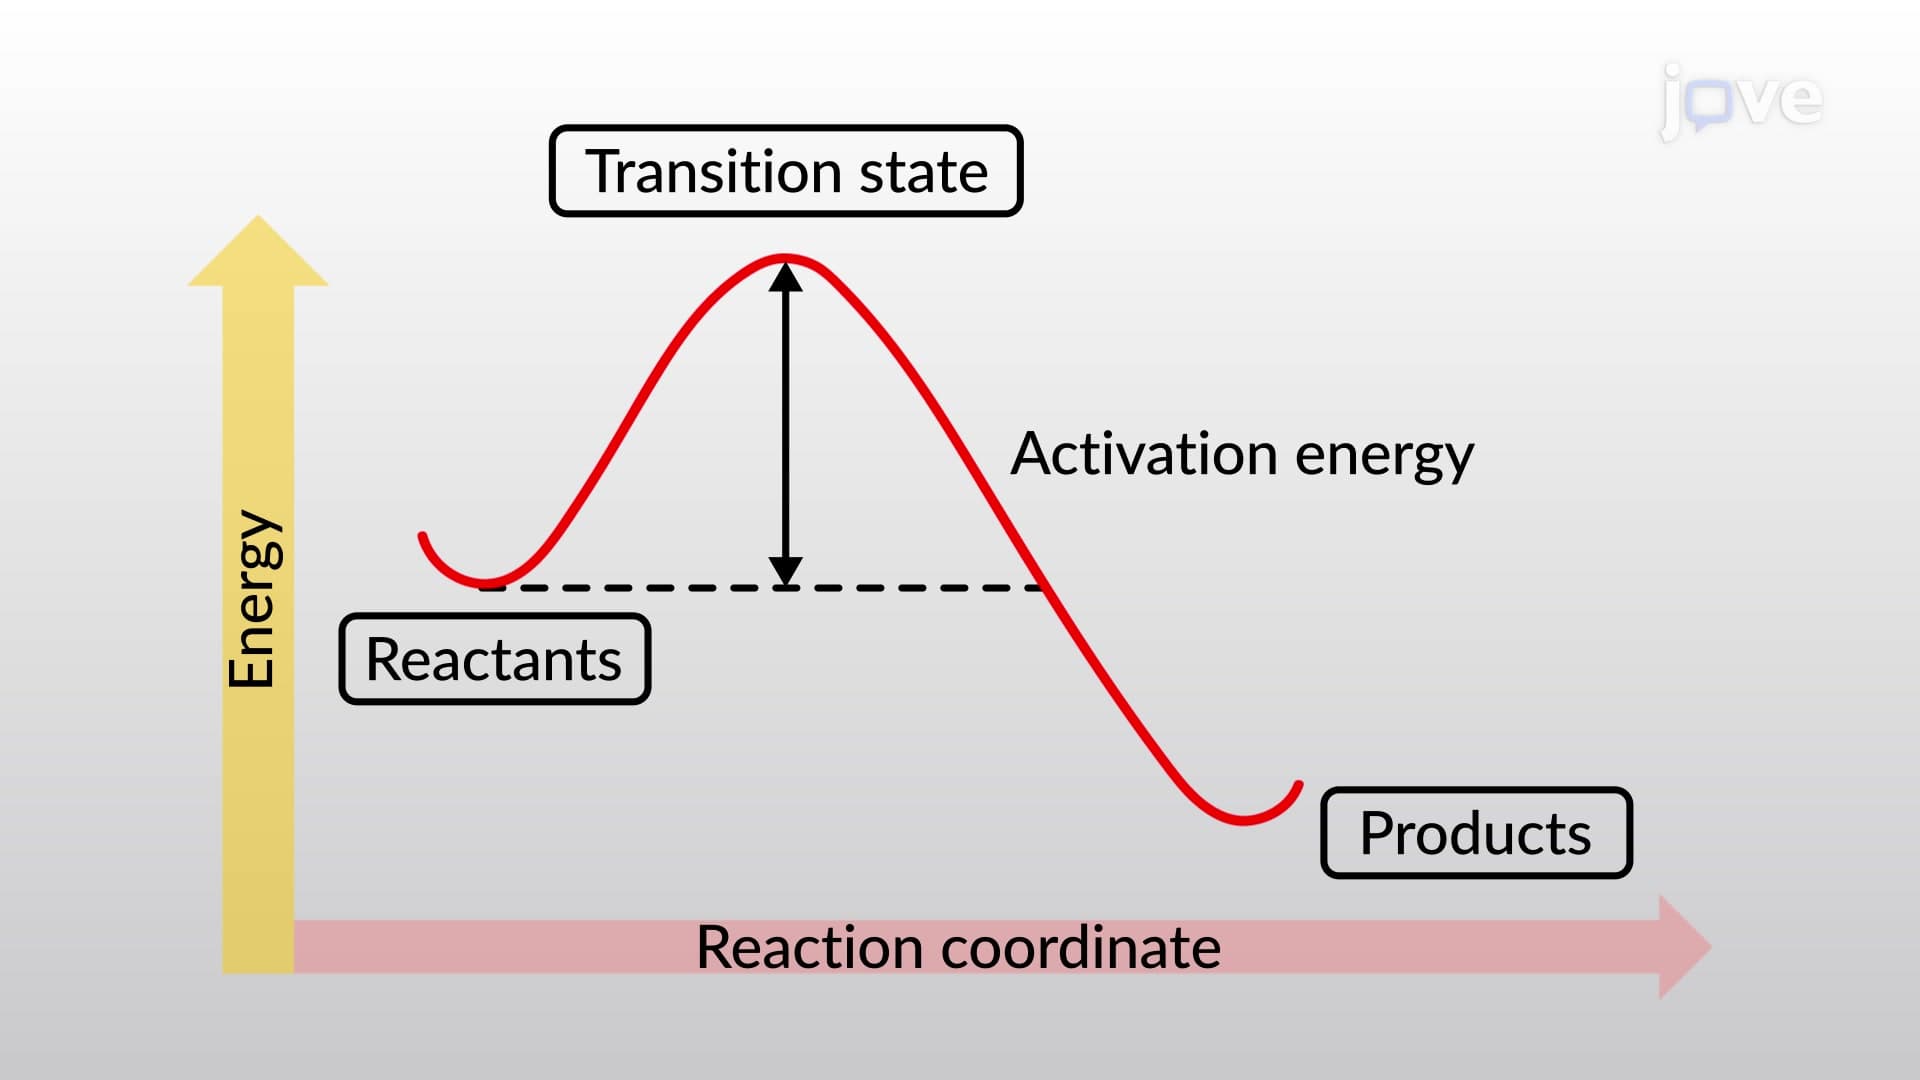
Bond Dissociation Energy and Activation Energy
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...