Related Experiment Video
Updated: Jan 18, 2026

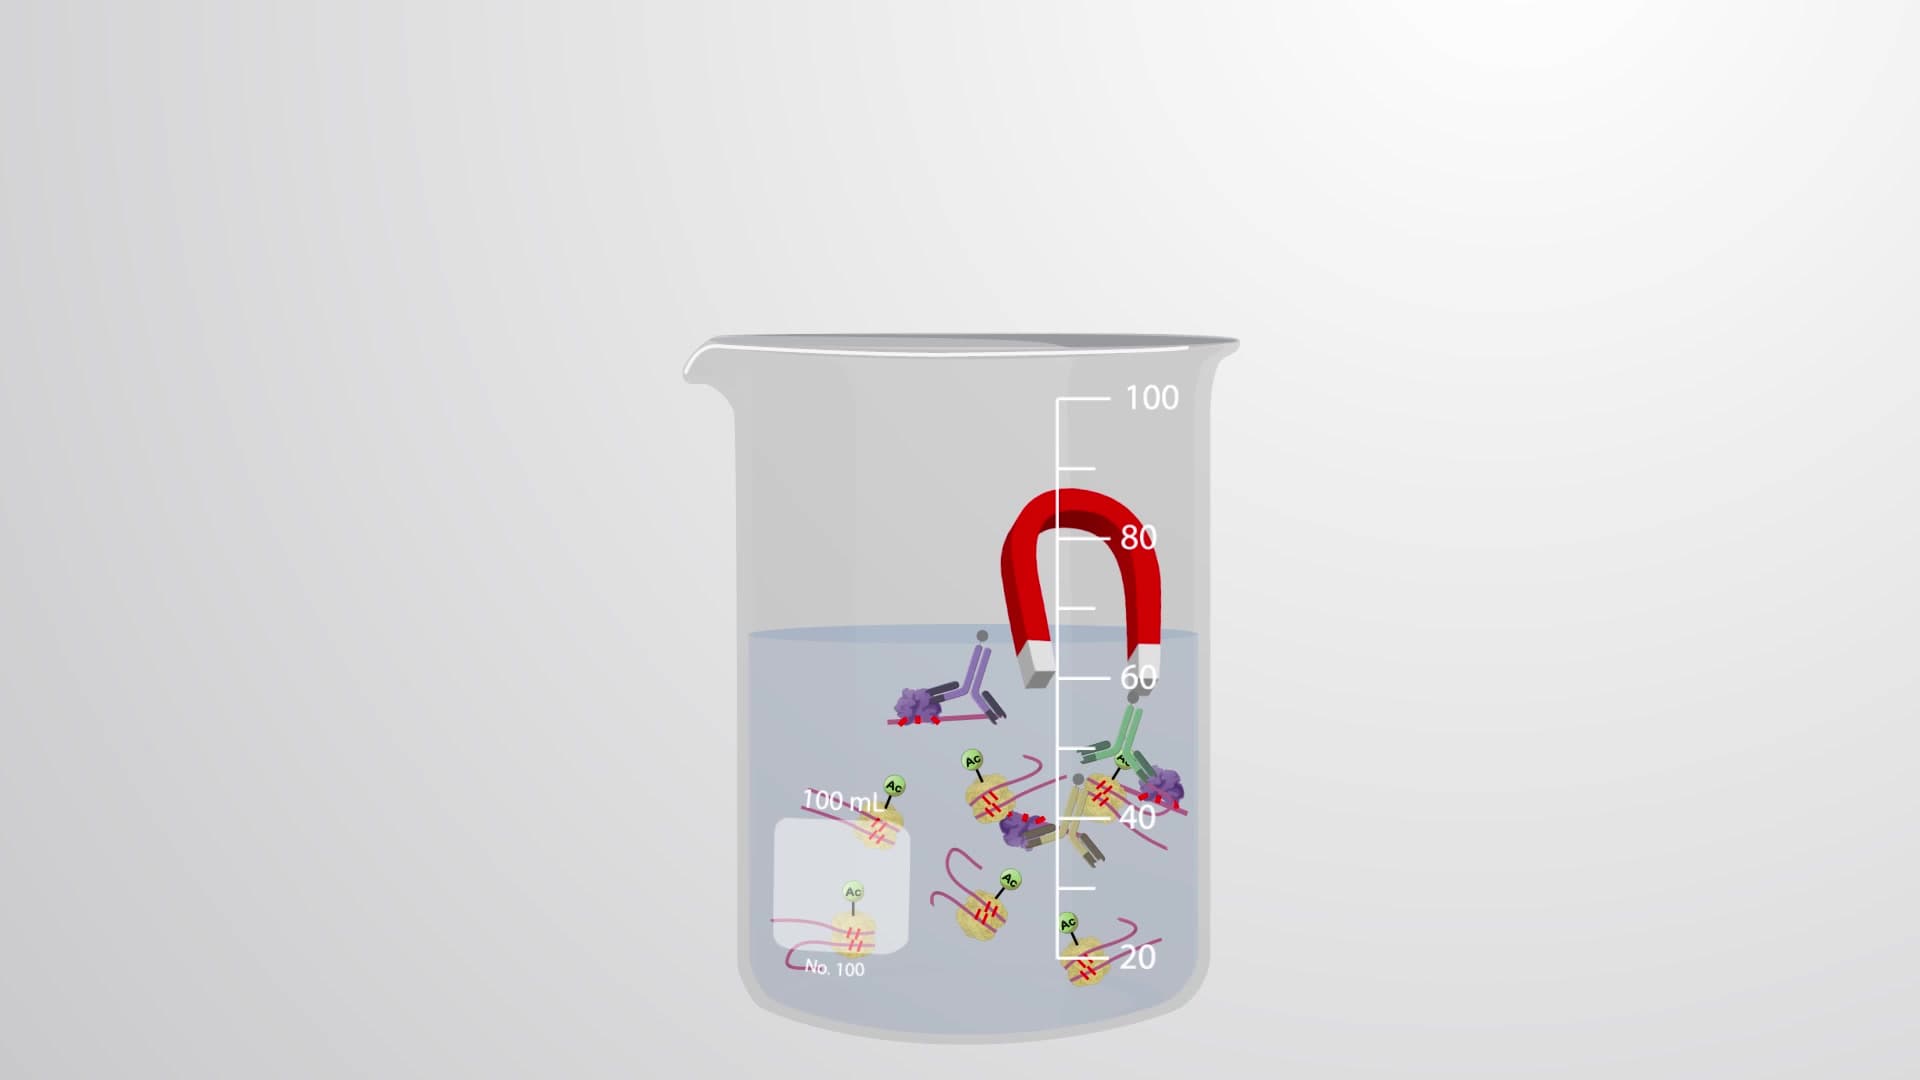
The ChroP Approach Combines ChIP and Mass Spectrometry to Dissect Locus-specific Proteomic Landscapes of Chromatin
Published on: April 11, 2014
ChIPseqSpikeInFree: a ChIP-seq normalization approach to reveal global changes in histone modifications without
Hongjian Jin1, Lawryn H Kasper2, Jon D Larson2
1Department of Computational Biology, St. Jude Children's Research Hospital, Memphis, TN 38105, USA.

Motivation:
The traditional reads per million normalization method is inappropriate for the evaluation of ChIP-seq data when treatments or mutations have global effects. Changes in global levels of histone modifications can be detected with exogenous reference spike-in controls. However, most ChIP-seq studies overlook the normalization that must be corrected with spike-in. A method that retrospectively renormalizes datasets without spike-in is lacking.
Results:
ChIPseqSpikeInFree is a novel ChIP-seq normalization method to effectively determine scaling factors for samples across various conditions and treatments, which does not rely on exogenous spike-in chromatin or peak detection to reveal global changes in histone modification occupancy. Application of ChIPseqSpikeInFree on five datasets demonstrates that this in silico approach reveals a similar magnitude of global changes as the spike-in method does.
Availability And Implementation:
St. Jude Cloud (https://pecan.stjude.cloud/permalink/spikefree) and St. Jude Github ( https://github.com/stjude/ChIPseqSpikeInFree).
Supplementary Information:
Supplementary data are available at Bioinformatics online.