Related Experiment Video
Updated: Jan 6, 2026

Interactive Molecular Model Assembly with 3D Printing
Published on: August 13, 2020
Atom Type Independent Modeling of the Conformational Energy of Benzylic, Allylic, and Other Bonds Adjacent to
Candide Champion1, Stephen J Barigye1, Wanlei Wei1
1Department of Chemistry , McGill University , 801 Sherbrooke Street W. , Montréal , QC , Canada H3A 0B8.

Abstract:
Applications of computational methods to predict binding affinities for protein/drug complexes are routinely used in structure-based drug discovery. Applications of these methods often rely on empirical force fields (FFs) and their associated parameter sets and atom types. However, it is widely accepted that FFs cannot accurately cover the entire chemical space of drug-like molecules, due to the restrictive cost of parametrization and the poor transferability of existing parameters. To address these limitations, initiatives have been carried out to develop more transferable methods, in order to allow for more rigorous descriptions of any drug-like molecule. We have previously reported H-TEQ, a method which does not rely on atom types and incorporates well established chemical principles to assign parameters to organic molecules. The previous implementation of H-TEQ (a torsional barrier prediction method) only covered saturated and lone pair containing molecules; here, we report our efforts to incorporate conjugated systems into our model. The next step was the evaluation of the introduction of unsaturations. The developed model (H-TEQ3.0) has been validated on a wide variety of molecules containing heteroaromatic groups, alkyls, and fused ring systems. Our method performs on par with one of the most commonly used FFs (GAFF2), without relying on atom types or any prior parametrization.
Related Concept Videos
 01:18
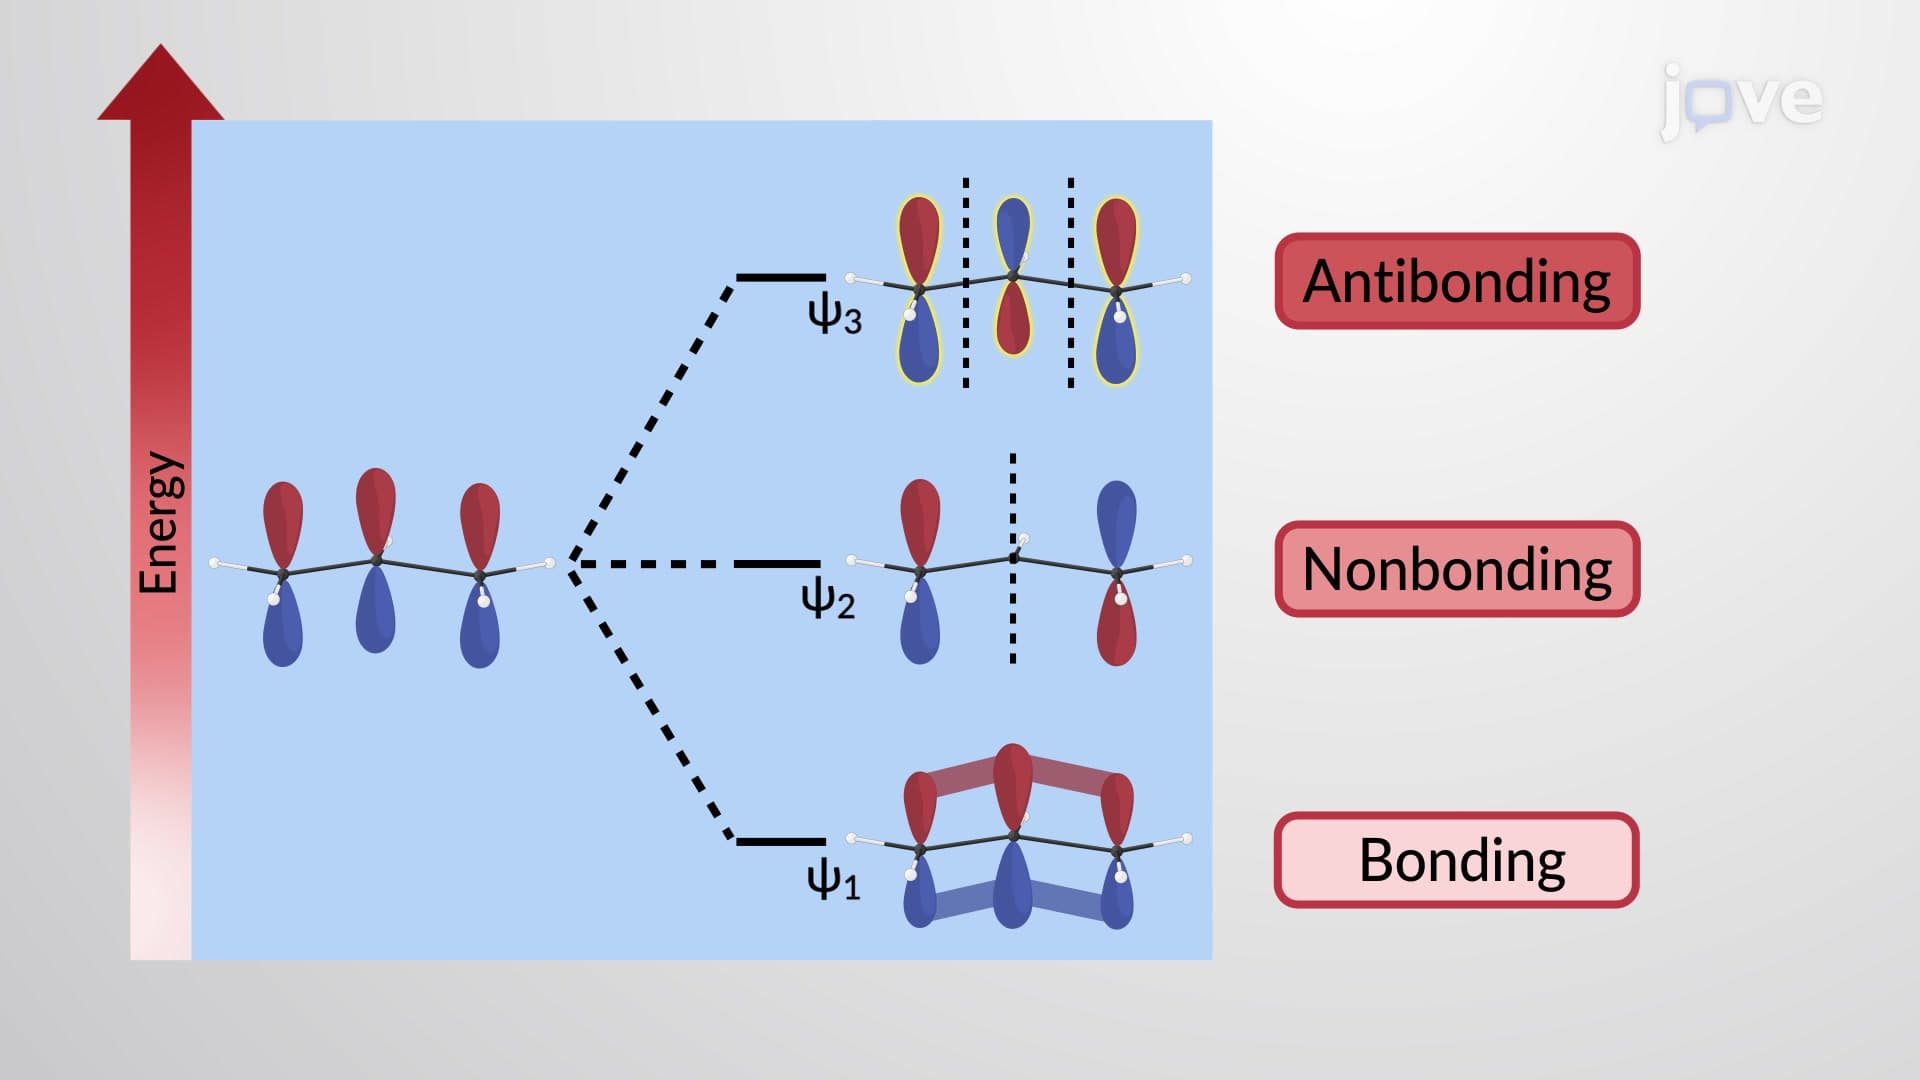
π Molecular Orbitals of the Allyl Cation and Anion
 03:51
Molecular Orbital Theory II
 01:18
Structure of Benzene: Molecular Orbital Model
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 02:18
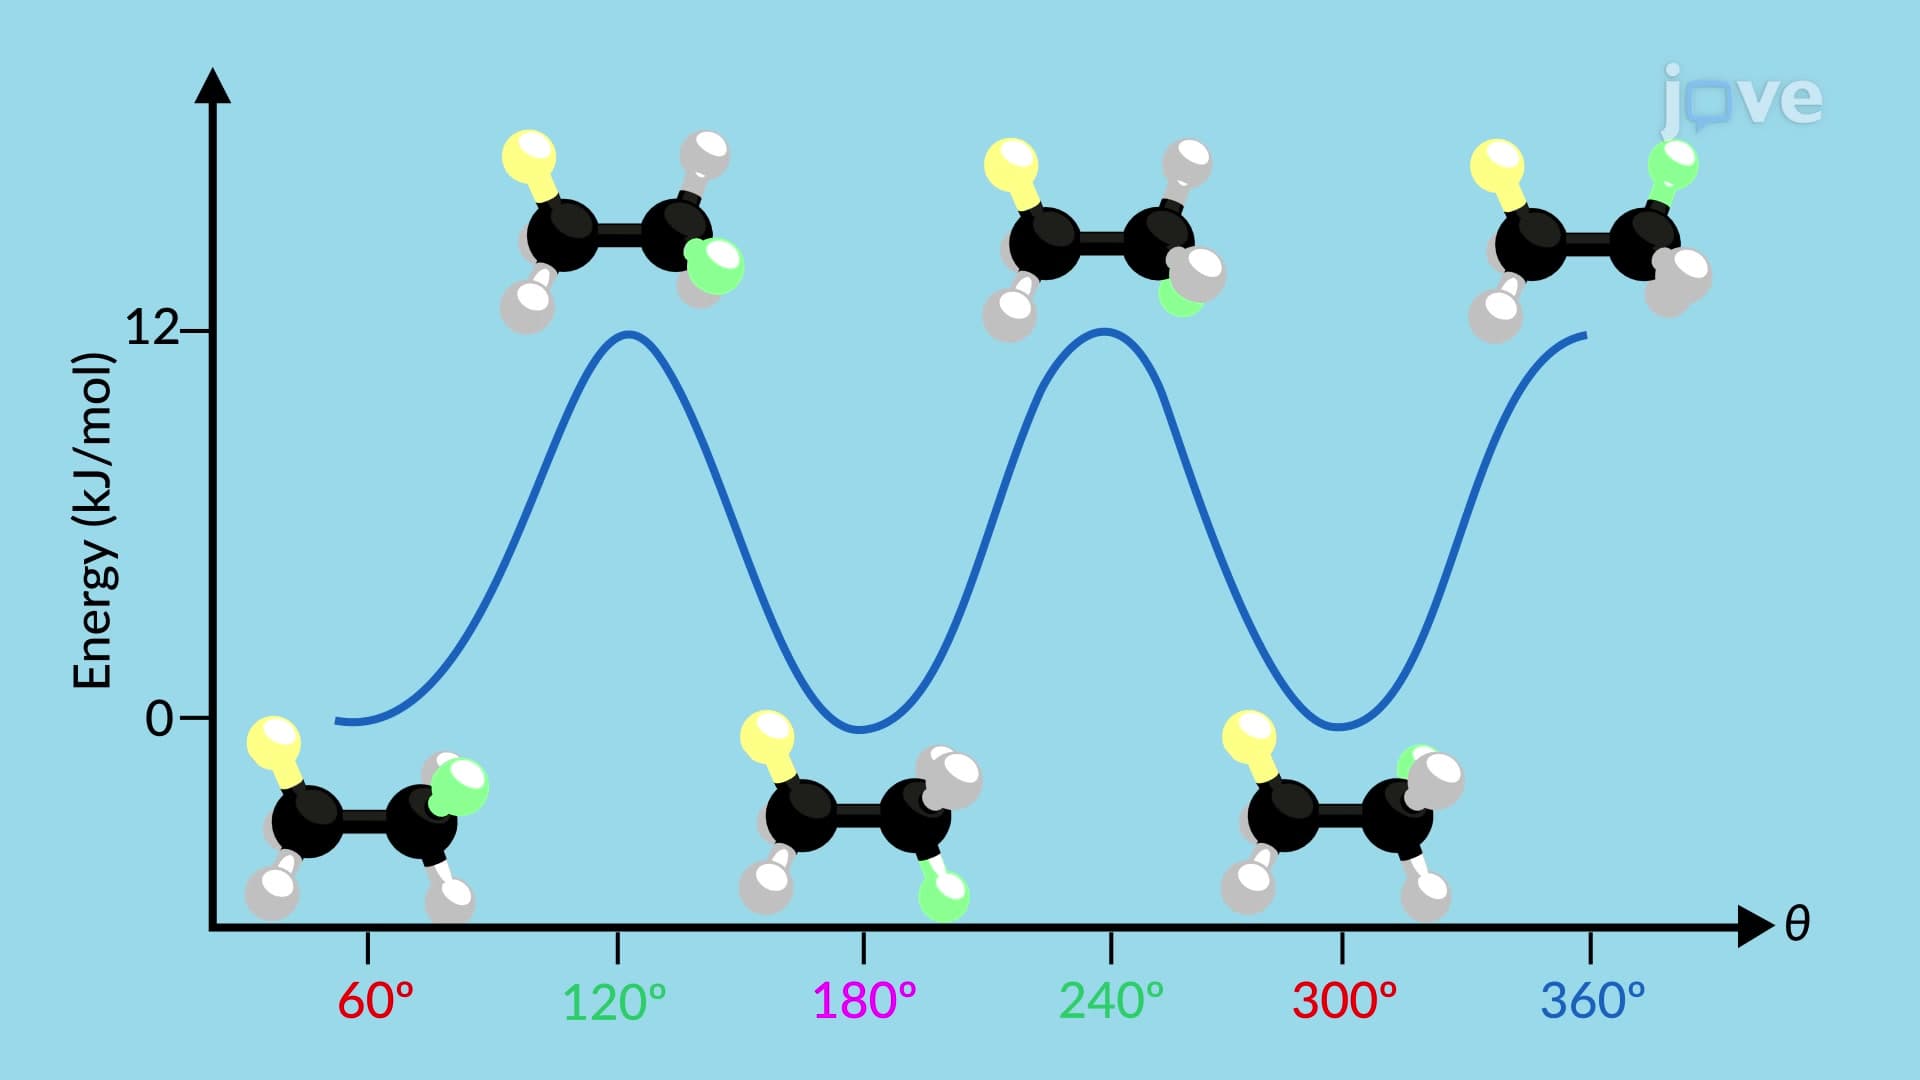
Conformations of Ethane and Propane
Staggered conformation is a low energy and more stable conformation with the C-H bonds on the front carbon placed at 60°dihedral angles relative to the C-H bonds on the back carbon, leading to a reduced torsional strain. In staggered...
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal...