Related Experiment Video
Updated: Jan 6, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Machine Learning the Physical Nonlocal Exchange-Correlation Functional of Density-Functional Theory
Jonathan Schmidt1, Carlos L Benavides-Riveros1, Miguel A L Marques1
1Institut für Physik , Martin-Luther-Universität Halle-Wittenberg , 06120 Halle (Saale) , Germany.

Abstract:
We train a neural network as the universal exchange-correlation functional of density-functional theory that simultaneously reproduces both the exact exchange-correlation energy and the potential. This functional is extremely nonlocal but retains the computational scaling of traditional local or semilocal approximations. It therefore holds the promise of solving some of the delocalization problems that plague density-functional theory, while maintaining the computational efficiency that characterizes the Kohn-Sham equations. Furthermore, by using automatic differentiation, a capability present in modern machine-learning frameworks, we impose the exact mathematical relation between the exchange-correlation energy and the potential, leading to a fully consistent method. We demonstrate the feasibility of our approach by looking at one-dimensional systems with two strongly correlated electrons, where density-functional methods are known to fail, and investigate the behavior and performance of our functional by varying the degree of nonlocality.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:35
Molecular Orbital Theory I
 03:51
Molecular Orbital Theory II
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 01:10
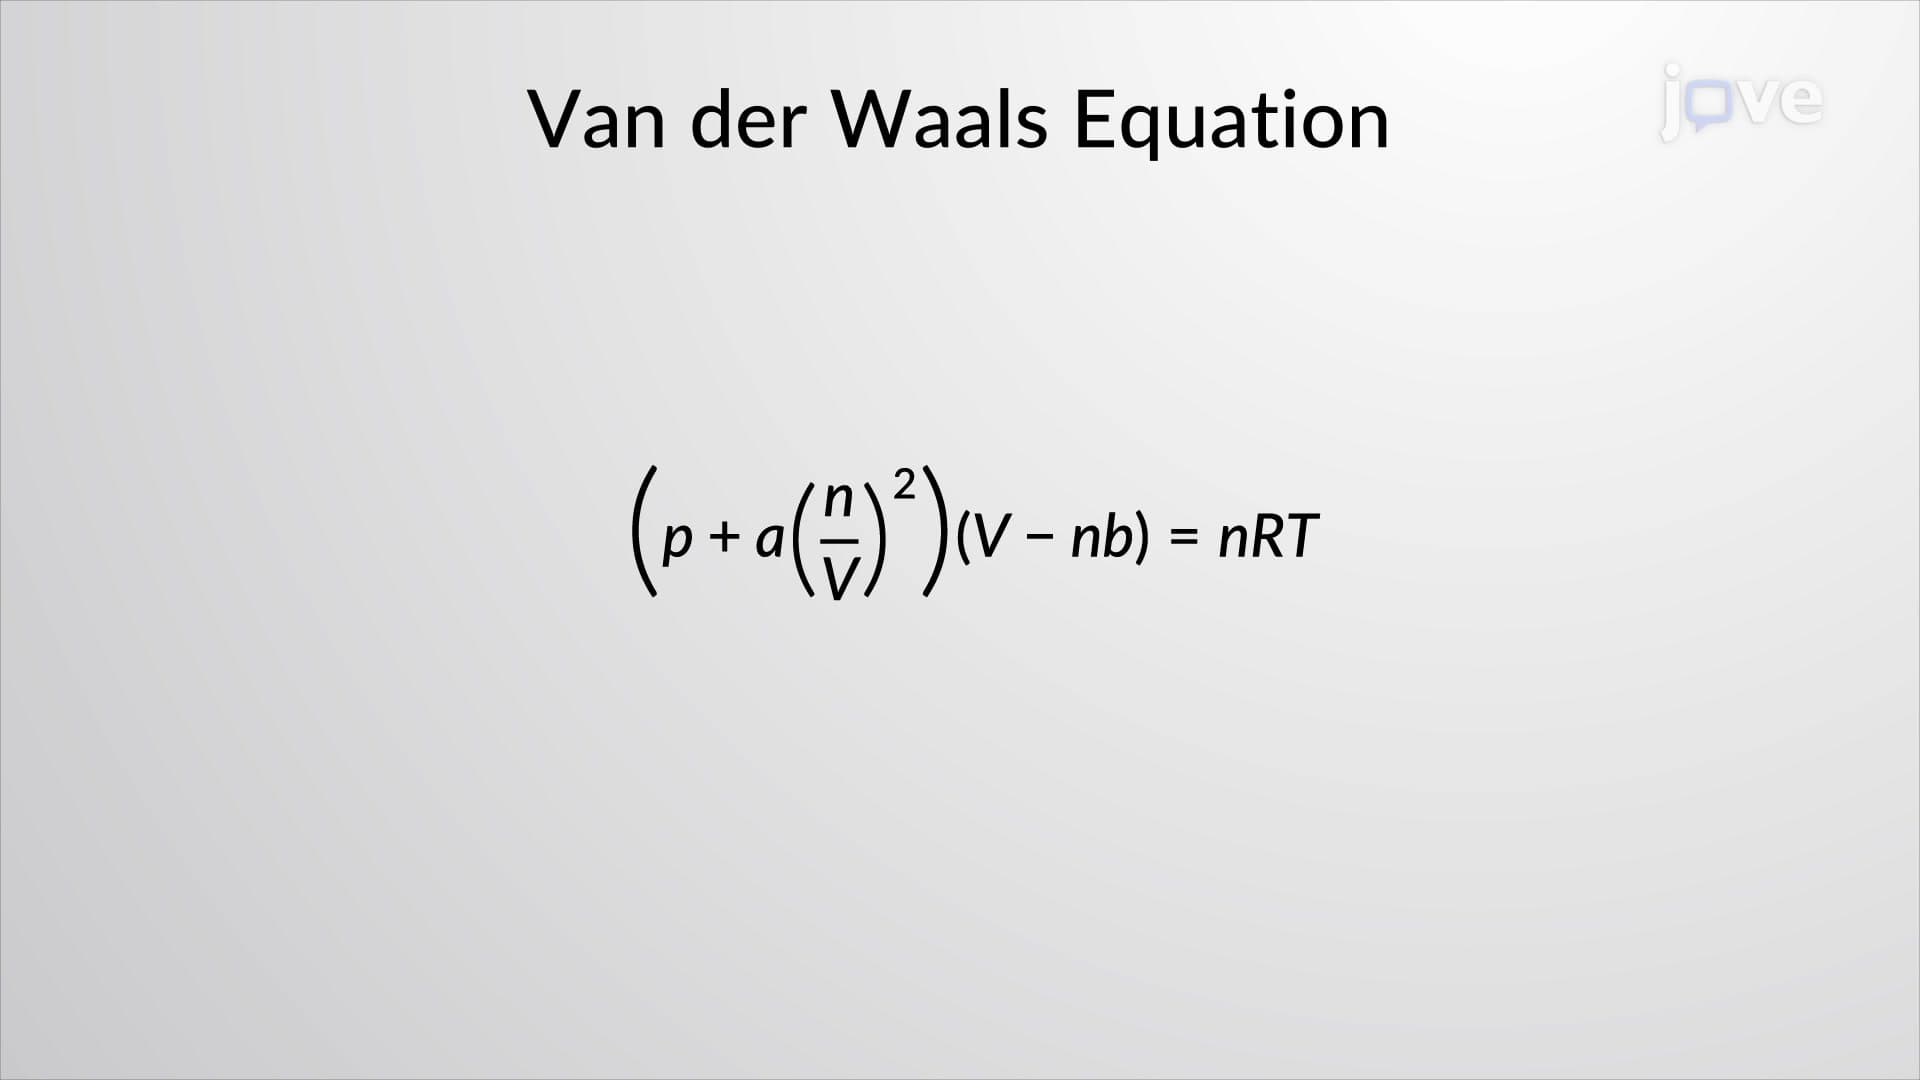
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...