Related Experiment Video
Updated: Jan 5, 2026


Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Performances of Density Functional Tight-Binding Methods for Describing Ground and Excited State Geometries of
Arnaud Fihey1, Denis Jacquemin2
1Institut des Sciences Chimiques de Rennes, UMR 6226 CNRS , Université de Rennes 1 , 263 Av. du Général Leclerc , 35042 Cedex Rennes , France.

Abstract:
We present a benchmark of the performances of the density functional tight-binding model (DFTB) and its time-dependent counterpart (TD-DFTB) in describing both the ground state (GS) and excited state (ES) geometries of a panel of 30 organic molecules. Thanks to high-level wave function reference calculations, we are able to quantitatively assess the strengths and weaknesses of four DFTB models, using either second- or third-order self-consistent charge procedures, as well as different sets of parameters. The performances of the different DFTB models are found to be largely dependent on the type of bond considered, but the global mean absolute error remains acceptable for such "cheap" calculations, as it slightly exceeds the one obtained with DFT (PBE and B3LYP) or CC2 models for the GS of the same set of compounds. When considering the ES of the molecules, the TD-DFTB errors are surprisingly not systematically larger than their GS counterparts. However, the trends, either when going from the GS to the ES or within a homologous chemical series, are less consistently reproduced with TD-DFTB than with the considered ab initio models. This work therefore validates the use of TD-DFTB for describing ES geometries while highlighting that care has to be applied when looking at subtle variations.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
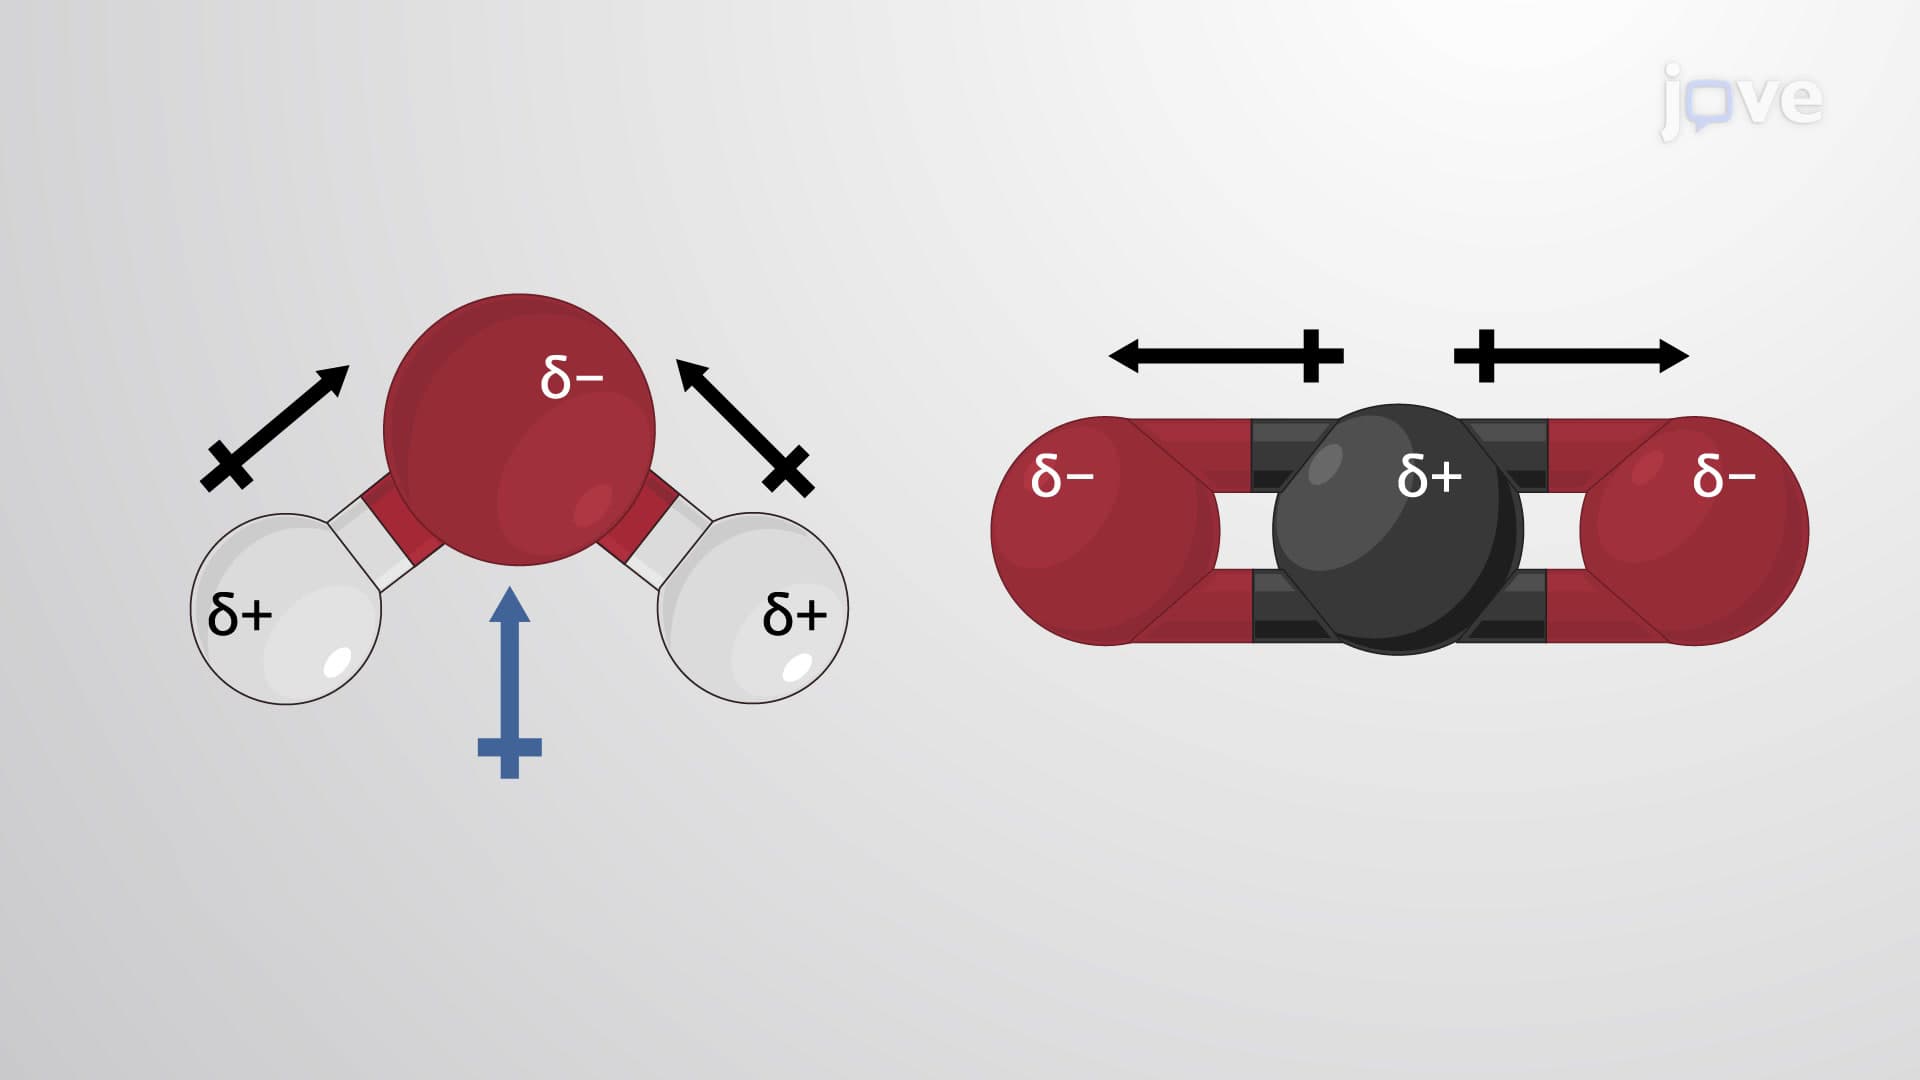 02:36
Molecular Geometry and Dipole Moments
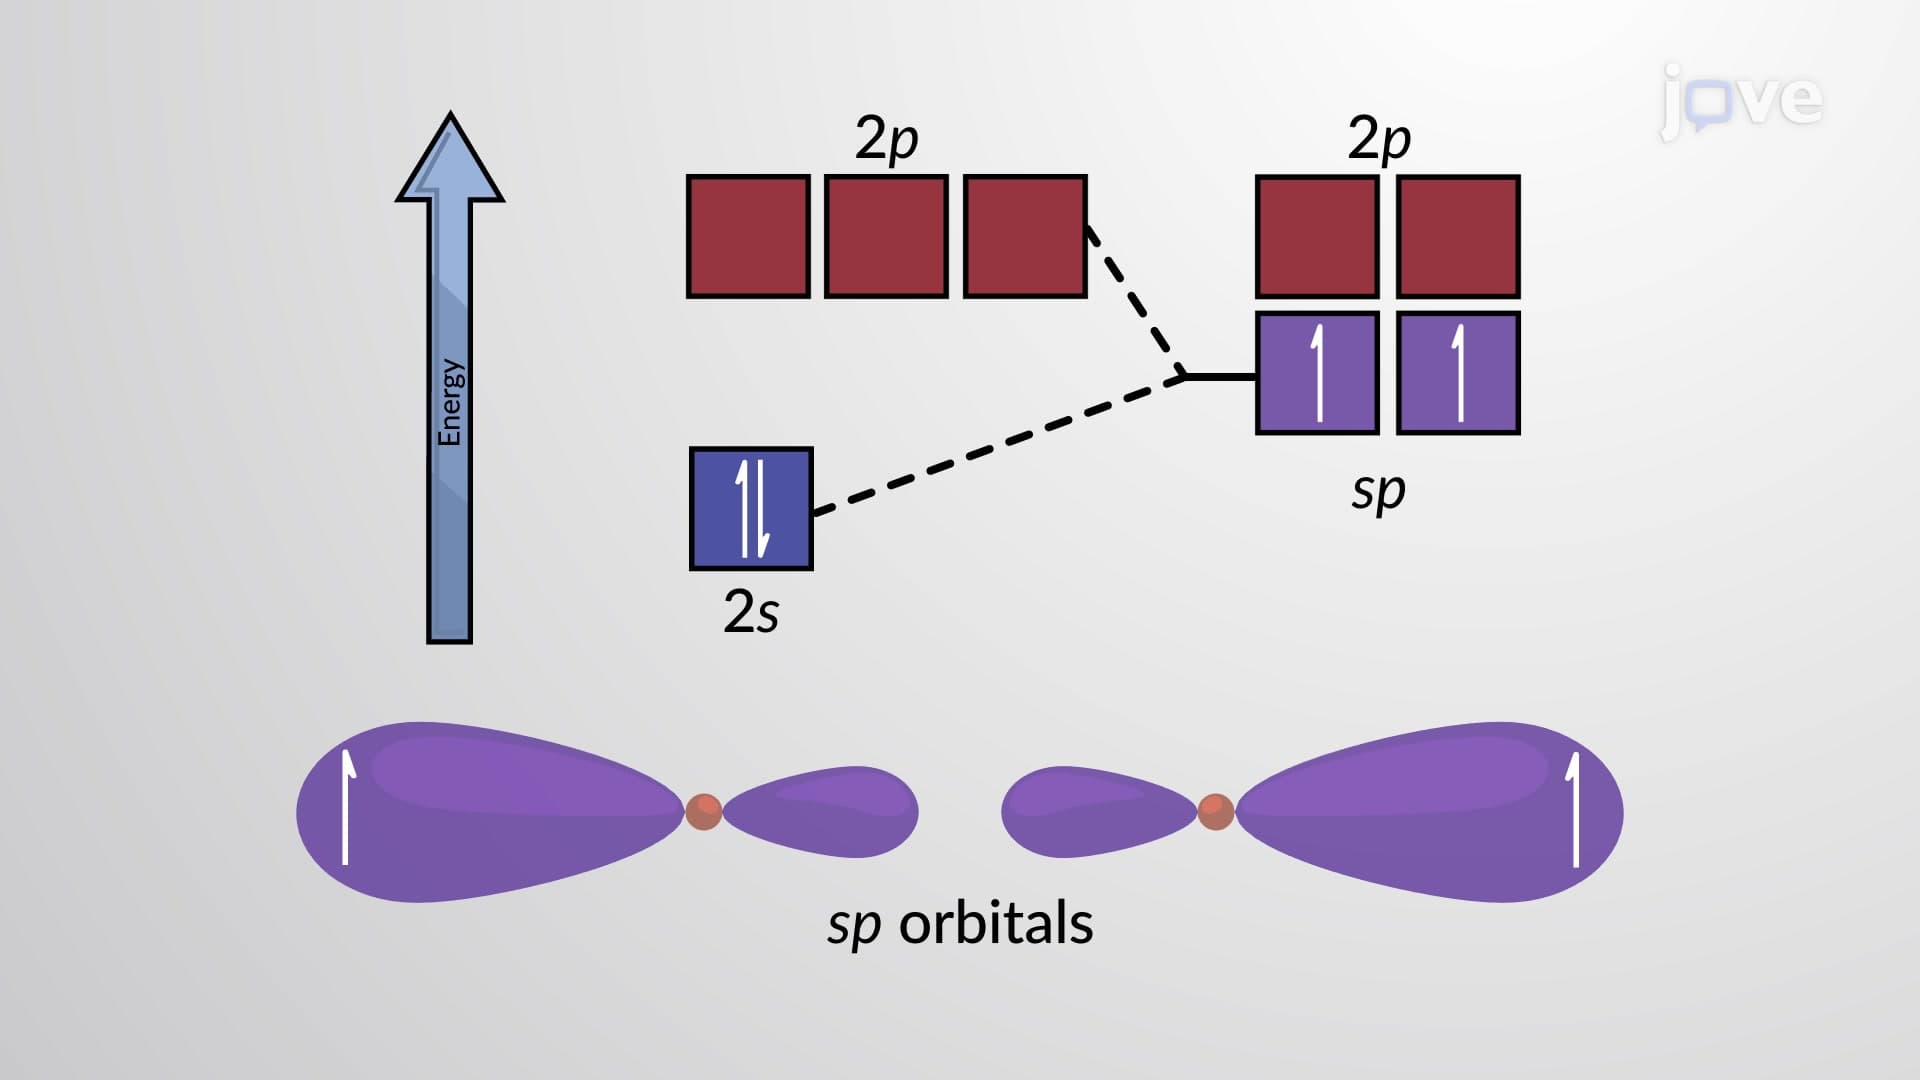 02:38
Valence Bond Theory and Hybridized Orbitals
A σ bond (single bond in a Lewis structure) is a covalent bond in which the electron density is...
 03:51
Molecular Orbital Theory II
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
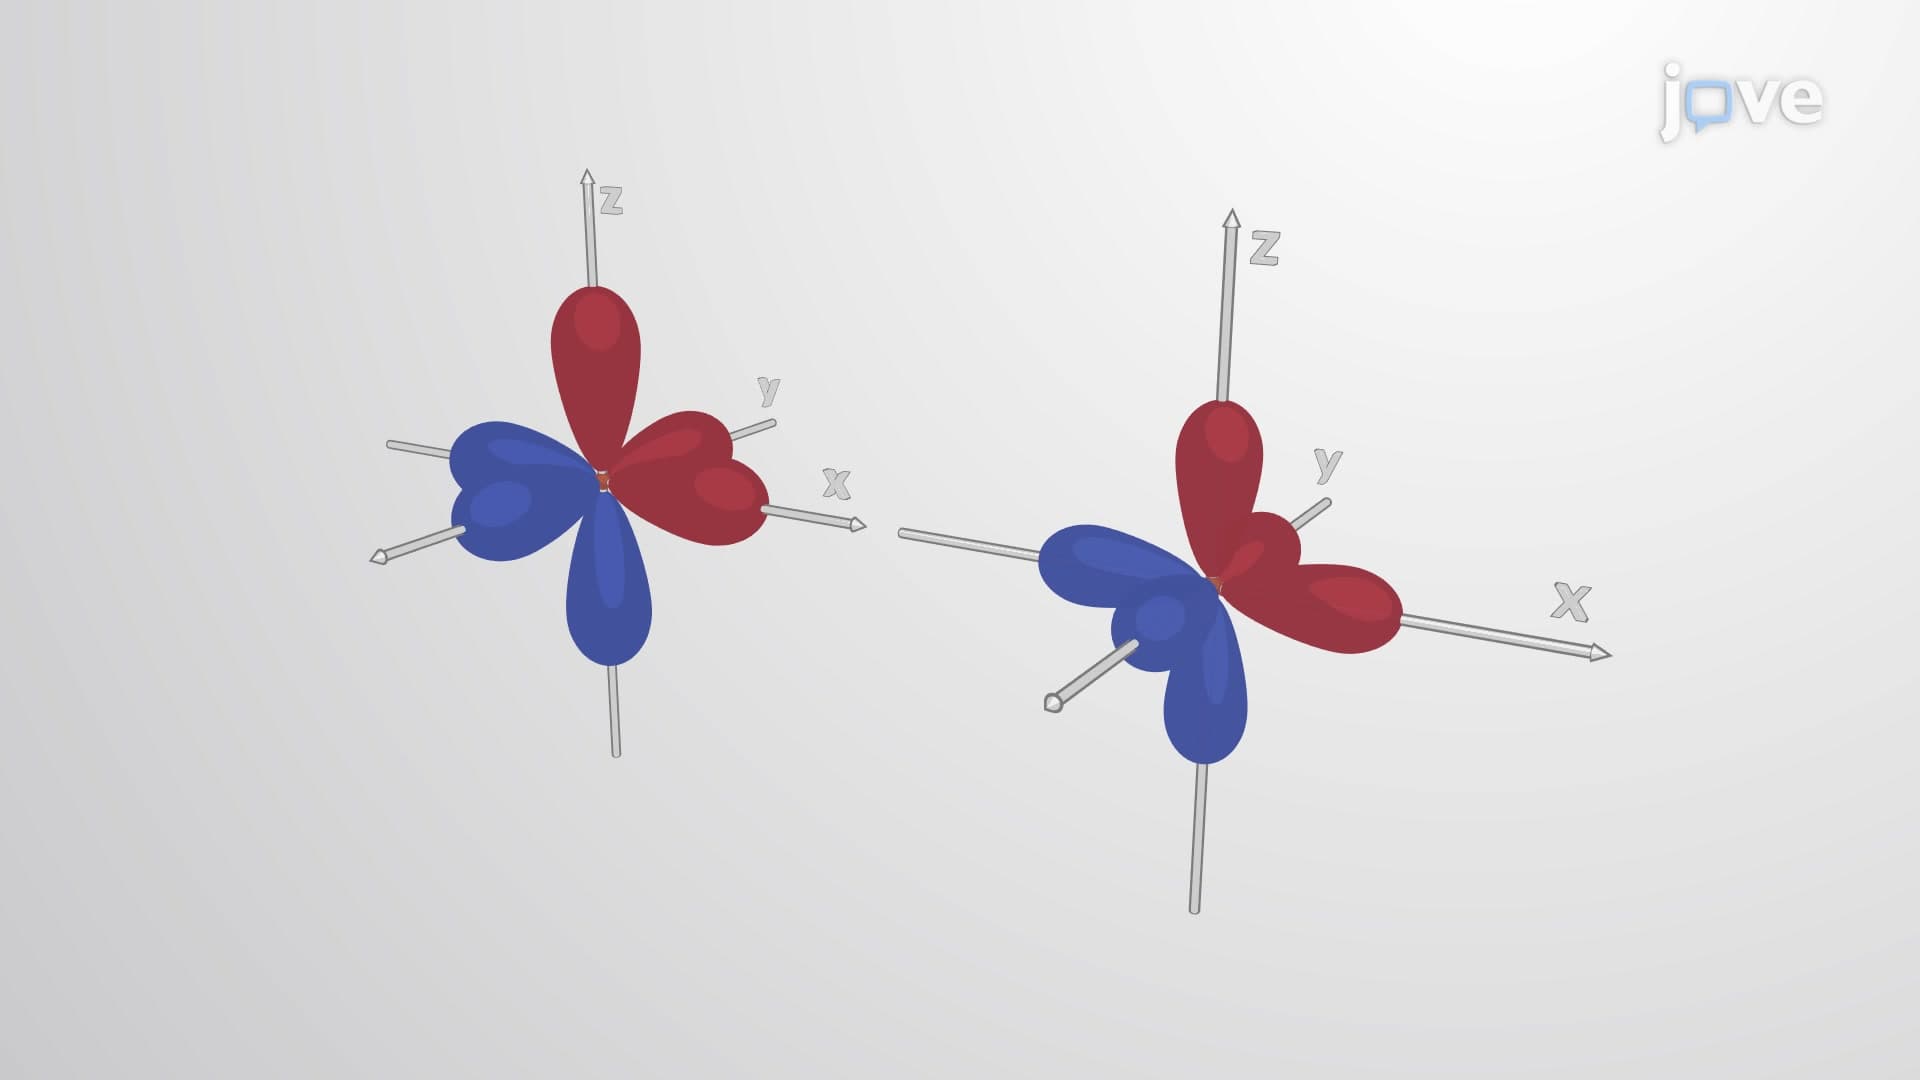 02:35
Molecular Orbital Theory I
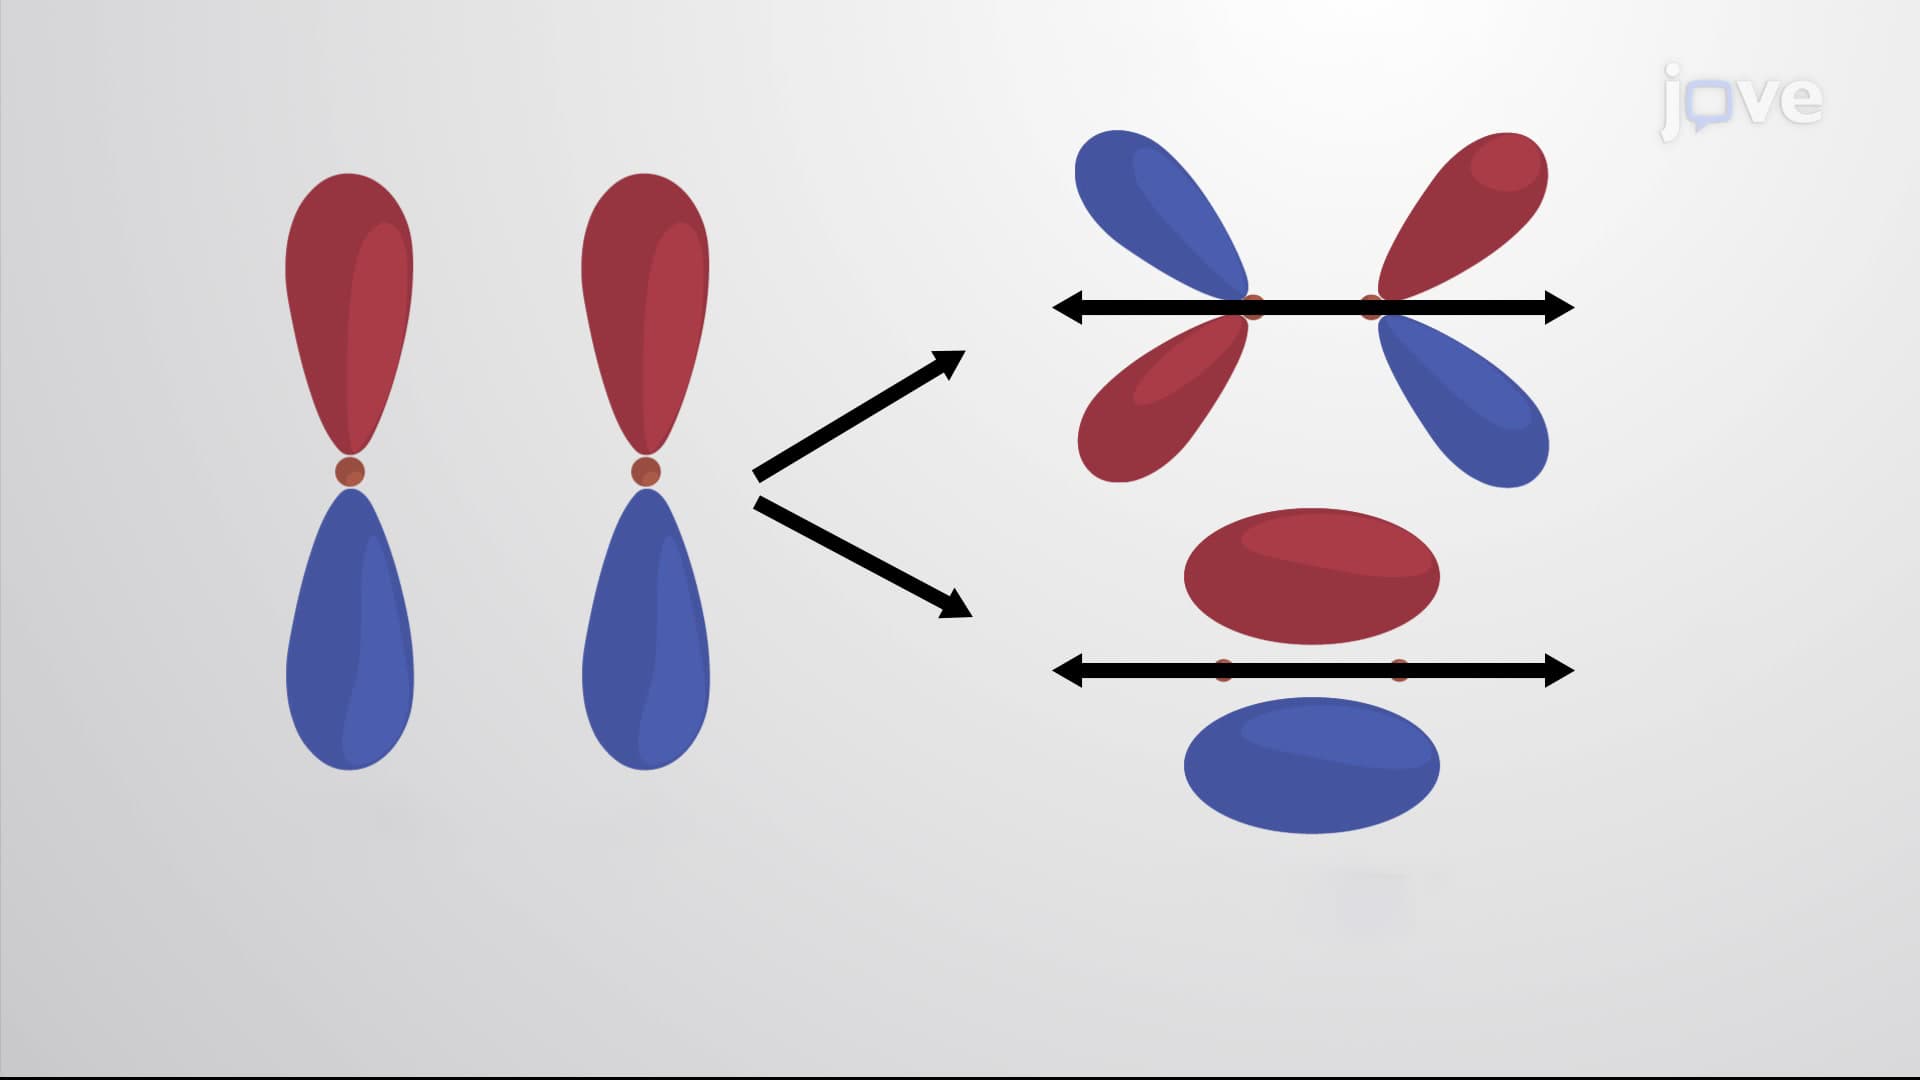 02:40
MO Theory and Covalent Bonding