
Expanding the Clinical Spectrum of LONP1-Related Mitochondrial Cytopathy
Fady Hannah-Shmouni1,2, Lauren MacNeil3,4, Lauren Brady5
1Clinical Biochemical Genetics, The Hospital for Sick Children, University of Toronto, Toronto, ON, Canada.

Abstract:
Pathogenic variants in the LONP1 gene have been associated with CODAS syndrome (Cerebral, Ocular, Dental, Auricular, and Skeletal Anomalies Syndrome). A recent report identified the first newborn case with LONP1-related mitochondrial cytopathy due to a compound heterozygous pathogenic variant in LONP1 without features of CODAS. The proband had manifested with severe congenital lactic acidosis and profound multiple respiratory chain complex activity deficiencies associated with the quantitative loss of mtDNA copy number in muscle. A subsequent report identified two siblings with regression during infancy, profound hypotonia and muscle weakness, severe intellectual disability, progressive cerebellar atrophy, where muscle biopsy showed an electron dense mitochondrial inclusions without ragged-red fibers and normal electron transport chain enzyme activities. Here, we report an additional case of autosomal recessive mitochondrial cytopathy due to a homozygous missense variant in LONP1 that was identified on whole exome sequencing (c.810G > A; p.D463N). The proband, a 20-year-old male born to consanguineous parents, presented with global developmental delay, emotional outbursts, speech and swallowing difficulties, hypotonia, and ataxia since childhood. Muscle biopsy showed massive granular bodies, increased oxidative stress, and autophagic block and reduced mitochondrial state 3 respiration. We have identified another case of LONP1-related mitochondrial cytopathy further confirming a neurological phenotype without CODAS features.
Related Concept Videos
 01:22
Lysosomal Hydrolases
 02:59
Animal Mitochondrial Genetics
 01:39
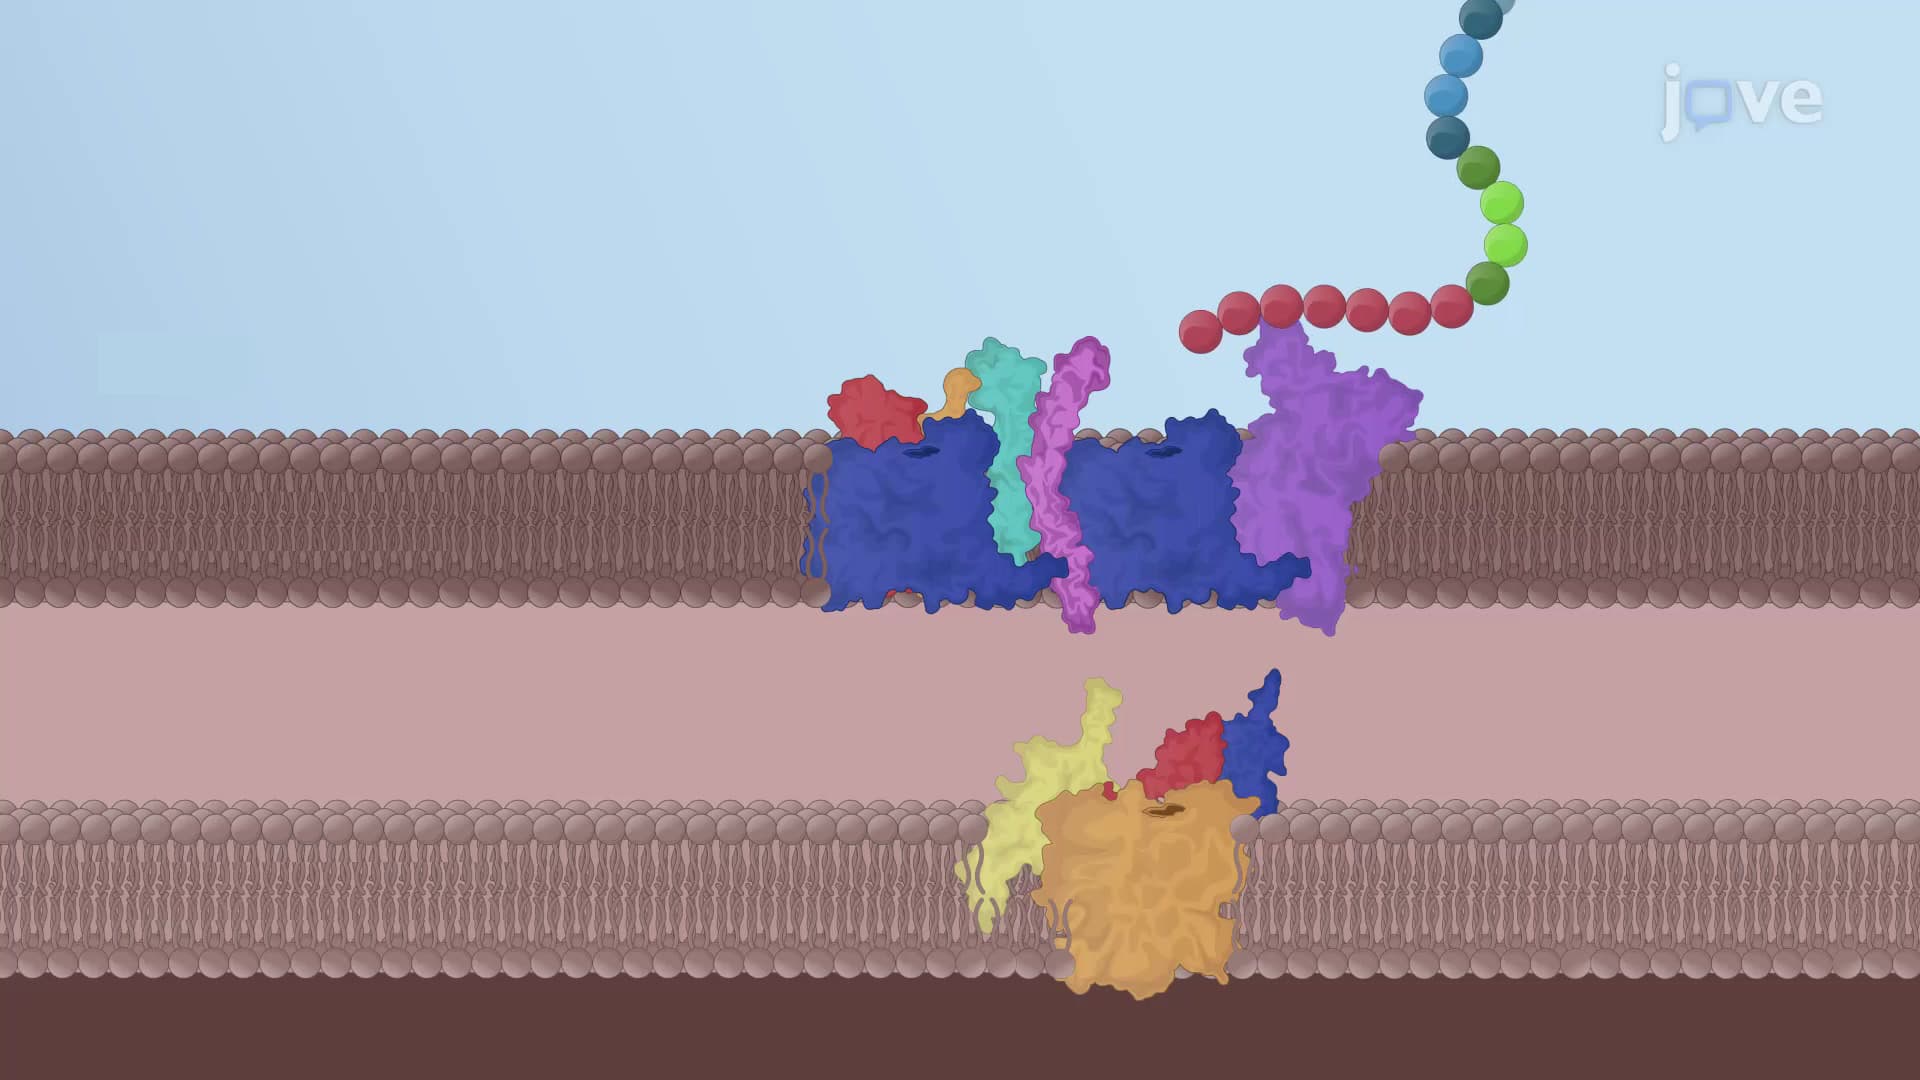
Mitochondrial Precursor Proteins
Most of the mitochondrial...
 01:18
ATP Synthase: Structure
 01:45
Mitochondrial Membranes
 01:12
Porin Insertion in the Outer Mitochondrial Membrane
Three models describe the assembly of porins by the SAM complex and their insertion into the outer membrane. Model 1 suggests that porins are assembled outside the SAM channel as the...